Related Experiment Video
Updated: Jul 3, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Rotationally invariant ab initio evaluation of Coulomb and exchange parameters for DFT+U calculations
Nicholas J Mosey1, Peilin Liao, Emily A Carter
1Department of Mechanical and Aerospace Engineering and Program in Applied and Computational Mathematics, Princeton University, Princeton, New Jersey 08544-5263, USA.

Abstract:
Conventional density functional theory (DFT) fails for strongly correlated electron systems due to large intra-atomic self-interaction errors. The DFT+U method provides a means of overcoming these errors through the use of a parametrized potential that employs an exact treatment of quantum mechanical exchange interactions. The parameters that enter into this potential correspond to the spherically averaged intra-atomic Coulomb (U) and exchange (J) interactions. Recently, we developed an ab initio approach for evaluating these parameters on the basis of unrestricted Hartree-Fock (UHF) theory, which has the advantage of being free of self-interaction errors and does not require experimental input [Mosey and Carter, Phys. Rev. B 76, 155123 (2007)]. In this work, we build on that method to develop a more robust and convenient ab initio approach for evaluating U and J. The new technique employs a relationship between U and J and the Coulomb and exchange integrals evaluated using the entire set of UHF molecular orbitals (MOs) for the system. Employing the entire set of UHF MOs renders the method rotationally invariant and eliminates the difficulty in selecting unambiguously the MOs that correspond to localized states. These aspects overcome two significant deficiencies of our earlier method. The new technique is used to evaluate U and J for Cr(2)O(3), FeO, and Fe(2)O(3). The resulting values of U-J are close to empirical estimates of this quantity for each of these materials and are also similar to results of constrained DFT calculations. DFT+U calculations using the ab initio parameters yield results that are in good agreement with experiment. As such, this method offers a means of performing accurate and fully predictive DFT+U calculations of strongly correlated electron materials.
More Related Videos


Related Concept Videos
 02:36
Molecular Geometry and Dipole Moments
 01:30
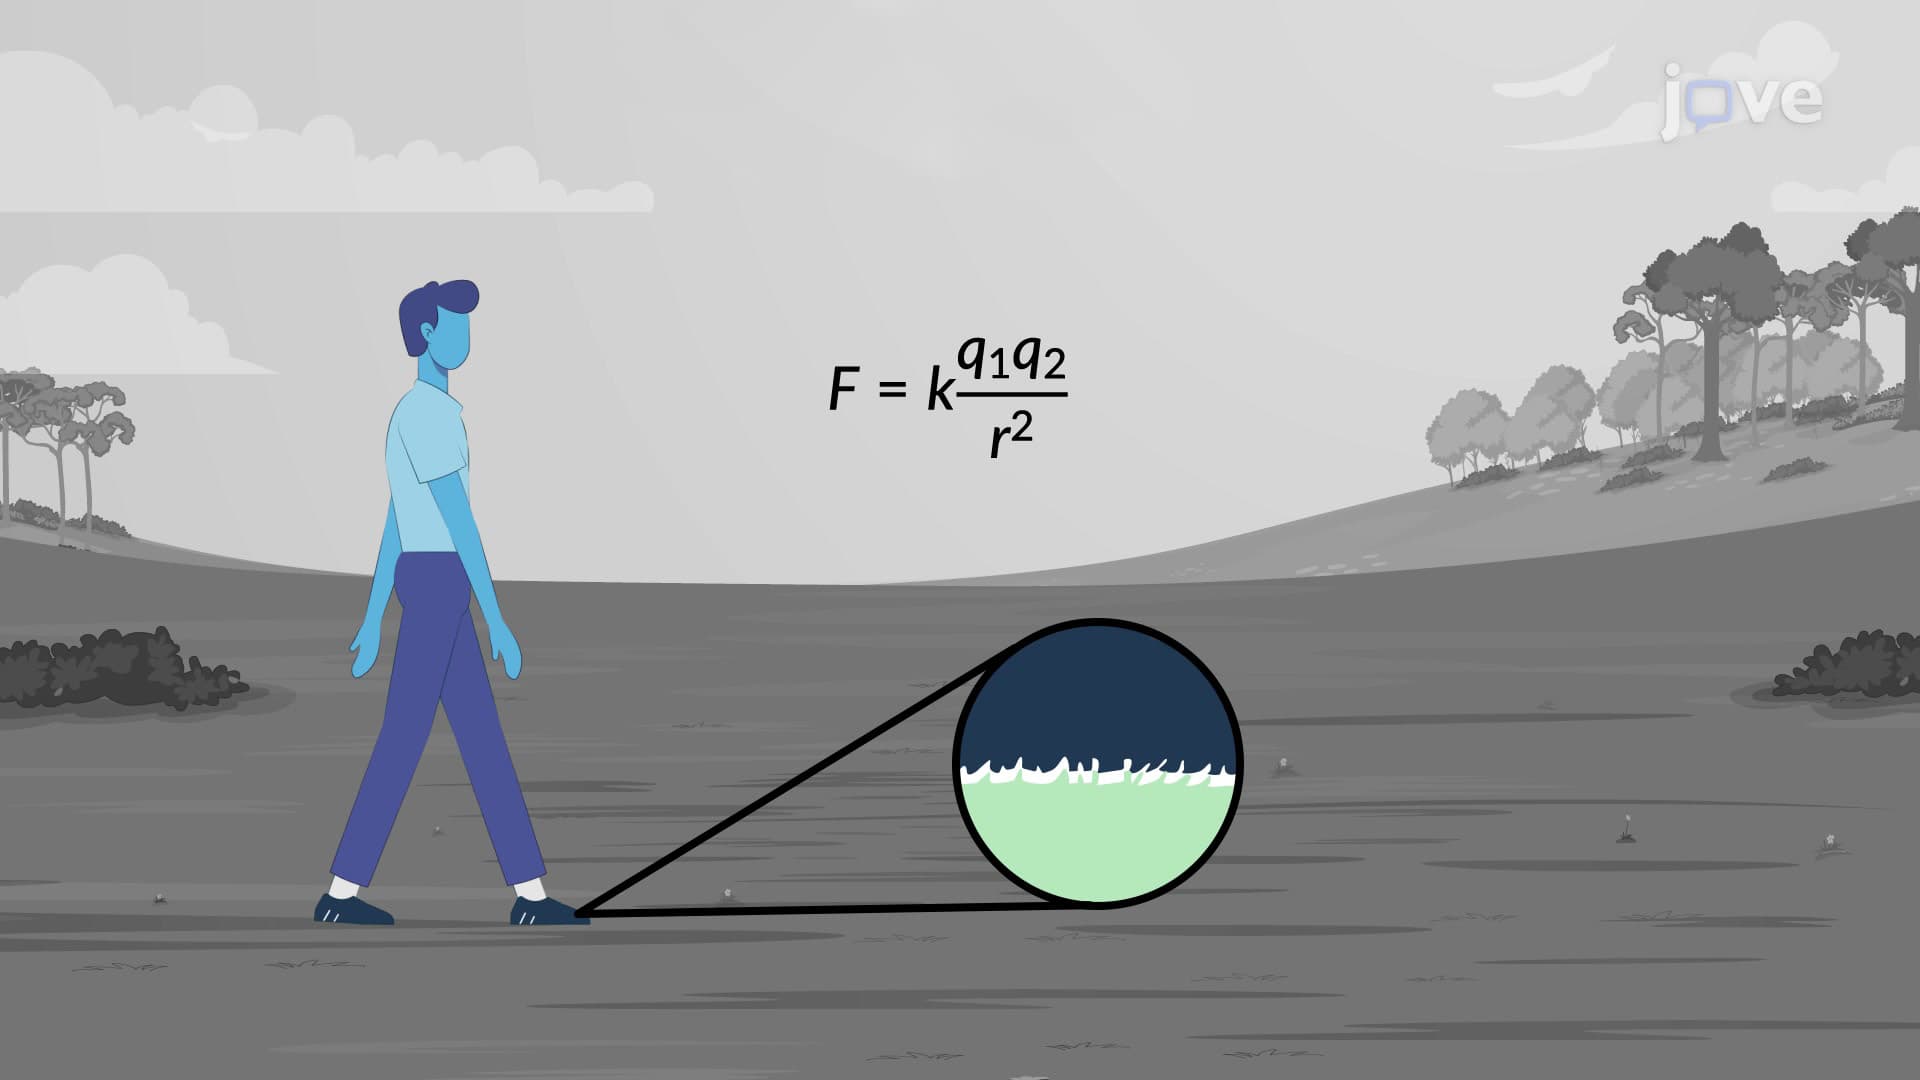
Coulomb's Law
Newton's third law applies to the Coulomb force — the force on...
 02:28
Electronic Structure of Atoms
An atom comprises protons and neutrons, which are contained inside the dense, central core called the nucleus, with electrons present around the nucleus. Taking into account the wave–particle duality of electrons and the uncertainty in position around the nucleus, quantum mechanics provides a more accurate model for the atomic structure. It describes atomic orbitals as the regions around the nucleus where electrons of discrete energy exist, characterized by four quantum numbers: n, l, ml, and...
 01:30
Electric Dipoles and Dipole Moment
Theoretically, studying electric dipoles leads to understanding why the resultant electric forces around us are weak. Since electric forces are strong, remnant net charges are rare. Hence, the interaction between dipoles helps us understand electrical interactions in...
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 01:10
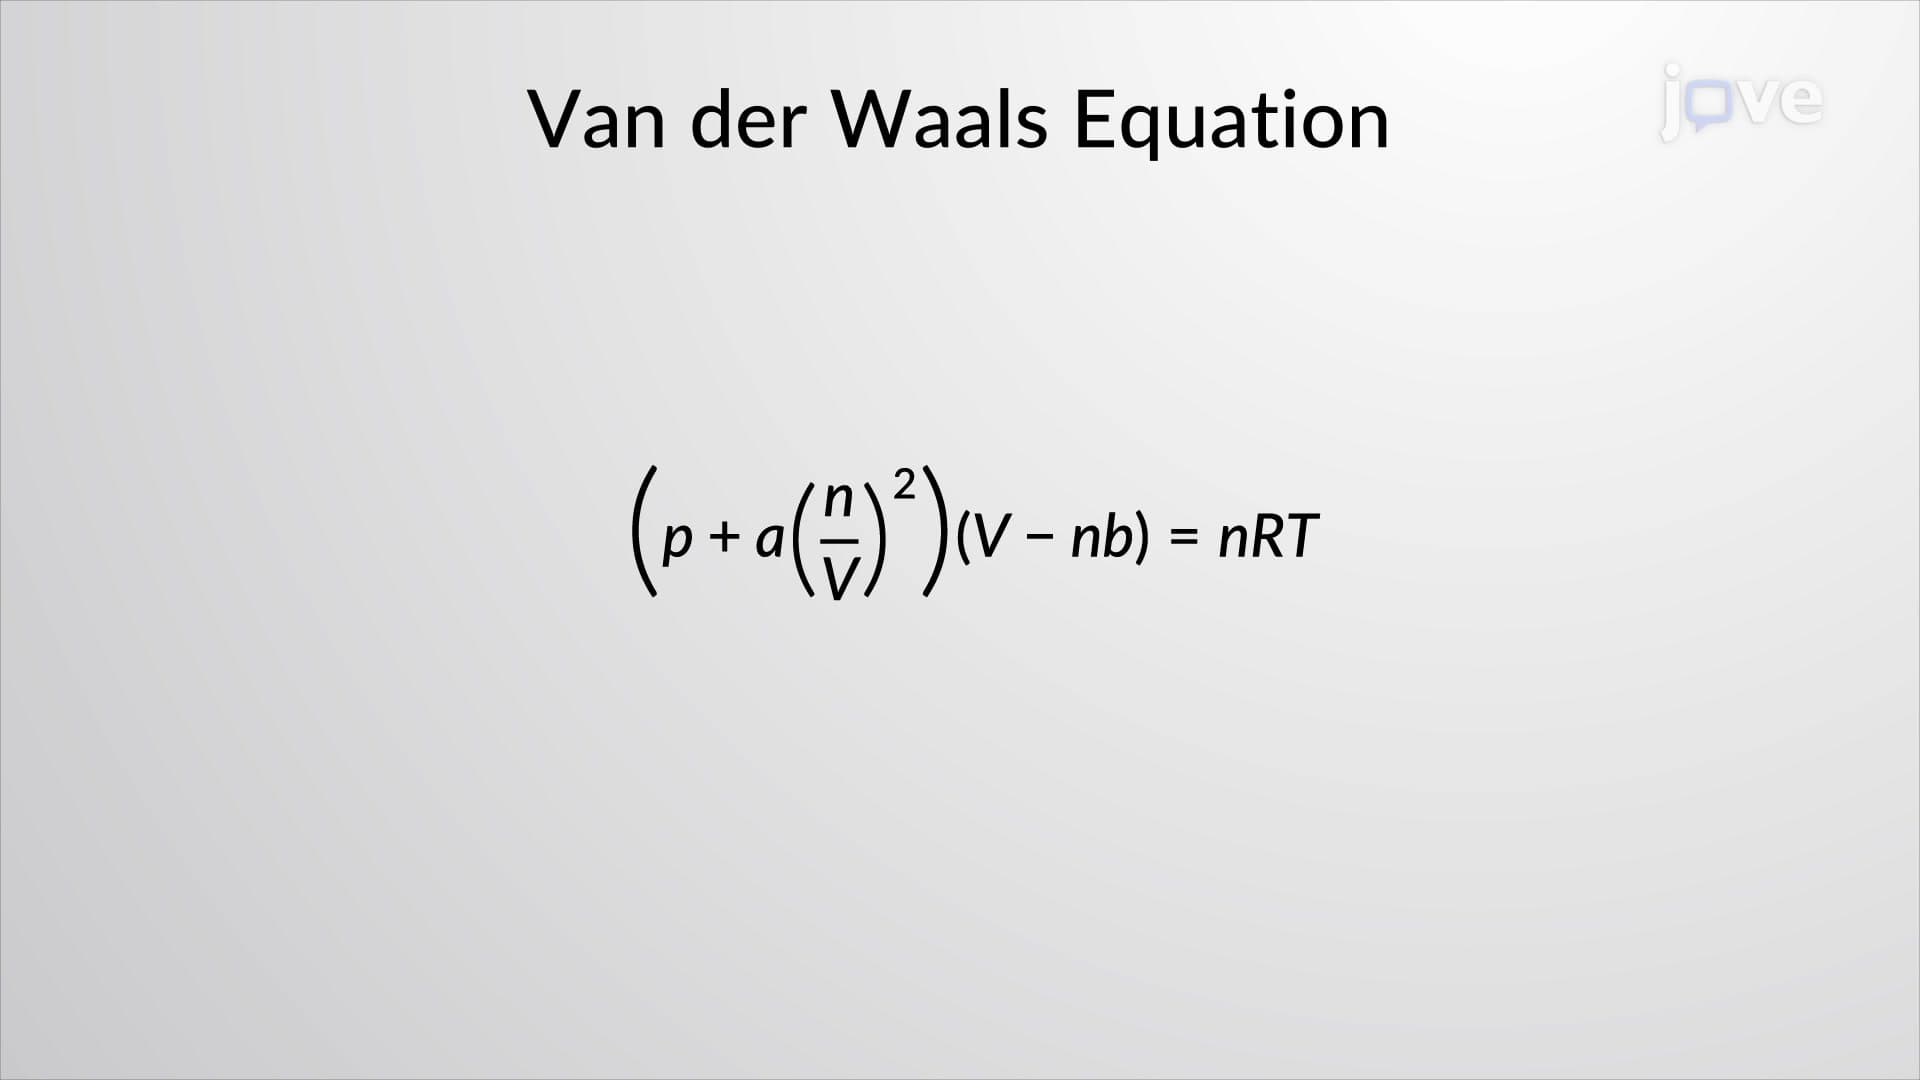
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...