Efficient quantum calculations of vibrational states of vinylidene in full dimensionality: a scheme with combination of methods
1Beijing National Laboratory for Molecular Sciences, State Key Laboratory of Molecular Reaction Dynamics, Institute of Chemistry, Chinese Academy of Sciences, Beijing 100190, People's Republic of China.

Abstract:
Full-dimensional quantum calculations of vibrational states of C(2)H(2) and C(2)D(2) are performed in the high-energy region (above 20,400 cm(-1) relative to the acetylene minimum). The theoretical scheme is a combination of several methods. To exploit the full parity and permutation symmetry, the CC-HH diatom-diatom Jacobi coordinates are chosen; phase space optimization in combination with physical considerations is used to obtain an efficient radial discrete variable representation, whereas a basis contraction scheme is applied for angular coordinates. The preconditioned inexact spectral transform method combined with an efficient preconditioner is employed to compute eigenstates within a desired spectral window. The computation is efficient. More definite assignments on vinylidene states than previous studies are acquired using the normal mode projection; in particular, a consistent analysis of the nu(1) (symmetric CH stretch) state is provided. The computed vinylidene vibrational energy levels are in general good agreement with experiment, and several vinylidene states are reported for the first time.
More Related Videos
 13:58
Probing C84-embedded Si Substrate Using Scanning Probe Microscopy and Molecular Dynamics
Published on: September 28, 2016
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 01:29
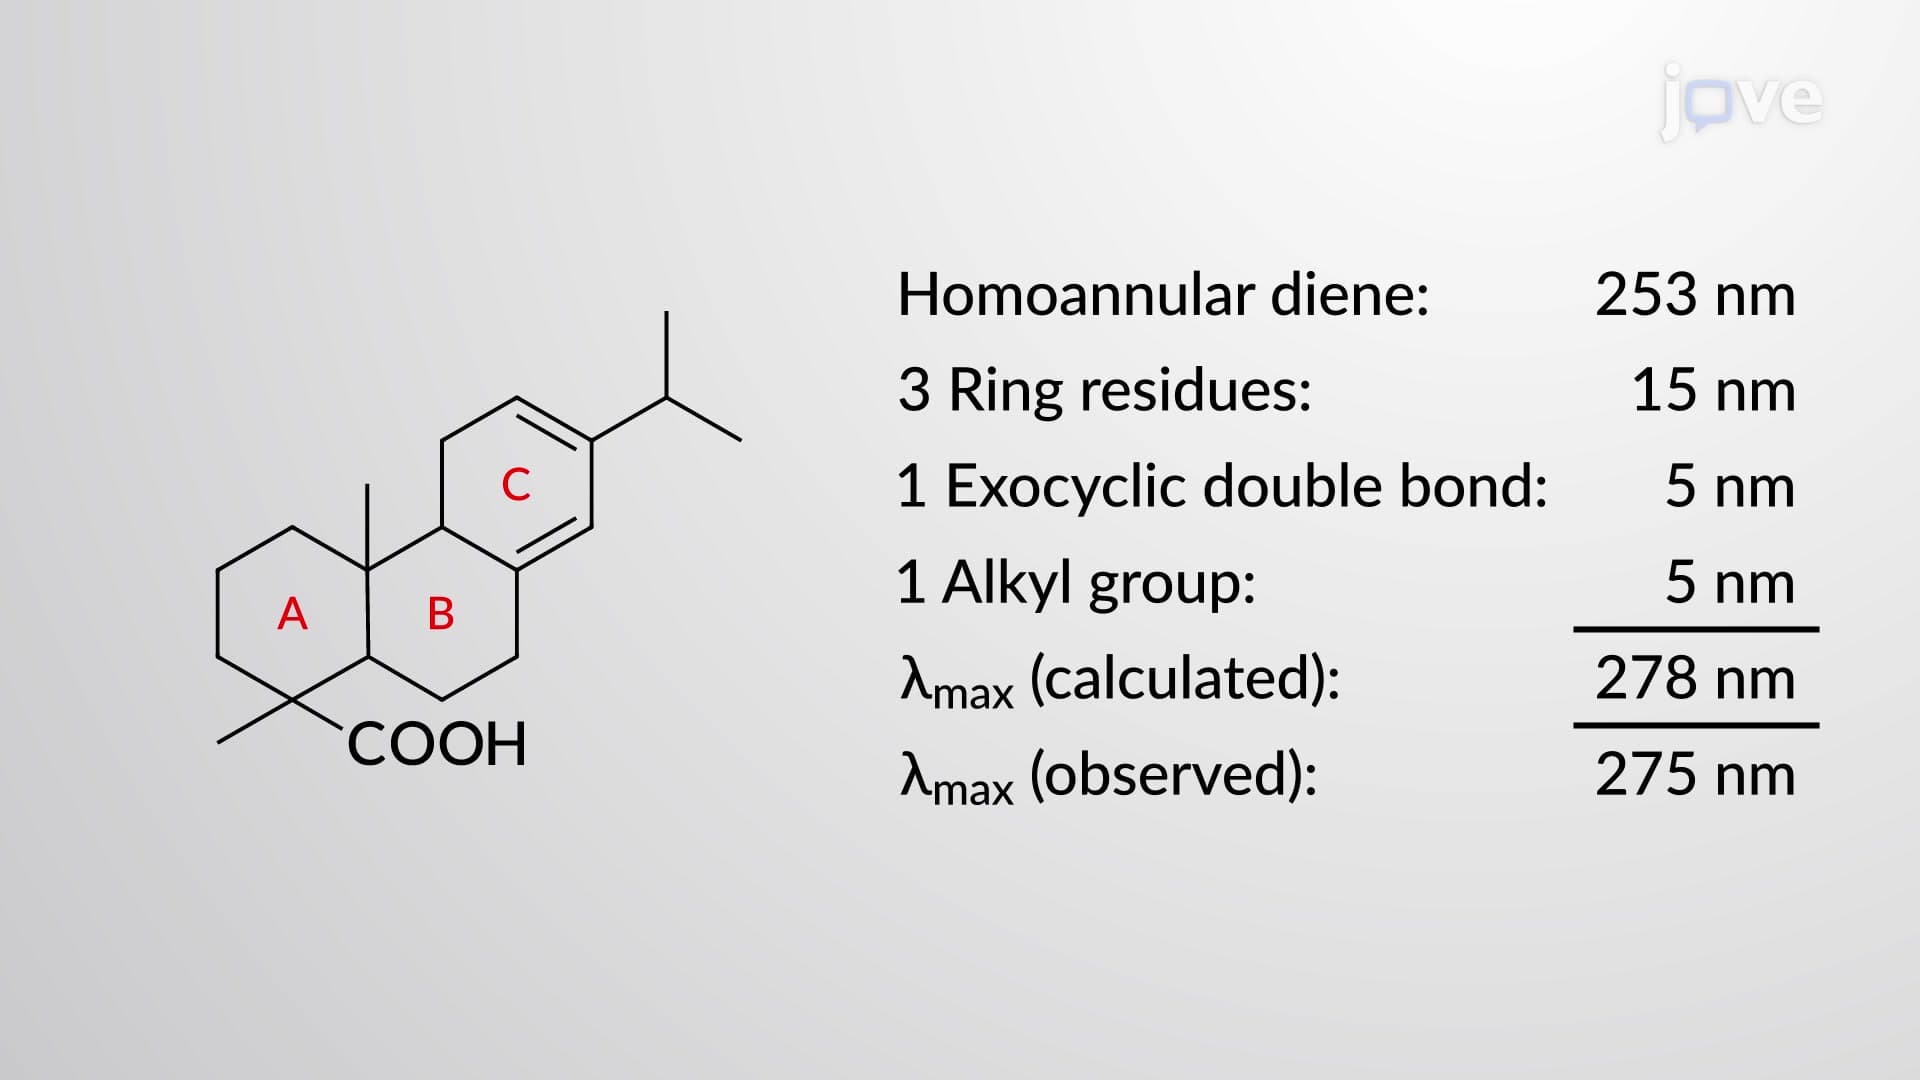
UV–Vis Spectroscopy: Woodward–Fieser Rules
 02:38
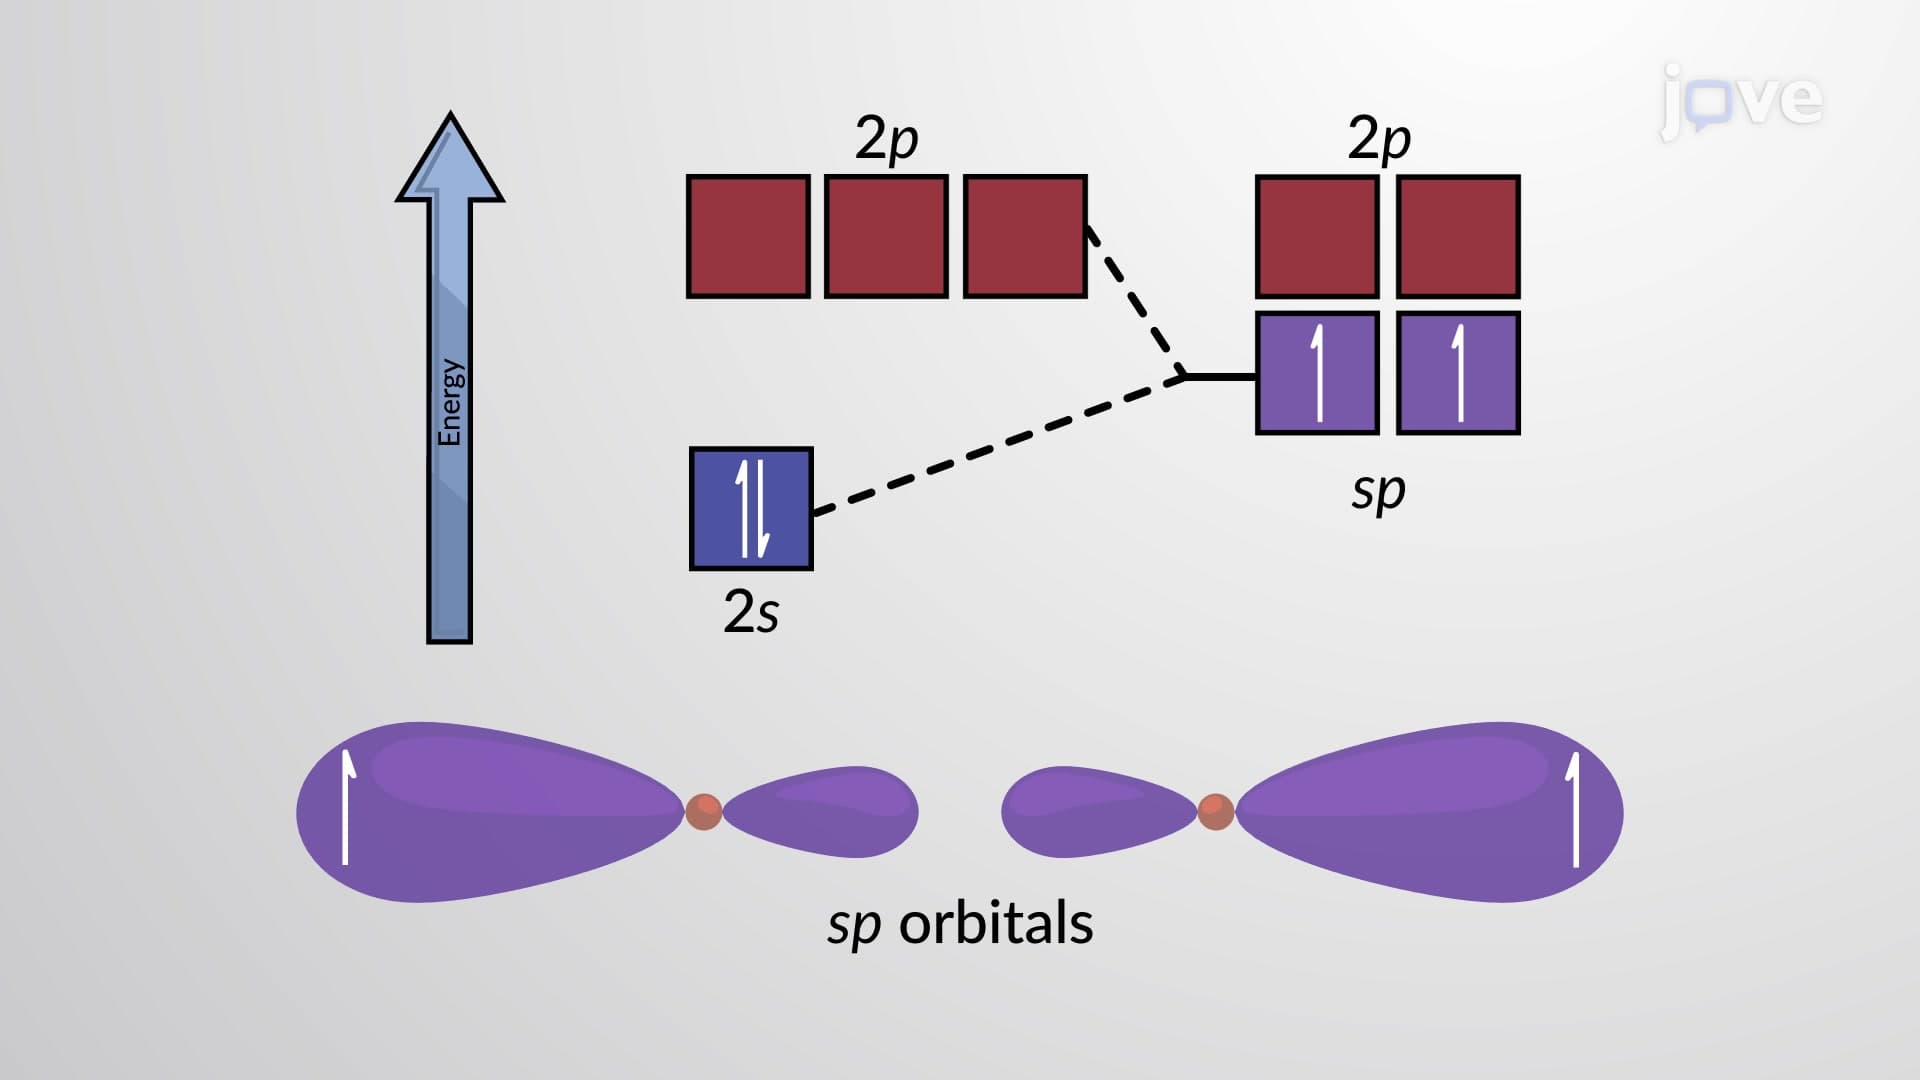
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to the...
 01:16
UV–Vis Spectroscopy: Molecular Electronic Transitions
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...