An efficient algorithm for energy gradients and orbital optimization in valence bond theory
Related Experiment Video
Updated: Jul 3, 2026

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Lingchun Song1, Jinshuai Song, Yirong Mo
1The State Key Laboratory of Physical Chemistry of Solid Surfaces and Department of Chemistry, College of Chemistry and Chemical Engineering, Xiamen University, Xiamen, Fujian 361005, China.

Abstract:
An efficient algorithm for energy gradients in valence bond theory with nonorthogonal orbitals is presented. A general Hartree-Fock-like expression for the Hamiltonian matrix element between valence bond (VB) determinants is derived by introducing a transition density matrix. Analytical expressions for the energy gradients with respect to the orbital coefficients are obtained explicitly, whose scaling for computational cost is m(4), where m is the number of basis functions, and is thus approximately the same as in HF method. Compared with other existing approaches, the present algorithm has lower scaling, and thus is much more efficient. Furthermore, the expression for the energy gradient with respect to the nuclear coordinates is also presented, and it provides an effective algorithm for the geometry optimization and the evaluation of various molecular properties in VB theory. Test applications show that our new algorithm runs faster than other methods.
More Related Videos
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Related Concept Videos
 02:38
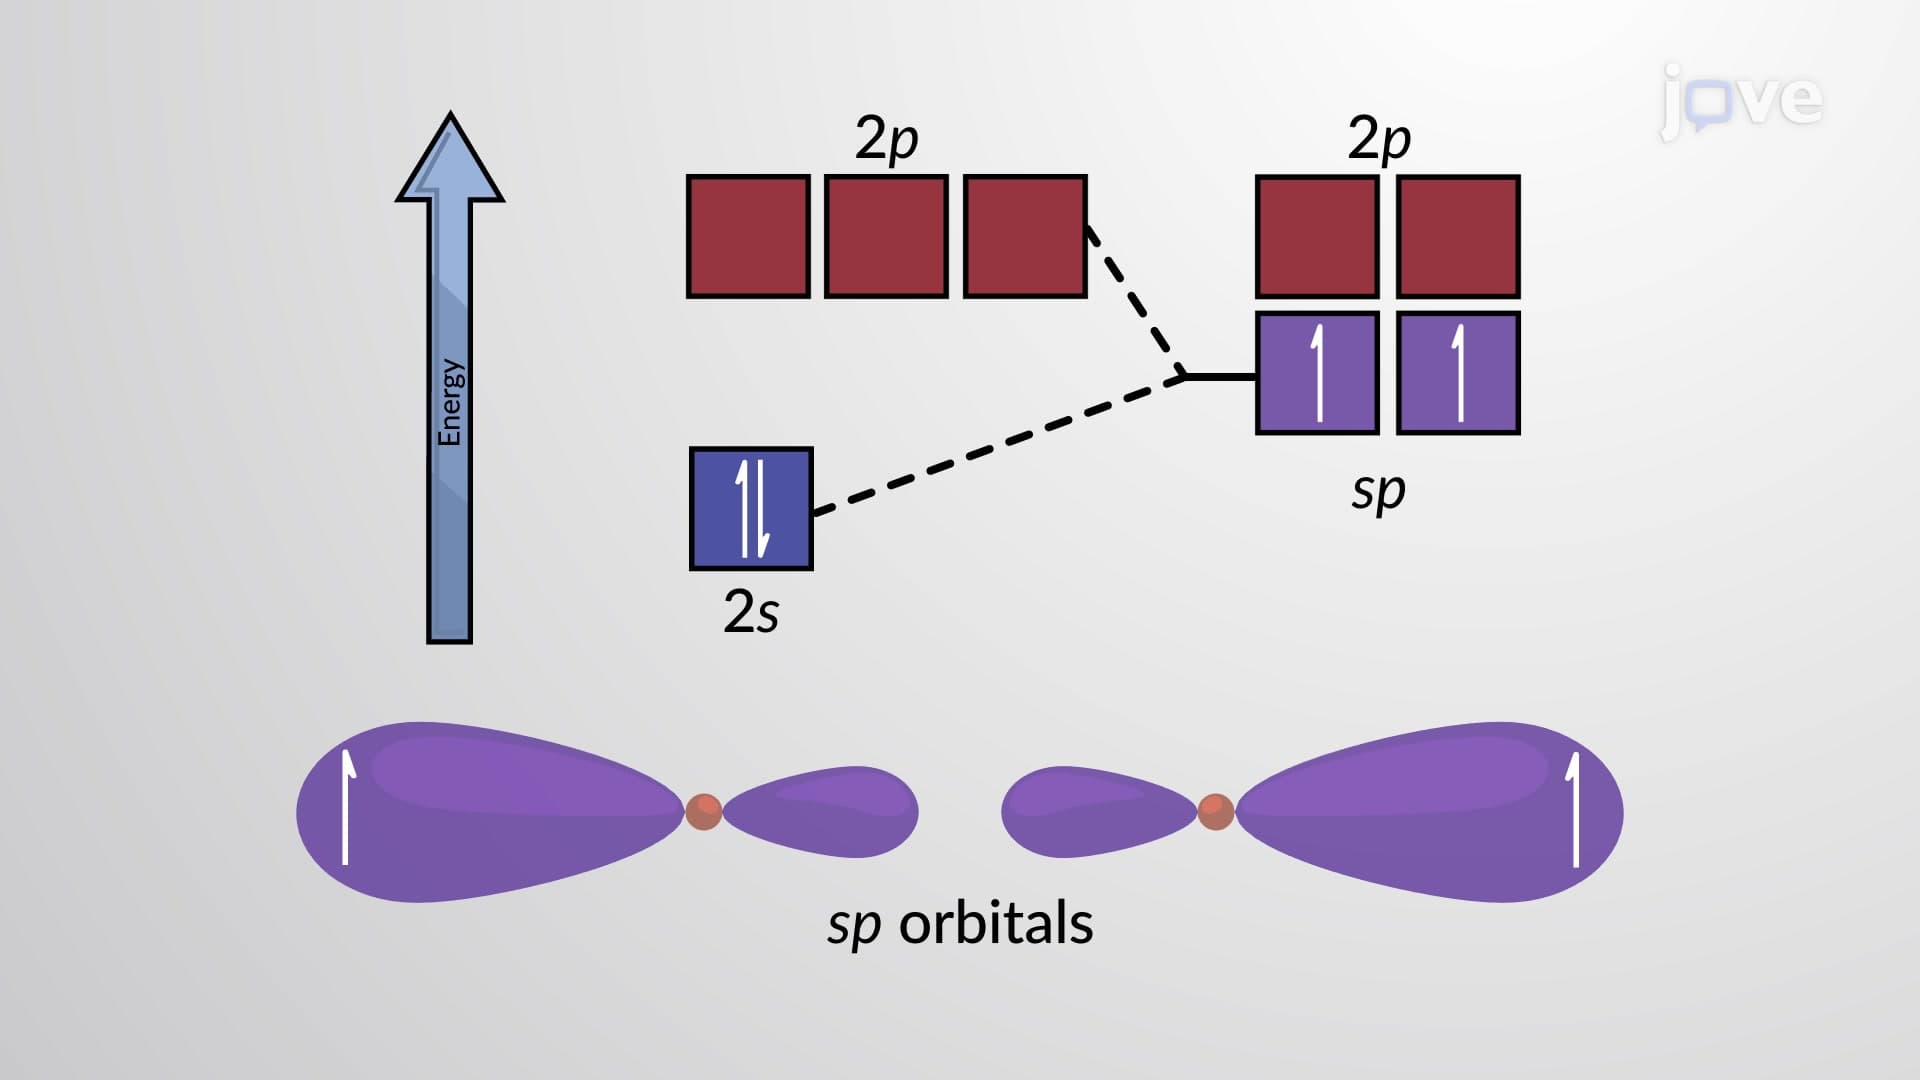
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 03:51
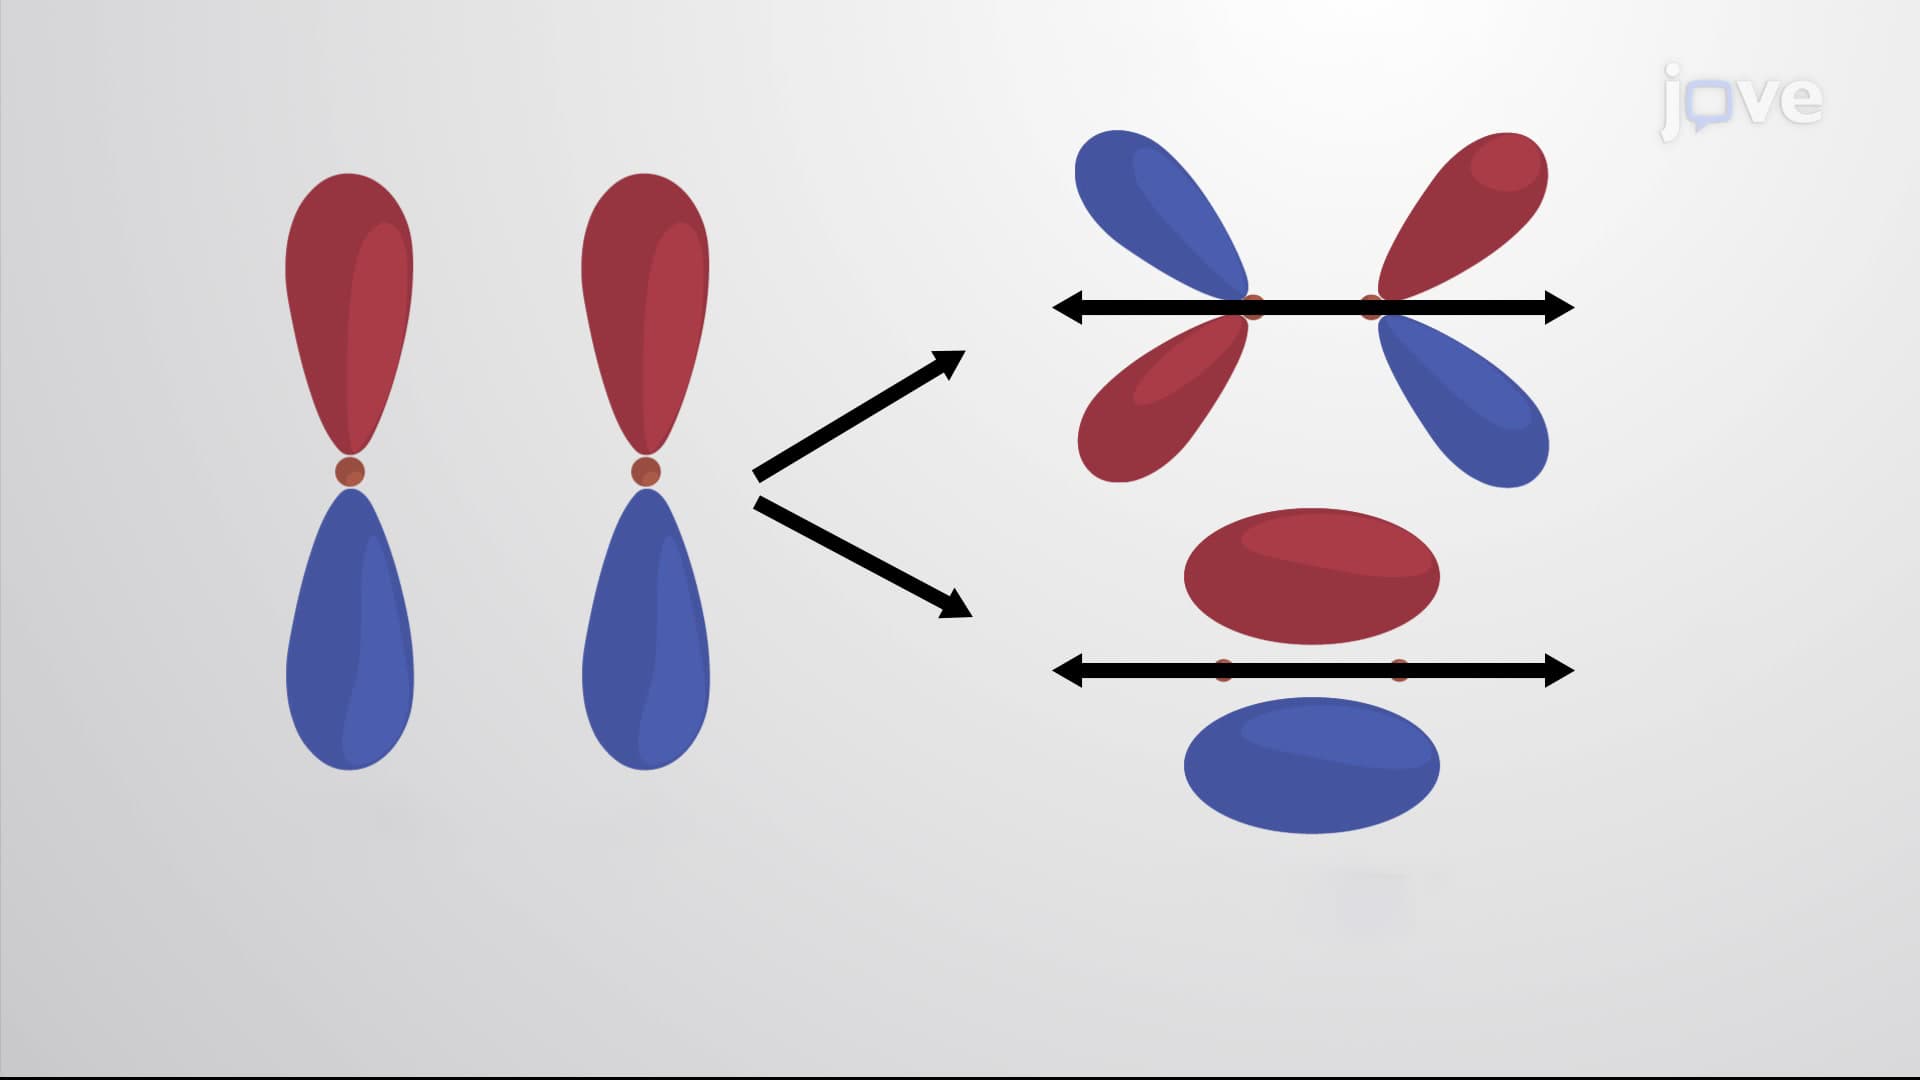
Molecular Orbital Theory II
 03:21
The Energies of Atomic Orbitals
 02:35
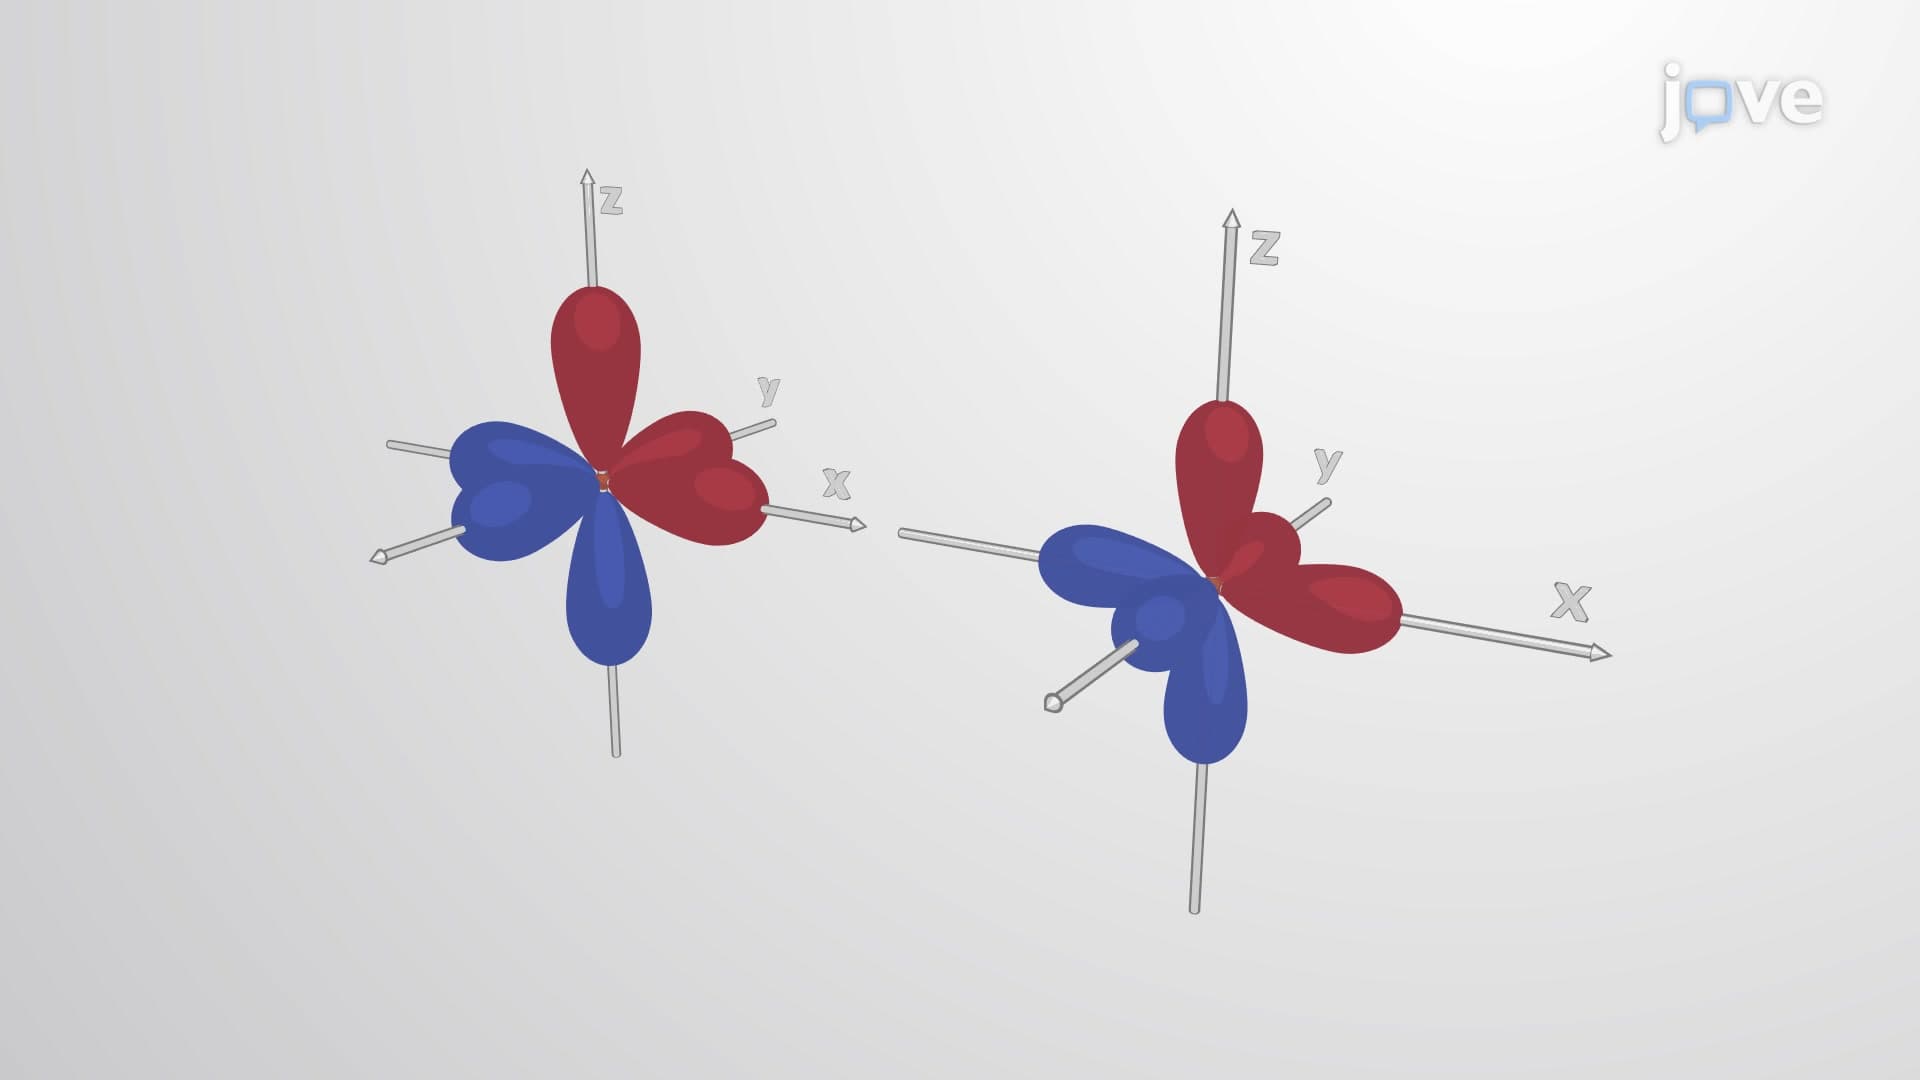
Molecular Orbital Theory I
 03:35
Hybridization of Atomic Orbitals II
 02:40
MO Theory and Covalent Bonding