Related Experiment Video
Updated: Jul 3, 2026

A Technique for Serial Collection of Cerebrospinal Fluid from the Cisterna Magna in Mouse
Published on: November 10, 2008
Parkin deletion causes cerebral and systemic amyloidosis in human mutated tau over-expressing mice
Jose A Rodríguez-Navarro1, Ana Gómez, Izaskun Rodal
1Department of Neurobiology, Hospital Ramón y Cajal, Madrid, Spain.

Abstract:
Deposition of proteins leading to amyloid takes place in some neurodegenerative diseases such as Alzheimer's disease and Huntington's disease. Mutations of tau and parkin proteins produce neurofibrillary abnormalities without deposition of amyloid. Here we report that mature, parkin null, over-expressing human mutated tau (PK(-/-)/Tau(VLW)) mice have altered behaviour and dopamine neurotransmission, tau pathology in brain and amyloid deposition in brain and peripheral organs. PK(-/-)/Tau(VLW) mice have abnormal behaviour and severe drop out of dopamine neurons in the ventral midbrain, up to 70%, at 12 months and abundant phosphorylated tau positive neuritic plaques, neuro-fibrillary tangles, astrogliosis, microgliosis and plaques of murine beta-amyloid in the hippocampus. PK(-/-)/Tau(VLW) mice have organomegaly of the liver, spleen and kidneys. The electron microscopy of the liver confirmed the presence of a fibrillary protein deposits with amyloid characteristics. There is also accumulation of mouse tau in hepatocytes. These mice have lower levels of CHIP-HSP70, involved in the proteosomal degradation of tau, increased oxidative stress, measured as depletion of glutathione which, added to lack of parkin, could trigger tau accumulation and amyloidogenesis. This model is the first that demonstrates beta-amyloid deposits caused by over-expression of tau and without modification of the amyloid precursor protein, presenilins or secretases. PK(-/-)/Tau(VLW) mice provide a link between the two proteins more important for the pathogenesis of Alzheimer disease.
Related Concept Videos
 01:23
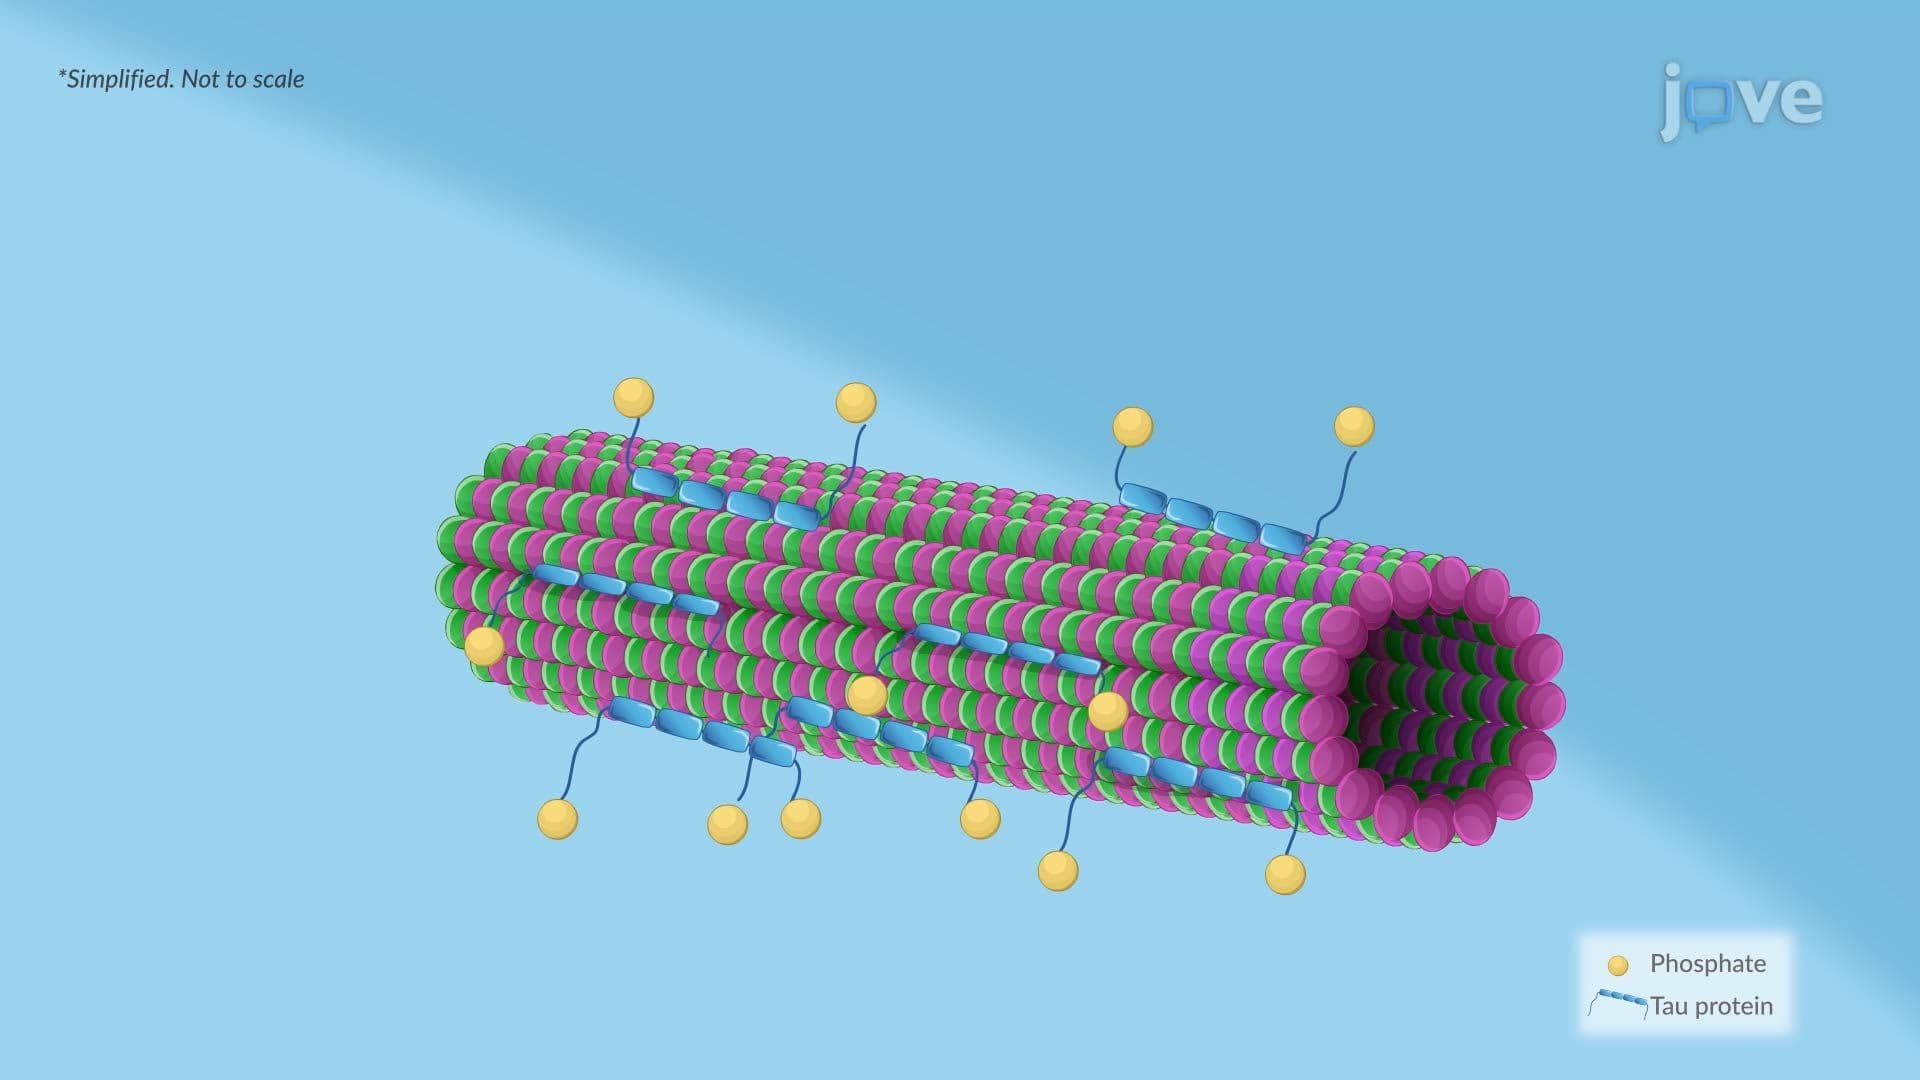
Alzheimer Disease ll: Pathophysiology
 01:24
Parkinson Disease ll: Pathophysiology