Related Experiment Video
Updated: Jul 3, 2026

Photoelectron Imaging of Anions Illustrated by 310 Nm Detachment of F−
Published on: July 27, 2018
Accurate ab initio structure, dissociation energy, and vibrational spectroscopy of the F(-)-CH4 anion complex
Gábor Czakó1, Bastiaan J Braams, Joel M Bowman
1Cherry L. Emerson Center for Scientific Computation and Department of Chemistry, Emory University, Atlanta, Georgia 30322, USA. czako@chem.elte.hu

Abstract:
Accurate equilibrium structure, dissociation energy, global potential energy surface (PES), dipole moment surface (DMS), and the infrared vibrational spectrum in the 0-3000 cm(-1) range of the F(-)-CH4 anion complex have been obtained. The equilibrium electronic structure calculations employed second-order Møller-Plesset perturbation theory (MP2) and coupled-cluster (CC) method up to single, double, triple, and perturbative quadruple excitations using the aug-cc-p(C)VXZ [X = 2(D), 3(T), 4(Q), and 5] correlation-consistent basis sets. The best equilibrium geometry has been obtained at the all-electron CCSD(T)/aug-cc-pCVQZ level of theory. The dissociation energy has been determined based on basis set extrapolation techniques within the focal-point analysis (FPA) approach considering (a) electron correlation beyond the all-electron CCSD(T) level, (b) relativistic effects, (c) diagonal Born-Oppenheimer corrections (DBOC), and (d) variationally computed zero-point vibrational energies. The final D(e) and D0 values are 2398 +/- 12 and 2280 +/- 20 cm(-1), respectively. The global PES and DMS have been computed at the frozen-core CCSD(T)/aug-cc-pVTZ and MP2/aug-cc-pVTZ levels of theory, respectively. Variational vibrational calculations have been performed for CH4 and F(-)-CH4 employing the vibrational configuration interaction (VCI) method as implemented in Multimode.
More Related Videos
 08:22
Measurement of Ultrafast Vibrational Coherences in Polyatomic Radical Cations with Strong-Field Adiabatic Ionization
Published on: August 6, 2018
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 01:29
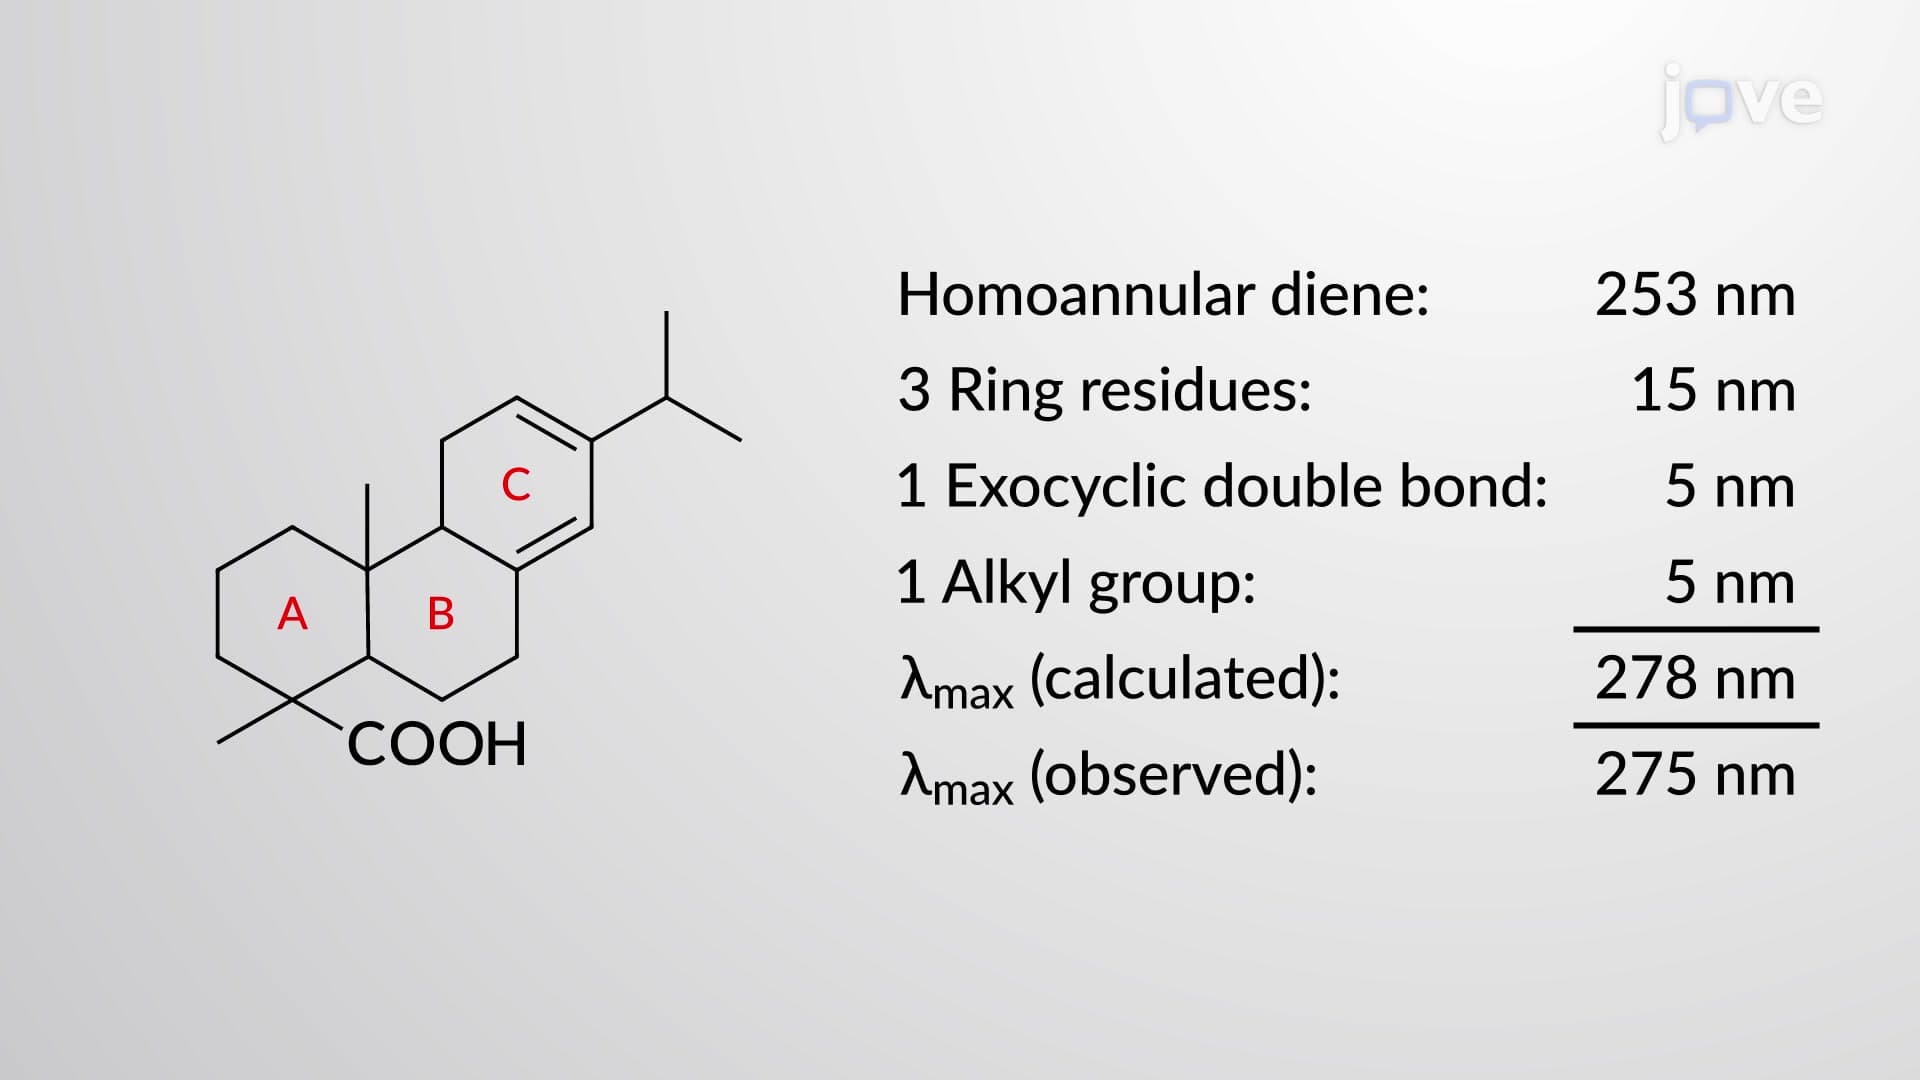
UV–Vis Spectroscopy: Woodward–Fieser Rules
 03:24
Hybridization of Atomic Orbitals I
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to the...
 01:24
IR Spectroscopy: Molecular Vibration Overview
Stretching vibrations are vibrational motions that occur along the bond line, changing the bond length or distance between two bonded atoms. They are further distinguished as symmetric or asymmetric. In symmetric stretching, the...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...