Related Experiment Video
Updated: Jul 2, 2026

Methods for the Discovery of Novel Compounds Modulating a Gamma-Aminobutyric Acid Receptor Type A Neurotransmission
Published on: August 16, 2018
Drug binding to the inactivated state is necessary but not sufficient for high-affinity binding to human
Mark J Perrin1, Philip W Kuchel, Terence J Campbell
1Victor Chang Cardiac Research Institute, 384 Victoria Street, Darlinghurst, NSW 2010, Australia.

Insights
Drug block of the human ether-à-go-go-related gene K(+) channel (hERG) causes acquired long QT syndrome. Researchers found that preferential binding to the inactivated state is crucial for high-affinity hERG channel block, impacting cardiac repolarization.
Area of Science:
- Cardiovascular Pharmacology
- Molecular Cardiology
- Ion Channel Physiology
Background:
- Drug block of the human ether-à-go-go-related gene K(+) channel (hERG) is a primary cause of acquired long QT syndrome.
- Acquired long QT syndrome can lead to dangerous ventricular arrhythmias and sudden cardiac death due to disrupted cardiac repolarization.
Purpose of the Study:
- To investigate the state-dependent block of hERG channels by drugs, specifically examining the open versus inactivated states.
- To determine the relationship between drug affinity and the channel's inactivation properties using specific hERG mutants.
Main Methods:
- Utilized hERG channel mutants (N588K, N588E, S620T) with altered inactivation properties to study drug binding.
- Assessed the affinity of high- and low-affinity drugs for wild-type and mutant hERG channels.
- Employed computer modeling to analyze drug block kinetics relative to channel inactivation kinetics.
Main Results:
- High-affinity drugs showed reduced affinity for inactivation-deficient hERG mutants (N588K) compared to wild-type and N588E mutants.
- Most low-affinity drugs did not exhibit state-dependent block, with dl-sotalol being an exception.
- All tested state-dependent blockers had lower affinity for the non-inactivating S620T mutant compared to the N588K mutant.
- Calculated relative affinities for the inactivated versus open states, ranging from 4- to 70-fold for tested drugs.
Conclusions:
- Preferential binding to the inactivated state of the hERG channel is a necessary, but not sufficient, condition for high-affinity drug block.
- Understanding state-dependent block mechanisms is critical for predicting and mitigating drug-induced cardiac risks.
Abstract:
Drug block of the human ether-à-go-go-related gene K(+) channel (hERG) is the most common cause of acquired long QT syndrome, a disorder of cardiac repolarization that may result in ventricular tachycardia and sudden cardiac death. We investigated the open versus inactivated state dependence of drug block by using hERG mutants N588K and N588E, which shift the voltage dependence of inactivation compared with wild-type but in which the mutated residue is remote from the drug-binding pocket in the channel pore. Four high-affinity drugs (cisapride, dofetilide, terfenadine, and astemizole) demonstrated lower affinity for the inactivation-deficient N588K mutant hERG channel compared with N588E and wild-type hERG. Three of four low-affinity drugs (erythromycin, perhexiline, and quinidine) demonstrated no preference for N588E over N588K channels, whereas dl-sotalol was an example of a low-affinity state-dependent blocker. All five state-dependent blockers showed an even lower affinity for S620T mutant hERG (no inactivation) compared with N588K mutant hERG (greatly reduced inactivation). Computer modeling indicates that the reduced affinity for S620T compared with N588K and wild-type channels can be explained by the relative kinetics of drug block and unblock compared with the kinetics of inactivation and recovery from inactivation. We were also able to calculate, for the first time, the relative affinities for the inactivated versus the open state, which for the drugs tested here ranged from 4- to 70-fold. Our results show that preferential binding to the inactivated state is necessary but not sufficient for high-affinity binding to hERG channels.
More Related Videos
 10:41
Capturing the Interaction Kinetics of an Ion Channel Protein with Small Molecules by the Bio-layer Interferometry Assay
Published on: March 7, 2018
 07:51
Inhibitory Synapse Formation in a Co-culture Model Incorporating GABAergic Medium Spiny Neurons and HEK293 Cells Stably Expressing GABAA Receptors
Published on: November 14, 2014
Related Concept Videos
 01:30
Ligand-Gated Ion Channel Receptor: Gating Mechanism
 01:29
Drug-Receptor Interactions
Several parameters, such as the drug's affinity for its receptor and its efficacy, which is its ability to activate the receptor, determine the drug's effect on the tissue.
 01:29
The Two-State Receptor Model
The binding affinity of a drug determines its interaction with one...
 01:19
Ligand-gated Ion Channels
Three Subfamilies of Ligand-gated Ion Channels
Ligand-gated ion channels fall into three subfamilies. The 'Cys-loop' includes the nicotinic acetylcholine receptors, γ-aminobutyric acid (GABA), glycine, and 5-hydroxytryptamine receptors. The second one is the 'Pore-loop' channels that include the...
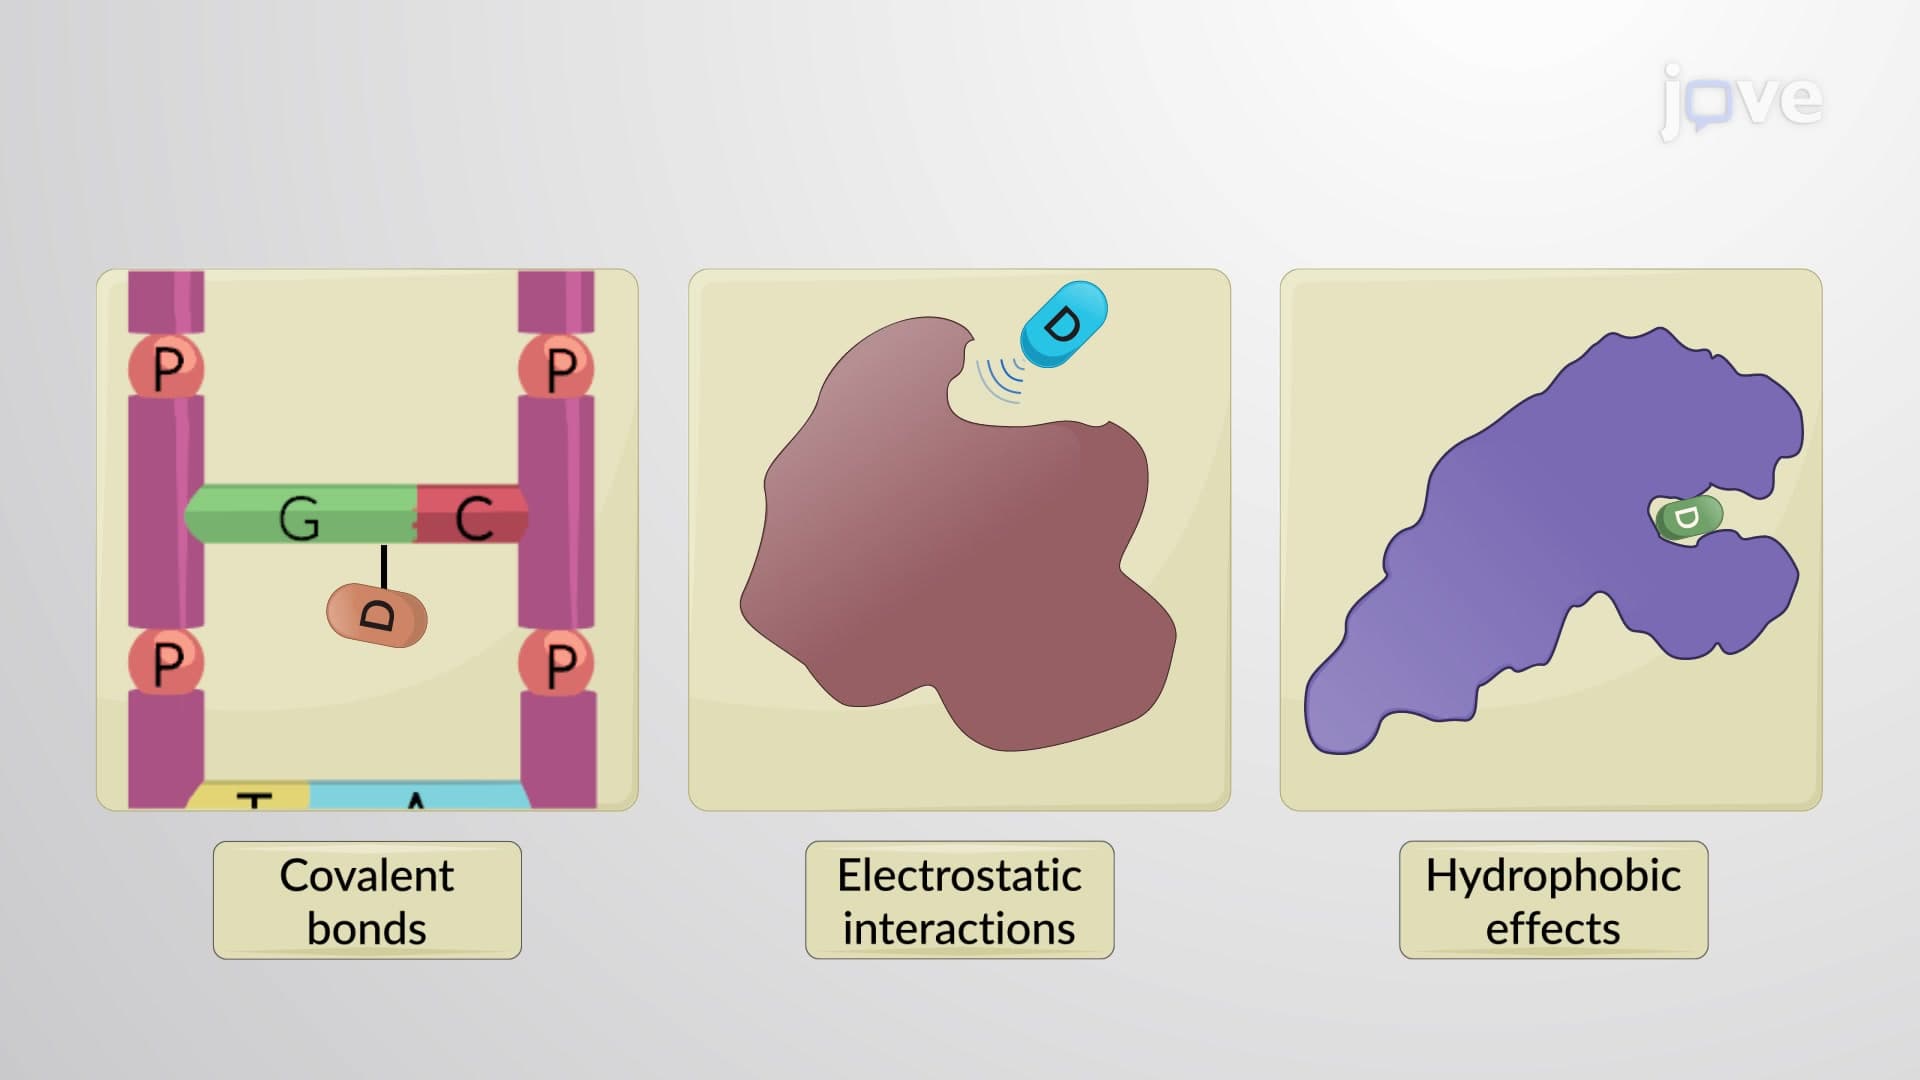 01:25
Drug-Receptor Bonds
In...
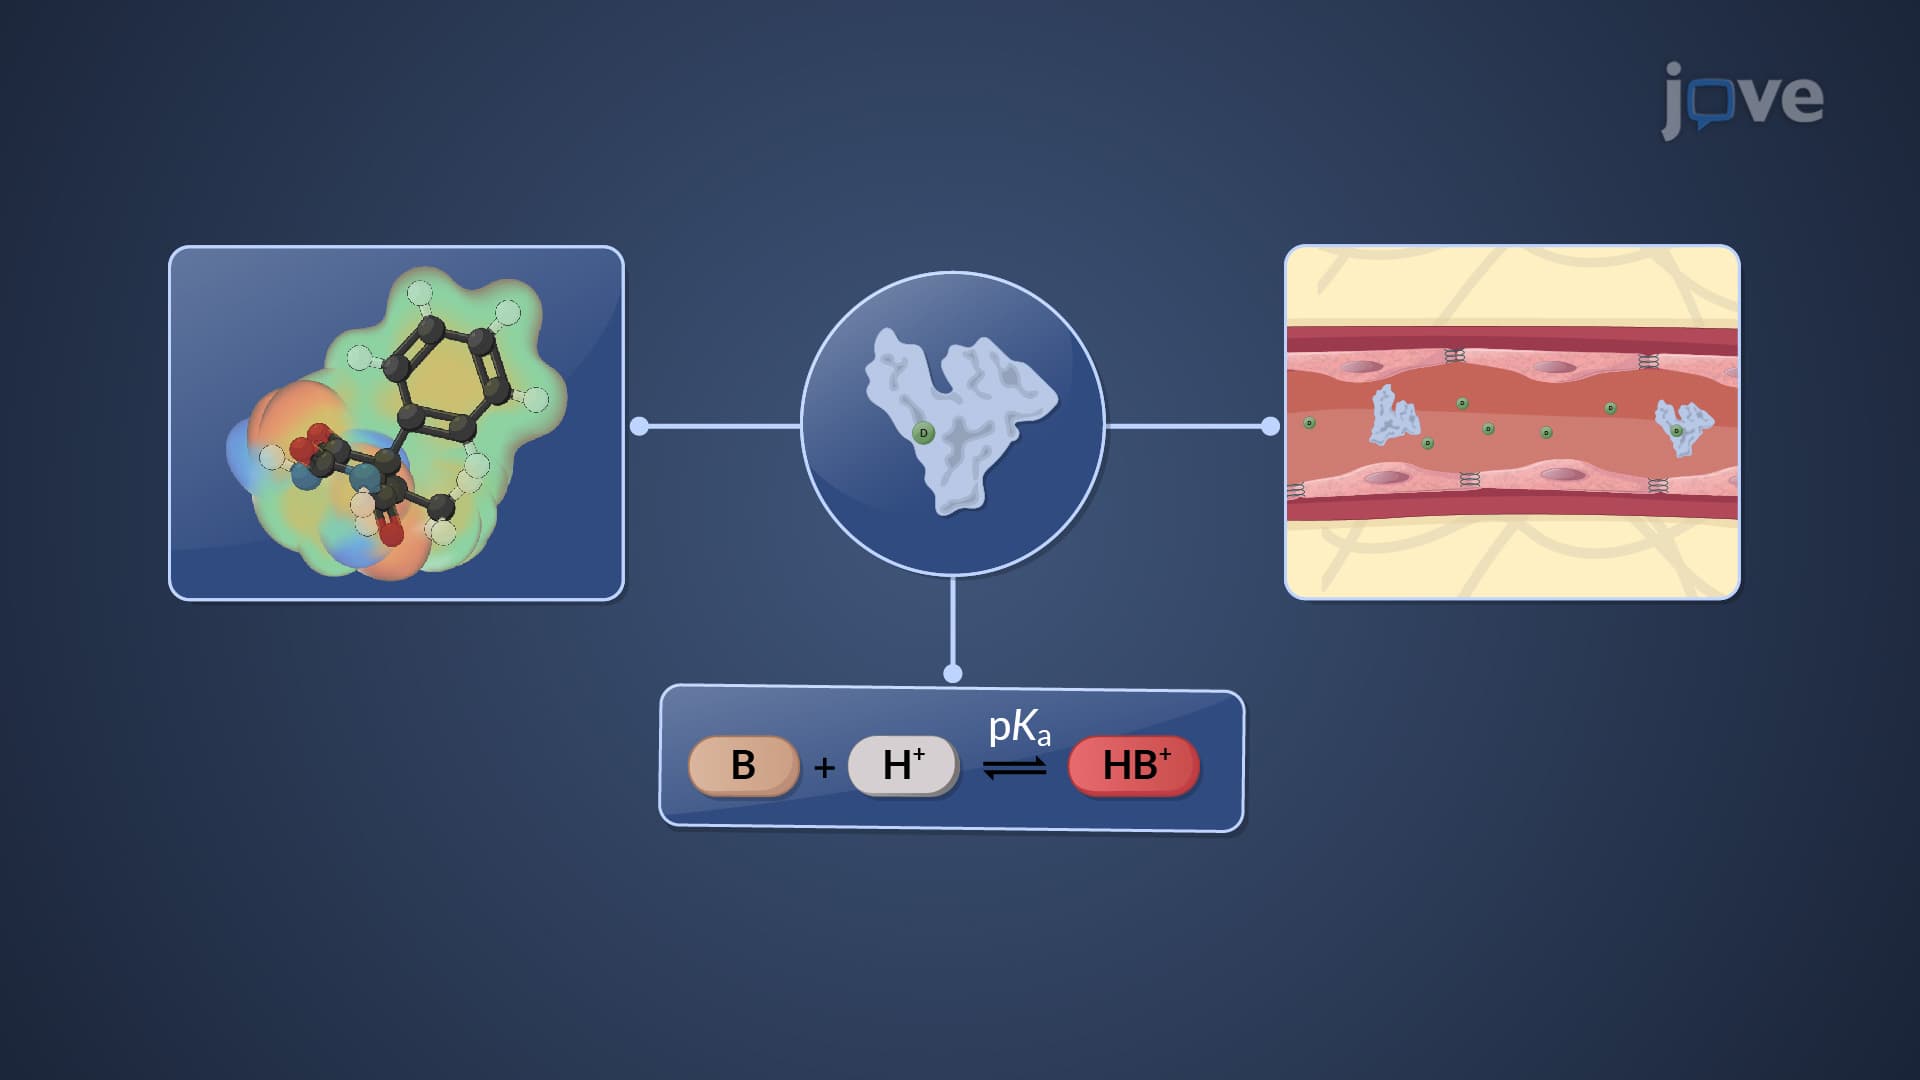 01:18
Factors Affecting Protein-Drug Binding: Drug-Related Factors
One crucial factor in drug-protein binding is the drug's lipophilicity or its affinity for fat. More lipophilic drugs tend to have higher binding extents. For example, highly lipophilic drugs like cloxacillin exhibit substantial protein binding, with as much as 95% of the drug binding to proteins. In contrast,...