Related Experiment Video
Updated: Jul 2, 2026


Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Conformational selection of protein kinase A revealed by flexible-ligand flexible-protein docking
1Department of Chemistry and Biochemistry and Center for Nanoscience, University of Missouri-Saint Louis, One University Boulevard, St. Louis, Missouri 63121, USA.

Abstract:
Protein kinases have high structural plasticity: their structure can change significantly, depending on what ligands are bound to them. Rigid-protein docking methods are not capable of describing such effects. Here, we present a new flexible-ligand flexible-protein docking model in which the protein can adopt conformations between two extremes observed experimentally. The model utilized a molecular dynamics-based simulated annealing cycling protocol and a distance-dependent dielectric model to perform docking. By testing this model on docking four diverse ligands to protein kinase A, we found that the ligands were able to dock successfully to the protein with the proper conformations of the protein induced. By imposing relatively soft conformational restraints to the protein during docking, this model reduced computational costs yet permitted essential conformational changes that were essential for these inhibitors to dock properly to the protein. For example, without adequate movement of the glycine-rich loop, it was difficult for the ligands to move from the surface of the protein to the binding site. In addition, these simulations called for better ways to compare simulation results with experiment other than using the popular root-mean-square deviation between the structure of a ligand in a docking pose and that in experiment because the structure of the protein also changed. In this work, we also calculated the correlation coefficient between protein-ligand/protein-protein distances in the docking structure and those in the crystal structure to check how well a ligand docked into the binding site of the protein and whether the proper conformation of the protein was induced.
More Related Videos
 10:01
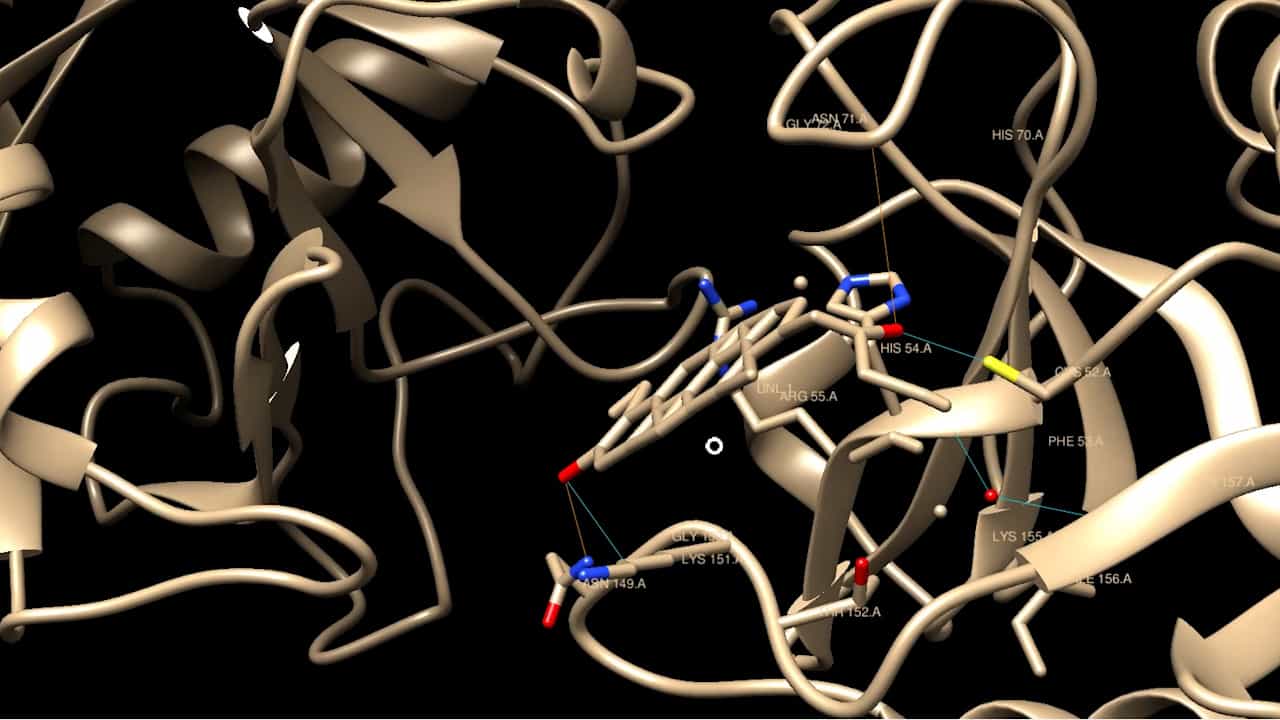
Structure-Guided Design and Development of Novel Cyclophilin A Inhibitors and Ganoderiol-F Derivatives: An In-Silico Approach
Published on: June 23, 2026
 14:34
A Bilingual Computational Workflow for Identifying Potential PLK1 Inhibitors in American Sign Language and English
Published on: April 3, 2026
Related Concept Videos
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 00:49
Ligand Binding and Linkage
 01:58
Cooperative Allosteric Transitions
01:58Cooperative Allosteric Transitions
 01:30
Assembly of Signaling Complexes
Interaction domains in cell signaling
Interaction domains recognize exposed features of their binding partners containing post-translationally modified sequences,...