ProTx-II, a selective inhibitor of NaV1.7 sodium channels, blocks action potential propagation in nociceptors
Related Experiment Video
Updated: Jul 2, 2026
Establishing a Mouse Model of a Pure Small Fiber Neuropathy with the Ultrapotent Agonist of Transient Receptor Potential Vanilloid Type 1
Published on: February 13, 2018
William A Schmalhofer1, Jeffrey Calhoun, Rachel Burrows
1Department of Ion Channels, Merck Research Laboratories, Rahway, NJ 07065-0900, USA.

Abstract:
Voltage-gated sodium (Na(V)1) channels play a critical role in modulating the excitability of sensory neurons, and human genetic evidence points to Na(V)1.7 as an essential contributor to pain signaling. Human loss-of-function mutations in SCN9A, the gene encoding Na(V)1.7, cause channelopathy-associated indifference to pain (CIP), whereas gain-of-function mutations are associated with two inherited painful neuropathies. Although the human genetic data make Na(V)1.7 an attractive target for the development of analgesics, pharmacological proof-of-concept in experimental pain models requires Na(V)1.7-selective channel blockers. Here, we show that the tarantula venom peptide ProTx-II selectively interacts with Na(V)1.7 channels, inhibiting Na(V)1.7 with an IC(50) value of 0.3 nM, compared with IC(50) values of 30 to 150 nM for other heterologously expressed Na(V)1 subtypes. This subtype selectivity was abolished by a point mutation in DIIS3. It is interesting that application of ProTx-II to desheathed cutaneous nerves completely blocked the C-fiber compound action potential at concentrations that had little effect on Abeta-fiber conduction. ProTx-II application had little effect on action potential propagation of the intact nerve, which may explain why ProTx-II was not efficacious in rodent models of acute and inflammatory pain. Mono-iodo-ProTx-II ((125)I-ProTx-II) binds with high affinity (K(d) = 0.3 nM) to recombinant hNa(V)1.7 channels. Binding of (125)I-ProTx-II is insensitive to the presence of other well characterized Na(V)1 channel modulators, suggesting that ProTx-II binds to a novel site, which may be more conducive to conferring subtype selectivity than the site occupied by traditional local anesthetics and anticonvulsants. Thus, the (125)I-ProTx-II binding assay, described here, offers a new tool in the search for novel Na(V)1.7-selective blockers.
Related Concept Videos
 01:17
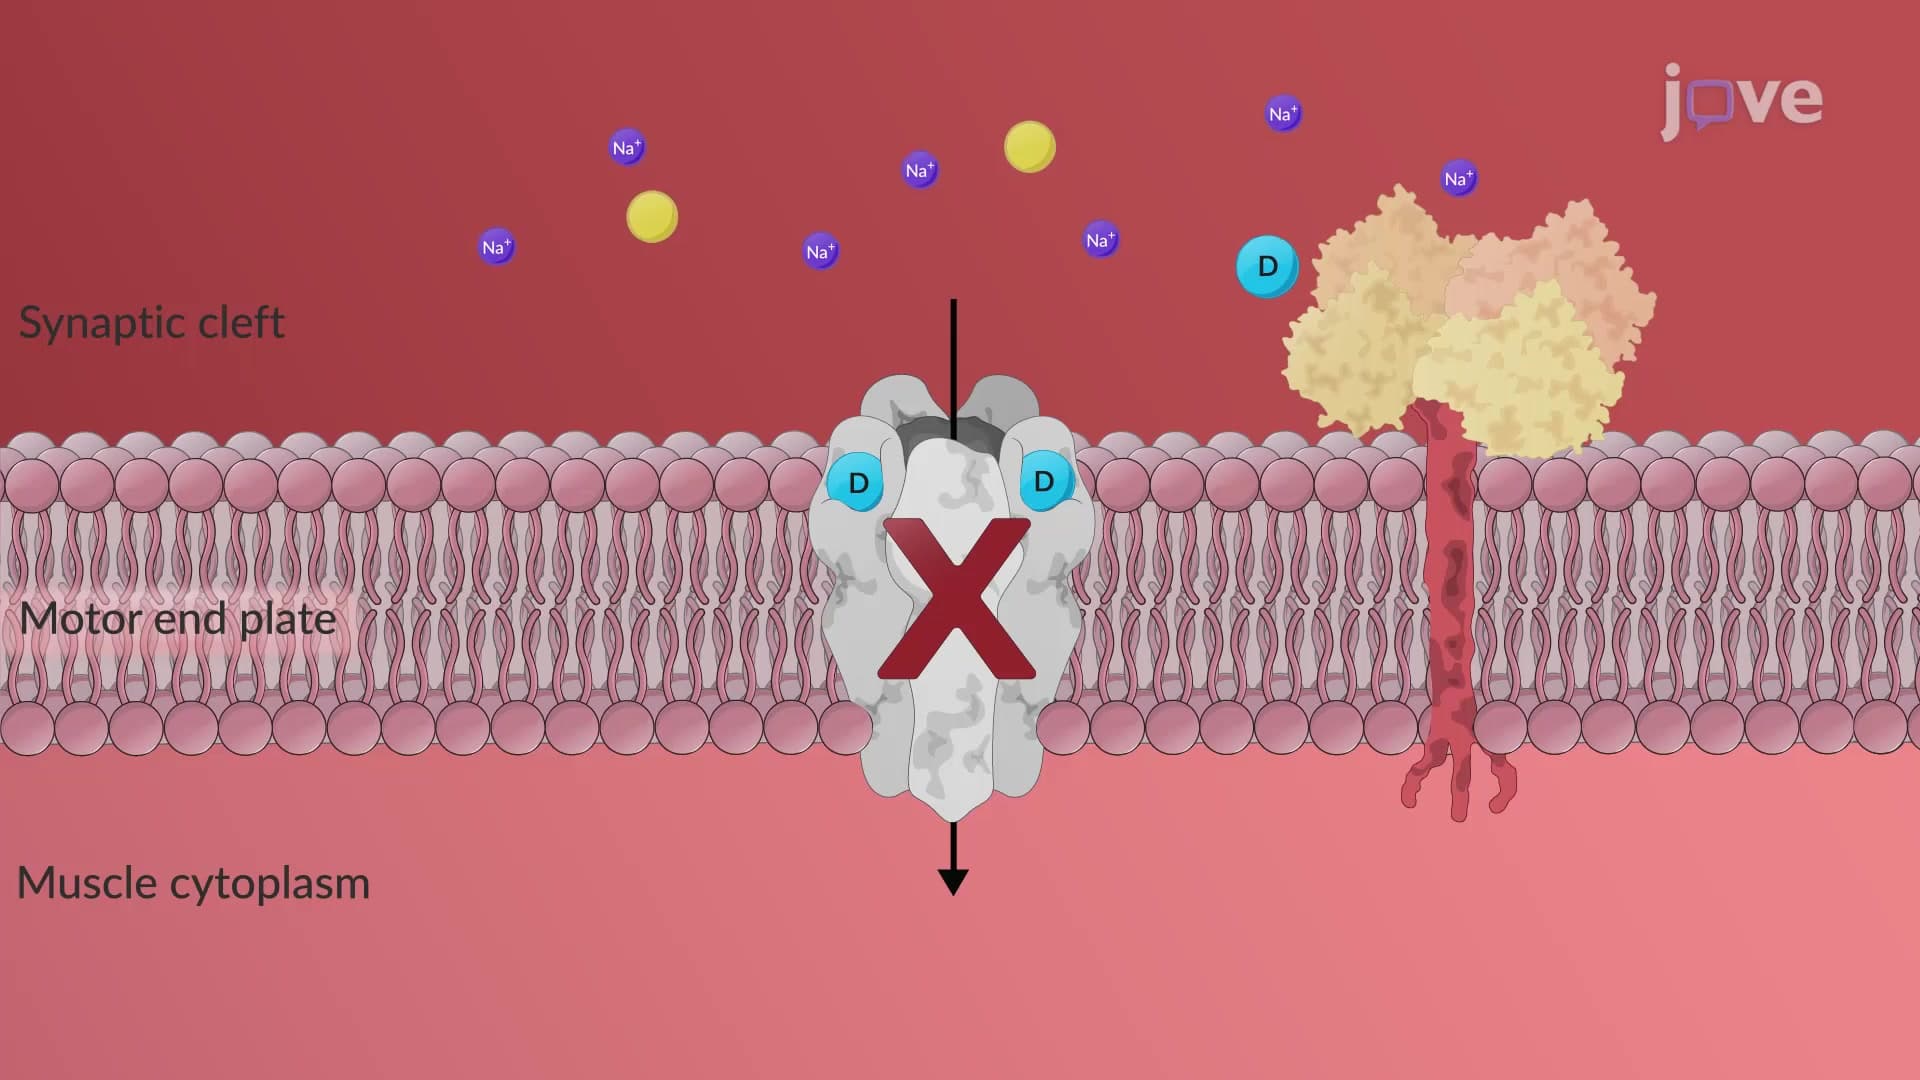
Nondepolarizing (Competitive) Neuromuscular Blockers: Mechanism of Action
Competitive antagonists prevent acetylcholine from binding to its receptor, inhibiting membrane depolarization. Without conformational changes or intrinsic...
 01:22
Antiarrhythmic Drugs: Class I Agents as Sodium Channel Blockers
Class 1A Antiarrhythmic Drugs: These drugs work by moderately blocking sodium channels,...
 01:27
Nondepolarizing (Competitive) Neuromuscular Blockers: Pharmacological Actions
Although all competitive neuromuscular blockers are designed...
 01:30
Ligand-Gated Ion Channel Receptor: Gating Mechanism
 01:28
Depolarizing Blockers: Mechanism of Action
Succinylcholine is the most commonly used depolarizing blocker. Chemically, it constitutes two molecules of acetylcholine joined together by an acetate methyl group. They act on the receptors in the same way as acetylcholine. Because succinylcholine...
 01:28
Chemotherapy-Induced Nausea and Vomiting: Neurokinin-1 Receptor Antagonists