Related Experiment Video
Updated: Jul 2, 2026
Quantification of Proteins Using Peptide Immunoaffinity Enrichment Coupled with Mass Spectrometry
Published on: July 31, 2011
Isotopic peak intensity ratio based algorithm for determination of isotopic clusters and monoisotopic masses of
Kunsoo Park1, Joo Young Yoon, Sunho Lee
1School of Computer Science and Engineering, Seoul National University, Seoul, Korea. kpark@theory.snu.ac.kr

Abstract:
Determining isotopic clusters and their monoisotopic masses is a first step in interpreting complex mass spectra generated by high-resolution mass spectrometers. We propose a mathematical model for isotopic distributions of polypeptides and an effective interpretation algorithm. Our model uses two types of ratios: intensity ratio of two adjacent peaks and intensity ratio product of three adjacent peaks in an isotopic distribution. These ratios can be approximated as simple functions of a polypeptide mass, the values of which fall within certain ranges, depending on the polypeptide mass. Given a spectrum as a peak list, our algorithm first finds all isotopic clusters consisting of two or more peaks. Then, it scores clusters using the ranges of ratio functions and computes the monoisotopic masses of the identified clusters. Our method was applied to high-resolution mass spectra obtained from a Fourier transform ion cyclotron resonance (FTICR) mass spectrometer coupled to reverse-phase liquid chromatography (RPLC). For polypeptides whose amino acid sequences were identified by tandem mass spectrometry (MS/MS), we applied both THRASH-based software implementations and our method. Our method was observed to find more masses of known peptides when the numbers of the total clusters identified by both methods were fixed. Experimental results show that our method performed better for isotopic mass clusters of weak intensity where the isotopic distributions deviate significantly from their theoretical distributions. Also, it correctly identified some isotopic clusters that were not found by THRASH-based implementations, especially those for which THRASH gave 1 Da mismatches. Another advantage of our method is that it is very fast, much faster than THRASH that calculates the least-squares fit.
More Related Videos
Related Concept Videos
 01:13
Mass Spectrometry: Isotope Effect
 01:15
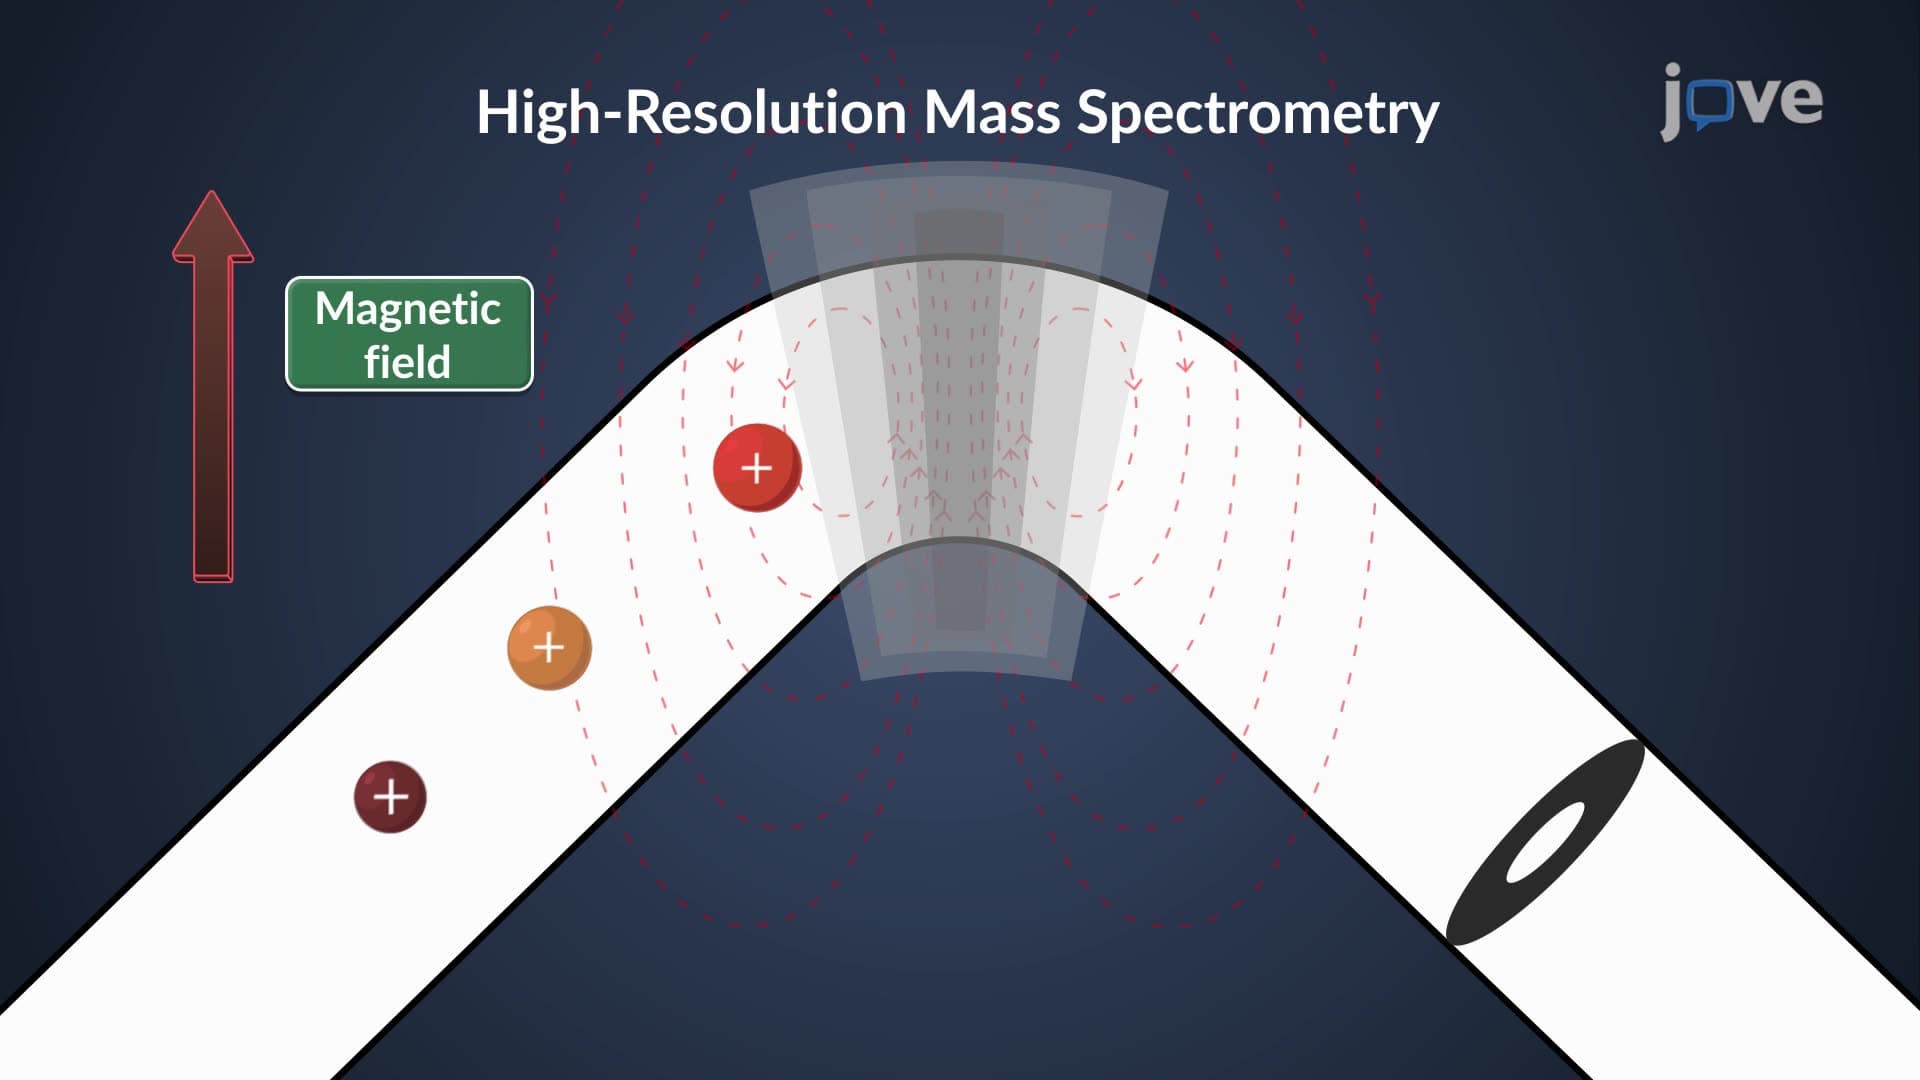
High-Resolution Mass Spectrometry (HRMS)
 01:33
Peptide Identification Using Tandem Mass Spectrometry
This technique helps gather information regarding the protein from which the peptide was obtained and to study the peptides’ amino acid sequence. Identifying peptides from a complex mixture is an important component of the growing field of...
 01:24
Mass Spectrum: Interpretation
 01:01
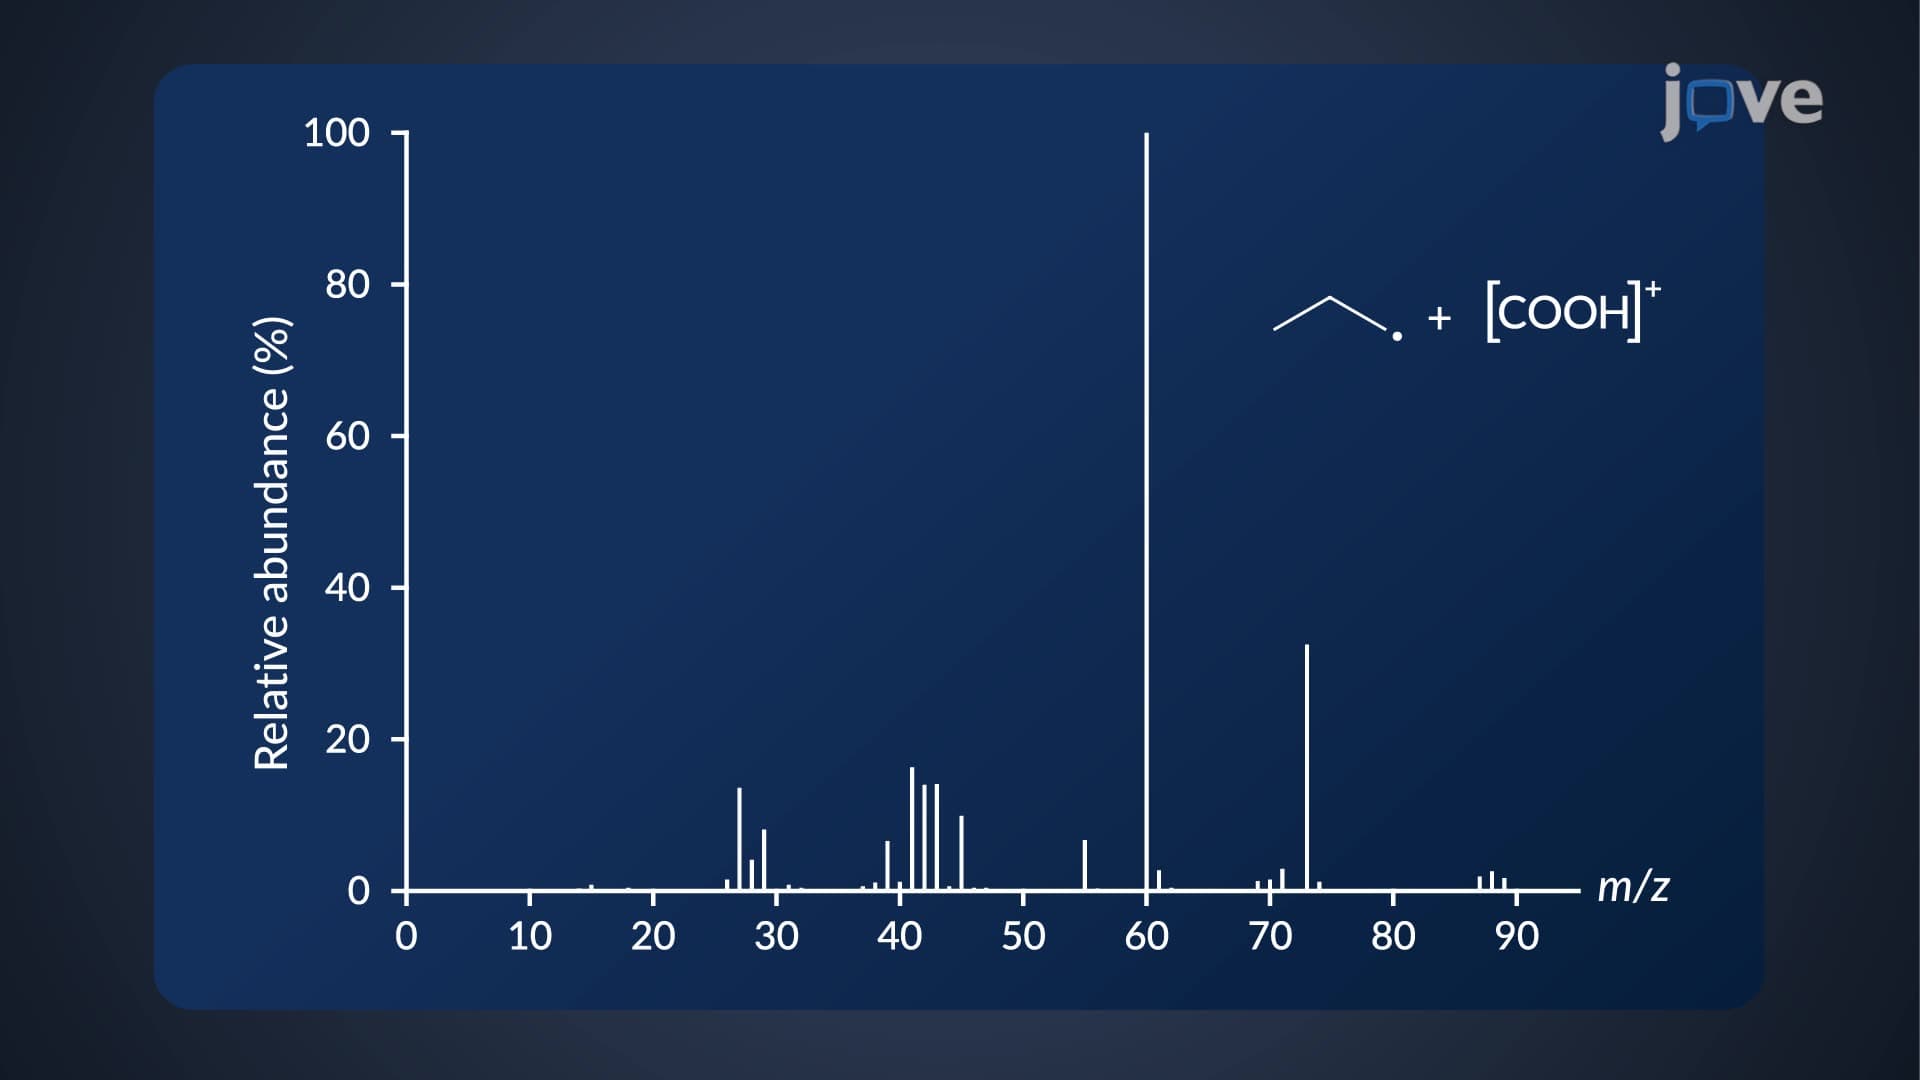
Mass Spectrometry: Carboxylic Acid, Ester, and Amide Fragmentation
For example, the fragmentation of...
 01:23
Mass Spectrum