Related Experiment Video
Updated: Jul 2, 2026

Millisecond Hydrogen/Deuterium-Exchange Mass Spectrometry for the Study of Alpha-Synuclein Structural Dynamics Under Physiological Conditions
Published on: June 23, 2022
Density functional study of intradimer proton transfers in hydrated adenine dimer ions, A2(+)(H2O)n (n = 0-2)
Hye Sun Park1, Sang Hwan Nam, Jae Kyu Song
1Department of Chemistry, Kyunghee University, Seoul 130-701, Republic of Korea.

Abstract:
Structures of mono- and dihydrated adenine dimers and their cations were calculated using B3LYP density functional theory with the 6-31+G(d,p) basis set, in order to help understand photofragmentation experiments of hydrated adenine dimers from the energetics point of view. Several important pathways leading to the major fragmentation product, protonated adenine ion (AH(+)), thermodynamically at minimum costs were investigated at the ground-state electronic potential surface of hydrated adenine dimer cations. Our calculations suggest that the proton transfer from one adenine moiety to the other in hydrated dimer ions readily occurs with negligible barriers in normal hydration conditions. In asymmetrically hydrated ions, however, the proton transfer to more hydrated adenine moieties is kinetically hindered due to heightened transition-state barriers, while the other way is still barrierless. Such directional preference in proton transfer may be characterized as a unique dimer ion property, stemming from the difference in basicity of the two nitrogen atoms involved in the double hydrogen bond that would be equivalent without hydration. We also found that dimer cleavage requires about 4 times larger energy than evaporation of individual water molecules, so it is likely that most solvent molecules evaporate before the eventual dimer cleavage when available internal energy is limited.
More Related Videos
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 00:48
¹H NMR of Labile Protons: Deuterium (²H) Substitution
 01:20
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
 01:22
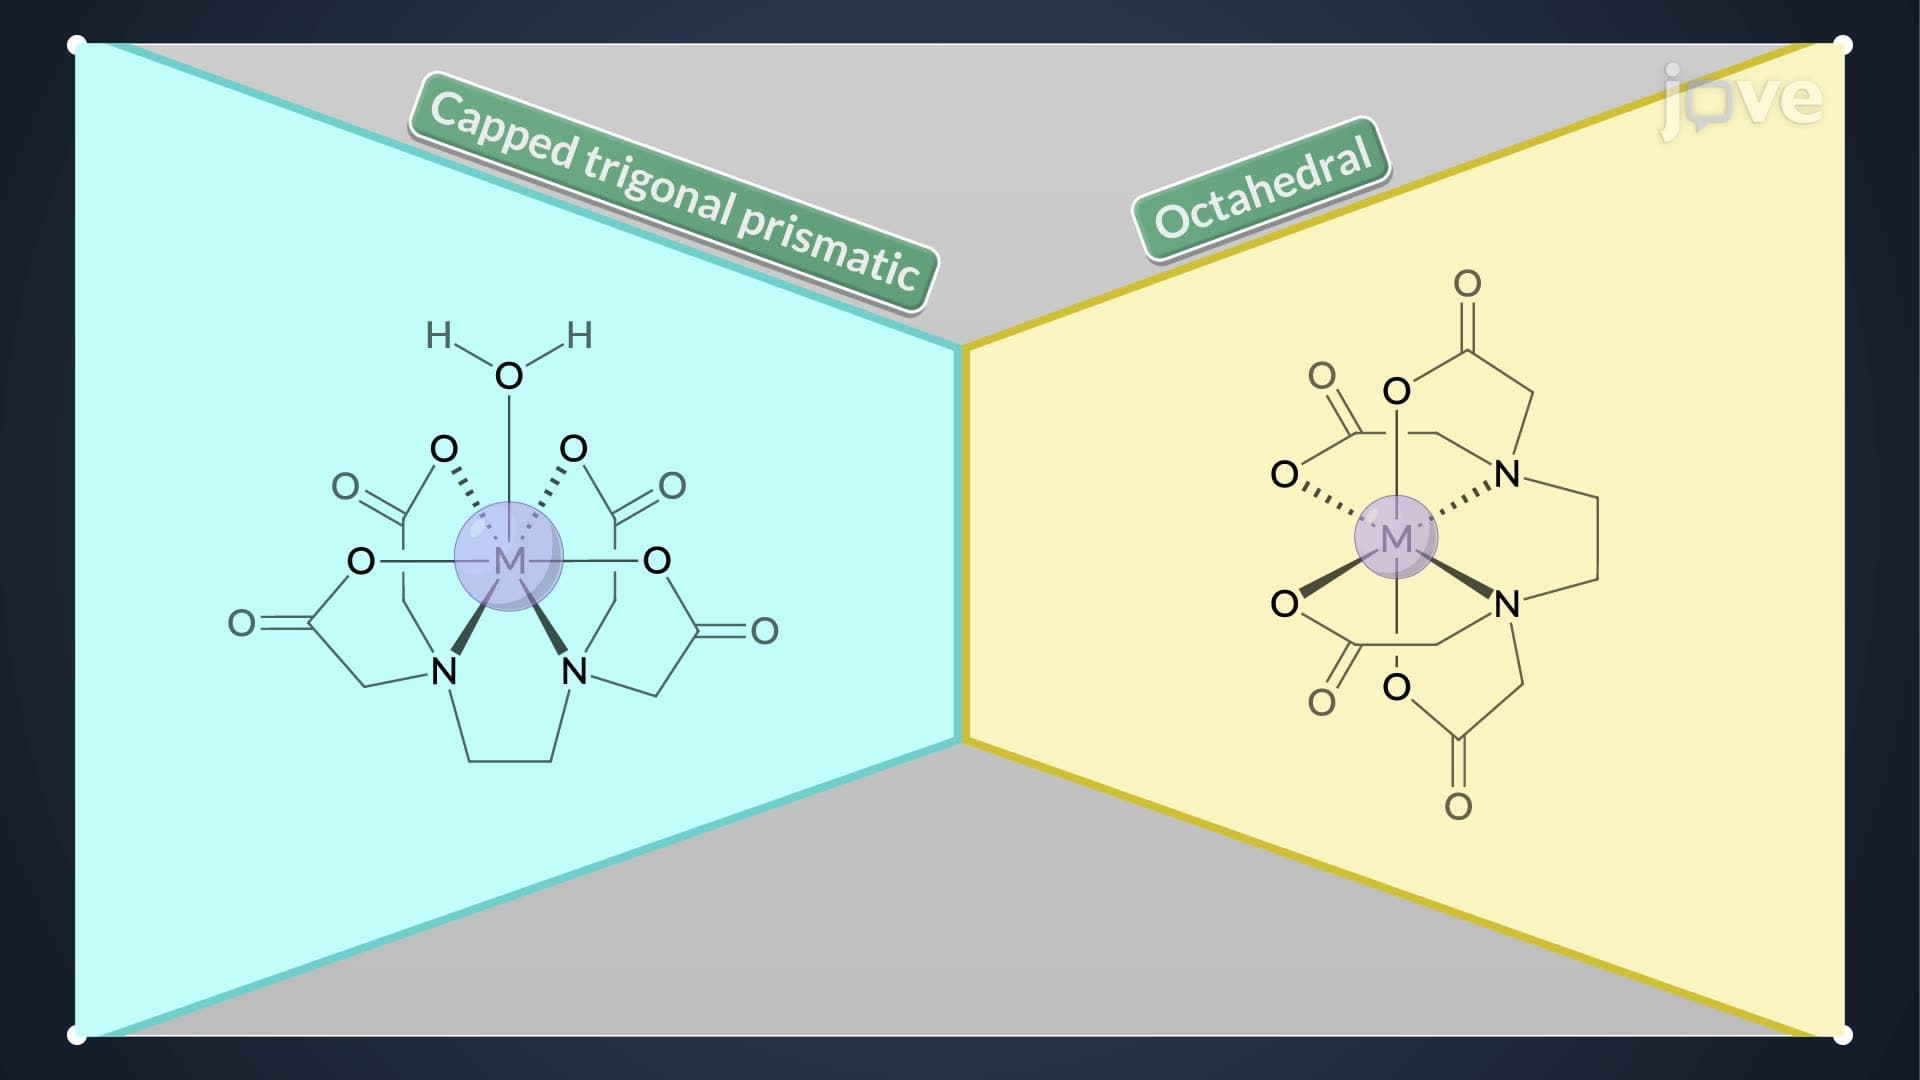
EDTA: Chemistry and Properties
 01:21
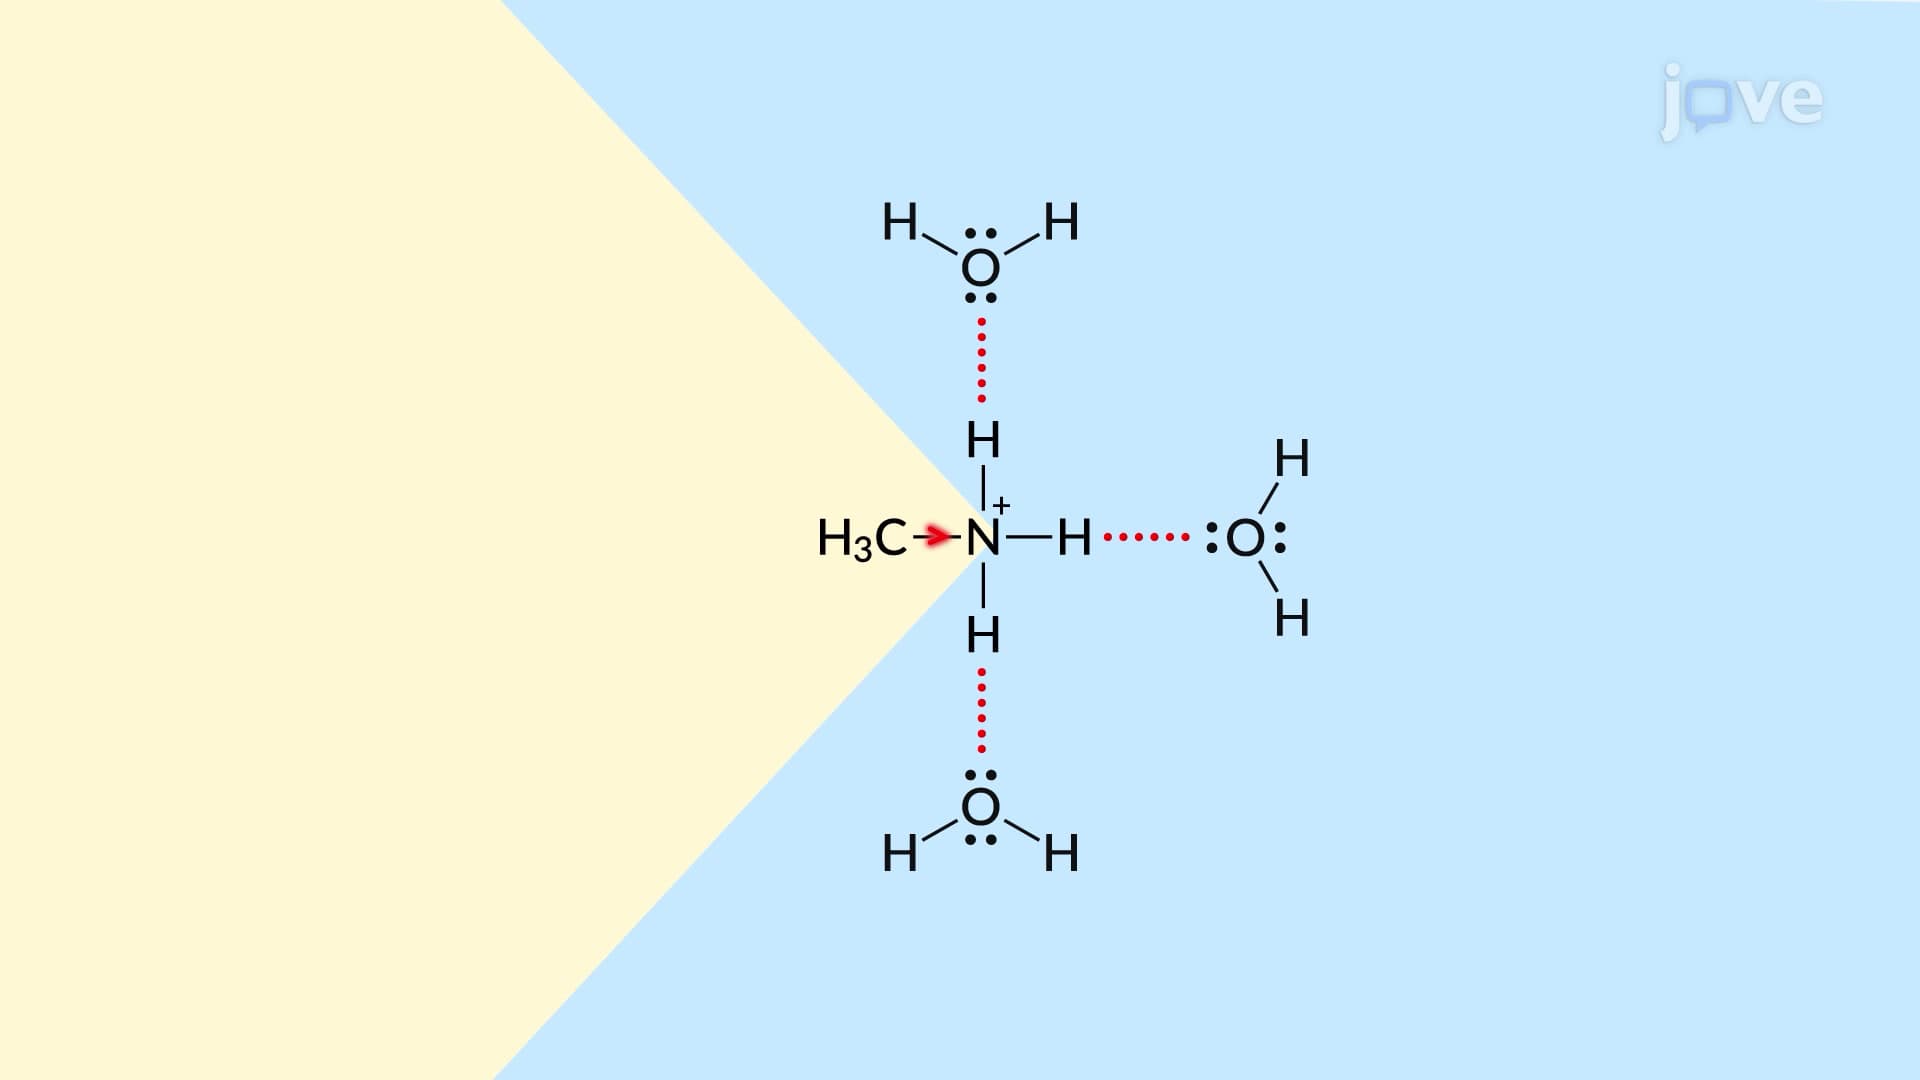
Basicity of Aliphatic Amines
To measure the basicity of amines, two conventions are generally used. The first defines Kb as the basicity constant for the deprotonation reaction of water by the amine, as presented in Figure 1. Conventionally, lower Kb indicates higher...
 00:53
Extraction: Effects of pH