Related Experiment Video
Updated: Jul 1, 2026

Quantifying Agonist Activity at G Protein-coupled Receptors
Published on: December 26, 2011
Quantitative structure-activity relationships for PPAR-gamma binding and gene transactivation of tyrosine-based
Costas Giaginis1, Stamatios Theocharis, Anna Tsantili-Kakoulidou
1Department of Pharmaceutical Chemistry, School of Pharmacy, University of Athens, Panepistimiopolis, Zografou, Athens 15771, Greece.

Abstract:
Peroxisome proliferator-activated receptor-gamma offers a molecular target for drugs aimed to treat type II diabetes mellitus, while its therapeutic potency against cancer disease is currently being explored in preclinical studies. Tyrosine derivatives constitute a major class of peroxisome proliferator-activated receptor-gamma agonists attracting considerable research interest in drug discovery. Thus, the establishment of adequate QSAR models would serve as a guide for further molecular design. In the present study, multivariate data analysis was applied on a large set of tyrosine-based peroxisome proliferator-activated receptor-gamma agonists for modelling binding affinity, expressed as pKi and gene transactivation, expressed as pEC(50). A pool of descriptors based on physicochemical and molecular properties as well as on specific structural characteristics was used and two PLS models with satisfactory statistics were produced for binding data. According to them, molecular weight, rotatable bonds and lipophilicity were found to exert a considerable positive influence, while excess negative and positive charge created by additional acidic or basic groups in the molecules was unfavourable. With gene transactivation data, an adequate model was obtained only for the highly active compounds if considered separately. The higher complexity incorporated in gene transactivation data was further investigated by establishing a PLS model, which improved the inter-relationship between pEC(50) and pKi.
Insights
Quantitative Structure-Activity Relationship (QSAR) models guide the design of novel tyrosine-based agonists targeting peroxisome proliferator-activated receptor-gamma (PPARγ) for diabetes and cancer therapies. Key molecular properties influencing binding affinity and gene transactivation were identified.
Area of Science:
- Medicinal Chemistry
- Computational Chemistry
- Pharmacology
Background:
- Peroxisome proliferator-activated receptor-gamma (PPARγ) is a key target for type II diabetes treatments and is being investigated for anti-cancer drug discovery.
- Tyrosine derivatives are a significant class of PPARγ agonists, prompting interest in developing Quantitative Structure-Activity Relationship (QSAR) models for drug design.
- QSAR models are crucial for predicting the activity of new compounds and guiding the molecular design of PPARγ agonists.
Purpose of the Study:
- To establish robust QSAR models for a large set of tyrosine-based PPARγ agonists.
- To identify key molecular descriptors influencing binding affinity (pKi) and gene transactivation (pEC50).
- To explore the relationship between binding affinity and gene transactivation for improved drug design.
Main Methods:
- Multivariate data analysis, specifically Partial Least Squares (PLS) regression, was employed.
- A comprehensive set of molecular descriptors, including physicochemical properties and structural characteristics, was generated.
- PLS models were developed for both binding affinity and gene transactivation data.
Main Results:
- Two PLS models for binding affinity demonstrated satisfactory statistical performance.
- Molecular weight, rotatable bonds, and lipophilicity positively influenced binding affinity.
- Excess positive or negative charges and gene transactivation data complexity required separate modeling for highly active compounds, with a PLS model improving the pEC50-pKi inter-relationship.
Conclusions:
- The study successfully developed QSAR models for tyrosine-based PPARγ agonists, identifying critical structural features for drug design.
- Molecular weight, lipophilicity, and rotatable bonds are favorable for PPARγ binding.
- Understanding the interplay between binding and transactivation is key for optimizing PPARγ-targeting therapeutics.
Related Concept Videos
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence its...
 01:31
Transducer Mechanism: Nuclear Receptors
About 48 different soluble family members of nuclear receptors are identified that can be divided into two main classes:
 01:29
The Two-State Receptor Model
The binding affinity of a drug determines its interaction with one...
 01:09
GPCRs Regulate Adenylyl Cylase Activity
Two...
 01:30
Quantitative Aspects of Drug-Receptor Interaction
 01:15
G Protein-coupled Receptors
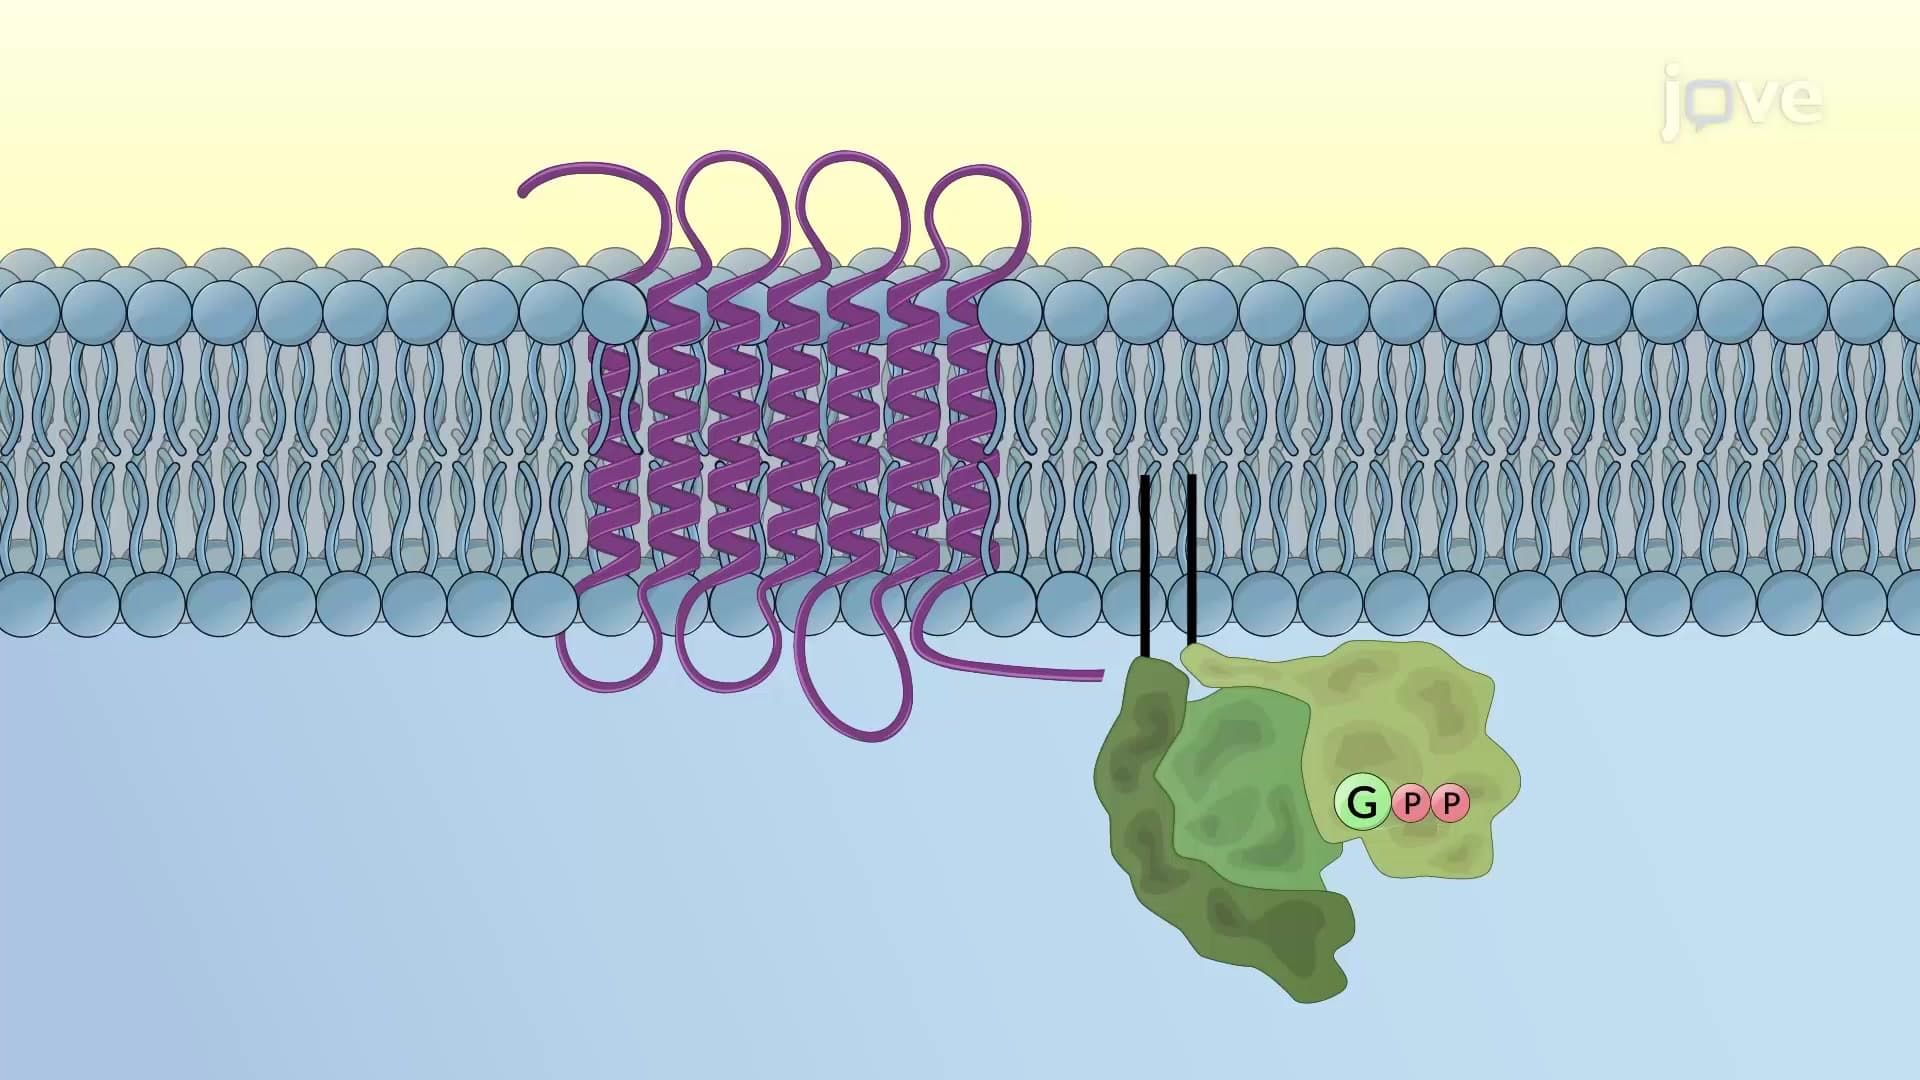
GPCRs are also called heptahelical, 7TM, or serpentine receptors, and consist of seven (H1-H7) transmembrane alpha-helices that span the bilayer to form a cylindrical core. The transmembrane helices are connected by three extracellular loops and three...