Primary hyperoxaluria type 1 with a novel mutation

Sidharth Kumar Sethi1, Hans R Waterham, Sonika Sharma

  • 1Departments of Clinical Chemistry & Pediatrics, Academic Medical Centre, Amsterdam, The Netherlands.

Indian Journal of Pediatrics
|September 24, 2008
PubMed

Related Concept Videos

Inborn Errors of Metabolism01:20

Inborn Errors of Metabolism

Phenylketonuria (PKU) is a protein metabolism disorder characterized by high blood levels of the amino acid phenylalanine. This results from a mutation in the gene responsible for phenylalanine hydroxylase, an enzyme that converts phenylalanine into tyrosine. When this enzyme is deficient, phenylalanine builds up in the blood, leading to symptoms such as vomiting, rashes, seizures, growth deficiency, and severe mental retardation. An early diagnosis and a diet restricting phenylalanine intake...
Protein Import into the Peroxisomes01:27

Protein Import into the Peroxisomes

Cells contain membrane-bound organelles called peroxisomes that oxidize organic molecules by transferring hydrogen atoms to oxygen, producing hydrogen peroxide. Peroxisomes enzymatically convert the released hydrogen peroxide into water and oxygen.
Peroxisomal Protein Import:
Peroxisomes lack the genetic machinery required to code for their own proteins. Hence, most peroxisomal membrane, lumenal and transmembrane proteins are synthesized in the cytoplasm or ER and transported to the peroxisome...
Lysosomal Hydrolases01:22

Lysosomal Hydrolases

Lysosomes are the site for the degradation of macromolecules and biological polymers released during membrane trafficking events such as secretory, endocytic, autophagic, and phagocytic pathways. The membrane-enclosed area of the lysosome, called the lumen, contains hydrolytic enzymes active in an acidic environment. These acid hydrolases are functional at a pH between 4.5 and 5 and are involved in cellular processes such as cell signaling, energy metabolism, restoration of the plasma membrane,...
Pharmacogenetics of Drug Transporters: P-Glycoprotein and Solute Carrier Transporters01:16

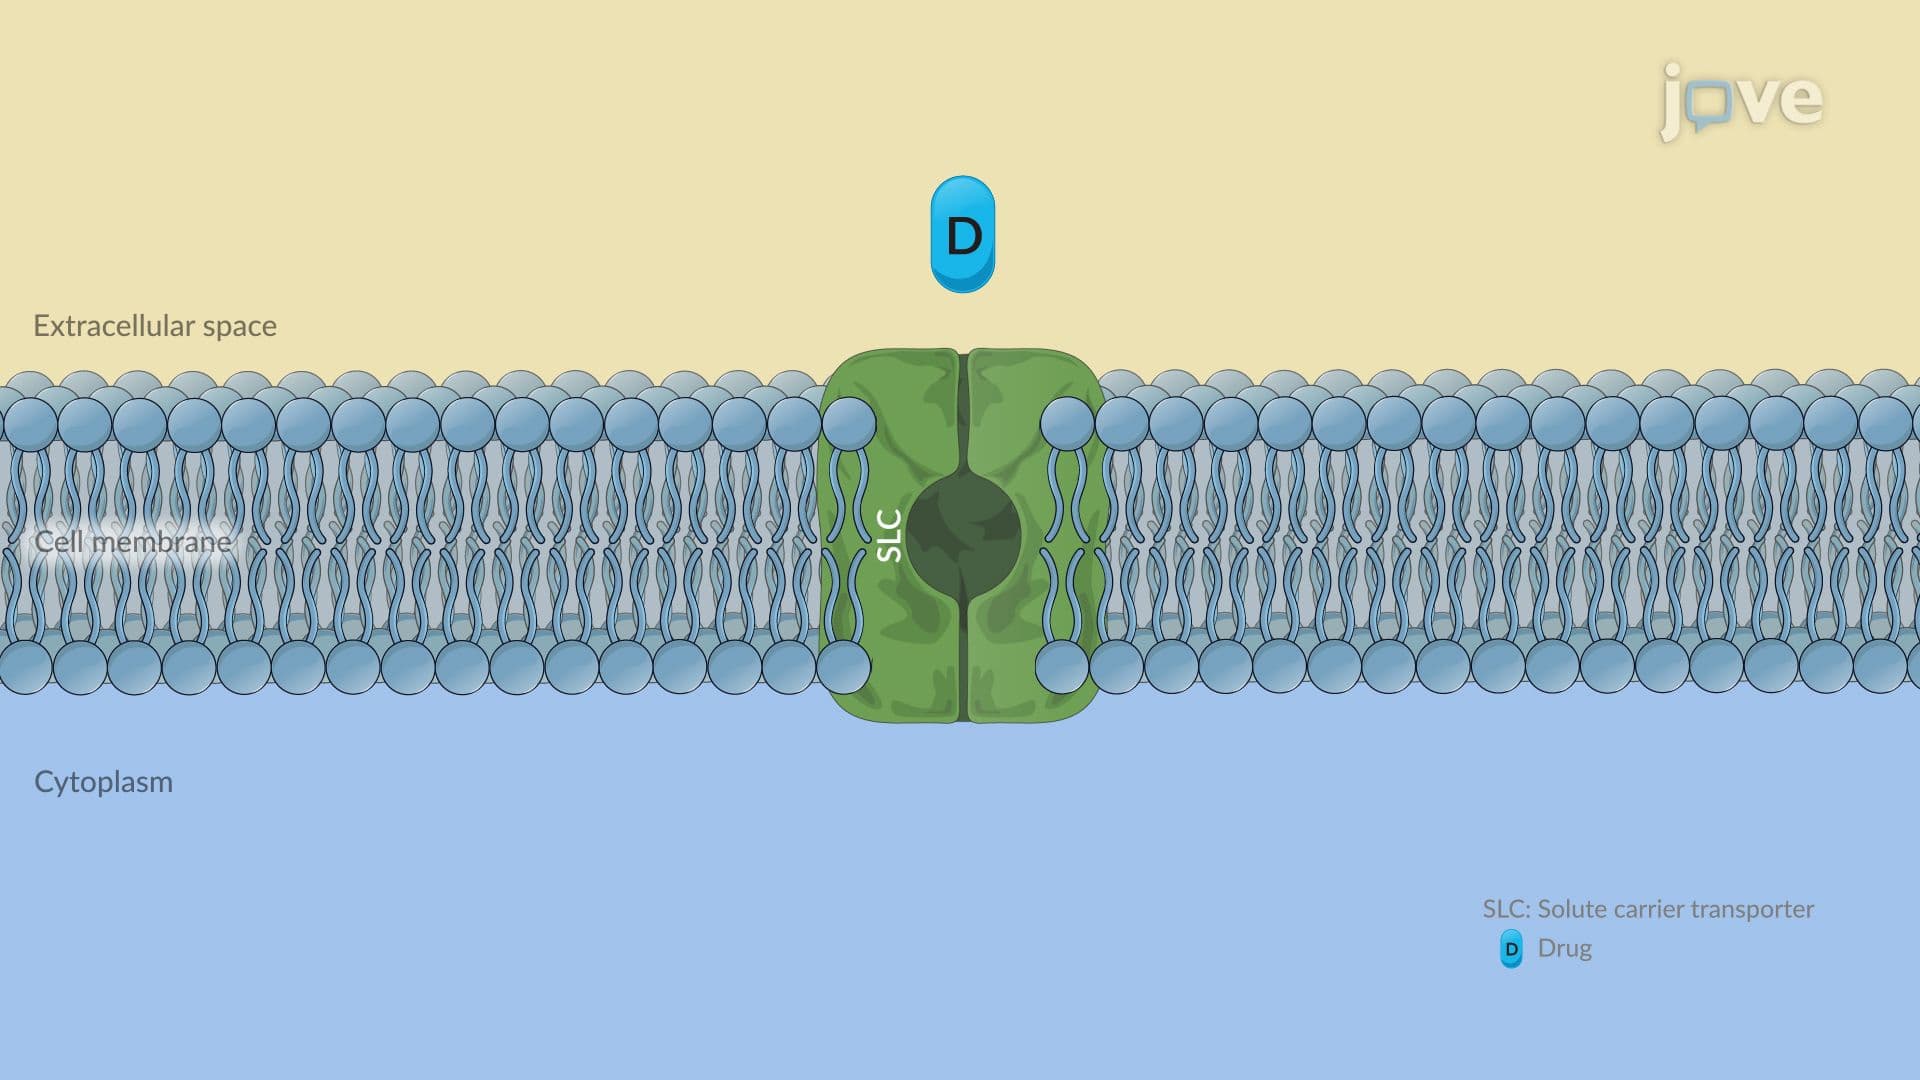
Pharmacogenetics of Drug Transporters: P-Glycoprotein and Solute Carrier Transporters

The pharmacogenetics of drug transporters is increasingly recognized as a critical factor influencing interindividual variability in drug absorption, distribution, and elimination. These membrane-bound proteins regulate drugs' movement across cellular barriers by actively pumping them out (efflux) or facilitating their uptake (influx). Among the major transporter families, ATP-binding cassette (ABC) and solute carrier (SLC) transporters play particularly prominent roles. Genetic polymorphisms...
Glucose Transporters01:27

Glucose Transporters

Glucose transporters facilitate the transport of glucose across the cell membrane. In addition to glucose, some glucose transporters can also aid the movement of other hexoses such as fructose, mannose, and galactose.
Facilitated diffusion-glucose transporters (GLUTs) are encoded by the solute-linked carrier (SLC) family 2, subfamily A gene family, or SLC2A. The 14 GLUT protein members are distributed into three classes:
Pharmacogenetics of Phase I Enzymes: Cytochrome P450 Isozymes01:28

Pharmacogenetics of Phase I Enzymes: Cytochrome P450 Isozymes

Cytochrome P450 (CYP450) enzymes are a superfamily of heme-containing monooxygenases that play a pivotal role in Phase I drug metabolism by catalyzing oxidation and reduction reactions.These enzymes transform lipophilic xenobiotics into more hydrophilic metabolites, facilitating subsequent Phase II conjugation and eventual excretion. The CYP450 family is classified into families (e.g., CYP1–CYP3) and subfamilies (e.g., CYP2A, CYP2C), based on amino acid sequence homology.CYP450 isoenzymes,...