Related Experiment Video
Updated: Jun 28, 2026
Versatile CO2 Transformations into Complex Products: A One-pot Two-step Strategy
Published on: November 9, 2019
Unraveling high precision stereocontrol in a triple cascade organocatalytic reaction
C B Shinisha1, Raghavan B Sunoj
1Department of Chemistry, Indian Institute of Technology Bombay, Powai, Mumbai 400076, India.

Abstract:
The mechanism and stereoselectivity in an organocatalyzed triple cascade reaction between an aldehyde, electron deficient olefin and an alpha,beta-unsaturated aldehyde are investigated for the first time using density functional theory. The factors responsible for high levels of observed stereoselectivity (Enders et al., Nature, 2006, 441, 861) towards the generation of cyclohexene carbaldehyde with four contiguous stereocentres are unravelled. The triple cascade reaction, comprising a Michael, Michael and aldol sequence as the key elementary reactions, is studied by identifying the corresponding transition states for the stereoselective C-C bond-formation. In the first Michael addition step between the enamine (derived from the chiral catalyst and propanal) and nitrostyrene, energetically the most preferred mode of addition is found to be between the si-face of (E)-anti-enamine on the si-face of nitrostyrene. The addition of the si-face of the nitroalkane anion on the re-face of the iminium ion (formed between the enal and the catalyst) is the lowest energy pathway for the second Michael addition step. The high level of asymmetric induction is rationalized with the help of relative activation barriers associated with the competitive diastereomeric pathways. Interesting weak interactions, along with the steric effects offered by the bulky alpha-substituent on the pyrrolidine ring, are identified as critical to the stereoselectivity in this triple cascade reaction. The predicted stereoselectivities using computed energetics are found to be in perfect harmony with the experimental stereoselectivities.
More Related Videos
Related Concept Videos
 01:17
Thermal Electrocyclic Reactions: Stereochemistry
Selection Rules: Thermal Activation
Conjugated systems containing an even number of π-electron pairs undergo a conrotatory ring closure. For example, thermal electrocyclization of (2E,4E)-2,4-hexadiene, a conjugated diene containing two π-electron pairs, gives trans-3,4-dimethylcyclobutene.
 02:36
Regioselectivity and Stereochemistry of Hydroboration
Hydroboration proceeds in a concerted fashion with the attack of borane on the π bond, giving a cyclic four-centered transition state. The –BH2 group is bonded to the less substituted carbon and –H to the more substituted carbon. The concerted nature requires the simultaneous addition of –H and –BH2 across the same face of the alkene giving syn stereochemistry.
 01:26
Photochemical Electrocyclic Reactions: Stereochemistry
Selection Rules: Photochemical Activation
 01:28
Diels–Alder Reaction Forming Cyclic Products: Stereochemistry
 02:43
E2 Reaction: Stereochemistry and Regiochemistry
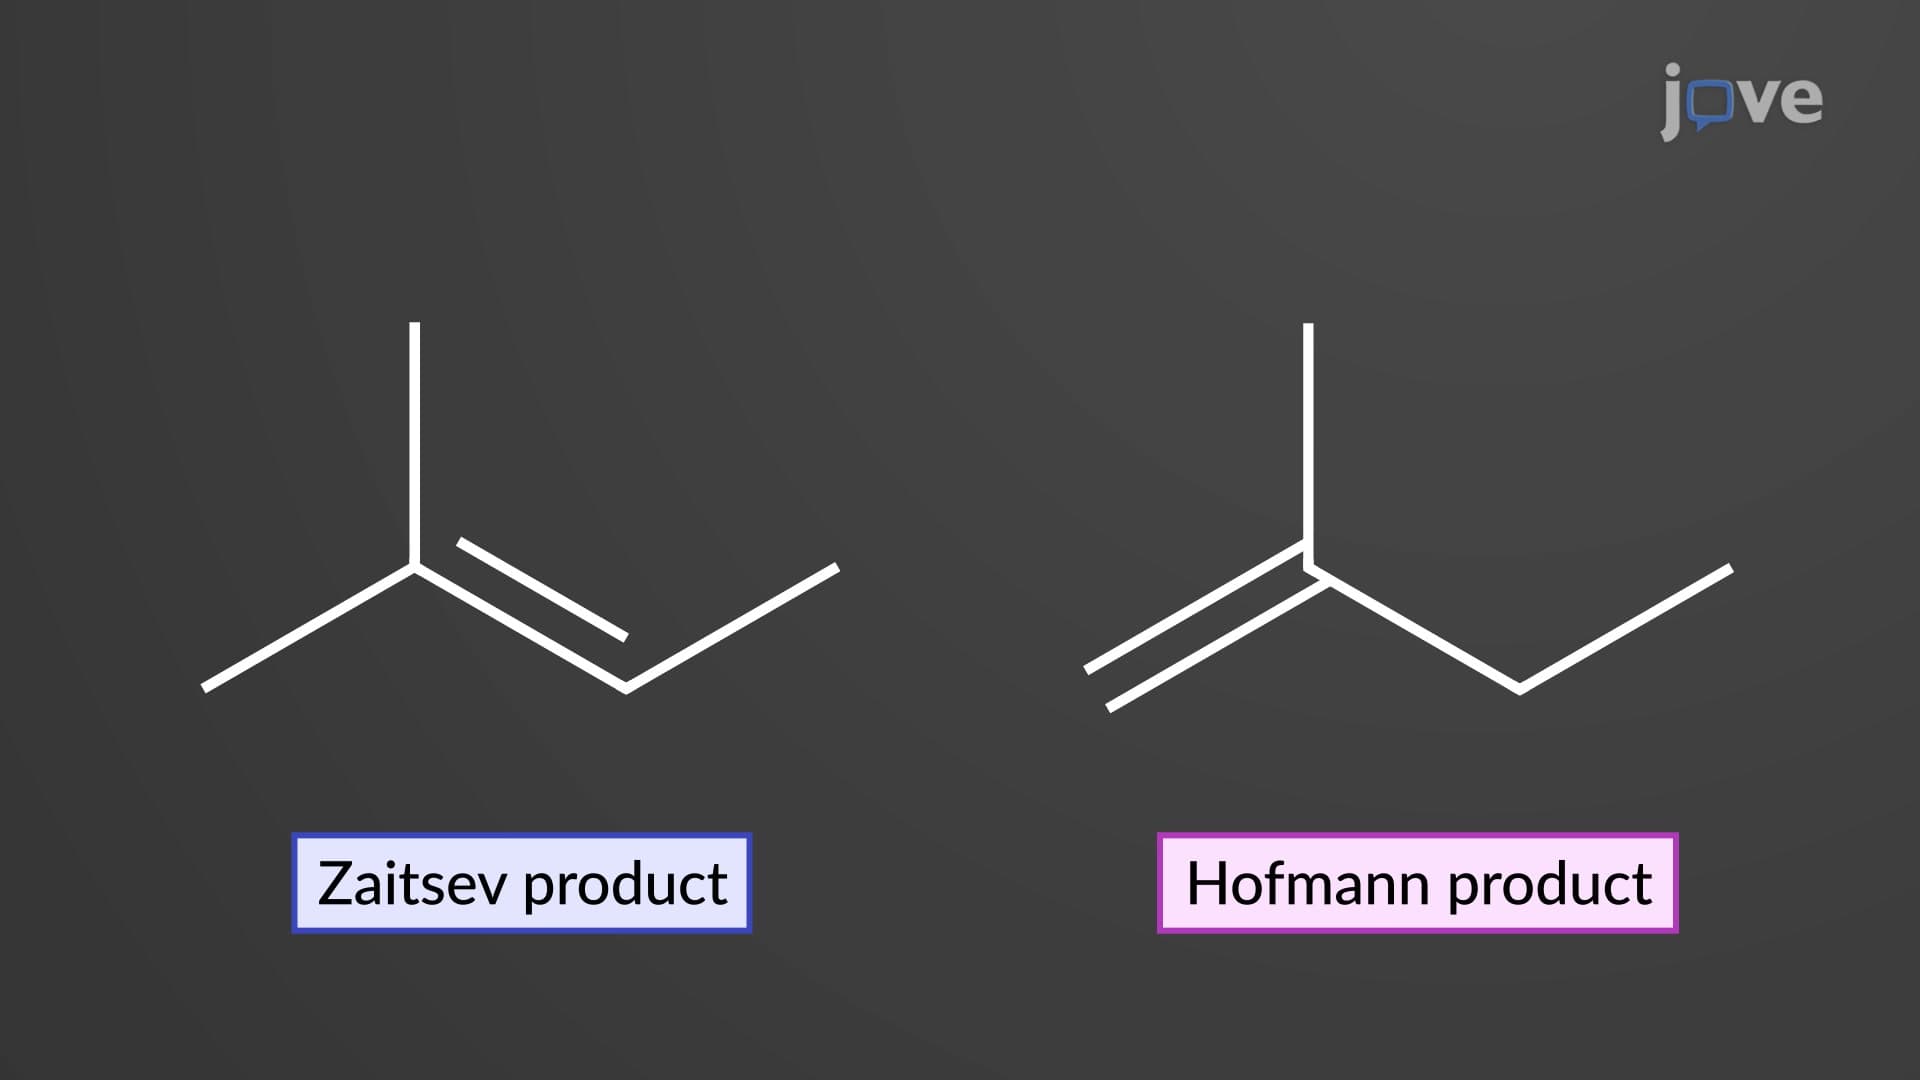
When a substrate with two different β hydrogens undergoes an E2 elimination, the presence of a strong base can yield two regioisomeric alkenes. The more-substituted alkene is the major product and...
 02:43
E1 Reaction: Stereochemistry and Regiochemistry