Related Experiment Video
Updated: Jun 28, 2026

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Not innocent: verdict from ab initio multiconfigurational second-order perturbation theory on the electronic
Björn O Roos1, Valera Veryazov, Jeanet Conradie
1Department of Theoretical Chemistry, Chemical Center, University of Lund, P.O. Box 124, S-221 00 Lund, Sweden.

Abstract:
From a suitably broad perspective, transition metal corroles may be viewed as stable, synthetic analogues of high-valent heme protein intermediates such as compounds I and II. Against this backdrop, the electronic structure of chloroiron corrole has provoked a lively debate in recent years. Thus, whereas NMR spectroscopy and DFT calculations suggest an S = 3/2 Fe(III) corrole (*2-) radical description, certain researchers have favored an Fe(IV) formulation. These two descriptions are indistinguishable as far as DFT calculations are concerned. Ab initio CASSCF/CASPT2 calculations provide unambiguous support for the former description. In addition, they rule out any Fe(IV) state, whether high- or low-spin, within 1.5 eV of the ground state.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 08:51
Coulomb Explosion Imaging as a Tool to Distinguish Between Stereoisomers
Published on: August 18, 2017
Related Concept Videos
 01:14
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds
 01:26
Photochemical Electrocyclic Reactions: Stereochemistry
Selection Rules: Photochemical Activation
 01:27
π Electron Effects on Chemical Shift: Overview
 01:27
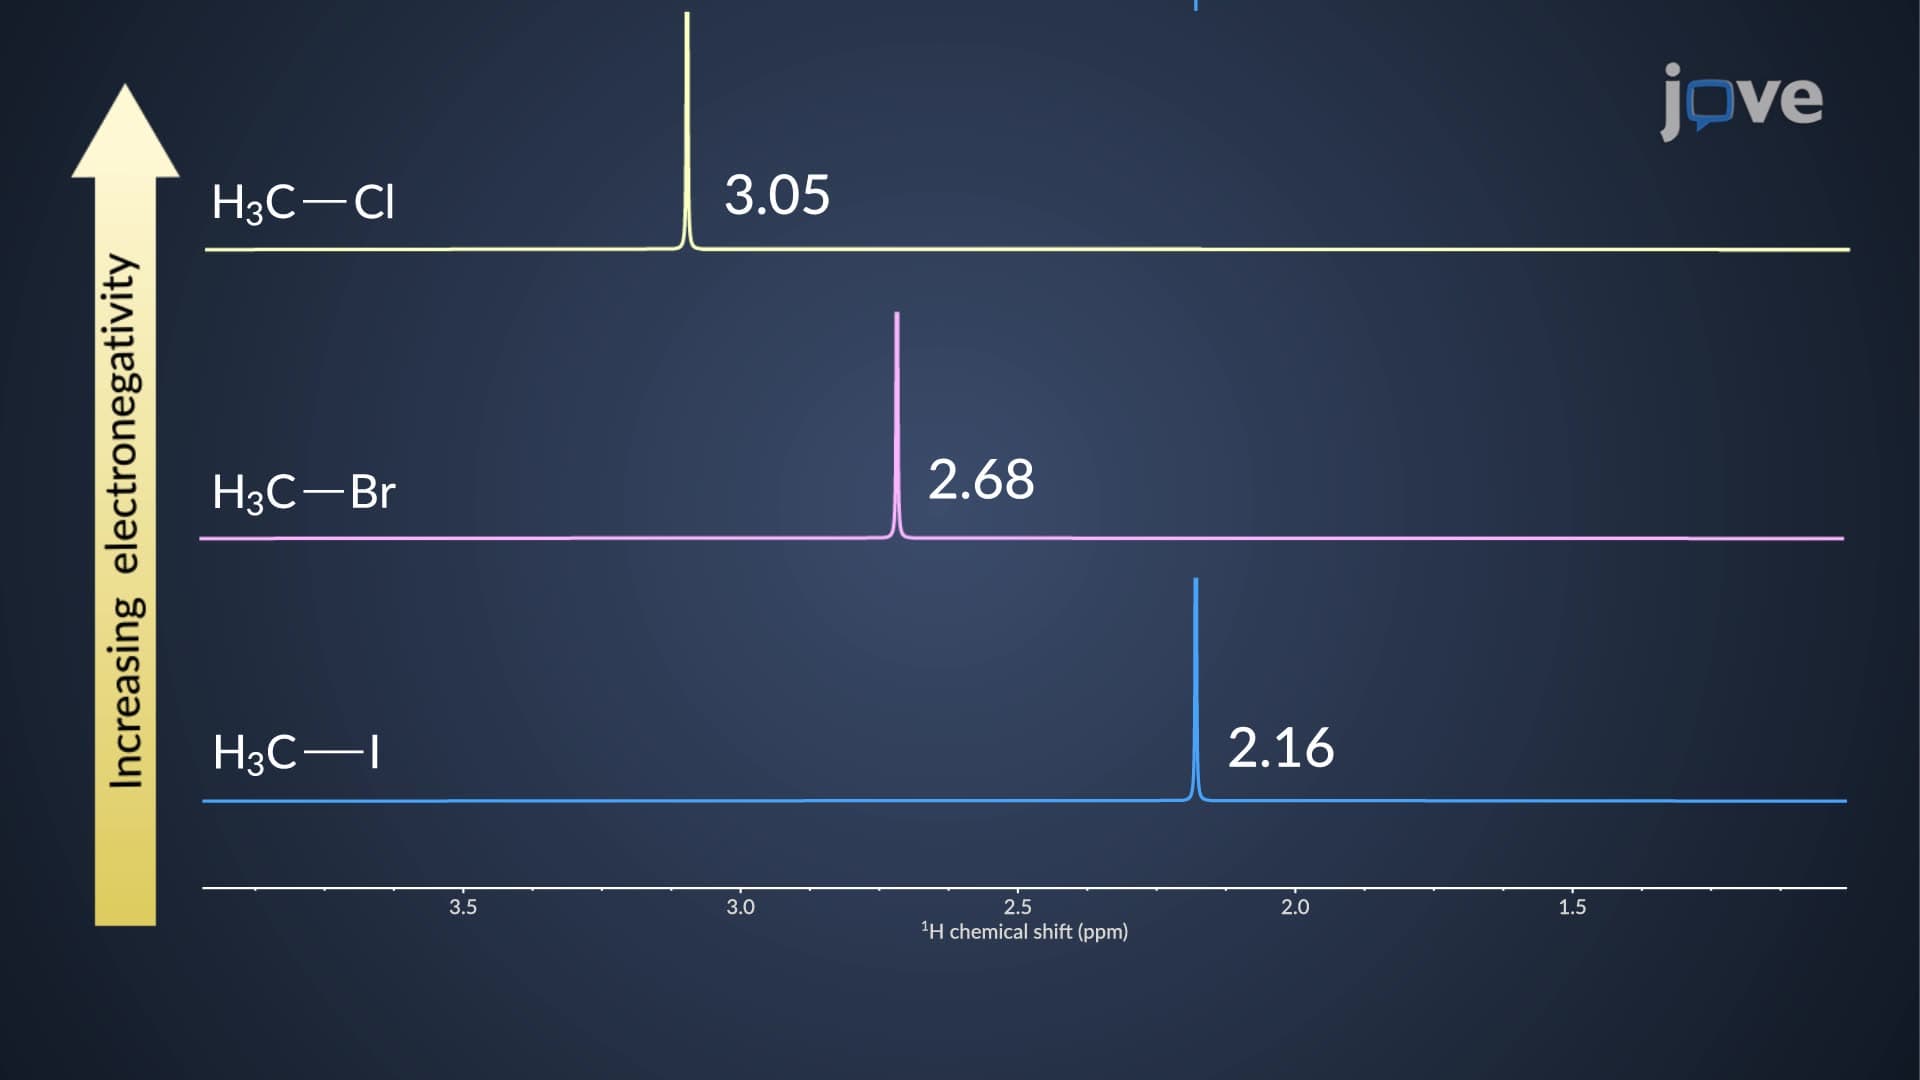
Inductive Effects on Chemical Shift: Overview
 03:51
Molecular Orbital Theory II
 02:35
Molecular Orbital Theory I