Related Experiment Video
Updated: Jun 28, 2026

Using Live Cell STED Imaging to Visualize Mitochondrial Inner Membrane Ultrastructure in Neuronal Cell Models
Published on: June 30, 2023
Mutant huntingtin and mitochondrial dysfunction
Ella Bossy-Wetzel1, Alejandra Petrilli, Andrew B Knott
1University of Central Florida, Burnett School of Biomedical Sciences, College of Medicine, 4000 Central Florida Boulevard, Orlando, FL 32816, USA. ebossywe@mail.ucf.edu

Insights
Huntington's disease (HD) involves a faulty huntingtin gene, leading to neurodegeneration. This study explores how mutant huntingtin impairs mitochondria, causing neuronal dysfunction and cell death in HD.
Area of Science:
- Neuroscience
- Genetics
- Cell Biology
Background:
- Huntington's disease (HD) is an inherited neurodegenerative disorder caused by a mutation in the huntingtin gene.
- Mitochondrial dysfunction is strongly implicated in HD pathogenesis.
Purpose of the Study:
- To elucidate the mechanisms by which mutant huntingtin (mtHtt) causes mitochondrial dysfunction in HD.
- To explore the role of mitochondrial impairment in the selective vulnerability of neurons in HD.
Main Methods:
- Review of existing research on mtHtt interactions with mitochondria.
- Analysis of proposed pathways linking mtHtt to mitochondrial dysfunction, including transcriptional effects and direct interactions.
- Examination of the impact on mitochondrial respiration, membrane potential, Ca(2+) buffering, and dynamics.
Main Results:
- Mutant huntingtin may disrupt mitochondrial function by affecting nuclear-encoded mitochondrial proteins or directly interacting with mitochondria.
- mtHtt can impair mitochondrial respiration, membrane potential, and calcium buffering.
- Defects in mitochondrial dynamics, trafficking, fission, and fusion are proposed mechanisms for mtHtt neurotoxicity.
Conclusions:
- Mitochondrial dysfunction is a central mechanism in Huntington's disease pathogenesis, driven by mutant huntingtin.
- mtHtt-induced bioenergetic failure and impaired mitochondrial dynamics contribute to neuronal dysfunction and cell death.
- Mitochondrial pathways may explain the selective vulnerability of specific neuronal populations, like medium spiny neurons, in HD.
Abstract:
Huntington's disease (HD) is a fatal, inherited neurodegenerative disorder that gradually robs affected individuals of memory, cognitive skills and normal movements. Although research has identified a single faulty gene, the huntingtin gene, as the cause of the disease, a cure remains elusive. Strong evidence indicates that mitochondrial impairment plays a key part in HD pathogenesis. Here, we highlight how mutant huntingtin (mtHtt) might cause mitochondrial dysfunction by either perturbing transcription of nuclear-encoded mitochondrial proteins or by direct interaction with the organelle and modulation of respiration, mitochondrial membrane potential and Ca(2+) buffering. In addition, we propose that mtHtt might convey its neurotoxicity by evoking defects in mitochondrial dynamics, organelle trafficking and fission and fusion, which, in turn, might result in bioenergetic failure and HD-linked neuronal dysfunction and cell death. Finally, we speculate how mitochondria might dictate selective vulnerability of long projection neurons, such as medium spiny neurons, which are particularly affected in HD.
Related Concept Videos
 01:21
Huntington Disease l: Introduction
 01:48
ATP Synthase: Mechanism
 01:19
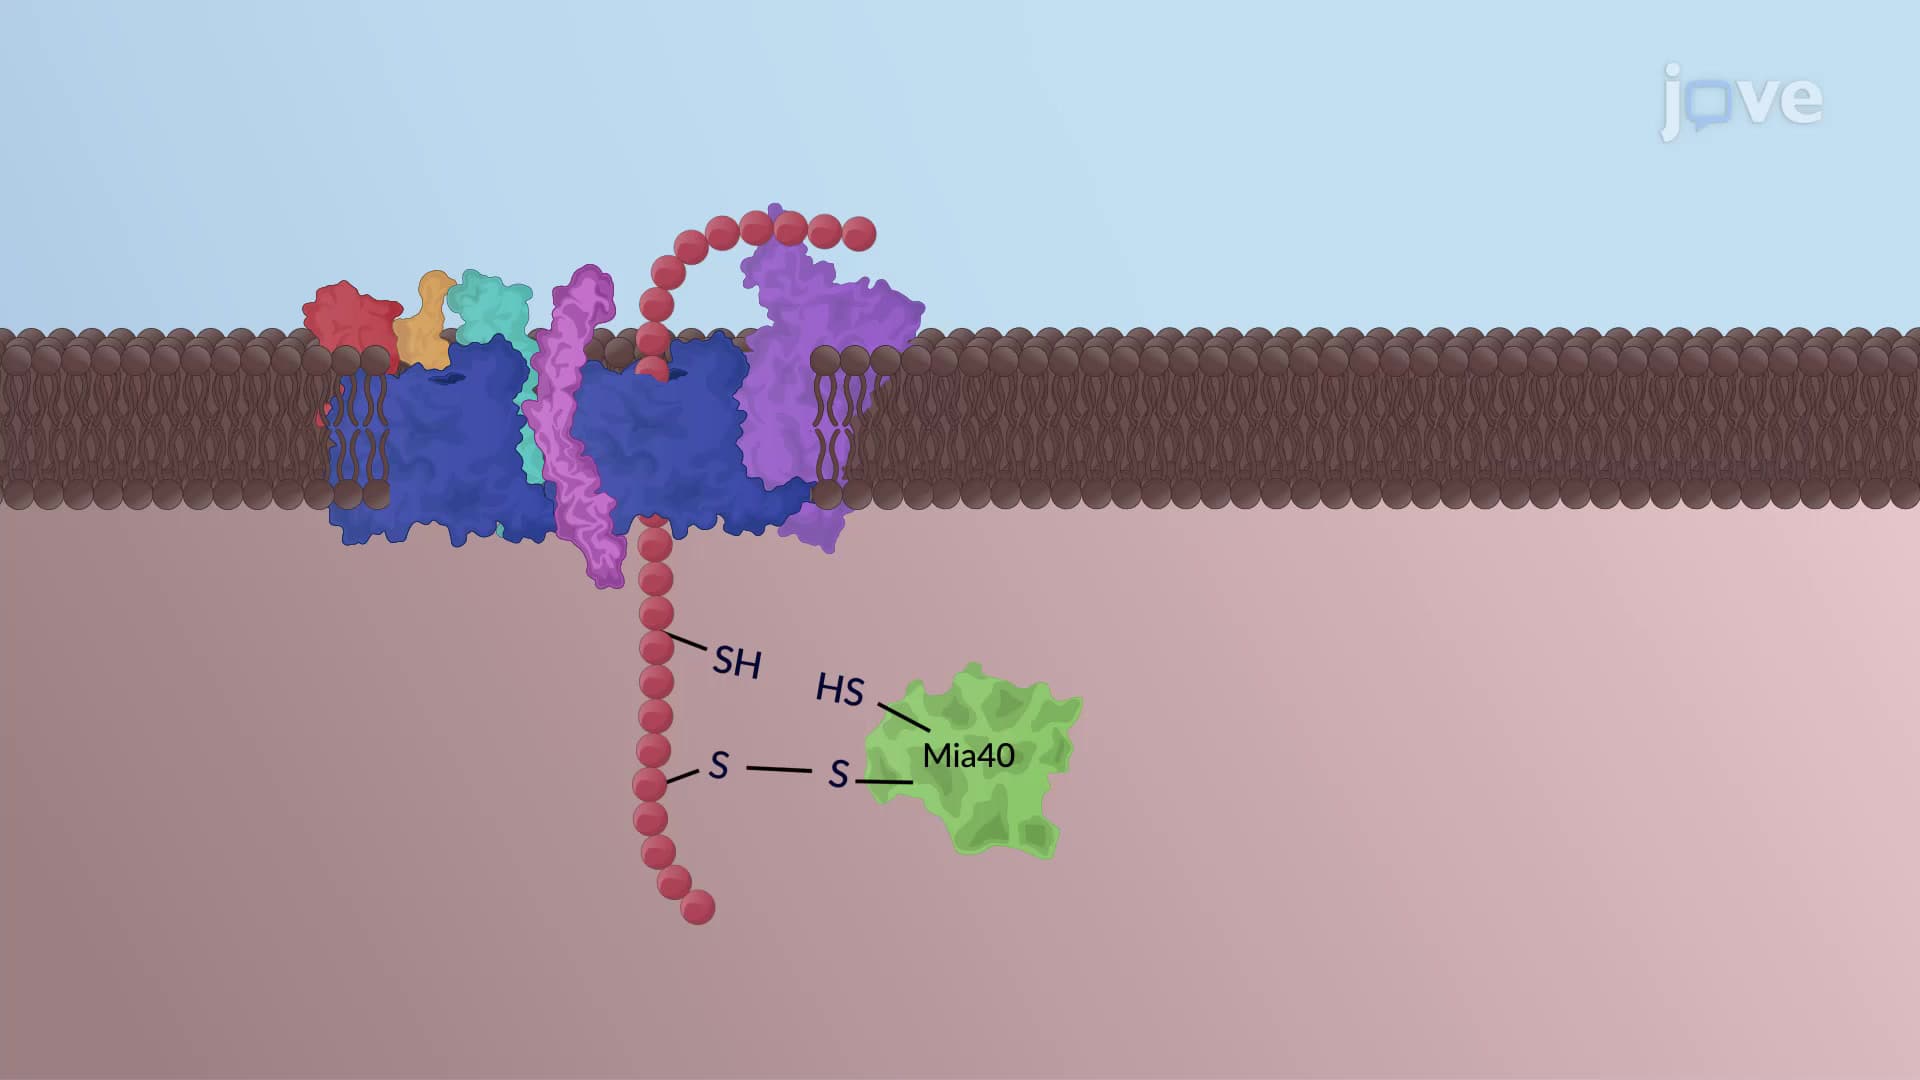
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...
 01:46
Electron Transport Chain: Complex I and II
ROS generation is regulated and maintained at moderate levels necessary...
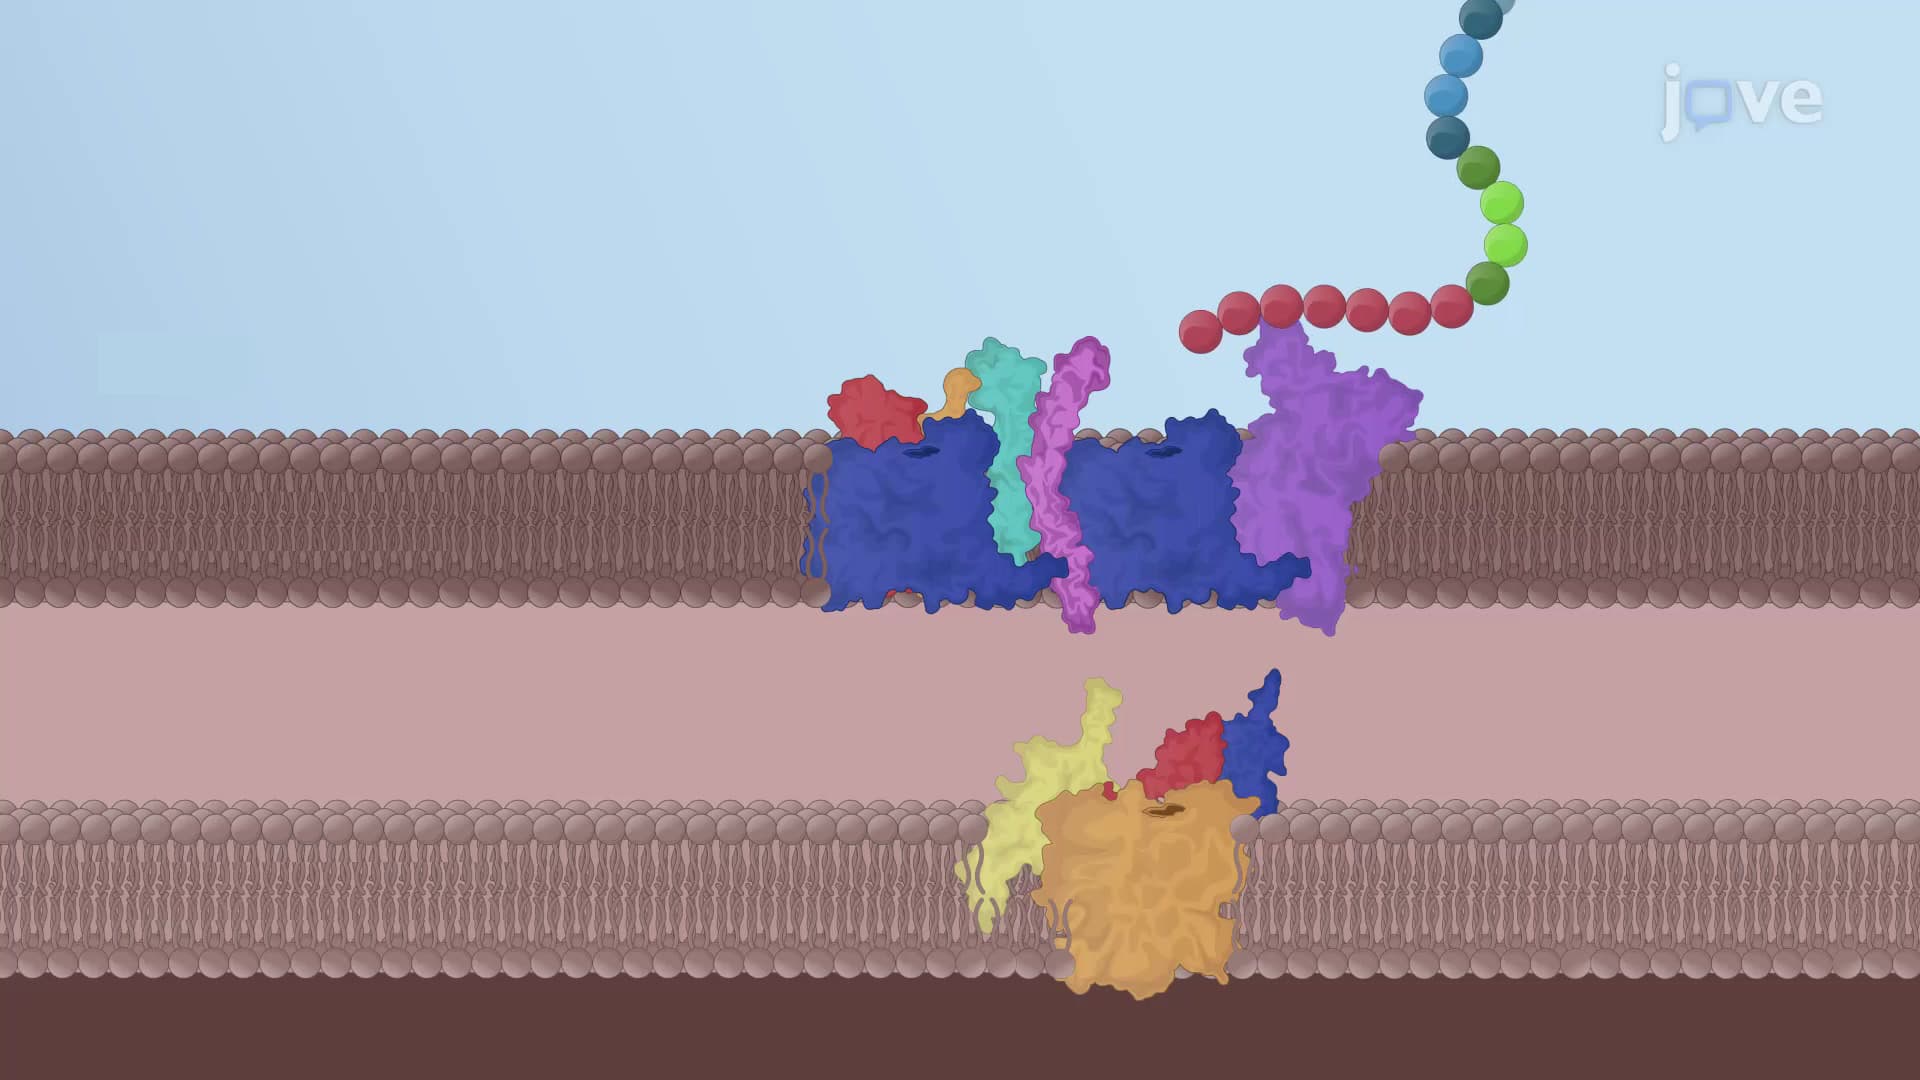 01:39
Mitochondrial Precursor Proteins
Most of the mitochondrial precursors...
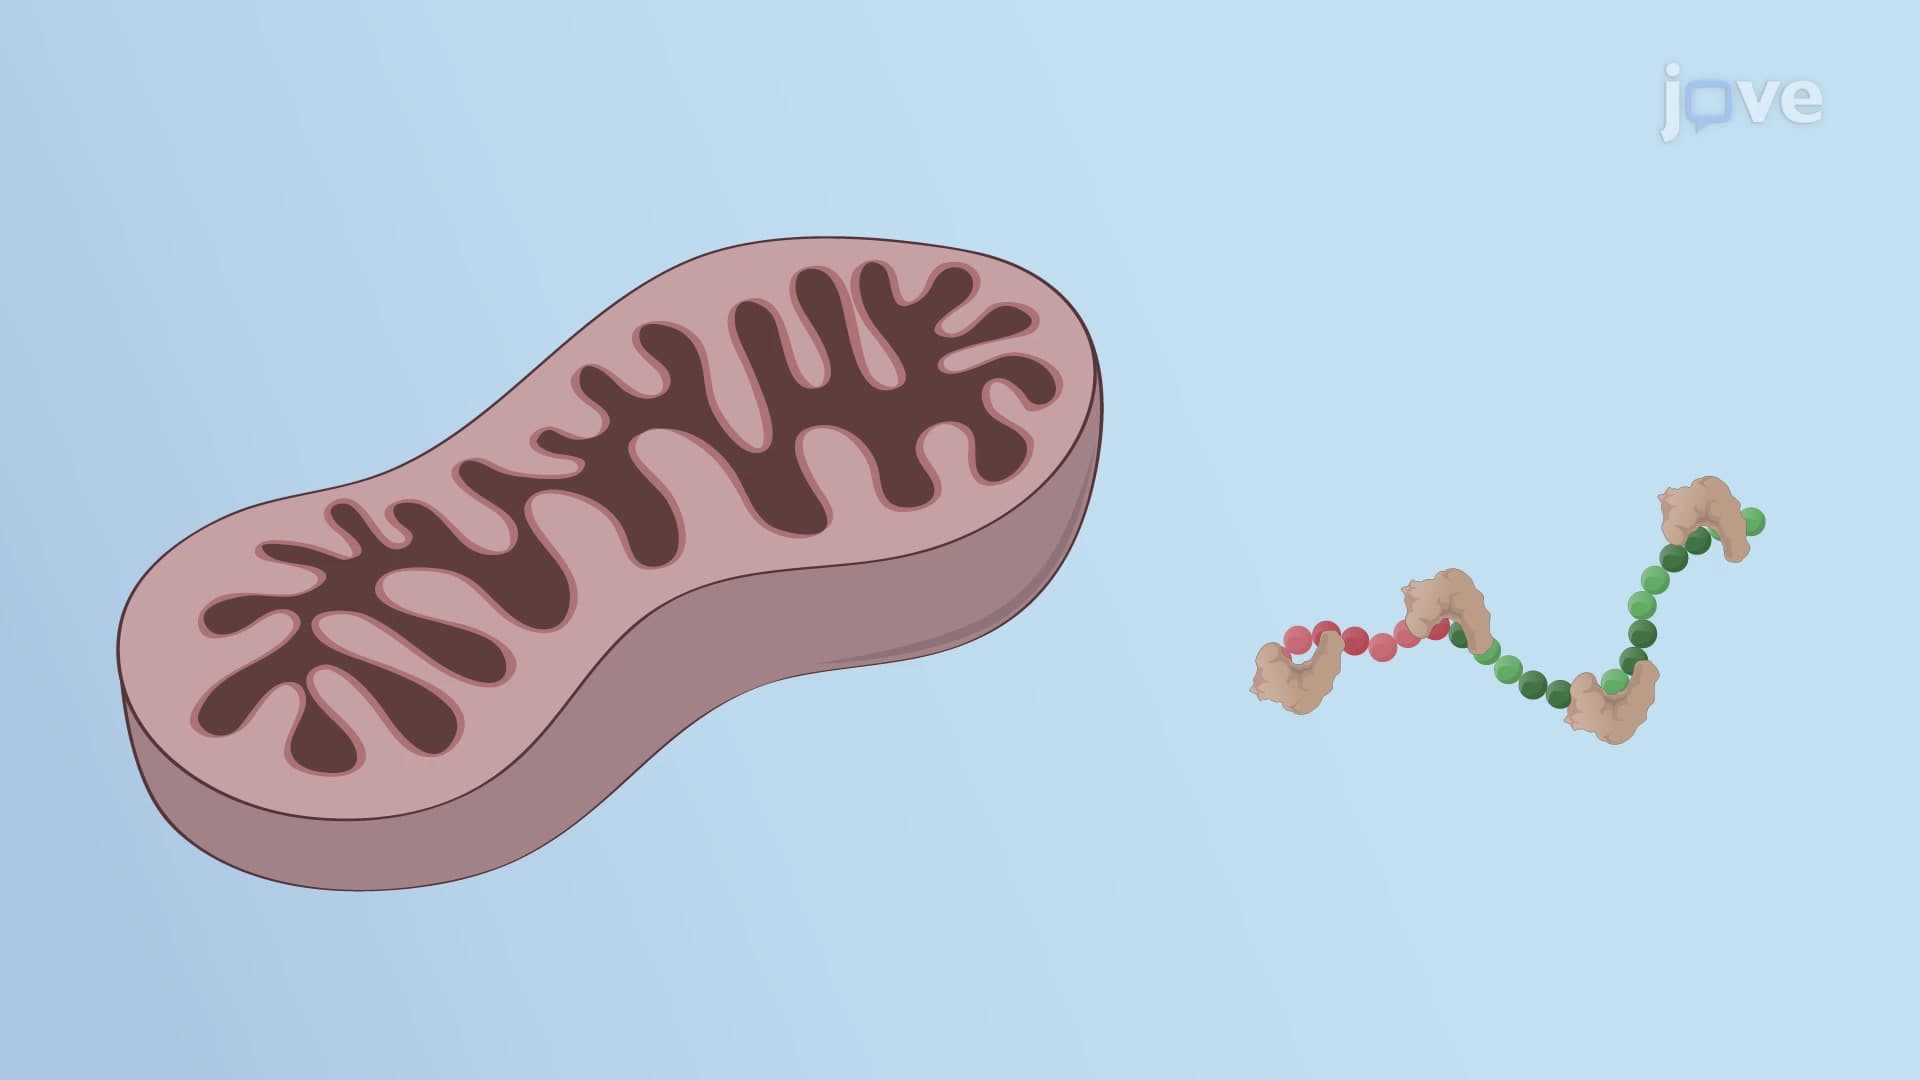 01:39
Mitochondrial Protein Sorting
Most of these mitochondrial proteins are encoded by the nucleus and imported to the mitochondria as unfolded or loosely folded precursors. Mitochondrial precursors...