Related Experiment Video
Updated: Feb 11, 2026

Biosensor-based High Throughput Biopanning and Bioinformatics Analysis Strategy for the Global Validation of Drug-protein Interactions
Published on: December 1, 2020
High-throughput analysis of drug binding interactions for the human cardiac channel, Kv1.5
Jerzy Karczewski1, Laszlo Kiss, Stefanie A Kane
1Department of Pain Research, Merck Research Laboratories, West Point, PA 19486, USA. jerzy_karczewski@merck.com

Abstract:
The voltage-gated potassium channel Kv1.5 is one of the key regulators of membrane potential repolarization in human atrial myocytes and is considered a potential drug target to treat atrial fibrillation. In this study we sought to determine molecular mechanism of action of DPO-1, a diphenylphosphine oxide derivative recently shown to terminate experimental atrial arrhythmia without affecting ventricular refractory period. In addition, we provided similar analysis for additional two small molecule blockers, representing different structural classes: cyclohexanones (PAC) and nor-triterpenoids (correolide). To rapidly identify the residues within the Kv1.5 channel critical for blocking activity of these molecules, two functional high-throughput ion channel assays were employed together with site-directed mutagenesis. Our study revealed that the residues critical for blocking activity of for DPO-1 include T480, localized at the outer mouth of the pore, and two residues along S6 helix: V505 and I508. The overlapping site was identified for PAC and included residues T480 and V505. In contrast to DPO-1, the I508A mutation resulted in only a modest reduction in the block of Kv1.5 by PAC (9-fold). Correolide, the largest molecule examined, made widespread interactions along the entire length of the pore (from T480 to V516). In summary, we have identified multiple residues involved in forming high affinity binding site for Kv1.5 blockers. Similar approaches of high-throughput ion channel technologies, combined with site-directed mutagenesis, may allow for parallel, rapid and accurate analysis of ion channel interactions with multiple compounds and could facilitate the design of more potent and selective ion channel blockers.
Related Concept Videos
 01:25
Pharmacokinetics: Drug–Drug Interactions
 01:23
Factors Affecting Protein-Drug Binding: Drug Interactions
Displacement interactions can have varying outcomes, ranging from toxicity to virtually...
 01:18
Factors Affecting Protein-Drug Binding: Drug-Related Factors
One crucial factor in drug-protein binding is the drug's lipophilicity or its affinity for fat. More lipophilic drugs tend to have higher binding extents. For example, highly lipophilic drugs like cloxacillin exhibit substantial protein binding, with as much as 95% of the drug binding to proteins. In...
 01:24
Tissue-Drug Binding: Localization of Drugs and its Significance
Drugs can bind to different tissue components, enhancing their distribution and localization. The factors influencing drug localization in tissues include the drug's lipophilicity, structural characteristics, tissue perfusion rate, and pH differences. These factors determine...
 01:21
Drug Distribution: Tissue Binding
For...
 01:30
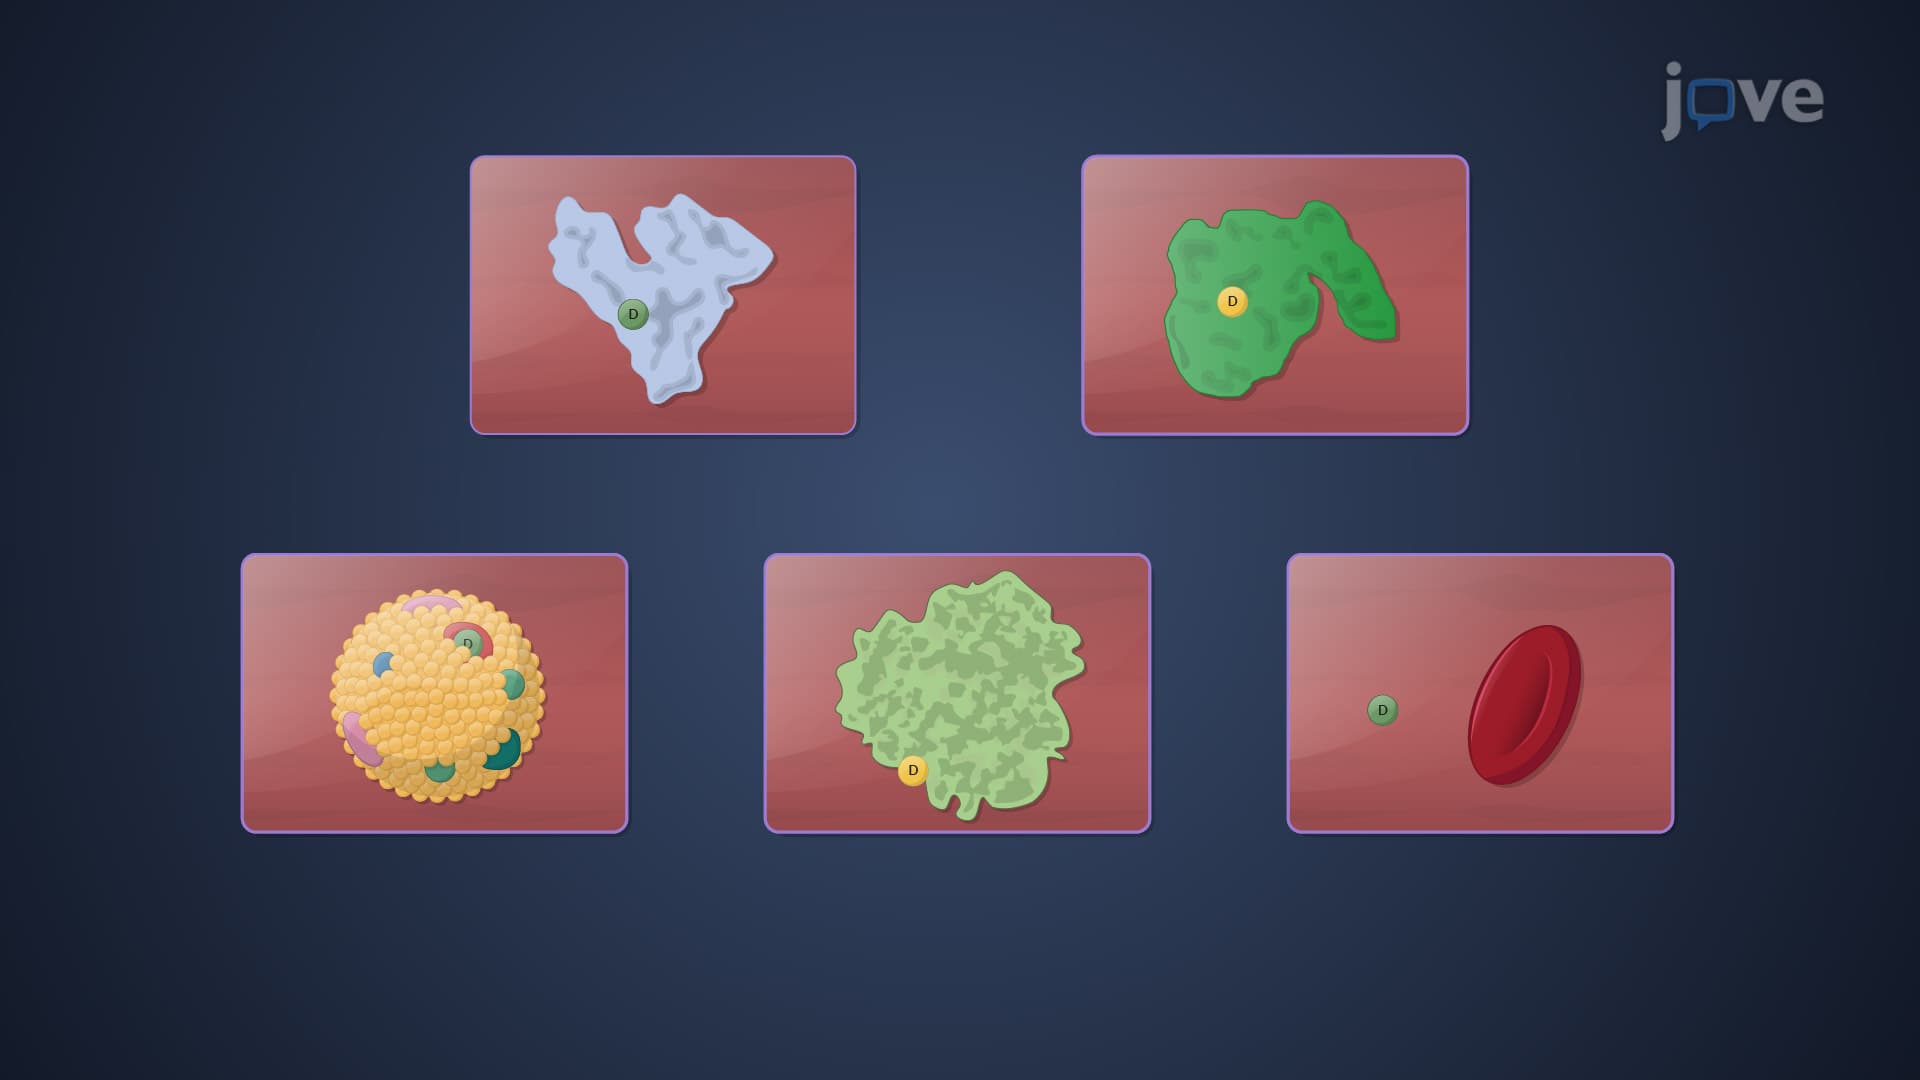
Drug Binding to Blood Components
HSA is the most abundant plasma protein and is vital in drug binding. It contains distinct drug-binding sites, with different drugs exhibiting affinity for specific sites. There are three main drug-binding domains for HSA: sites I, II, and III. These domains are...