Spectroscopic properties of polycyclic aromatic compounds Part 6. The nitromethane selective quenching rule visited

S Pandey1, W E Acree, B P Cho

  • 1Department of Chemistry, University of North Texas, Denton, Texas 76203-0068, USA.

Talanta
|March 1, 1997
PubMed

Related Concept Videos

meta-Directing Deactivators: –NO2, –CN, –CHO, –⁠CO2R, –COR, –CO2H01:13

meta-Directing Deactivators: –NO2, –CN, –CHO, –⁠CO2R, –COR, –CO2H

All meta-directing substituents are deactivating groups. These substituents withdraw electrons from the aromatic ring, making the ring less reactive toward electrophilic substitution. For example, the nitration of nitrobenzene is 100,000 times slower than that of benzene because of the deactivating effect of the nitro group. The first step in an electrophilic aromatic substitution is the addition of an electrophile to form a resonance-stabilized carbocation. The energy diagrams for the...
¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR01:15

¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR

The axial and equatorial protons in cyclohexane can be distinguished by performing a variable-temperature NMR experiment. In this process, except for one proton, the remaining eleven protons are replaced by deuterium. The deuterium substitution avoids the possible peak splitting caused by the spin-spin coupling between the adjacent protons. The remaining proton flips between the axial and equatorial positions.
Nucleophilic Aromatic Substitution of Aryldiazonium Salts: Aromatic SN101:14

Nucleophilic Aromatic Substitution of Aryldiazonium Salts: Aromatic SN1

Treating arylamines with nitrous acid gives aryldiazonium salts that are effective substrates in nucleophilic aromatic substitution reactions. The diazonio group in these salts can be easily displaced by different nucleophiles, yielding a wide variety of substituted benzenes. The leaving group departs as nitrogen gas, and this easy elimination is the driving force for the substitution reaction.
In the Sandmeyer reaction, for example, the diazonio group is replaced by a chloro, bromo, or cyano...
UV–Vis Spectroscopy: Woodward–Fieser Rules01:29

UV–Vis Spectroscopy: Woodward–Fieser Rules
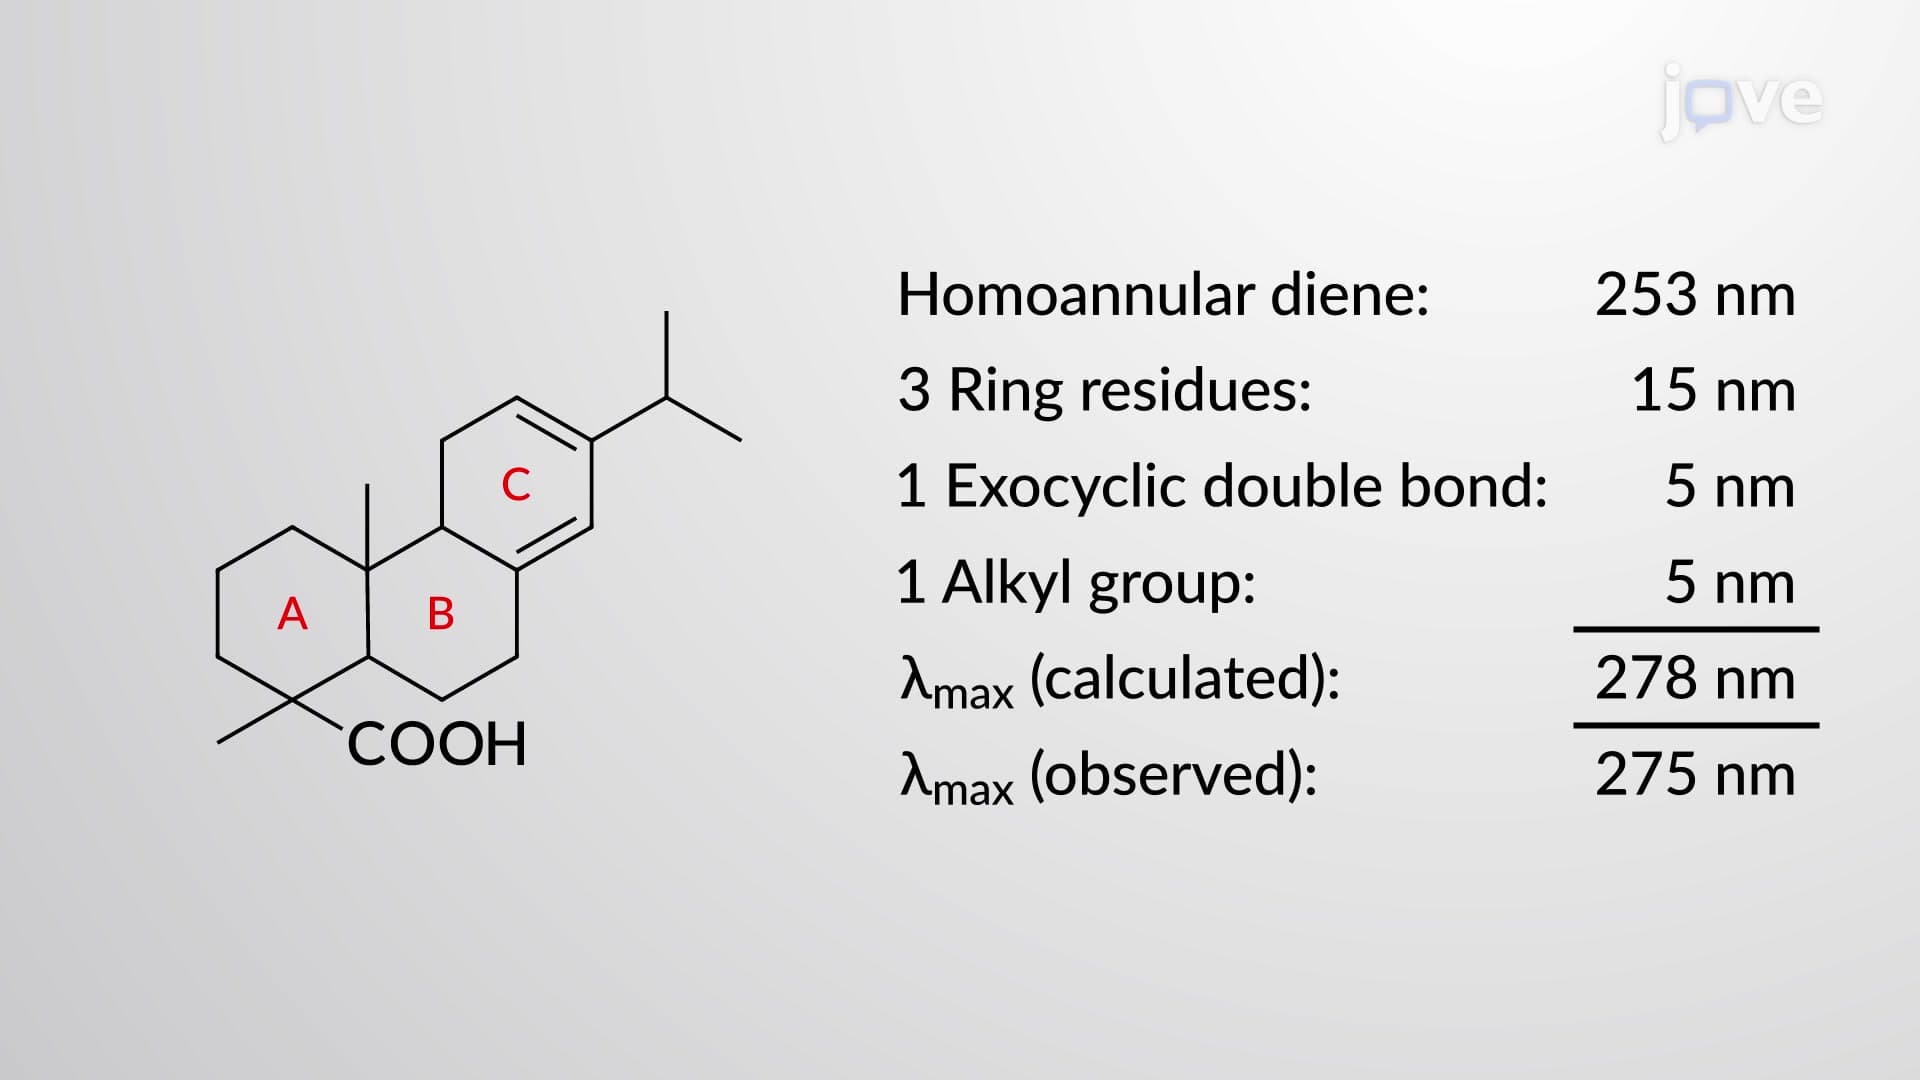
UV–Visible absorption spectra of conjugated dienes arise from the lowest energy π → π* transitions. The light-absorbing part of the molecule is called the chromophore, and the substituents directly attached to the chromophore are called auxochromes. A strong correlation exists between the absorption maxima, λmax, and the structure of a conjugated π system. The Woodward–Fieser rules predict the value of λmax for a given structure by adding the contributions...
NMR Spectroscopy of Aromatic Compounds01:14

NMR Spectroscopy of Aromatic Compounds

Aromatic compounds can be identified or analyzed using proton NMR and carbon‐13 NMR. Typically, aromatic hydrogens or hydrogens directly bonded to the aromatic rings are strongly deshielded by the aromatic ring current. Therefore, they absorb in the range of 6.5–8.0 ppm in proton NMR spectra. For instance, aromatic hydrogens directly bonded to the benzene ring absorb at 7.3 ppm. However, aromatic hydrogens of larger rings absorb farther upfield or downfield than the ideal range. Consider...
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds01:14

π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds

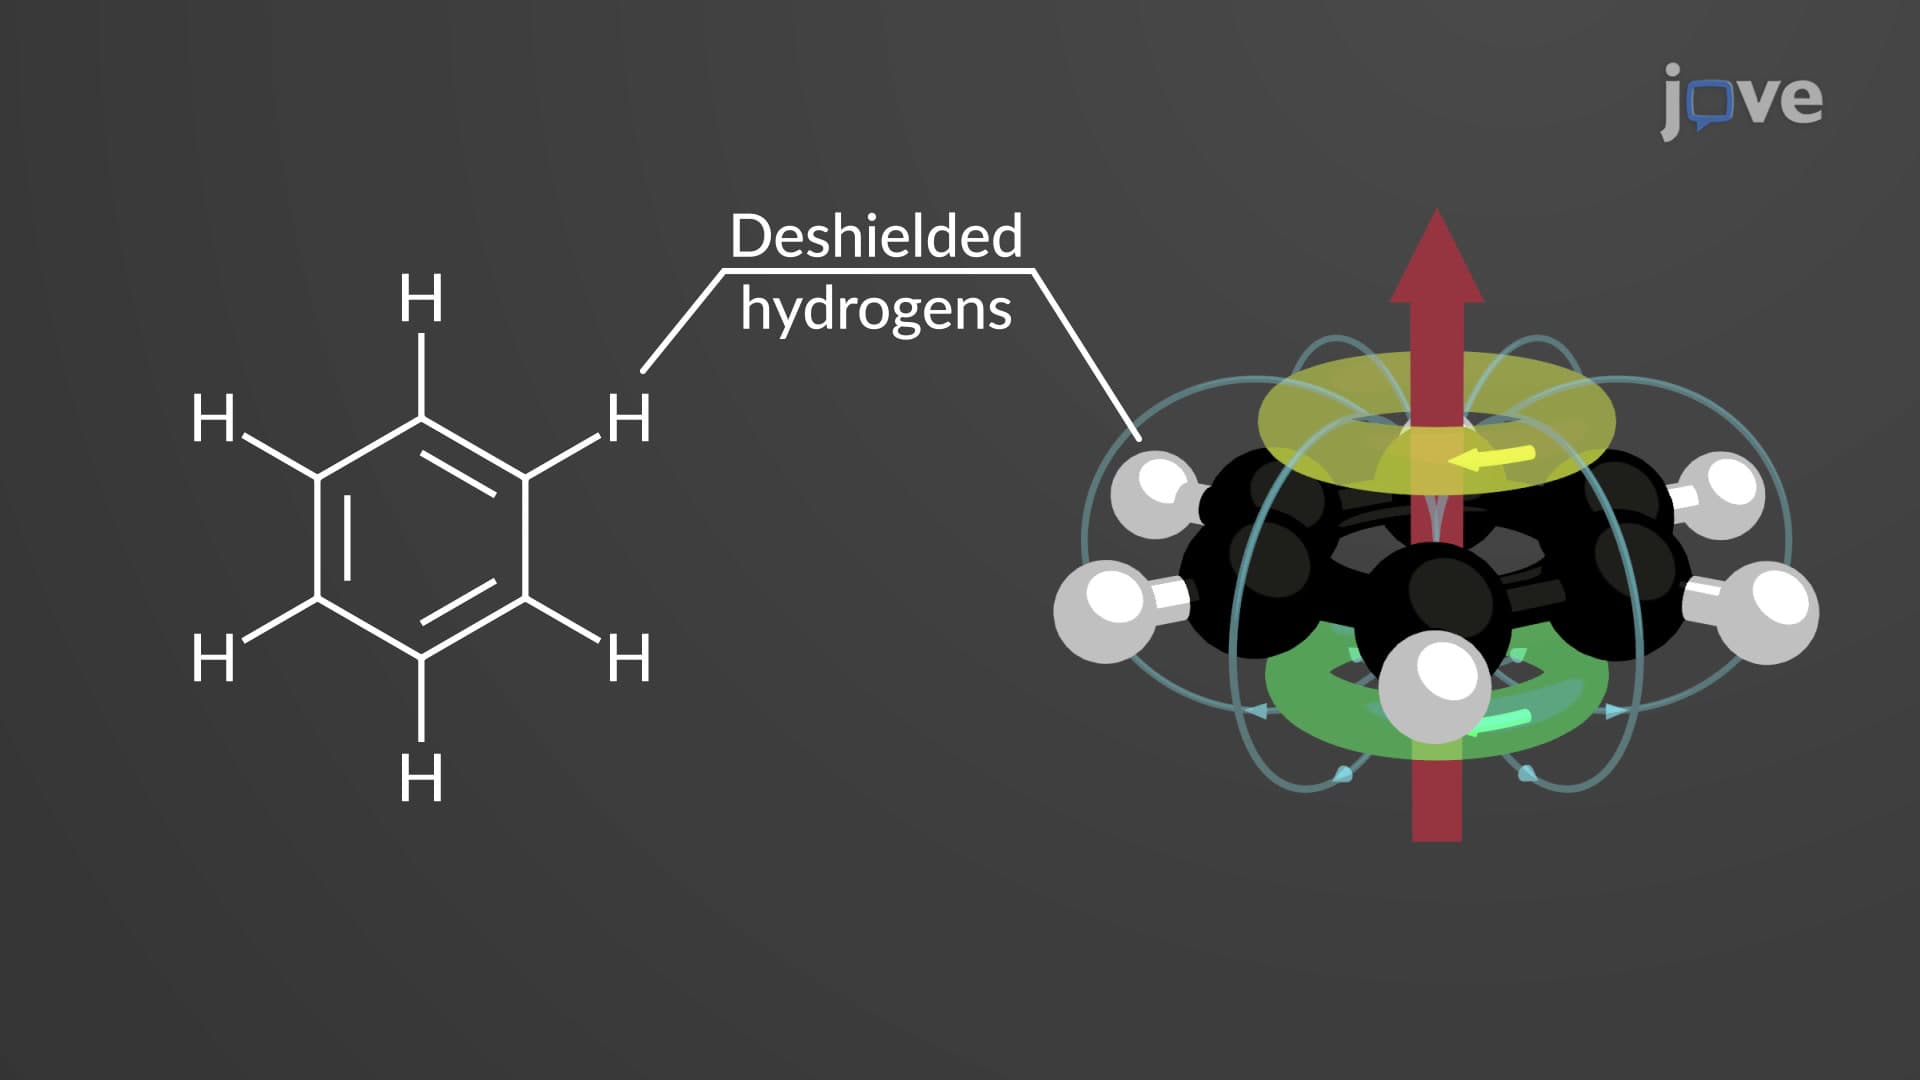
In aromatic compounds, such as benzene, the circulation of (4n + 2) π-electrons sets up a diamagnetic or diatropic ring current around the perimeter of the molecule. This current induces a magnetic field that opposes the external field inside the ring and reinforces it on the outside. The protons in benzene are deshielded and exhibit high chemical shifts in the range 6.5–8.5 ppm. The shielding effect at the center of the ring is evident in complex aromatic molecules, such as annulenes. In...