Related Experiment Video
Updated: Jun 28, 2026


Determining the Likelihood of Variant Pathogenicity Using Amino Acid-level Signal-to-Noise Analysis of Genetic Variation
Published on: January 16, 2019
Establishing an adjusted p-value threshold to control the family-wide type 1 error in genome wide association studies
Priya Duggal1, Elizabeth M Gillanders, Taura N Holmes
1Statistical Genetics Section, Inherited Disease Research Branch, National Human Genome Research Institute, National Institutes of Health, Baltimore, MD USA. pduggal@mail.nih.gov

Background:
By assaying hundreds of thousands of single nucleotide polymorphisms, genome wide association studies (GWAS) allow for a powerful, unbiased review of the entire genome to localize common genetic variants that influence health and disease. Although it is widely recognized that some correction for multiple testing is necessary, in order to control the family-wide Type 1 Error in genetic association studies, it is not clear which method to utilize. One simple approach is to perform a Bonferroni correction using all n single nucleotide polymorphisms (SNPs) across the genome; however this approach is highly conservative and would "overcorrect" for SNPs that are not truly independent. Many SNPs fall within regions of strong linkage disequilibrium (LD) ("blocks") and should not be considered "independent".
Results:
We proposed to approximate the number of "independent" SNPs by counting 1 SNP per LD block, plus all SNPs outside of blocks (interblock SNPs). We examined the effective number of independent SNPs for Genome Wide Association Study (GWAS) panels. In the CEPH Utah (CEU) population, by considering the interdependence of SNPs, we could reduce the total number of effective tests within the Affymetrix and Illumina SNP panels from 500,000 and 317,000 to 67,000 and 82,000 "independent" SNPs, respectively. For the Affymetrix 500 K and Illumina 317 K GWAS SNP panels we recommend using 10(-5), 10(-7) and 10(-8) and for the Phase II HapMap CEPH Utah and Yoruba populations we recommend using 10(-6), 10(-7) and 10(-9) as "suggestive", "significant" and "highly significant" p-value thresholds to properly control the family-wide Type 1 error.
Conclusion:
By approximating the effective number of independent SNPs across the genome we are able to 'correct' for a more accurate number of tests and therefore develop 'LD adjusted' Bonferroni corrected p-value thresholds that account for the interdepdendence of SNPs on well-utilized commercially available SNP "chips". These thresholds will serve as guides to researchers trying to decide which regions of the genome should be studied further.
Related Concept Videos
 01:13
Accuracy and Errors in Hypothesis Testing
In hypothesis testing, the probability of making a Type I error, denoted as α, is commonly set at 0.05. This significance level indicates a 5% chance...
 01:09
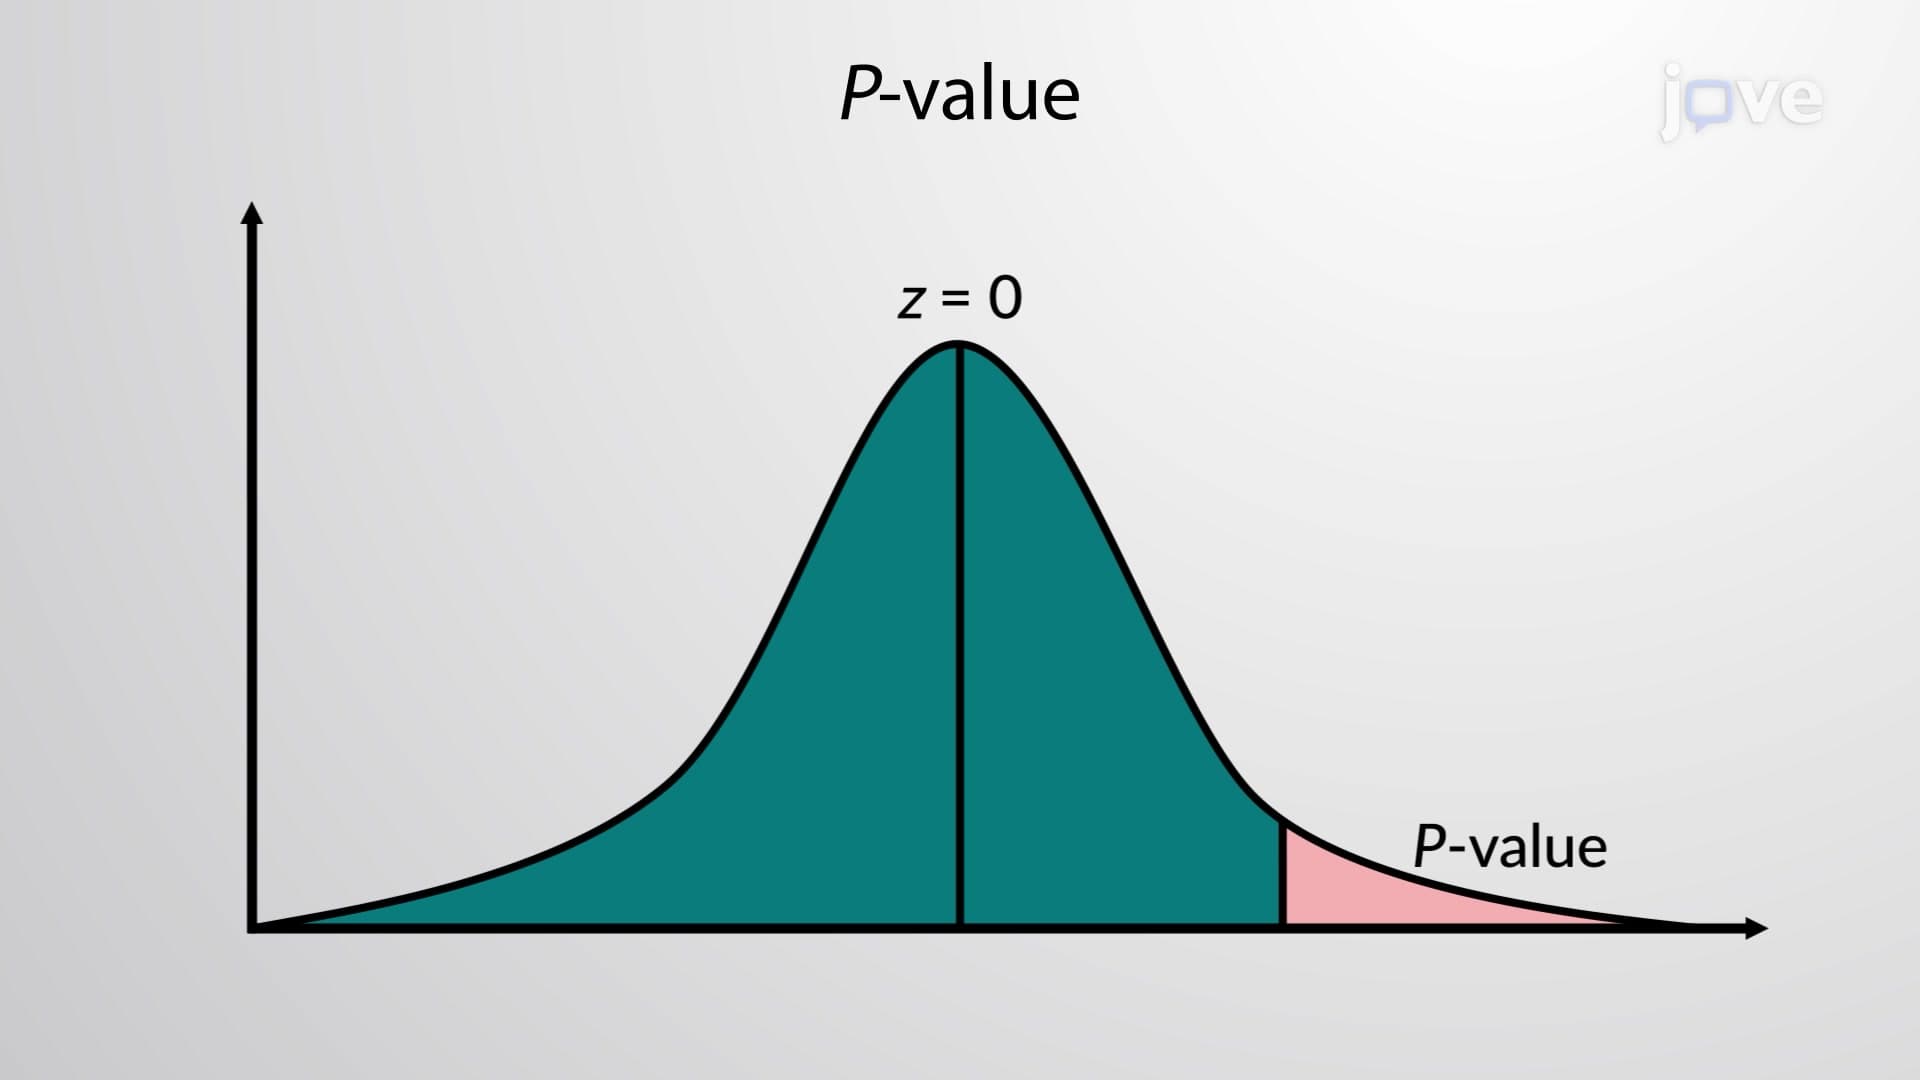
Decision Making: P-value Method
First, a specific claim about the population parameter is proposed. The claim is based on the research question and is stated in a simple form. Further, an opposing statement to the claim is also stated. These statements can act as null and alternative hypotheses: a null hypothesis would be a neutral statement while the alternative hypothesis can have a...
 01:11
Genome-wide Association Studies-GWAS
GWAS does not require the identification of the target gene involved in...
 01:10
Bonferroni Test
The means of different samples are first paired in all possible combinations.
The null hypothesis of the...
 01:14
Errors In Hypothesis Tests
 01:49
Hardy-Weinberg Principle