Related Experiment Video
Updated: Jun 28, 2026
Atomic Scale Structural Studies of Macromolecular Assemblies by Solid-state Nuclear Magnetic Resonance Spectroscopy
Published on: September 17, 2017
Vibrational assignments of six-coordinate ferrous heme nitrosyls: new insight from nuclear resonance vibrational
Florian Paulat1, Timothy C Berto, Serena DeBeer George
1Department of Chemistry, The University of Michigan, 930 N. University, Ann Arbor, Michigan 48109-1055, USA.

Abstract:
This Communication addresses a long-standing problem: the exact vibrational assignments of the low-energy modes of the Fe-N-O subunit in six-coordinate ferrous heme nitrosyl model complexes. This problem is addressed using nuclear resonance vibrational spectroscopy (NRVS) coupled to (15)N(18)O isotope labeling and detailed simulations of the obtained data. Two isotope-sensitive features are identified at 437 and 563 cm(-1). Normal coordinate analysis shows that the 437 cm(-1) mode corresponds to the Fe-NO stretch, whereas the 563 cm(-1) band is identified with the Fe-N-O bend. The relative NRVS intensities of these features determine the degree of vibrational mixing between the stretch and the bend. The implications of these results are discussed with respect to the trans effect of imidazole on the bound NO. In addition, a comparison to myoglobin-NO (Mb-NO) is made to determine the effect of the Mb active site pocket on the bound NO.
More Related Videos
Related Concept Videos
 01:08
IR Spectrum Peak Splitting: Symmetric vs Asymmetric Vibrations
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 01:02
¹H NMR: Interpreting Distorted and Overlapping Signals
As Δν decreases and the signals move closer, the doublets appear increasingly distorted. The intensities of the inner lines increase at the cost of those of the outer lines as the signals are slanted or...
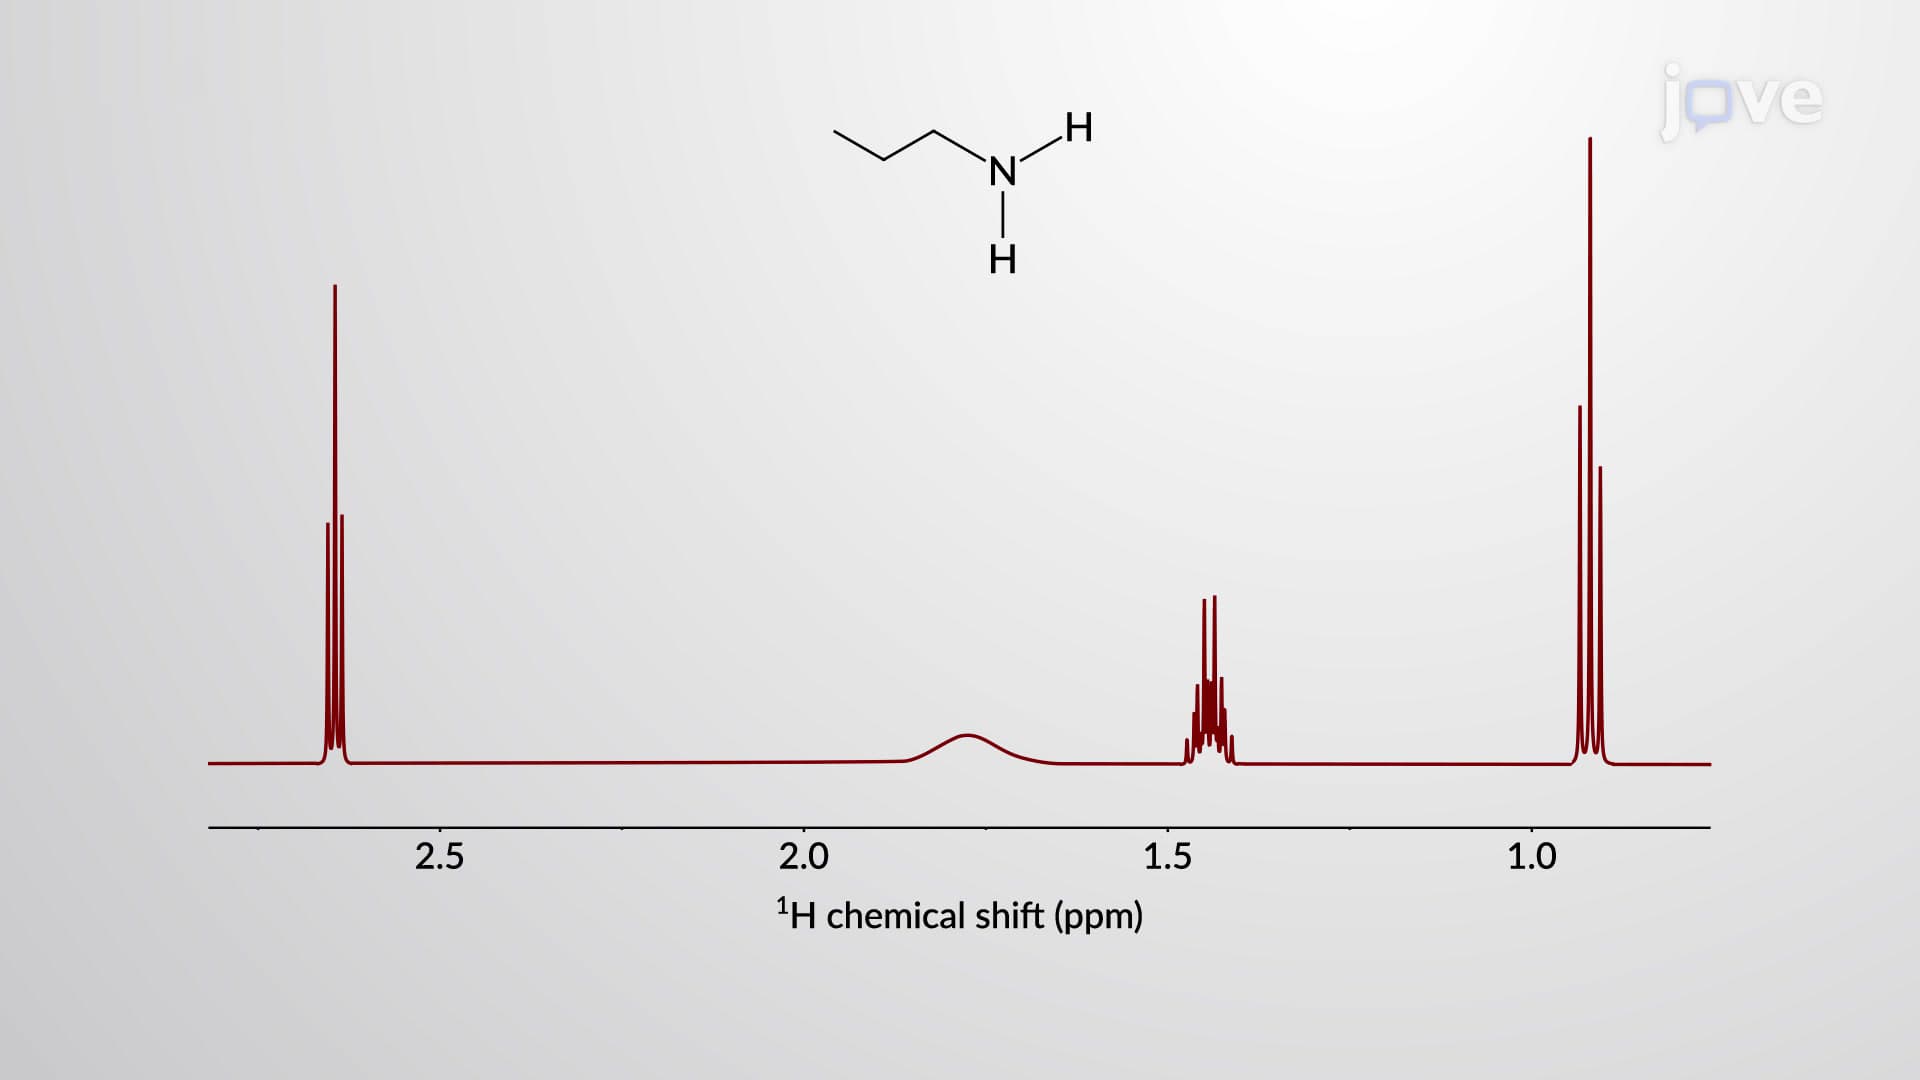 01:19
NMR Spectroscopy Of Amines
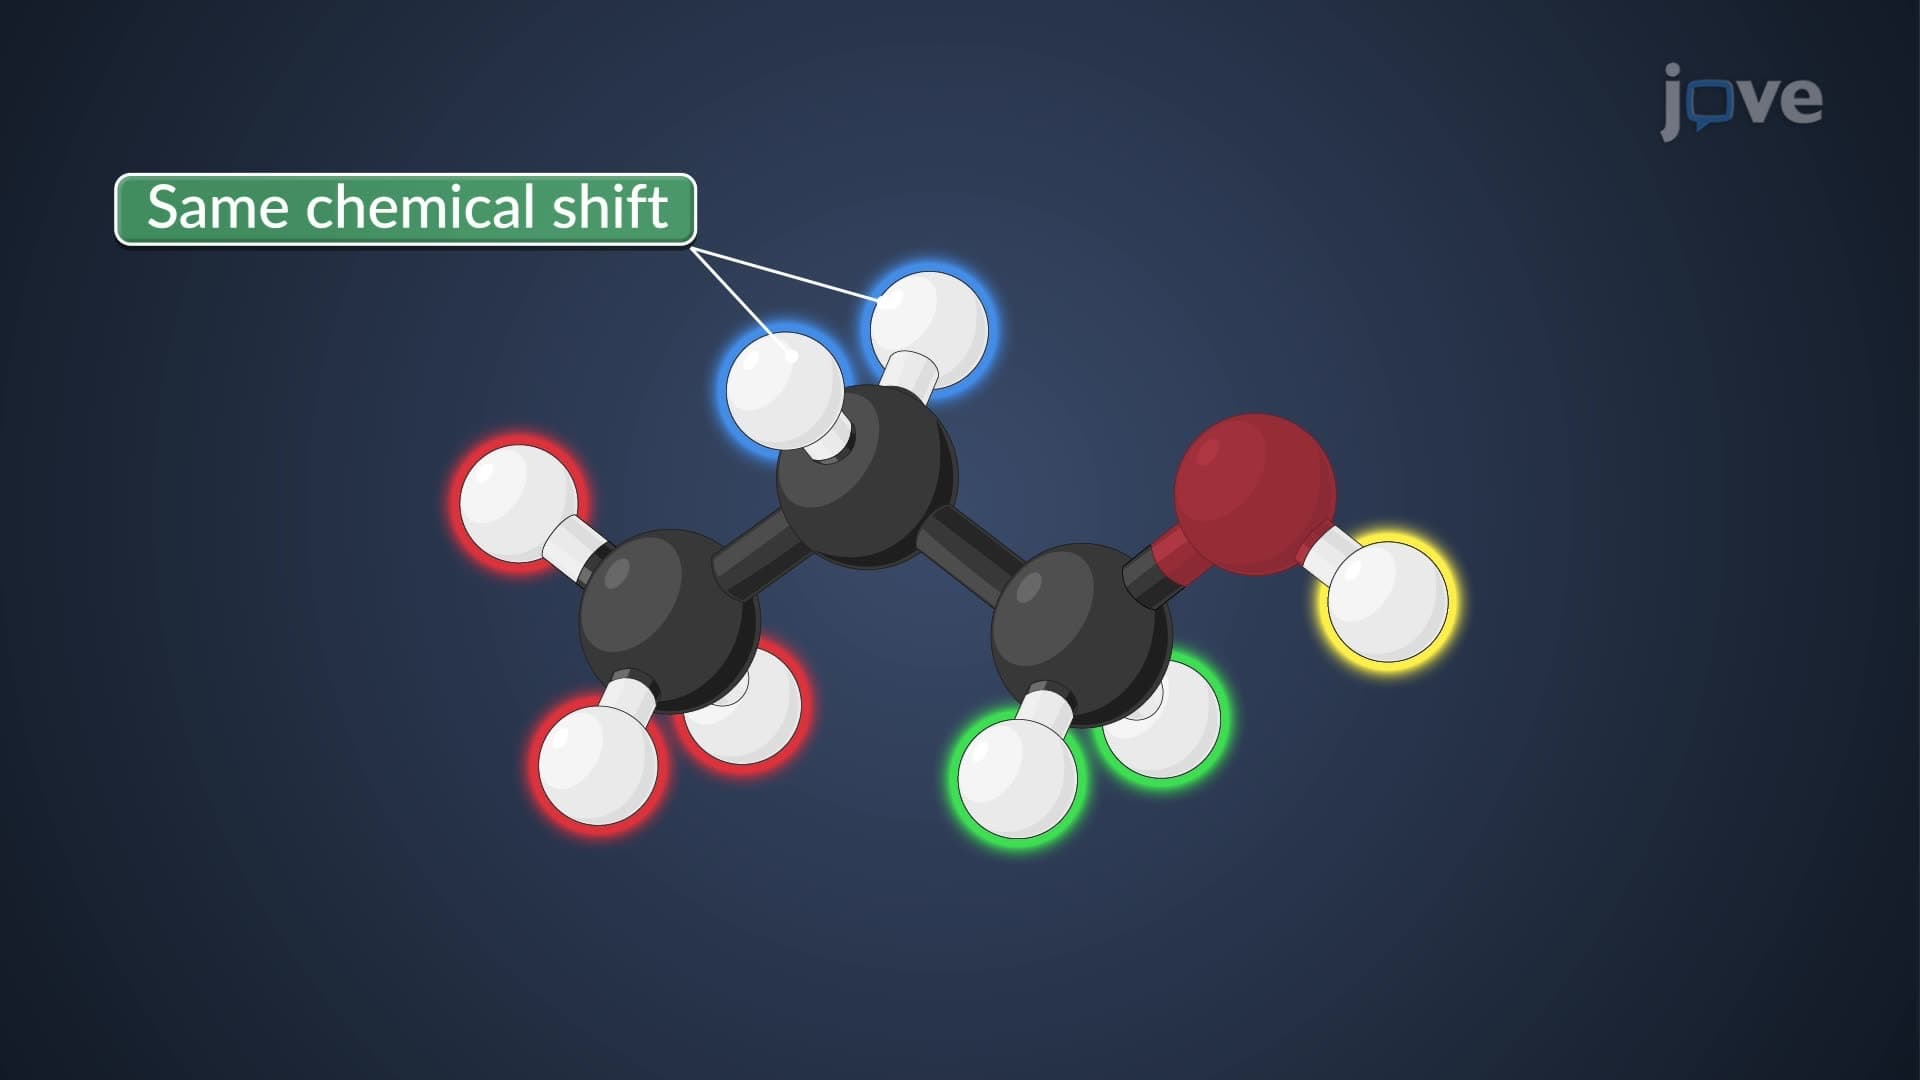 01:03
¹H NMR Chemical Shift Equivalence: Homotopic and Heterotopic Protons
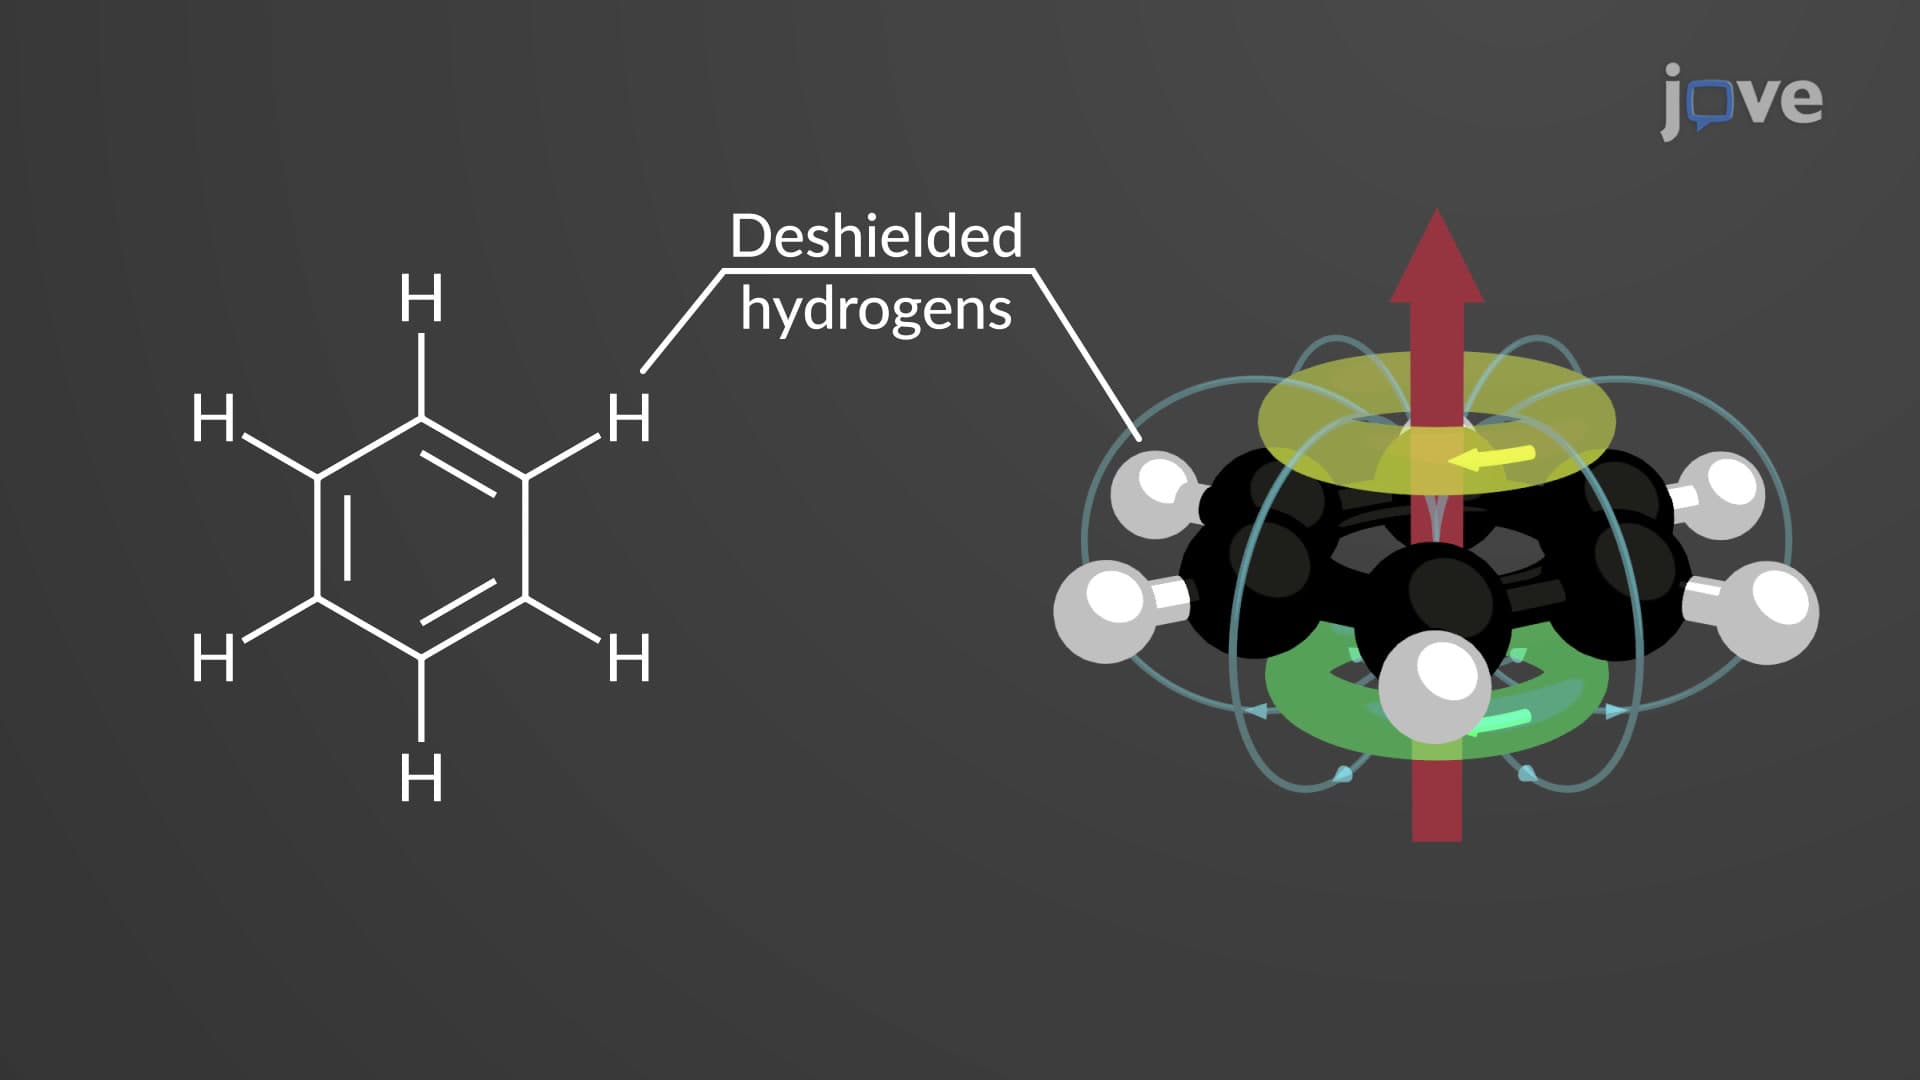 01:14
NMR Spectroscopy of Aromatic Compounds