Related Experiment Video
Updated: Jun 27, 2026
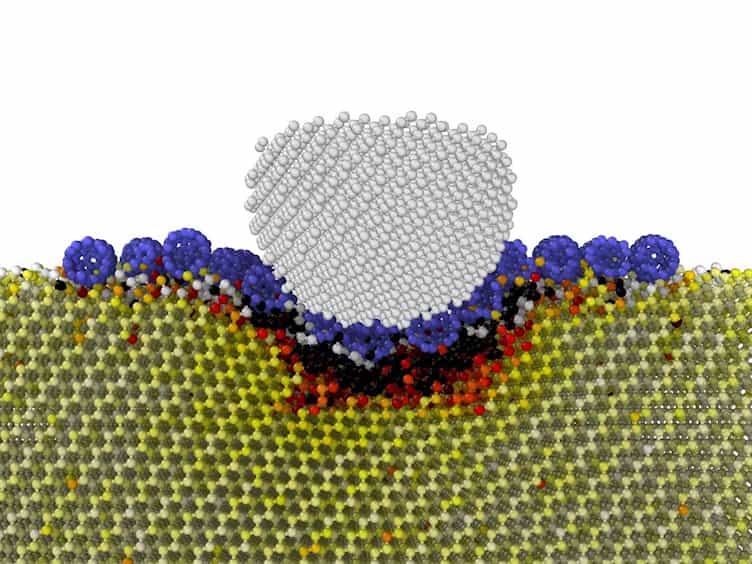

Probing C84-embedded Si Substrate Using Scanning Probe Microscopy and Molecular Dynamics
Published on: September 28, 2016
Dynamics and magnetic resonance properties of Sc3C2@C80 and its monoanion
Stefan Taubert1, Michal Straka, Teemu O Pennanen
1Department of Chemistry, (A.I. Virtanens plats 1), FIN-00014 University of Helsinki, Finland. stefan.taubert@helsinki.fi

Abstract:
We report density functional theory (DFT) studies on the endohedral scandium carbide fullerene Sc3C2@C80 and its monoanion [Sc3C2@C80](-). The system consisting of a Sc3C2 moiety inside the Ih C80 fullerene has been studied by using first principles molecular dynamics simulations at the DFT level. On the picosecond time scale, the triangle defined by the scandium atoms is seen to jump between orientations along the equatorial six-membered ring belt of the cage. The confined carbide unit, in turn, is engaged in a flipping motion through the Sc3 plane. In contrast to the equilibrium geometry optimisations using large basis sets that predict a trigonal bipyramidal structure, a planar Sc3C2-moiety is preferred during the finite-temperature simulation. In the molecular dynamics picture, Sc3C2@C80 is best described as an equilibrium between the two static minimum structures. Calculations of the vibrational frequencies show that the earlier predicted C2 and C2v symmetric isomers are in fact saddle points, with one imaginary normal mode frequency that is related to the flipping motion of the confined carbon dimer. Reoptimisation revealed two new minimum energy structures where the C2 unit is tilted with respect to its orientation in the earlier suggested higher-symmetry structures. The nature of the bonding in the static structures of the two isomers of Sc3C2@C80 has been investigated using the electron localisation function and natural population analysis. Some increased electron pair localisation is detected on the six-membered rings closest to the Sc atoms. 13C nuclear magnetic resonance (NMR) chemical shifts have been calculated for the closed-shell monoanion of Sc3C2@C80. The 13C shifts were also calculated for Sc2C2@C84, for further comparison to experimentally measured spectra. The confined carbon atoms are strongly deshielded in these metallofullerenes, implying an incorrect earlier interpretation of the experimental 13C NMR spectrum of Sc2C2@C84. The neutral Sc3C2@C80 system with one unpaired electron is further characterised by calculating the hyperfine coupling constants, the g tensor, as well as paramagnetic NMR (pNMR) 13C shifts for both static isomers. The chemical shifts of the confined carbon atoms and the hyperfine coupling constants of all the confined atoms are strongly dependent on the conformation of the Sc3C2 moiety. Consequently, dynamical effects are expected to be important in the modelling of the magnetic properties of endohedral scandium carbide fullerenes. The two low-lying isomers have rather different pNMR 13C shifts, implying the potential of this method in structural assignment.
Related Concept Videos
 02:42
Valence Bond Theory
 03:02
Colors and Magnetism
When atoms or molecules absorb light at the proper frequency, their electrons are excited to higher-energy orbitals. For many main group atoms and molecules, the absorbed photons are in the ultraviolet range of the electromagnetic spectrum, which cannot be detected by the human eye. For coordination compounds, the energy difference between the d orbitals often allows photons in the visible range to be absorbed and emitted, which is seen as colors by the human eye.
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:42
Ionic Crystal Structures
Most monatomic ions behave as charged spheres, and their attraction for ions of opposite charge is the same in every direction. Consequently, stable structures for ionic compounds result (1) when ions of one charge are surrounded by as many ions as possible of the opposite...
 02:48
Ionic Bonding and Electron Transfer