Related Experiment Video
Updated: Jun 27, 2026

Using Microarrays to Interrogate Microenvironmental Impact on Cellular Phenotypes in Cancer
Published on: May 21, 2019
Use of normalization methods for analysis of microarrays containing a high degree of gene effects
Terri T Ni1, William J Lemon, Yu Shyr
1Division of Cardiovascular Medicine, Department of Medicine, Vanderbilt University School of Medicine, Nashville, TN 37232, USA. terri.ni@vanderbilt.edu

Background:
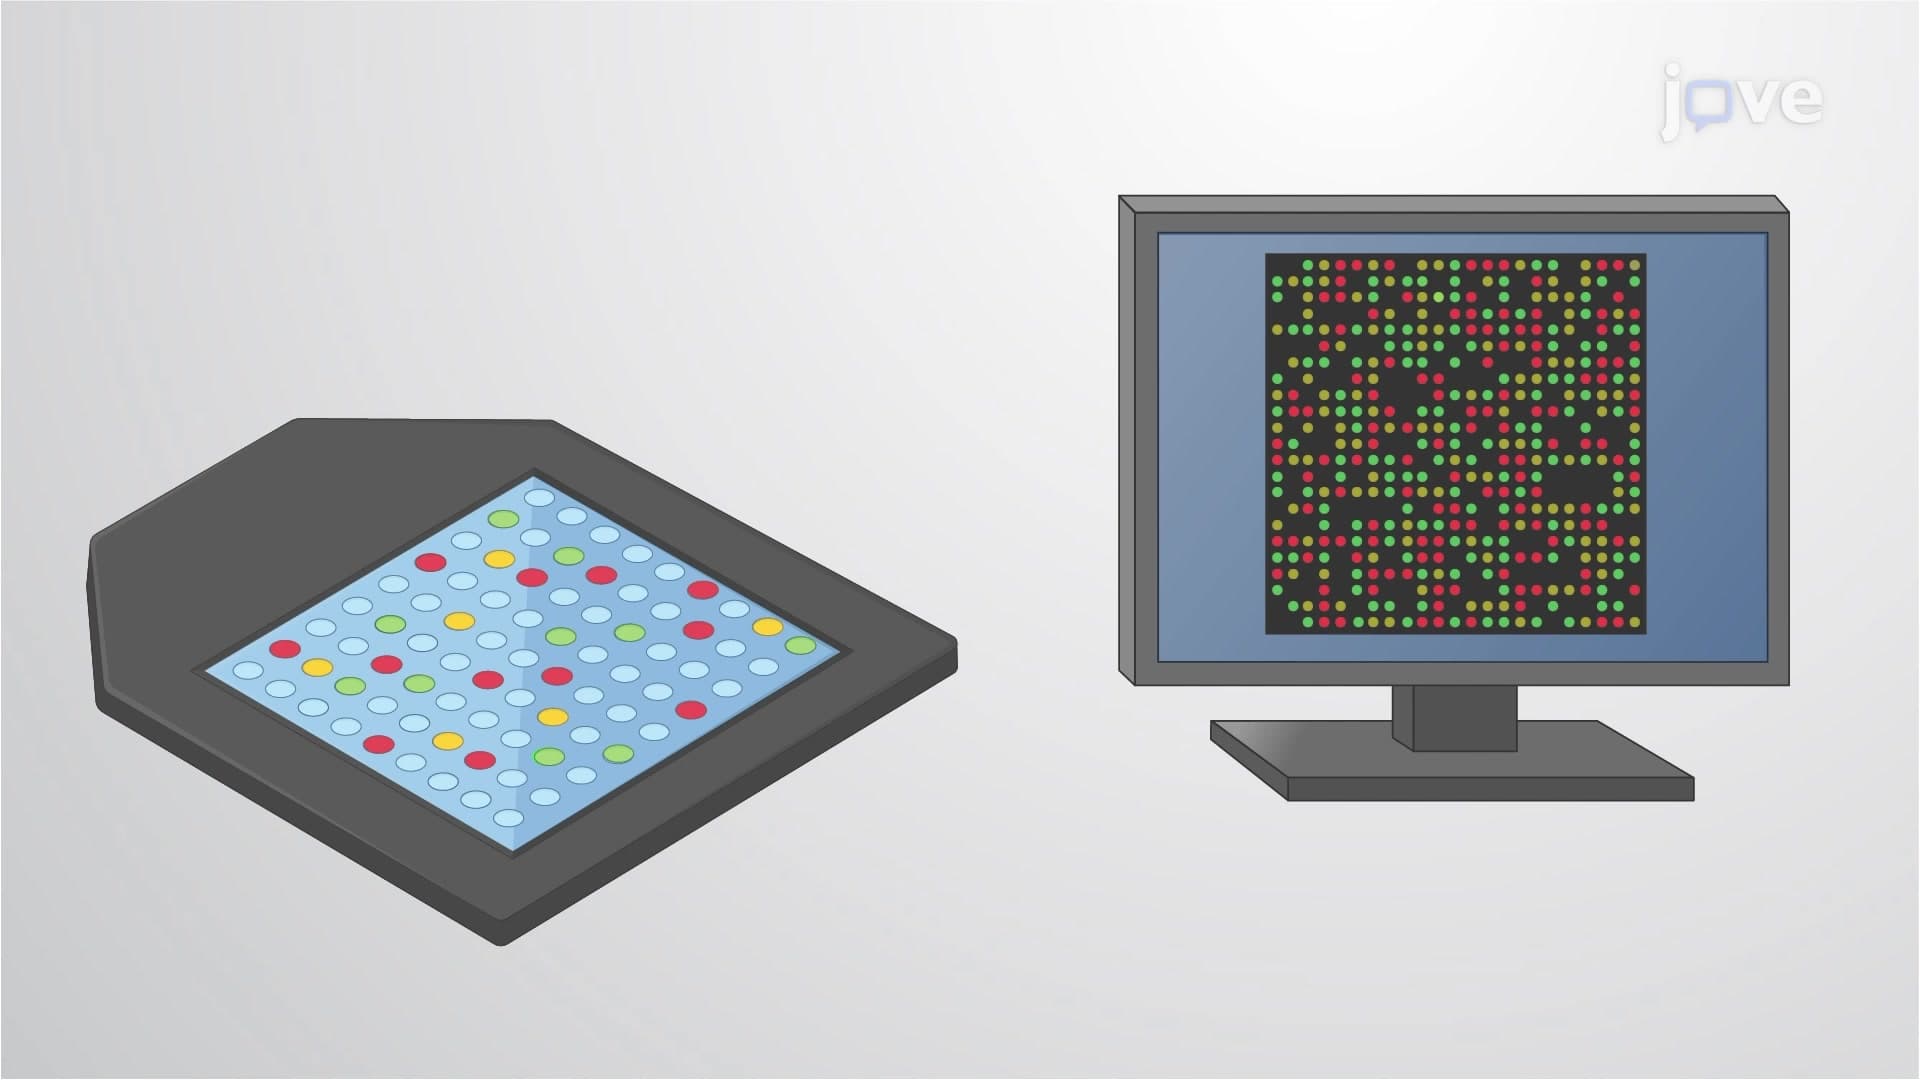
High-throughput microarrays are widely used to study gene expression across tissues and developmental stages. Analysis of gene expression data is challenging in these experiments due to the presence of significant percentages of differentially expressed genes (DEG) observed between tissues and developmental stages. Data normalization methods that are widely used today are not designed for data with a large proportion of tissue or gene effects.
Results:
In our current study, we describe a novel two-dimensional nonparametric normalization method for analyzing microarray data which functions well in the absence or presence of large numbers of gene effects. Rather than relying on an assumption of low variability among most genes, the method implements a unique peak selection strategy to distinguish DEG from genes that are invariant in expression, prior to nonlinear curve fitting. We compared the method under simulated and experimental conditions with five alternative nonlinear normalization approaches: quantile, lowess, robust lowess, invariant set, and cross-correlation (Xcorr). Simulations included various percentages of simulated DEG and the experimental data used is from publicly available datasets known to be difficult to analyze due to the presence of approximately 34% DEG.
Conclusion:
We have demonstrated that the new method provides considerable improvement in the accuracy of data normalization when large proportions of gene effects are present. The performance improvement is mostly attributed to its variable selection component, which is designed to separate expression invariant genes from DEG. Adding this key component of the new method to alternative normalization approaches rescues the most of the sensitivity of these methods to gene effects. The results indicate that our method may be used without prior knowledge of or assumptions about housekeeping genes to normalize microarrays that are quite different.
More Related Videos
 09:58
Mapping the Structure-Function Relationships of Disordered Oncogenic Transcription Factors Using Transcriptomic Analysis
Published on: June 27, 2020
 08:27
Large-Scale Multi-Omics Genome-Wide Association Studies (Mo-GWAS): Guidelines for Sample Preparation and Normalization
Published on: July 27, 2021