

Theoretical studies of potential energy surface and rotational spectra of Xe-H(2)O van der Waals complex
1State Key Laboratory of Magnetic Resonance and Atomic and Molecular Physics, Wuhan Institute of Physics and Mathematics, Chinese Academy of Sciences, Wuhan 430071, People's Republic of China.

Abstract:
In this work we report an ab initio intermolecular potential energy surface and theoretical spectroscopic studies for Xe-H(2)O complex. The ab initio energies are calculated with CCSD(T) method and large basis sets (aug-cc-pVQZ for H and O and aug-cc-pVQZ-PP for Xe) augmented by a {3s3p2d2f1g} set of bond functions. This potential energy surface has a global minimum corresponding to a planar and nearly linear hydrogen bonded configuration with a well depth of 192.5 cm(-1) at intermolecular distance of 4.0 A, which is consistent with the previous determined potential by Wen and Jager [J. Phys. Chem. A 110, 7560 (2006)]. The bound state calculations have been performed for the complex by approximating the water molecule as a rigid rotor. The theoretical rotational transition frequencies, isotopic shifts, nuclear quadrupole coupling constants, and structure parameters are in good agreement with the experimental observed values. The wavefunctions are analyzed to understand the dynamics of the ground and the first excited states.

Related Concept Videos
 01:24
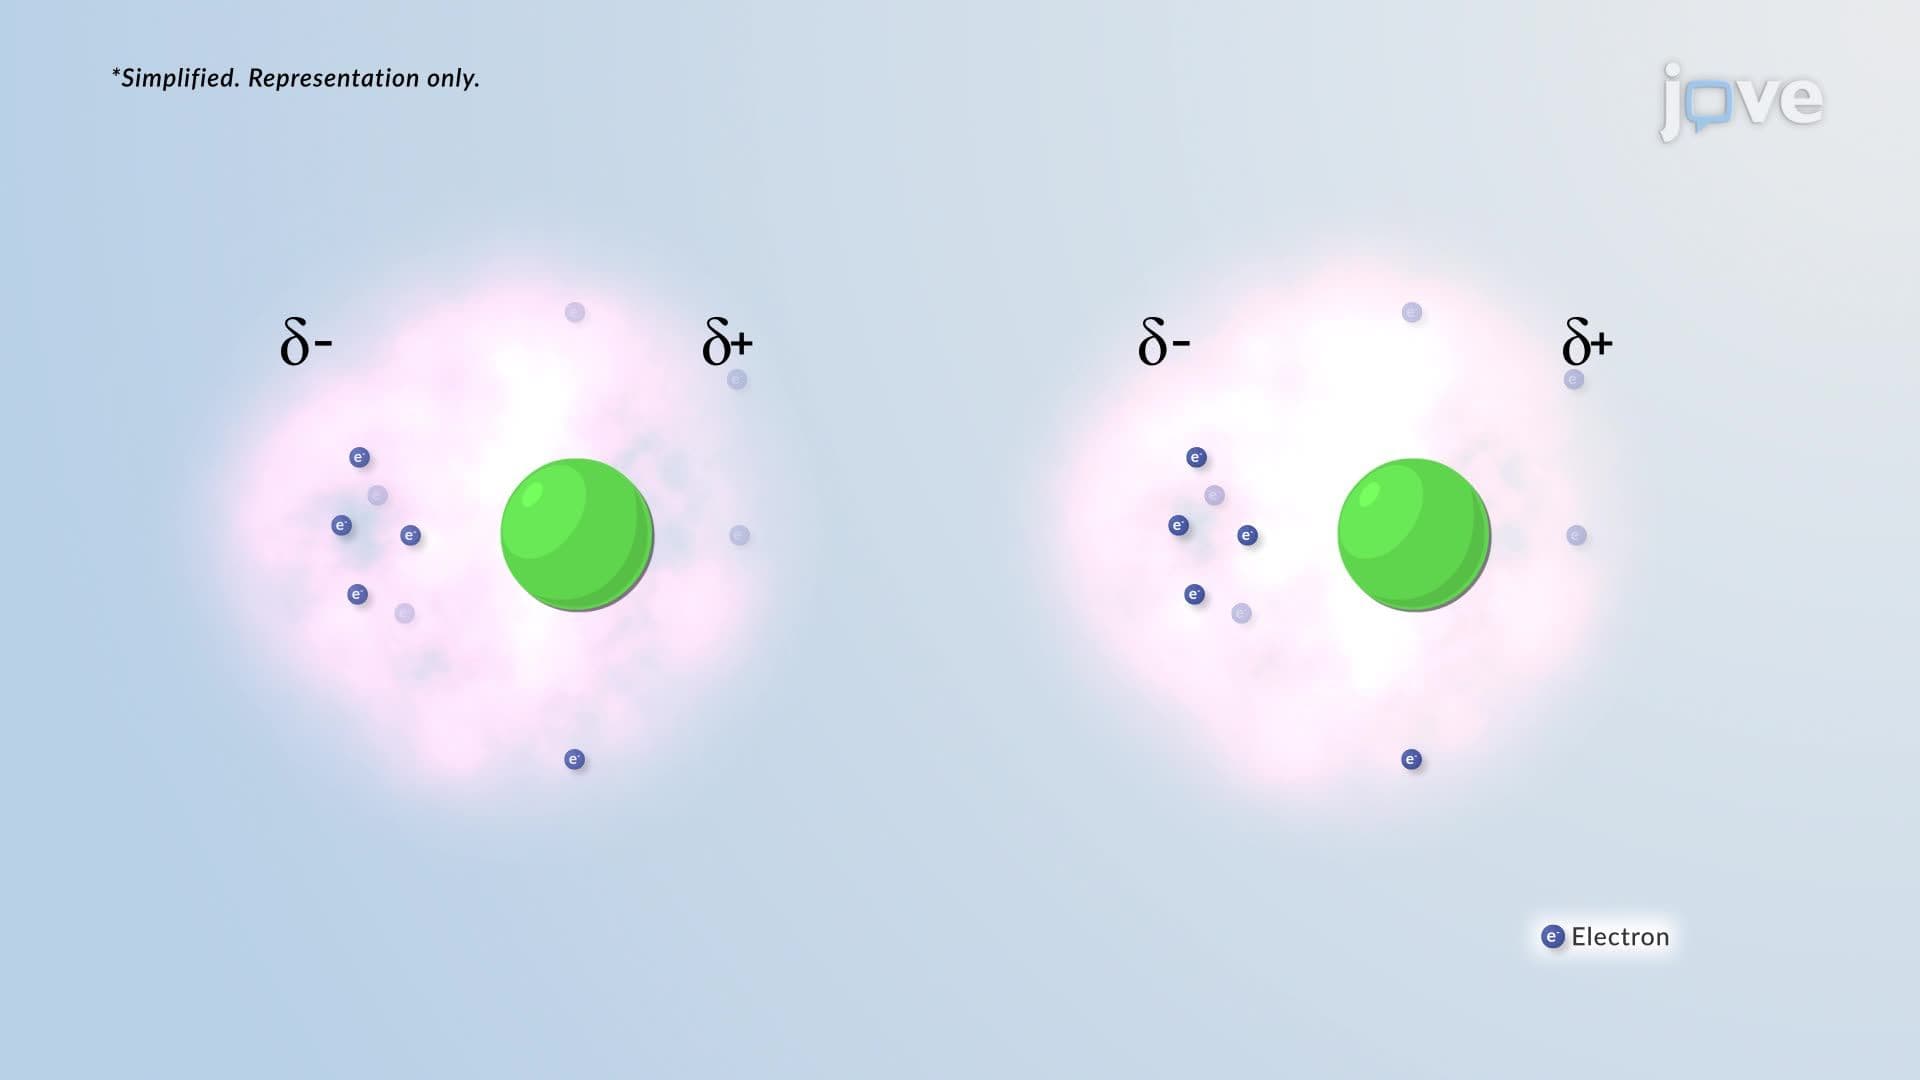
Van der Waals Interactions
 01:10
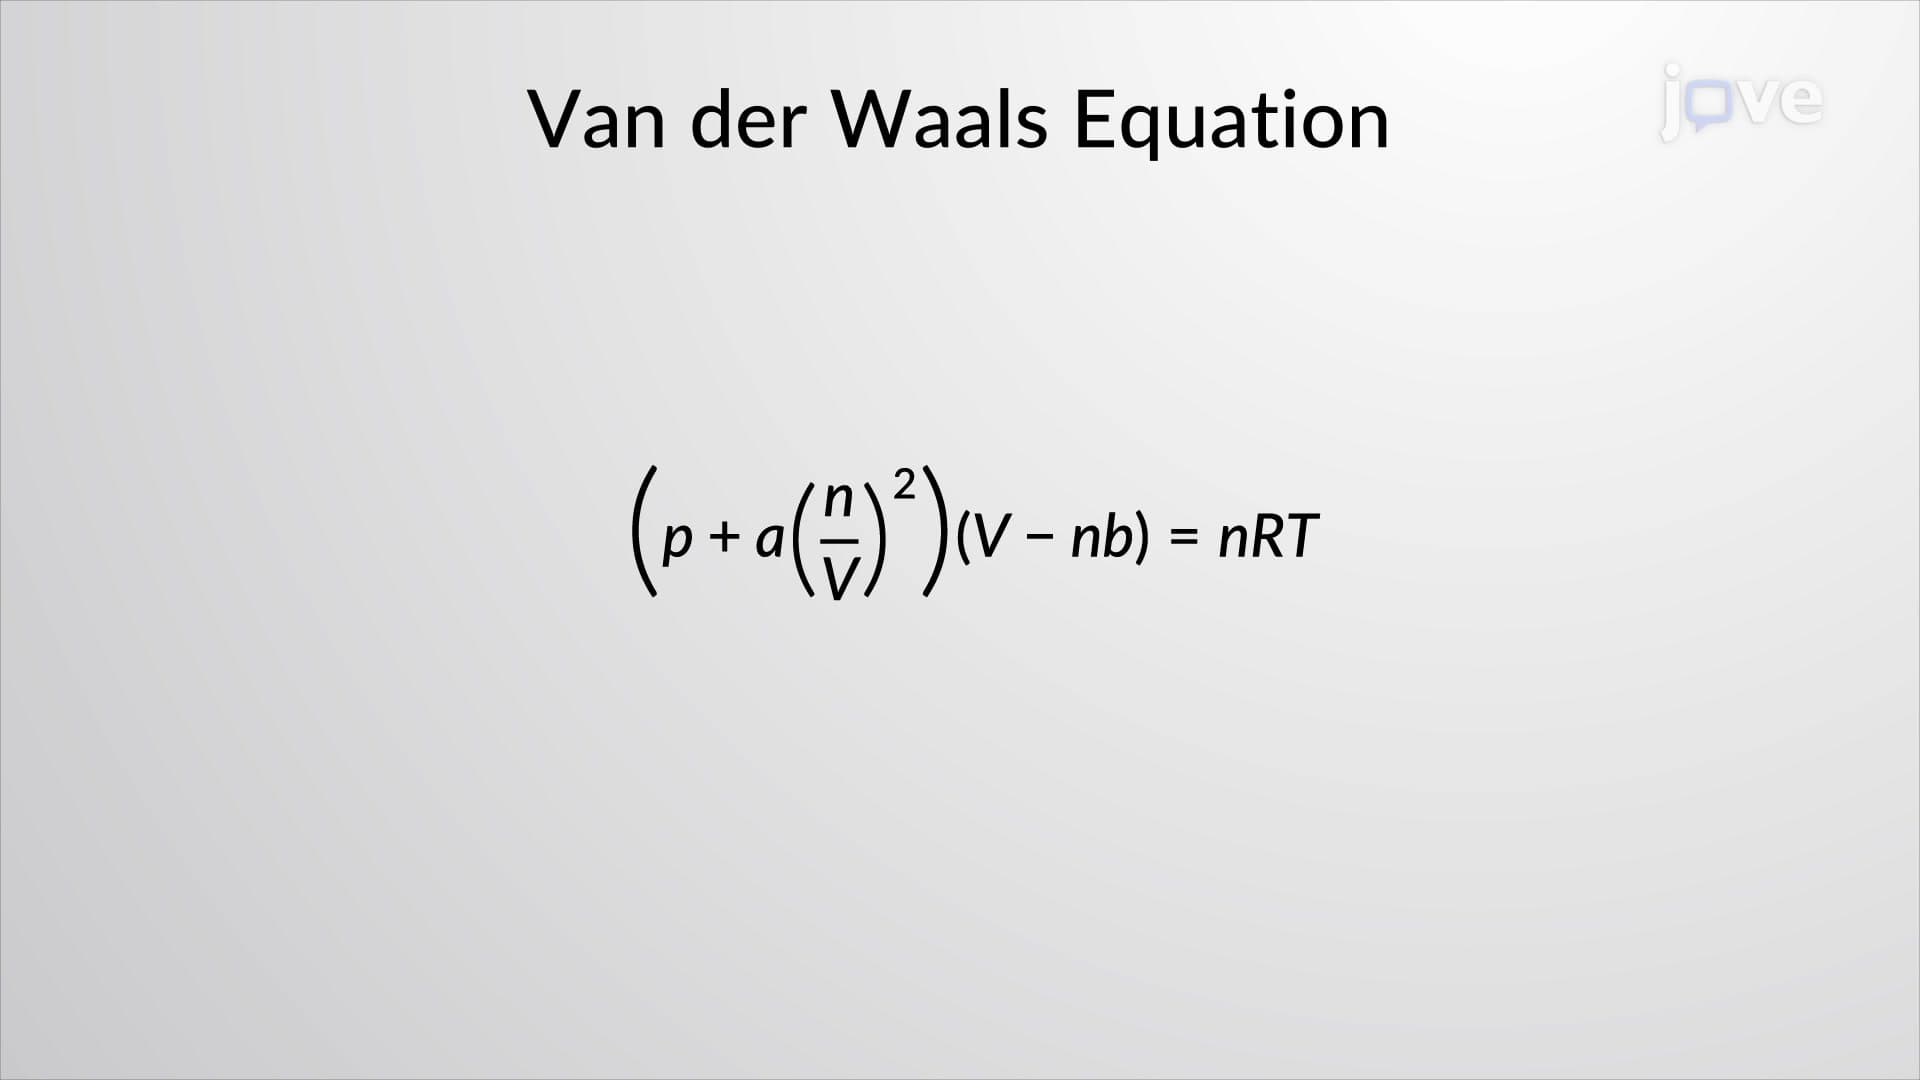
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 01:16
UV–Vis Spectroscopy: Molecular Electronic Transitions
 01:26
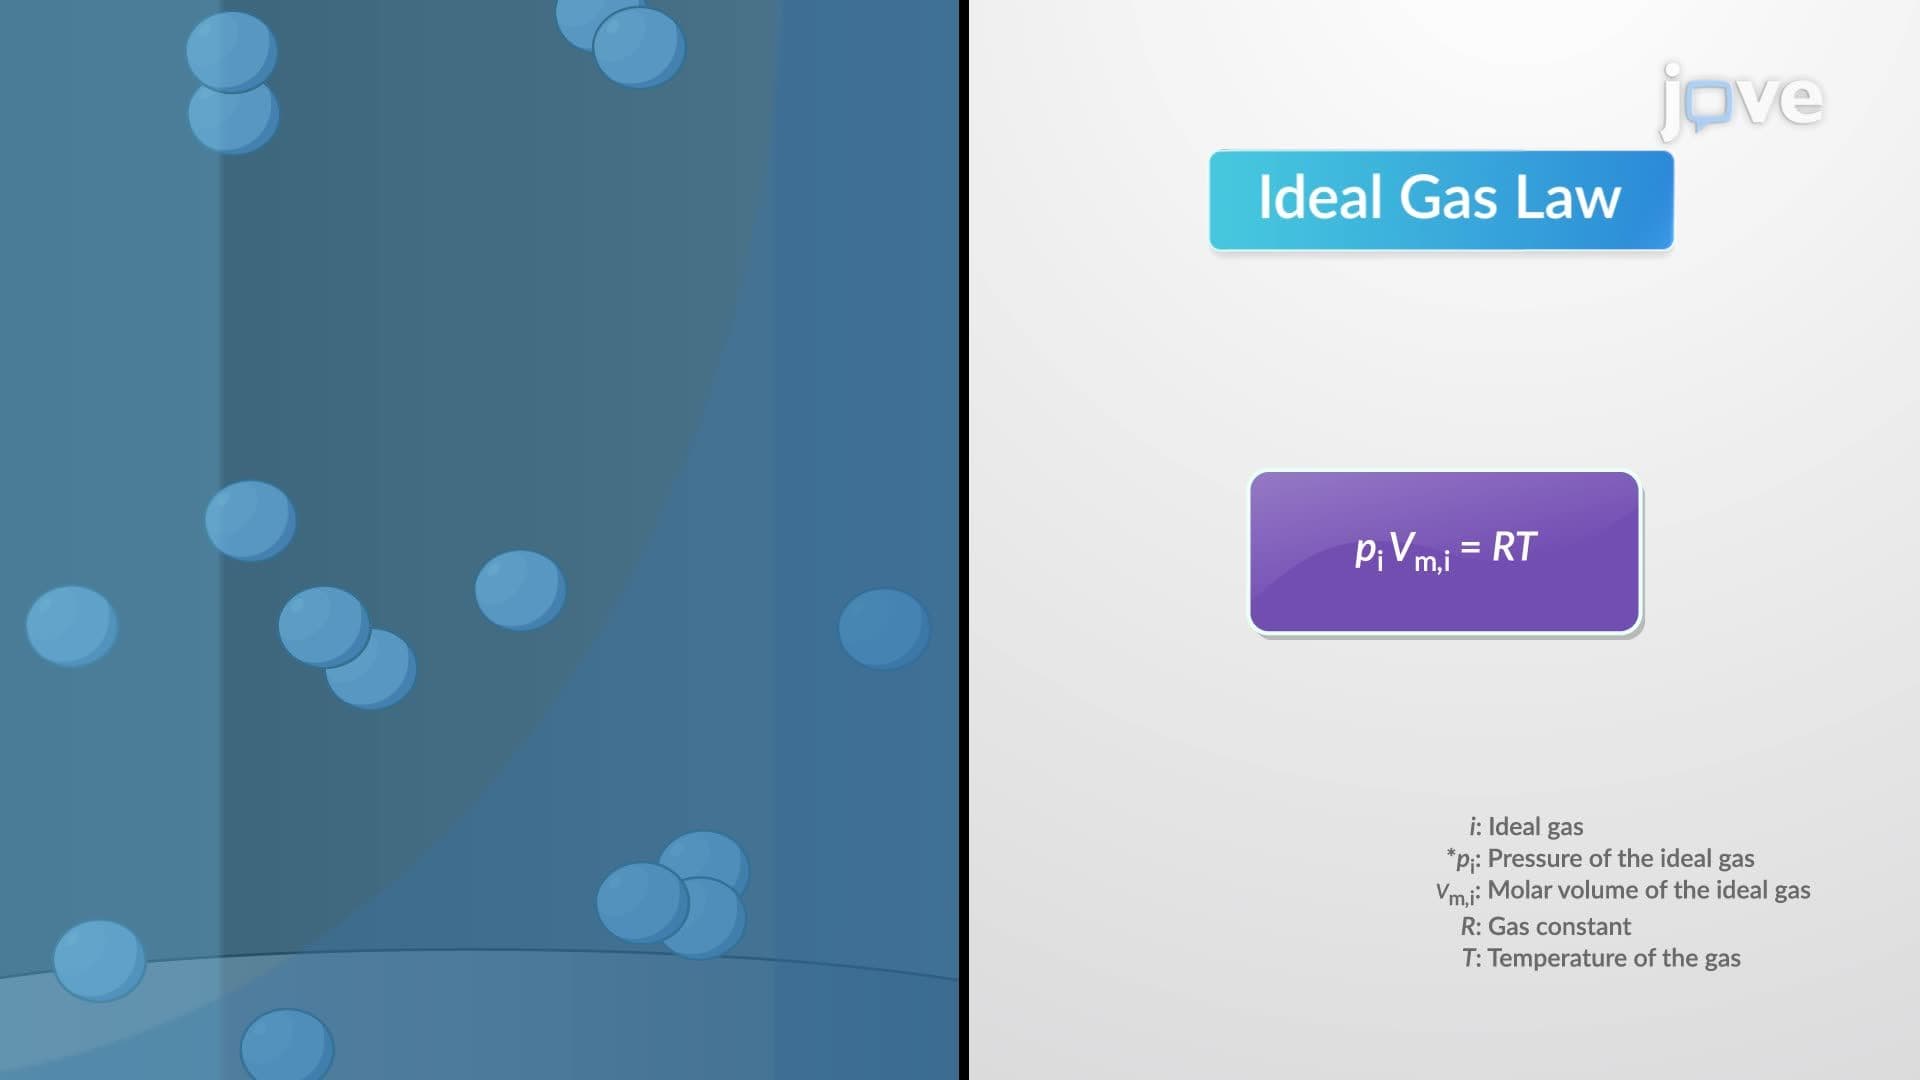
The Van der Waals Equation
 02:52
VSEPR Theory and the Basic Shapes
 01:14
Molecular Spectroscopy: Absorption and Emission