Related Experiment Video
Updated: Jun 27, 2026
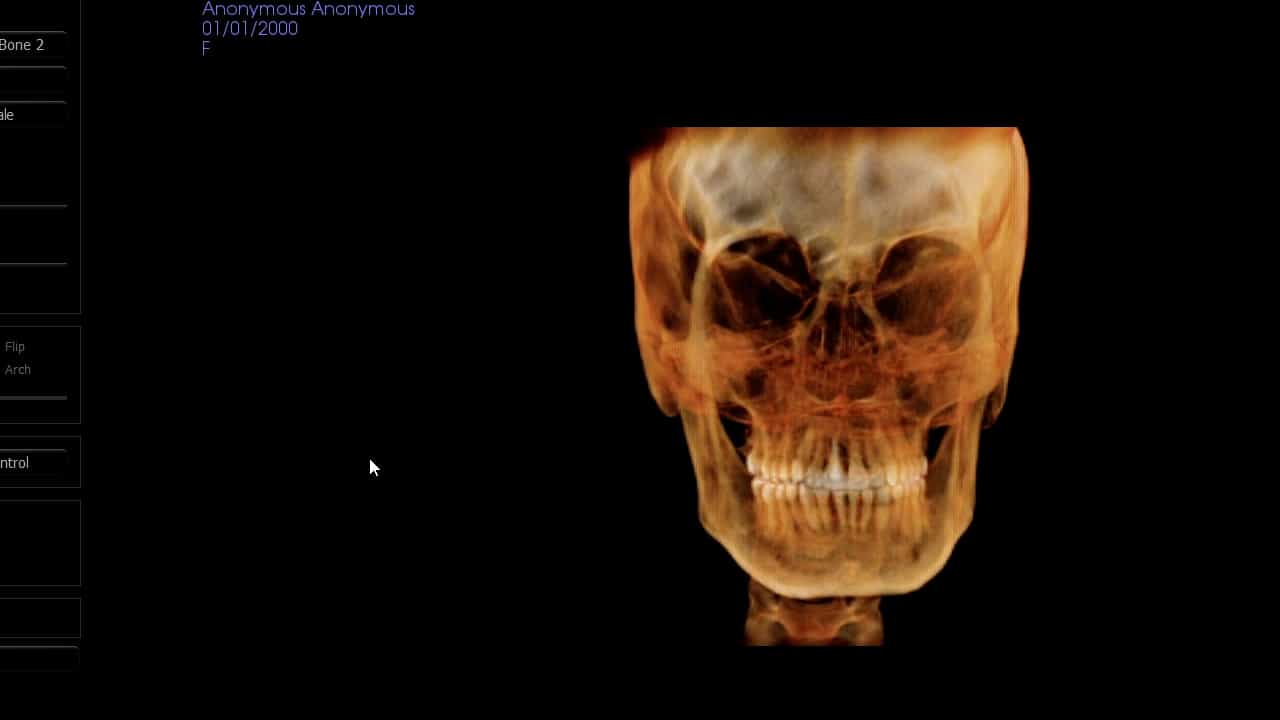

Three-Dimensional Cephalometric Landmark Annotation Demonstration on Human Cone Beam Computed Tomography Scans
Published on: September 8, 2023
Characterization of the first FGFRL1 mutation identified in a craniosynostosis patient
Thorsten Rieckmann1, Lei Zhuang, Christa E Flück
1Department of Clinical Research, University of Bern, 3010 Bern, Switzerland.

Abstract:
Fibroblast growth factor receptor-like 1 (FGFRL1) is a recently discovered transmembrane protein whose functions remain unclear. Since mutations in the related receptors FGFR1-3 cause skeletal malformations, DNA samples from 55 patients suffering from congenital skeletal malformations and 109 controls were searched for mutations in FGFRL1. One patient was identified harboring a frameshift mutation in the intracellular domain of this novel receptor. The patient showed craniosynostosis, radio-ulnar synostosis and genital abnormalities and had previously been diagnosed with Antley-Bixler syndrome. The effect of the FGFRL1 mutation was studied in vitro. In a reporter gene assay, the wild-type as well as the mutant receptor inhibited FGF signaling. However, the mutant protein differed from the wild-type protein in its subcellular localization. Mutant FGFRL1 was mainly found at the plasma membrane where it interacted with FGF ligands, while the wild-type protein was preferentially located in vesicular structures and the Golgi complex. Two motifs from the intracellular domain of FGFRL1 appeared to be responsible for this differential distribution, a tandem tyrosine based motif and a histidine-rich sequence. Deletion of either one led to the preferential redistribution of FGFRL1 to the plasma membrane. It is therefore likely that mutant FGFRL1 contributes to the skeletal malformations of the patient.
Insights
A novel mutation in Fibroblast Growth Factor Receptor-Like 1 (FGFRL1) was found in a patient with skeletal malformations. This FGFRL1 mutation alters protein localization, likely contributing to congenital skeletal abnormalities.
Area of Science:
- Genetics
- Molecular Biology
- Developmental Biology
Background:
- Fibroblast growth factor receptor-like 1 (FGFRL1) is a transmembrane protein with largely unknown functions.
- Mutations in related fibroblast growth factor receptors (FGFRs) are known to cause skeletal malformations.
Purpose of the Study:
- To investigate the role of FGFRL1 in congenital skeletal malformations by searching for mutations.
- To characterize the functional consequences of a identified FGFRL1 mutation in vitro.
Main Methods:
- Screening of FGFRL1 in DNA samples from patients with congenital skeletal malformations and controls.
- In vitro reporter gene assays to assess FGF signaling inhibition.
- Subcellular localization studies of wild-type and mutant FGFRL1 proteins using microscopy.
Main Results:
- A patient with craniosynostosis, radio-ulnar synostosis, and genital abnormalities (diagnosed with Antley-Bixler syndrome) was found to have a frameshift mutation in the FGFRL1 intracellular domain.
- Both wild-type and mutant FGFRL1 inhibited FGF signaling in reporter gene assays.
- Mutant FGFRL1 exhibited altered subcellular localization, predominantly residing at the plasma membrane, unlike wild-type FGFRL1 which localized to vesicular structures and the Golgi complex.
- Specific intracellular motifs (tandem tyrosine-based and histidine-rich sequences) were identified as responsible for the differential localization of FGFRL1.
Conclusions:
- The identified frameshift mutation in FGFRL1 is likely pathogenic and contributes to the patient's skeletal malformations.
- Altered subcellular localization of FGFRL1 due to the mutation may disrupt normal FGF signaling pathways involved in skeletal development.
More Related Videos
Related Concept Videos
 01:20
The Retinoblastoma Gene
The first-ever tumor suppressor gene called Rb was identified in retinoblastoma - a rare eye tumor in children. In inherited forms of the disease, a child inherits one defective copy of the Rb gene, which predisposes them to retinoblastoma. However,...
01:20The Retinoblastoma Gene
The first-ever tumor suppressor gene called Rb was identified in retinoblastoma - a rare eye tumor in children. In inherited forms of the disease, a child inherits one defective copy of the Rb gene, which predisposes them to retinoblastoma. However,...
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...