Efficient table lookup without inverse square roots for calculation of pair wise atomic interactions in classical simulations
Related Experiment Video
Updated: Jun 27, 2026

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
1Department of Biosciences and Nutrition, Karolinska Institutet, SE-141 57 Huddinge, Sweden. Lennart.Nilsson@ki.se

Abstract:
A major bottleneck in classical atomistic simulations of biomolecular systems is the calculation of the pair wise nonbonded (Coulomb, van der Waals) interactions. This remains an issue even when methods are used (e.g., lattice summation or spherical cutoffs) in which the number of interactions is reduced from O(N(2)) to O(NlogN) or O(N). The interaction forces and energies can either be calculated directly each time they are needed or retrieved using precomputed values in a lookup table; the choice between direct calculation and table lookup methods depends on the characteristics of the system studied (total number of particles and the number of particle kinds) as well as the hardware used (CPU speed, size and speed of cache, and main memory). A recently developed lookup table code, implemented in portable and easily maintained FORTRAN 95 in the CHARMM program (www.charmm.org), achieves a 1.5- to 2-fold speedup compared with standard calculations using highly optimized FORTRAN code in real molecular dynamics simulations for a wide range of molecular system sizes. No approximations other than the finite resolution of the tables are introduced, and linear interpolation in a table with the relatively modest density of 100 points/A(2) yields the same accuracy as the standard double precision calculations. For proteins in explicit water a less dense table (10 points/A(2)) is 10-20% faster than using the larger table, and only slightly less accurate. The lookup table is even faster than hand coded assembler routines in most cases, mainly due to a significantly smaller operation count inside the inner loop.

Related Concept Videos
 01:26
The Van der Waals Equation
 01:29
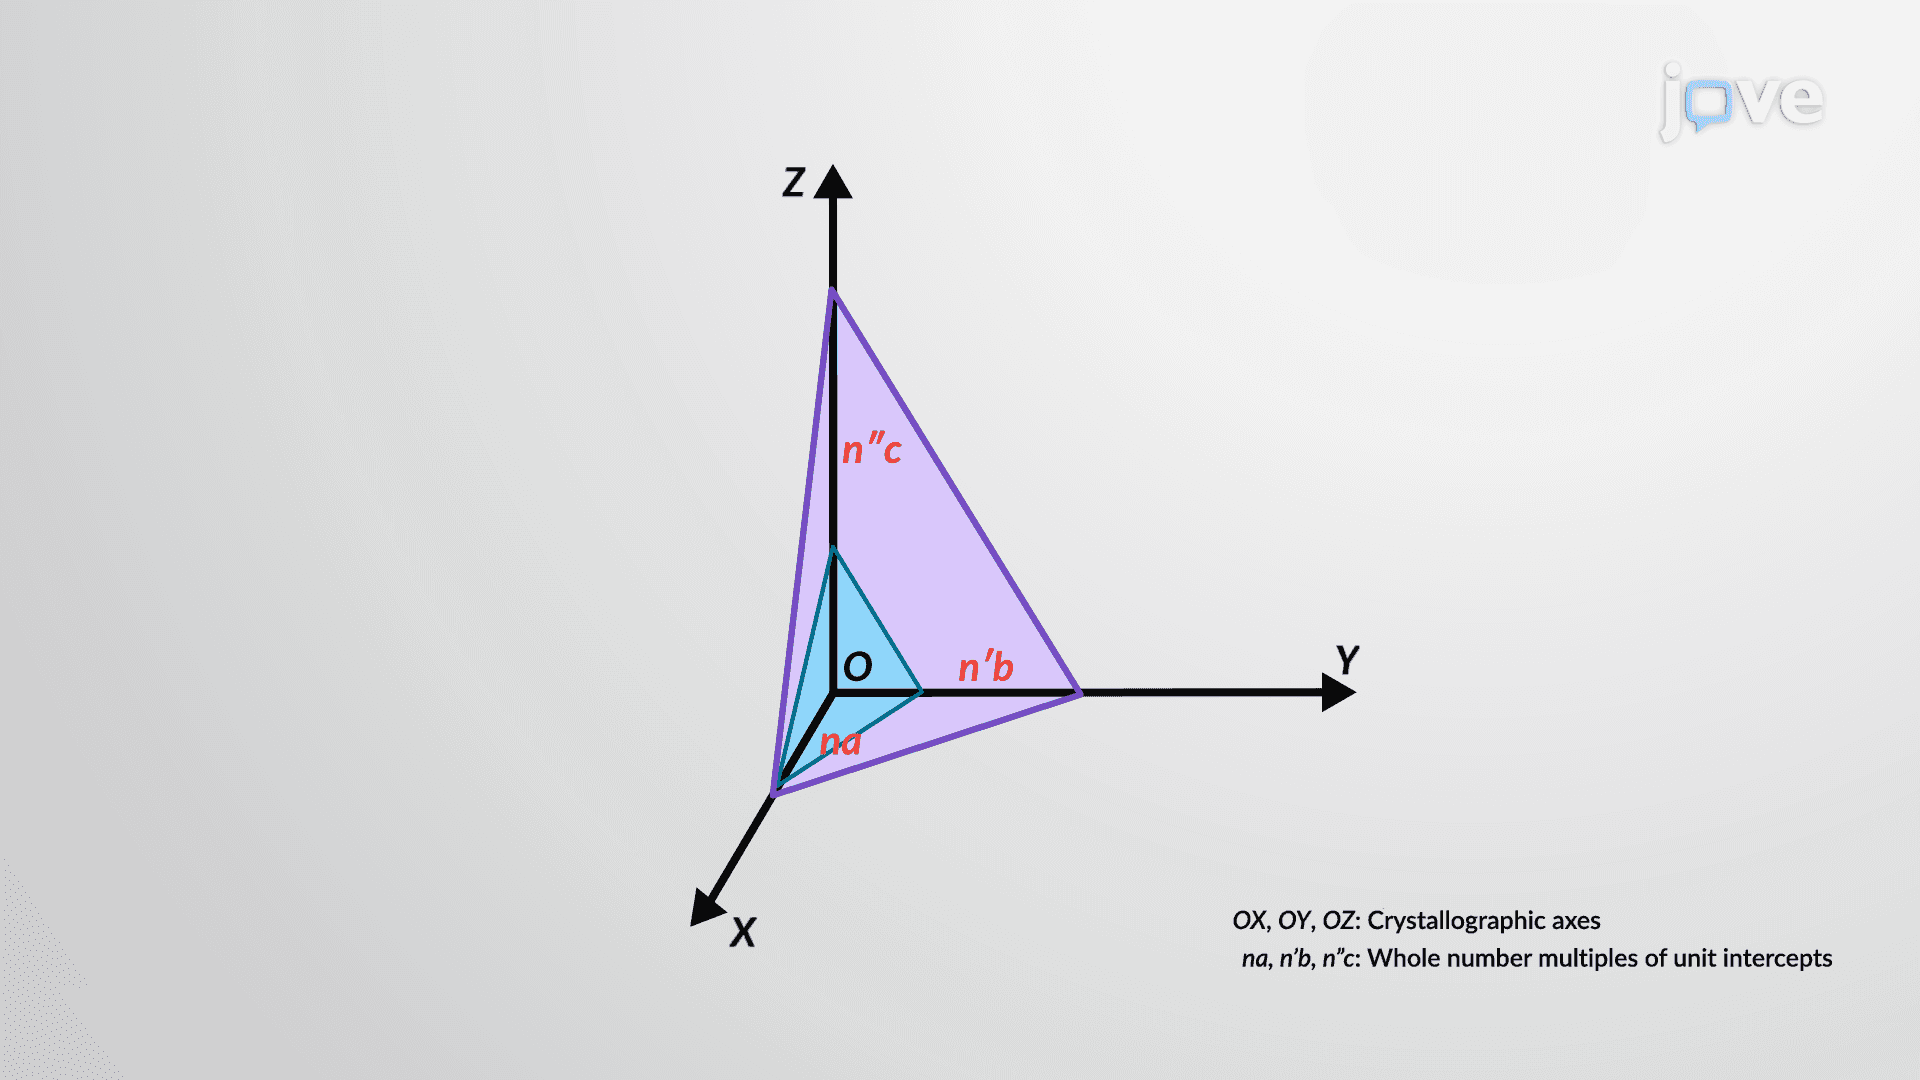
Law of Rational Indices
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:10
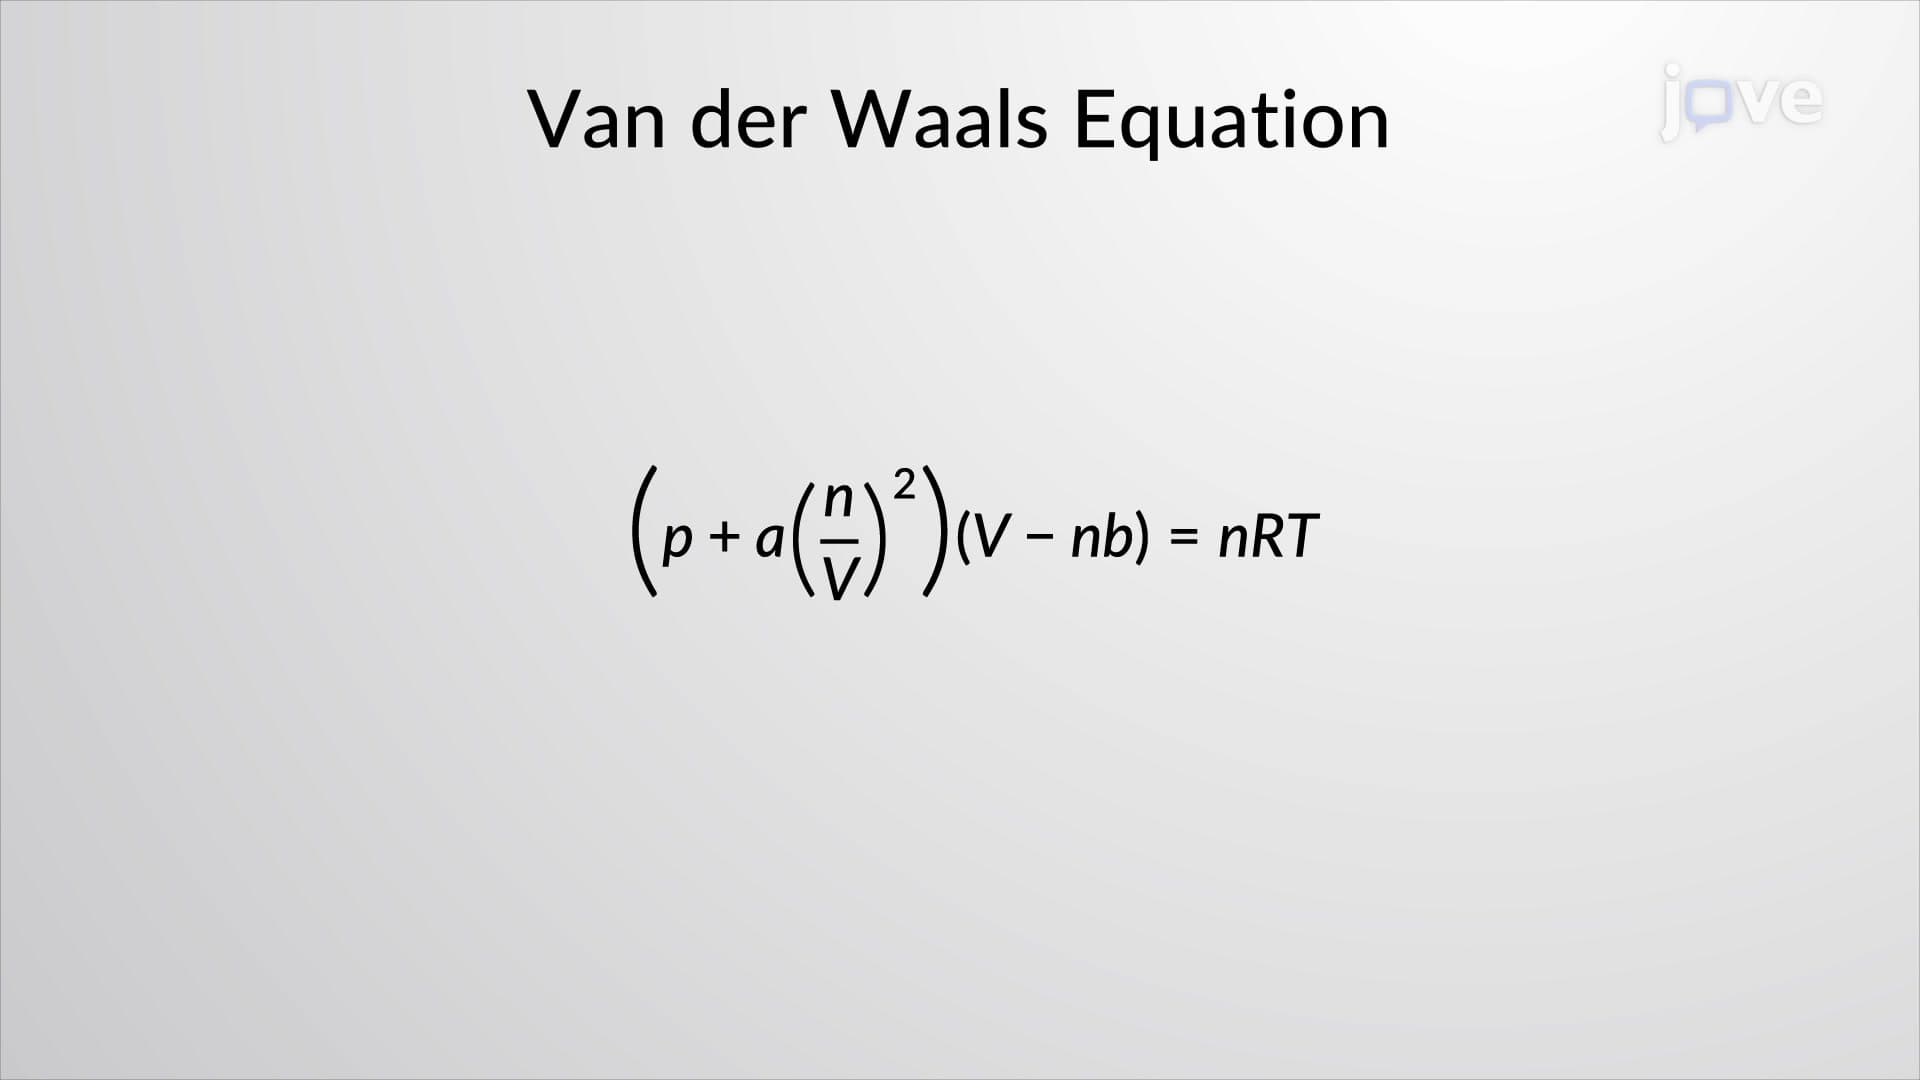
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 02:35
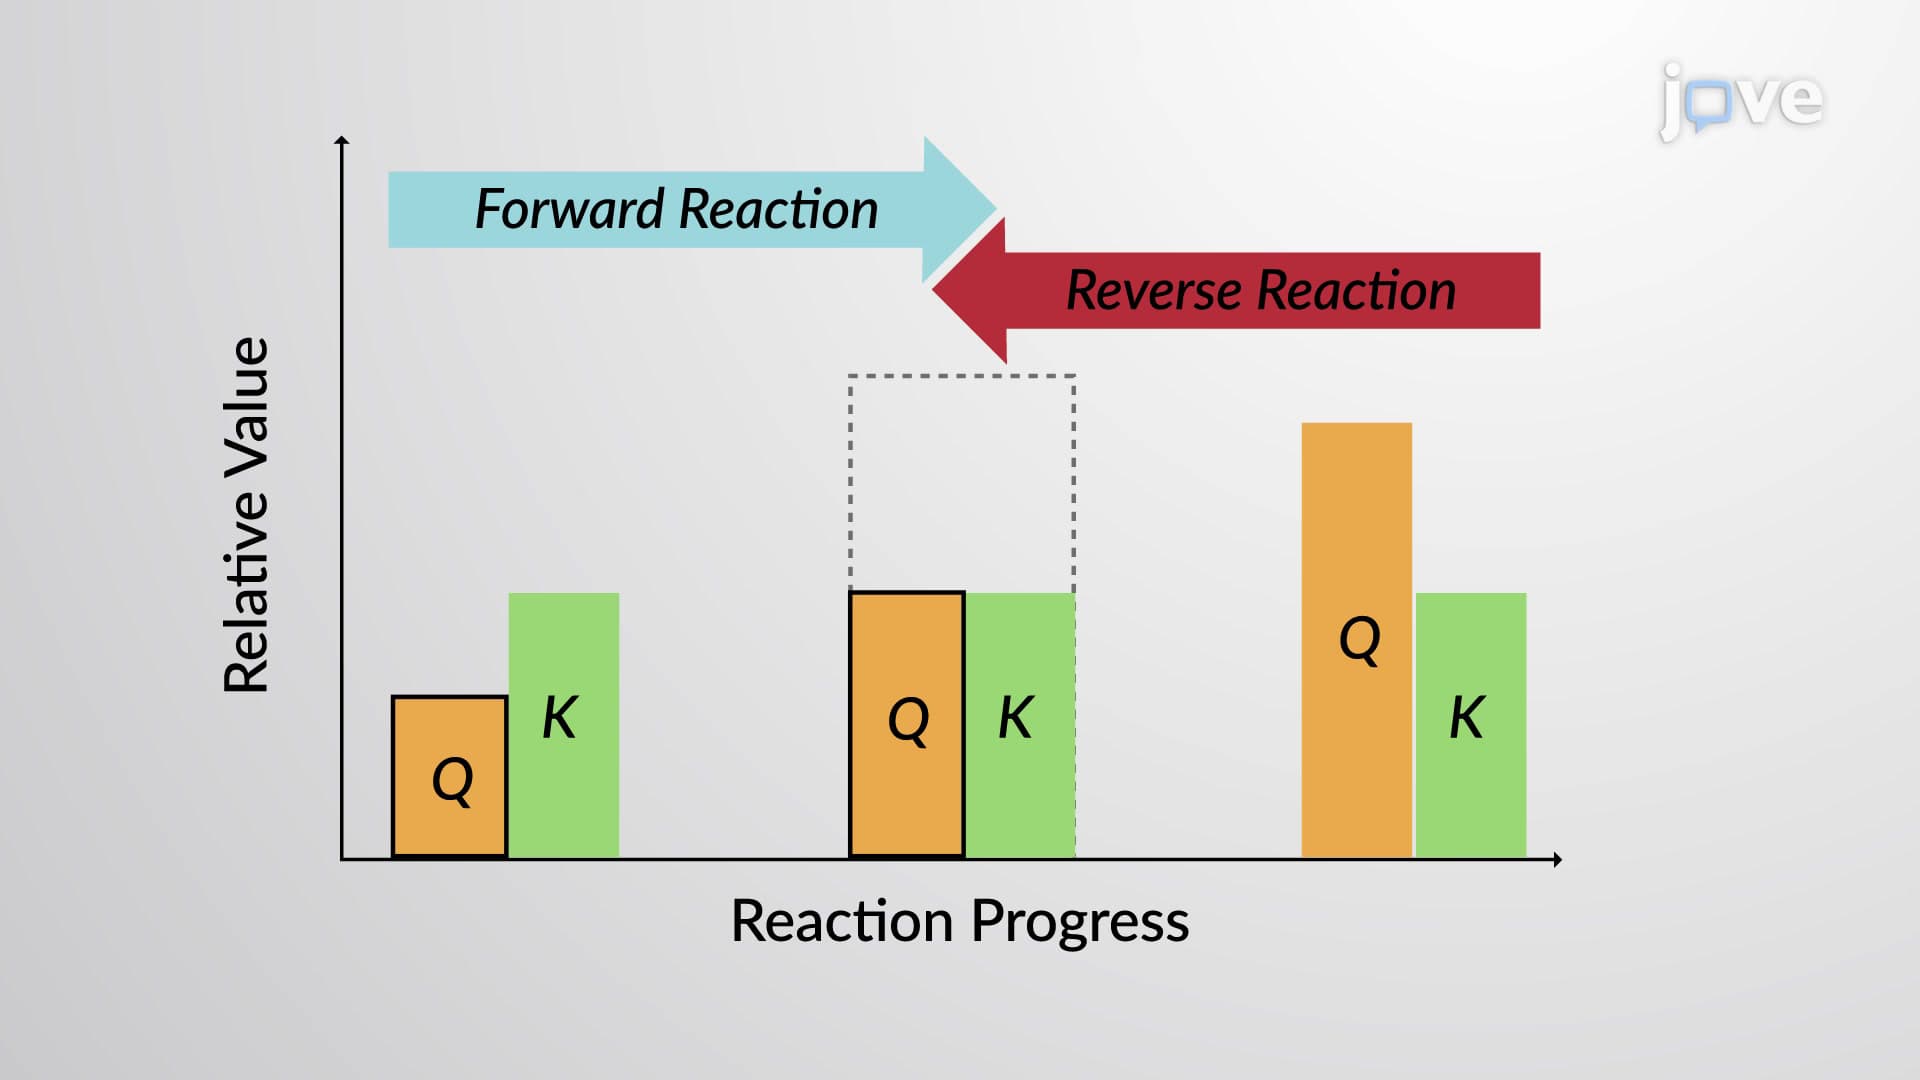
Reaction Quotient
 01:24
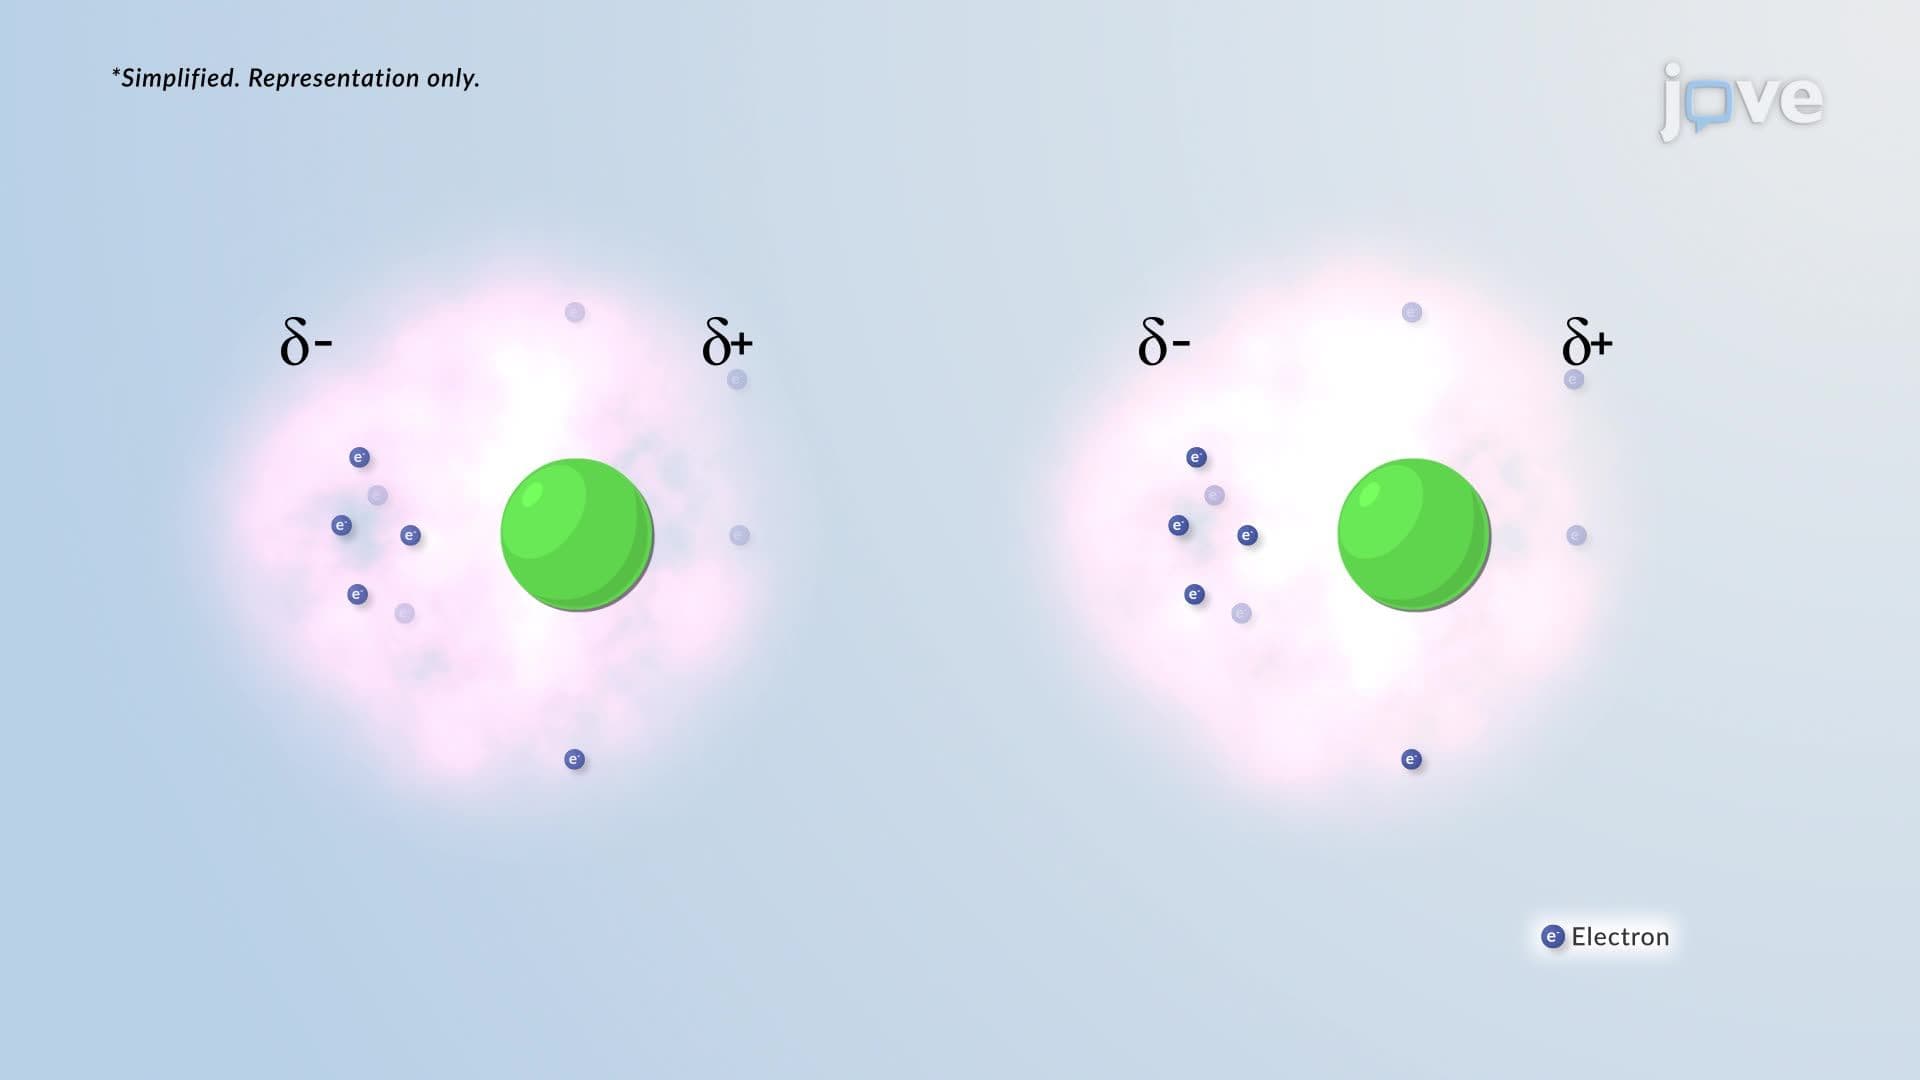
Van der Waals Interactions