Related Experiment Video
Updated: Jun 27, 2026
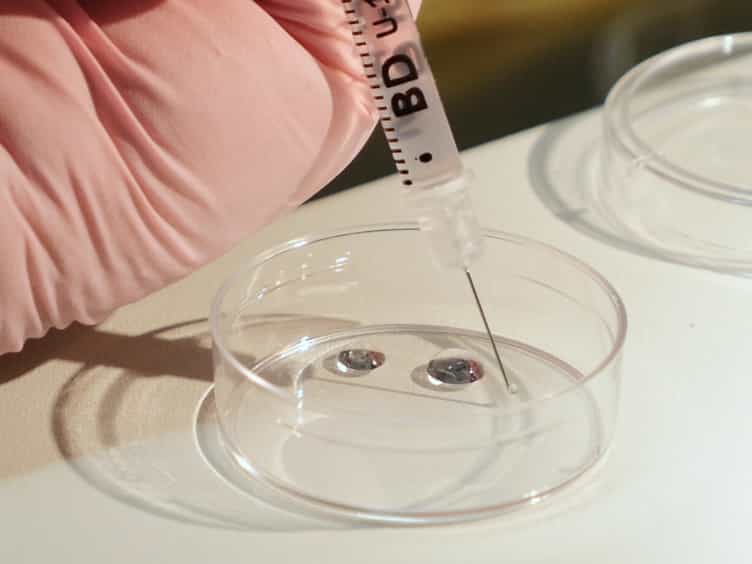

Generation of Genetically Modified Mice through the Microinjection of Oocytes
Published on: June 15, 2017
De novo generation of a transmissible spongiform encephalopathy by mouse transgenesis
Christina J Sigurdson1, K Peter R Nilsson, Simone Hornemann
1UniversitätsSpital Zürich, Institute of Neuropathology, CH-8091 Zürich, Switzerland.

Abstract:
Most transmissible spongiform encephalopathies arise either spontaneously or by infection. Mutations of PRNP, which encodes the prion protein, PrP, segregate with phenotypically similar diseases. Here we report that moderate overexpression in transgenic mice of mPrP(170N,174T), a mouse PrP with two point mutations that subtly affect the structure of its globular domain, causes a fully penetrant lethal spongiform encephalopathy with cerebral PrP plaques. This genetic disease was reproduced with 100% attack rate by intracerebral inoculation of brain homogenate to tga20 mice overexpressing WT PrP, and from the latter to WT mice, but not to PrP-deficient mice. Upon successive transmissions, the incubation periods decreased and PrP became more protease-resistant, indicating the presence of a strain barrier that was gradually overcome by repeated passaging. This shows that expression of a subtly altered prion protein, with known 3D structure, efficiently generates a prion disease.
Insights
Altered prion protein (PrP) expression in mice caused a fatal spongiform encephalopathy. This prion disease could be transmitted between mice, demonstrating how subtle PrP structural changes can generate infectious agents.
Area of Science:
- Neuroscience
- Molecular Biology
- Genetics
Background:
- Transmissible spongiform encephalopathies (TSEs) are typically spontaneous or infectious.
- Mutations in the prion protein gene (PRNP) are linked to similar diseases.
Purpose of the Study:
- To investigate if subtle structural alterations in mouse prion protein (mPrP) can induce spongiform encephalopathy.
- To determine the transmissibility and characteristics of the induced prion disease.
Main Methods:
- Generation of transgenic mice overexpressing mPrP with two specific point mutations (170N, 174T).
- Intracerebral inoculation of brain homogenates from affected mice into recipient mice (tga20 and wild-type).
- Analysis of disease progression, PrP plaque formation, and prion characteristics upon transmission.
Main Results:
- Moderate overexpression of the mutated mPrP(170N, 174T) in transgenic mice resulted in a fully penetrant, lethal spongiform encephalopathy with cerebral PrP plaques.
- The induced disease was 100% transmissible to tga20 mice overexpressing wild-type PrP, and subsequently to wild-type mice, but not to PrP-deficient mice.
- Repeated passaging led to decreased incubation periods and increased prion protease resistance, indicating the gradual overcoming of a strain barrier.
Conclusions:
- Subtle structural changes in the prion protein, even with a known 3D structure, are sufficient to efficiently generate a prion disease.
- The study demonstrates a novel mechanism for prion disease generation through altered protein expression and structure.
- The findings highlight the critical role of PrP structure and expression levels in prion pathogenesis and transmission.