All-atom chain-building by optimizing MODELLER energy function using conformational space annealing
Related Experiment Video
Updated: Jun 27, 2026
Exploring Caspase Mutations and Post-Translational Modification by Molecular Modeling Approaches
Published on: October 13, 2022
Keehyoung Joo1, Jinwoo Lee, Joo-Hyun Seo
1School of Computational Sciences, Korea Institute for Advanced Study, Seoul 130-722, Korea.

Abstract:
We have investigated the effect of rigorous optimization of the MODELLER energy function for possible improvement in protein all-atom chain-building. For this we applied the global optimization method called conformational space annealing (CSA) to the standard MODELLER procedure to achieve better energy optimization than what MODELLER provides. The method, which we call MODELLERCSA, is tested on two benchmark sets. The first is the 298 proteins taken from the HOMSTRAD multiple alignment set. By simply optimizing the MODELLER energy function, we observe significant improvement in side-chain modeling, where MODELLERCSA provides about 10.7% (14.5%) improvement for chi(1) (chi(1) + chi(2)) accuracy compared to the standard MODELLER modeling. The improvement of backbone accuracy by MODELLERCSA is shown to be less prominent, and a similar improvement can be achieved by simply generating many standard MODELLER models and selecting lowest energy models. However, the level of side-chain modeling accuracy by MODELLERCSA could not be matched either by extensive MODELLER strategies, side-chain remodeling by SCWRL3, or copying unmutated rotamers. The identical procedure was successfully applied to 100 CASP7 template base modeling domains during the prediction season in a blind fashion, and the results are included here for comparison. From this study, we observe a good correlation between the MODELLER energy and the side-chain accuracy. Our findings indicate that, when a good alignment between a target protein and its templates is provided, thorough optimization of the MODELLER energy function leads to accurate all-atom models.
More Related Videos
 15:05
Deciphering the Structural Effects of Activating EGFR Somatic Mutations with Molecular Dynamics Simulation
Published on: May 20, 2020
 05:57
Synthesizing Amino Acids Modified with Reactive Carbonyls in Silico to Assess Structural Effects Using Molecular Dynamics Simulations
Published on: April 26, 2024
Related Concept Videos
 02:20
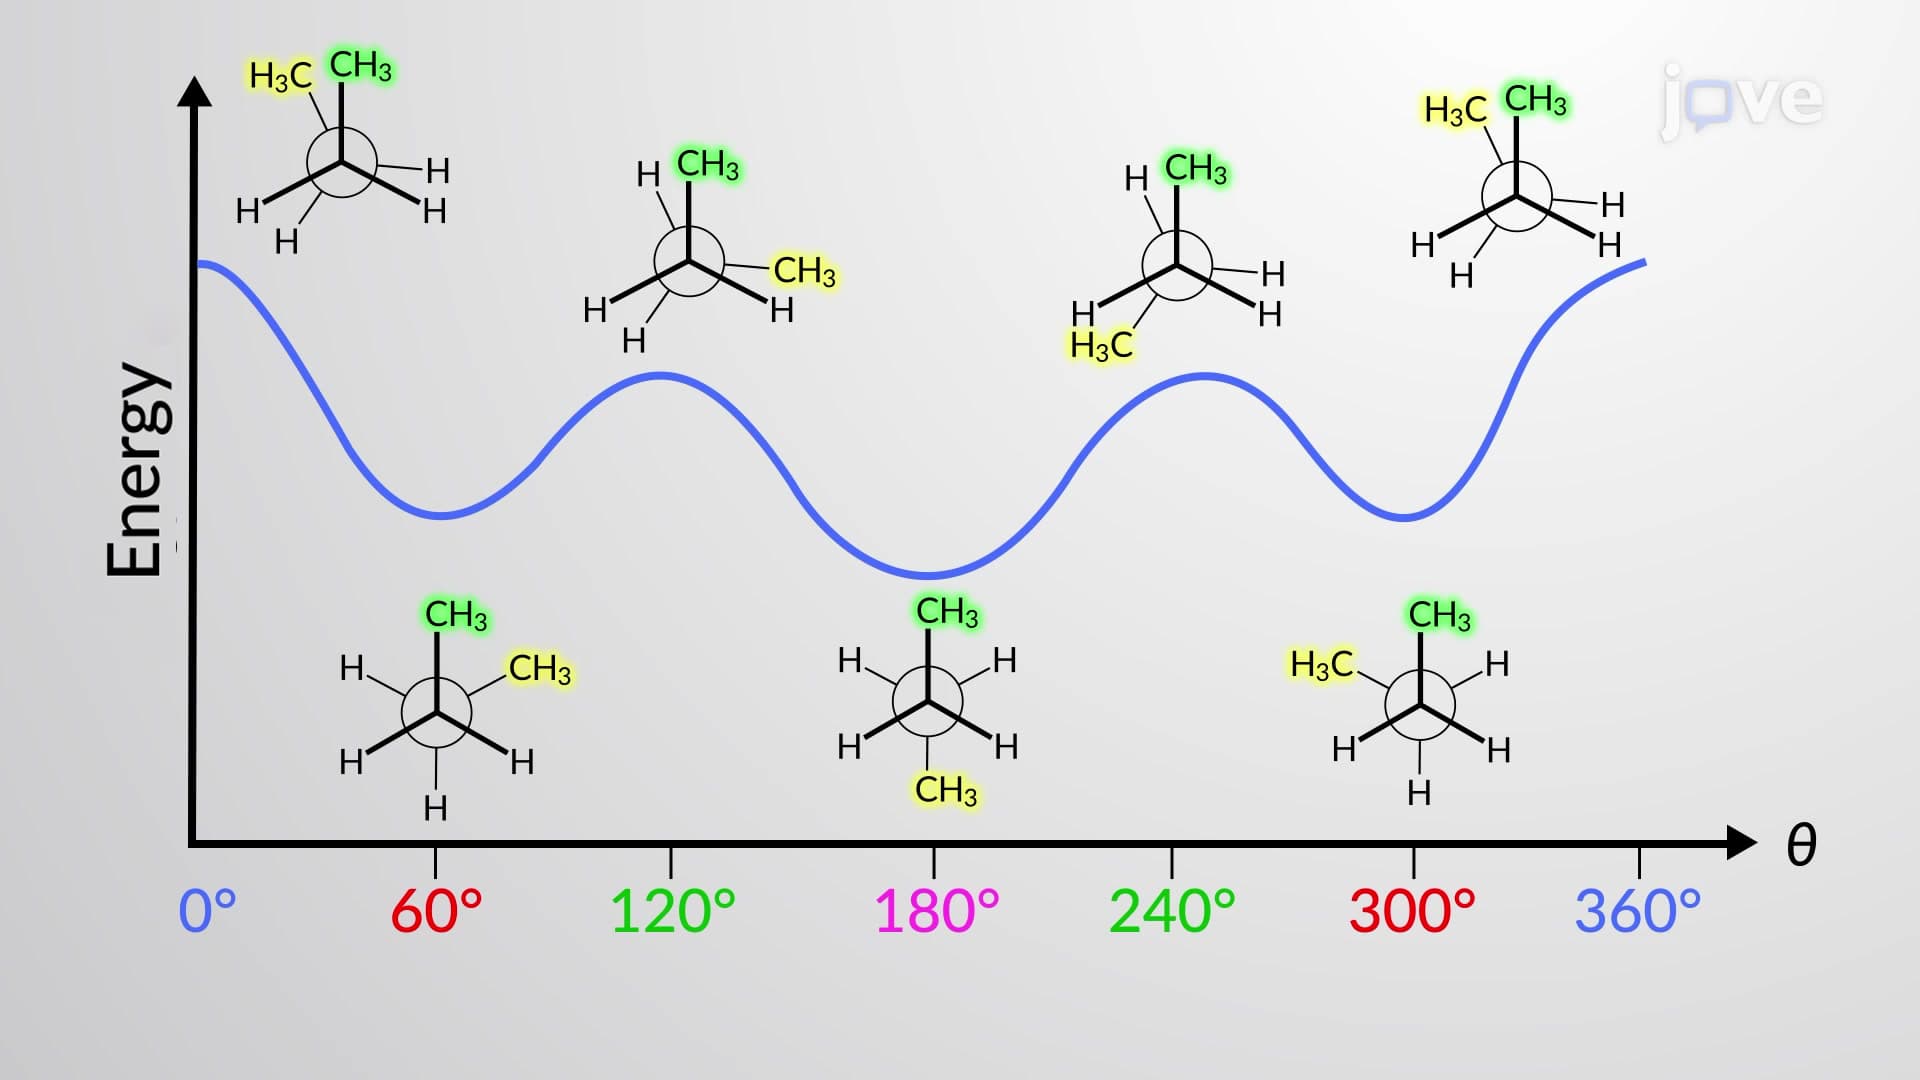
Conformations of Butane
 02:18
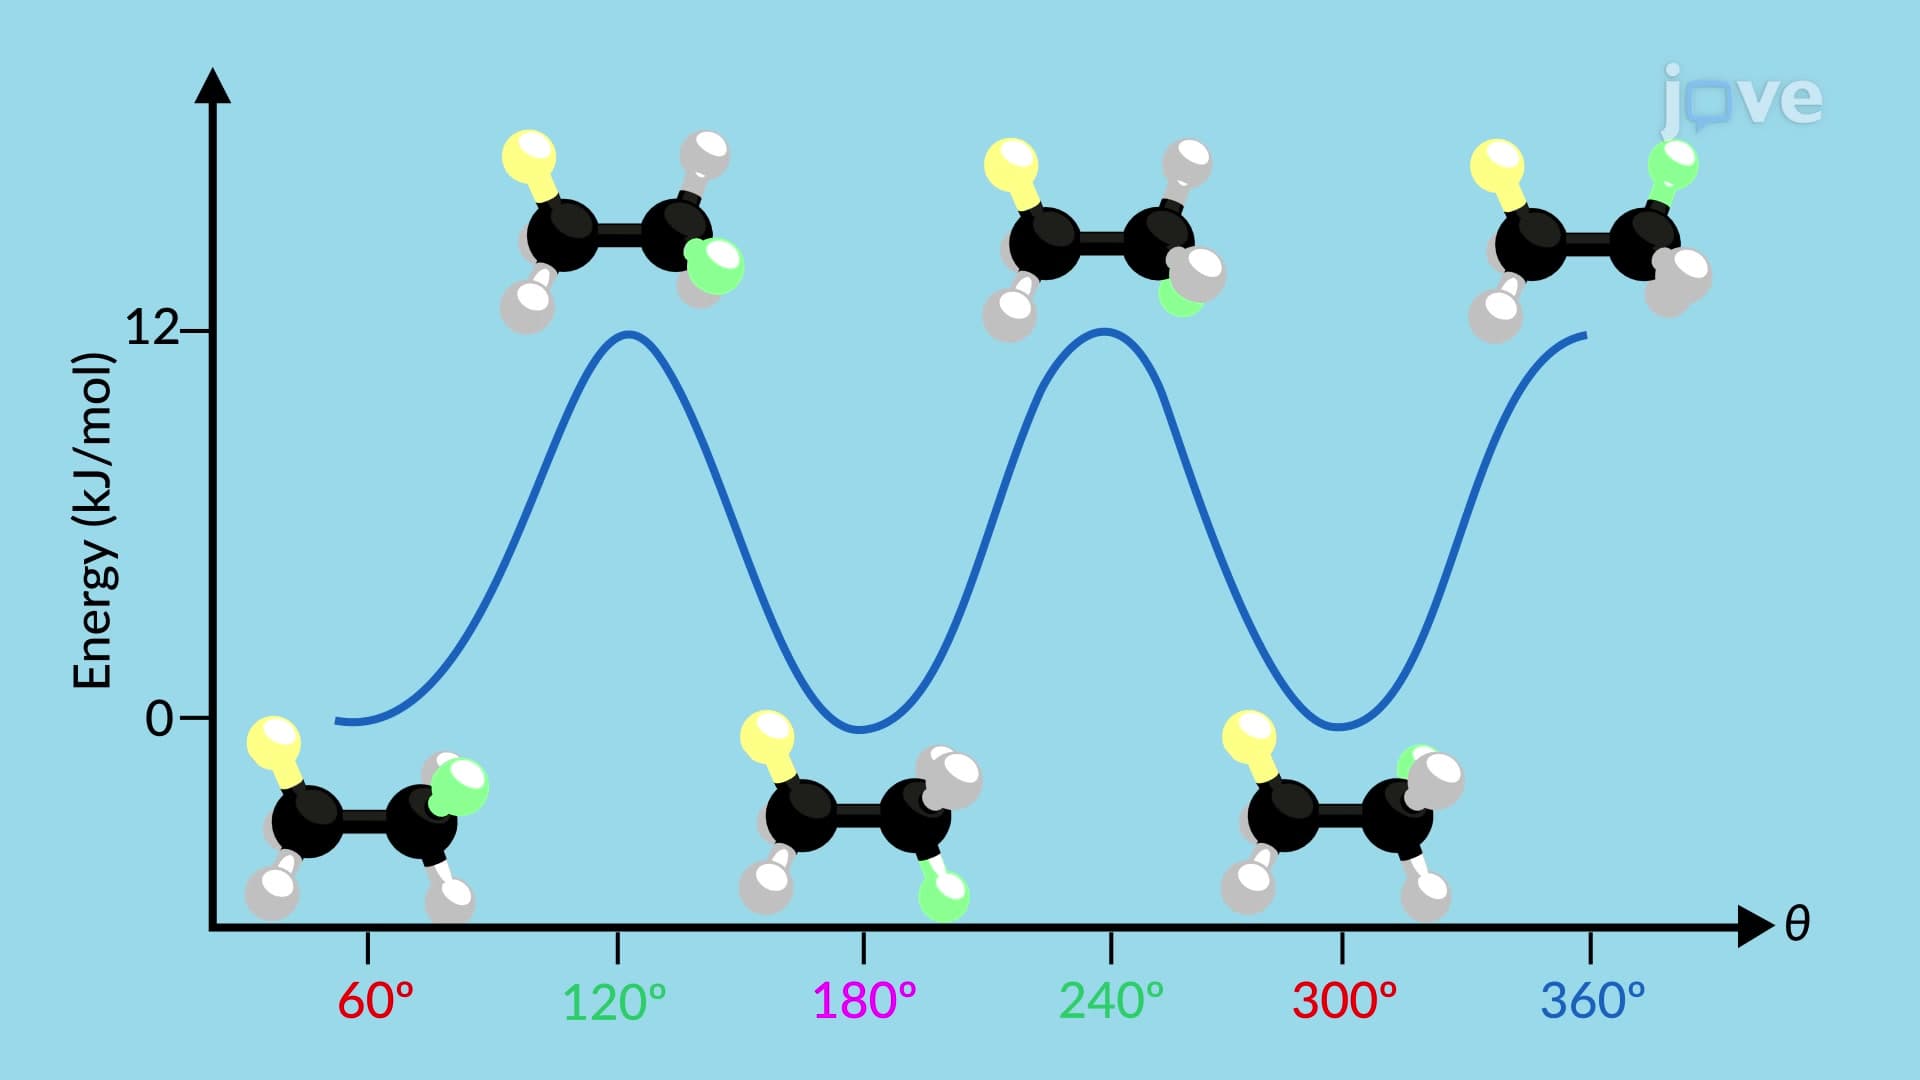
Conformations of Ethane and Propane
Staggered conformation is a low energy and more stable conformation with the C-H bonds on the front carbon placed at 60°dihedral angles relative to the C-H bonds on the back carbon, leading to a reduced torsional strain. In staggered ethane, the...
 02:00
Molecular Models
 02:02
Chair Conformation of Cyclohexane
The hydrogen atoms linked to carbons are arranged in two different axial and equatorial orientations to achieve this staggered...
 02:11
Conformations of Cyclohexane
The chair form is the most stable and derives its name from its resemblance to the “easy chair.” In the chair conformation, two carbon atoms are arranged out-of-plane — one above and one below, minimizing the torsional strain. In the chair form, the bond angle is very close to the ideal tetrahedral value,...
 01:15
¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR