Interaction of NO molecules with Pd clusters: ab initio density-functional study
Related Experiment Video
Updated: Jun 26, 2026

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Robert Grybos1, Lubomir Benco, Tomas Bucko
1Fakultät für Physik and Center for Computational Material Science, Universität Wien, Sensengasse, A-1090 Wien, Austria.

Abstract:
The adsorption of NO molecules on small Pd(n) (n = 1-6) clusters has been studied using first-principles density-functional theory. Three adsorption sites were considered: vertex (on-top), bridge, and hollow. Adsorption is strong, ranging from 2 to 3 eV. In all cases NO adsorbs in a bent configuration. Calculated shifts in N-O bond vibration frequencies (with anharmonic corrections) agree very well with available experimental data. In contrast to metallic Pd surfaces, adsorption of NO on palladium clusters causes considerable changes in geometry around adsorption site because palladium d-orbitals rehybridize to maximize the overlap with NO orbitals (mainly the antibonding pi*). Thus, the overall energetic effect of NO adsorption is the result of two competing processes: lowering of the total energy through tighter bonding with NO and rising the energy due to cluster deformation. The Pd(n)-NO bond creation is governed by electron transfer from Pd-d orbitals into the NO pi*. As a result, the Pd cluster becomes locally demagnetized (with total magnetic moment of 1 micro(B) located at Pd atoms not connected to NO) and the NO molecule is activated: the N-O bond length is increased and the vibration frequency is redshifted.

Related Concept Videos
 02:36
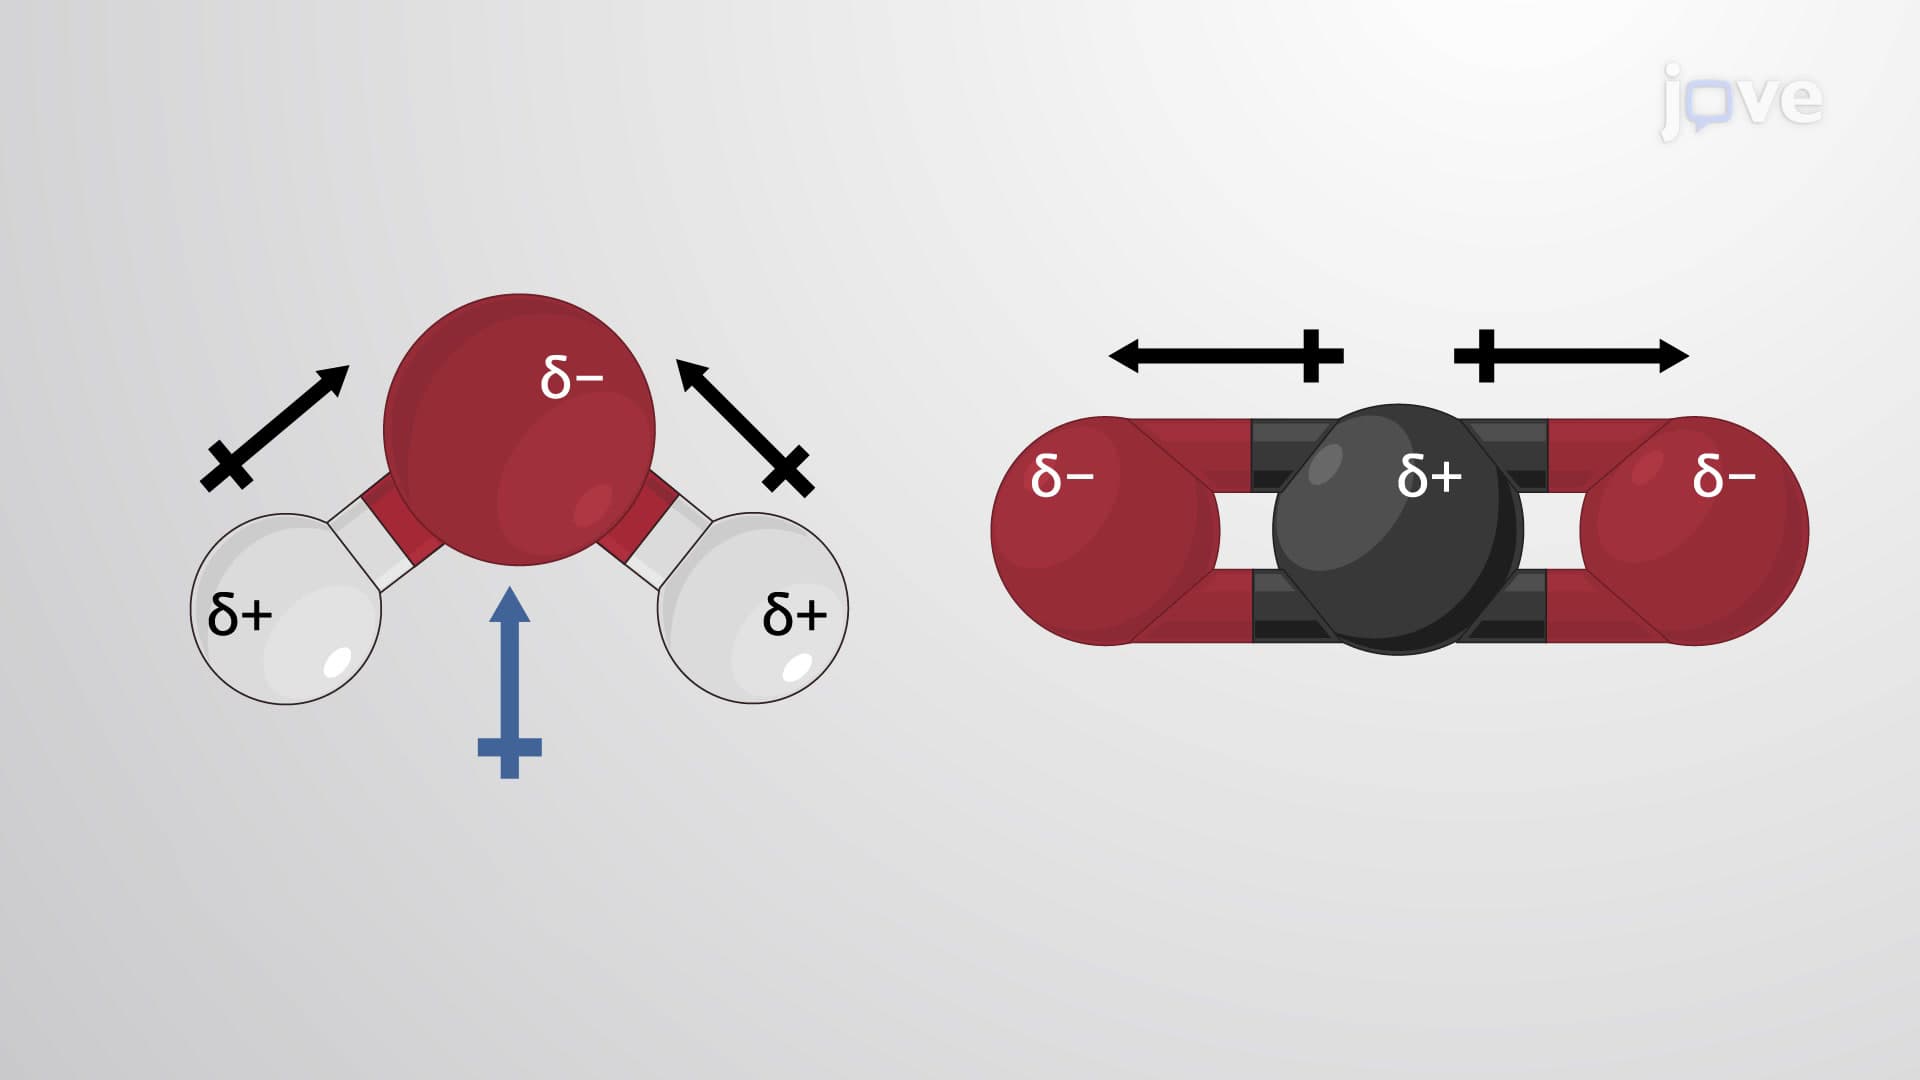
Molecular Geometry and Dipole Moments
 02:40
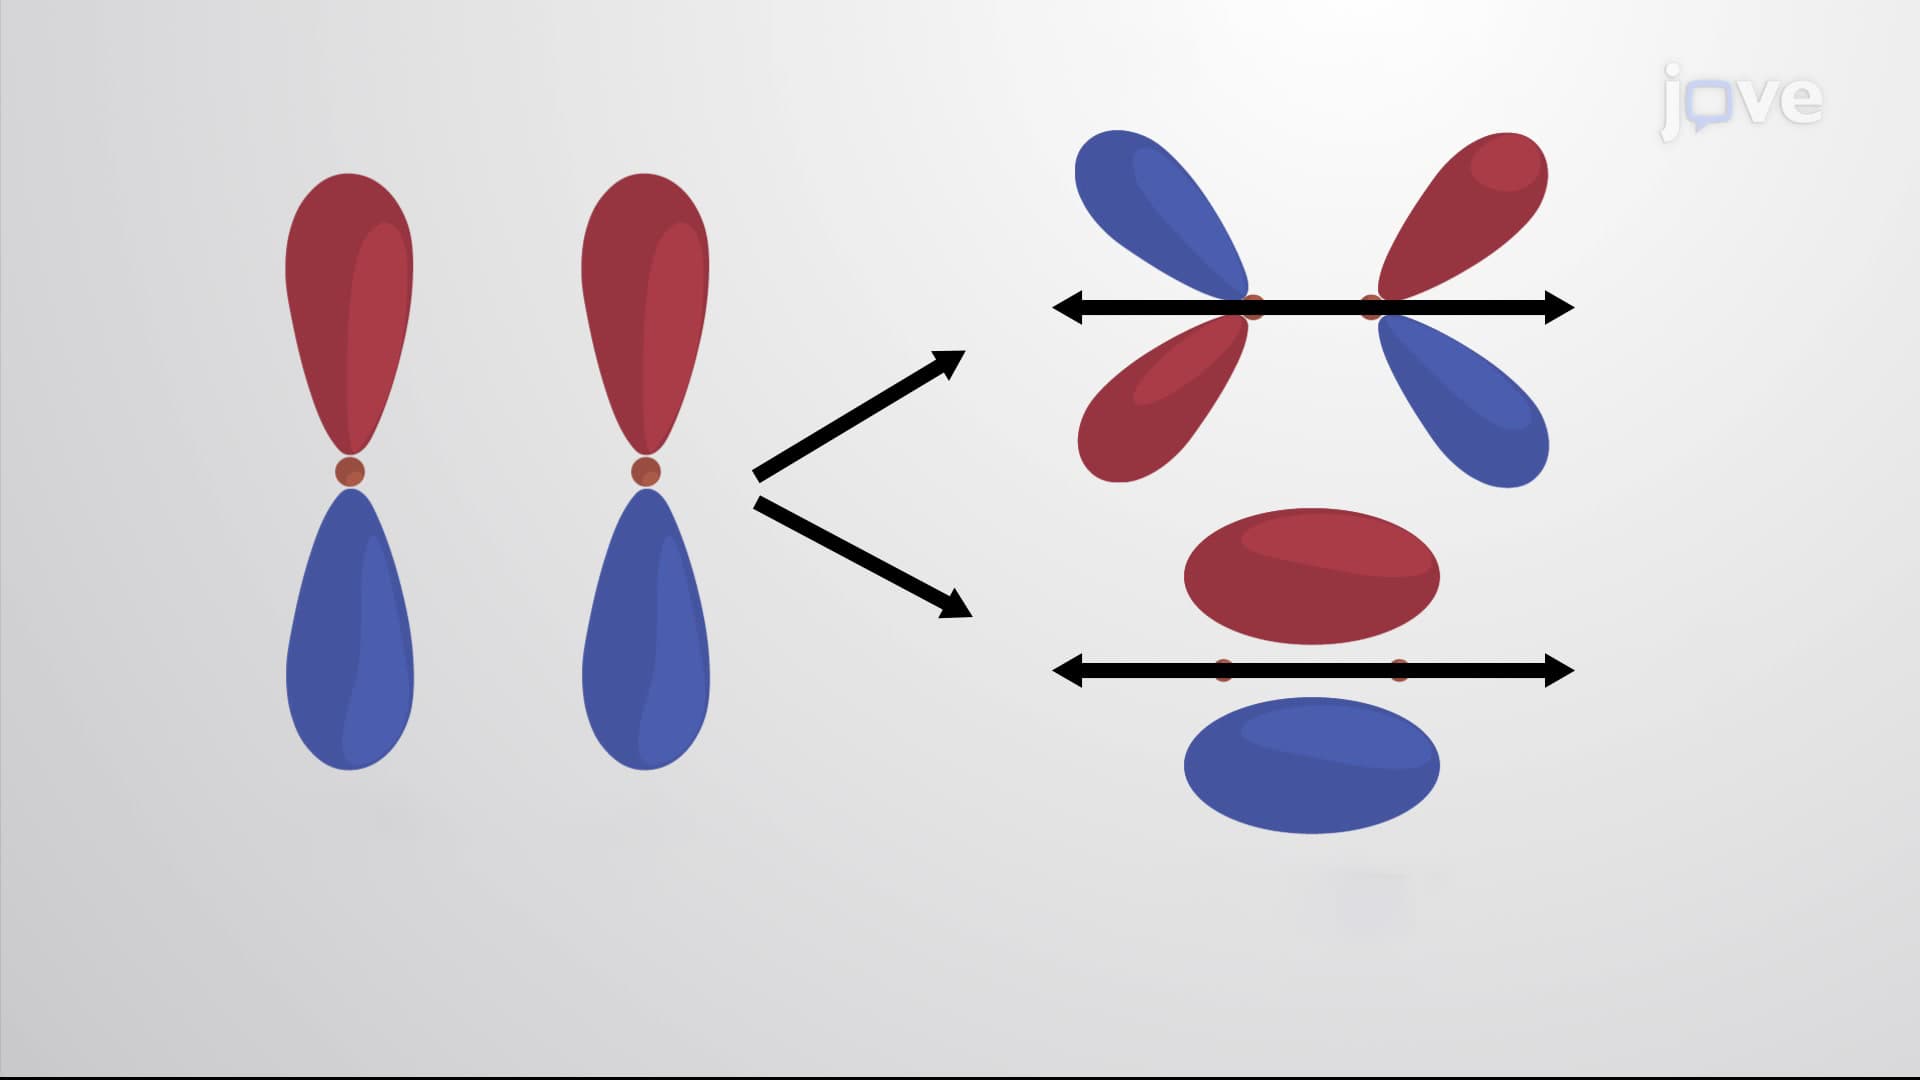
MO Theory and Covalent Bonding
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 03:51
Molecular Orbital Theory II
 01:24
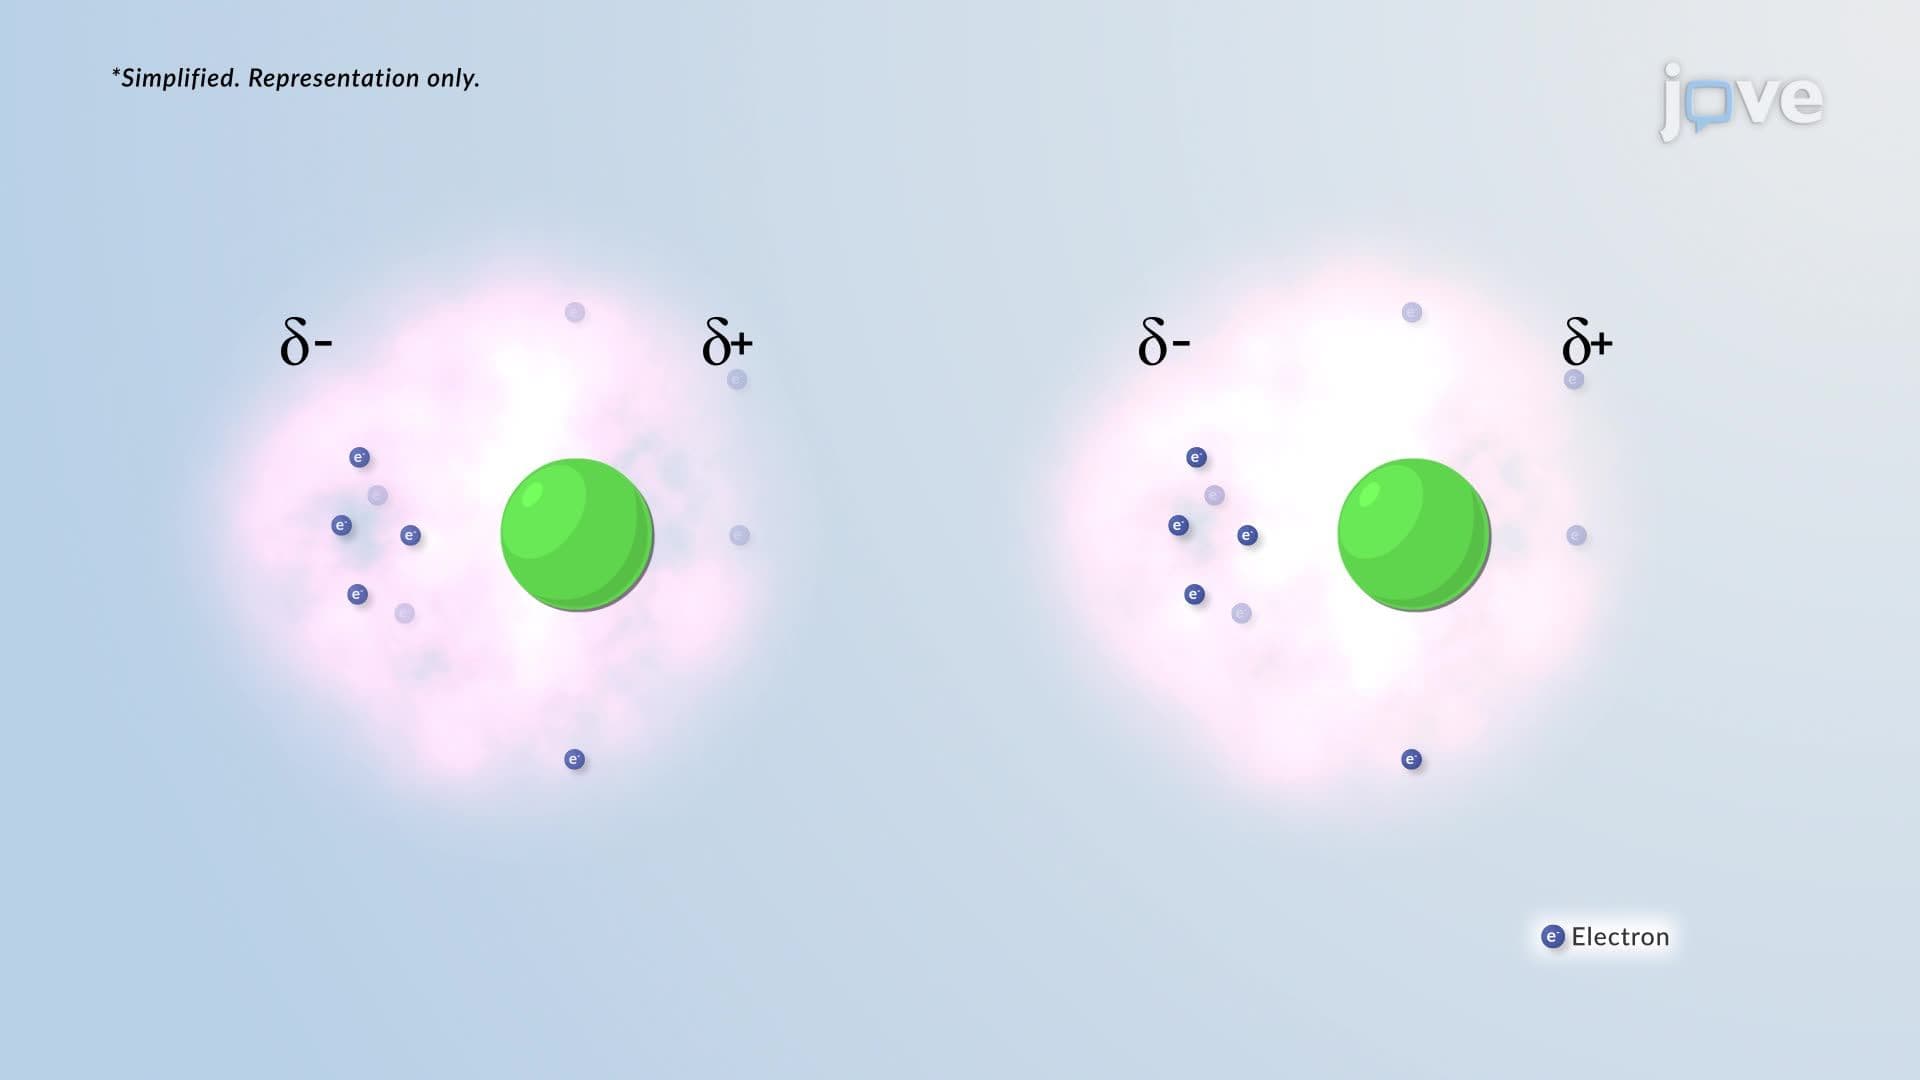
Van der Waals Interactions
 02:35
Molecular Orbital Theory I