Related Experiment Video
Updated: Jun 26, 2026

Implantation of Osmotic Pumps and Induction of Stress to Establish a Symptomatic, Pharmacological Mouse Model for DYT/PARK-ATP1A3 Dystonia
Published on: September 12, 2020
Paroxysmal non-kinesigenic dyskinesia is caused by mutations of the MR-1 mitochondrial targeting sequence
Daniele Ghezzi1, Carlo Viscomi, Alessandra Ferlini
1Pierfranco and Luisa Mariani Center for the Study of Children's Mitochondrial Disorders, National Neurological Institute Carlo Besta, Milan, Italy.

Abstract:
Paroxysmal non-kinesigenic dyskinesia (PNKD) is an autosomal-dominant movement disorder characterized by attacks of dystonia, chorea and athetosis. Myofibrillogenesis regulator-1 (MR-1), the gene responsible for PNKD, is transcribed into three alternatively spliced forms: long (MR-1L), medium (MR-1M) and small (MR-1S). Two mutations, A7V and A9V, were previously discovered in the N-terminal region common to MR-1L and MR-1S. We now found a third mutation, A33P, in a new PNKD patient in the same region. Contrary to previous reports, we show here that the mutation-free MR-1M is localized in the Golgi apparatus, ER and plasma membrane, whereas both MR-1L and MR-1S isoforms are mitochondrial proteins, imported into the organelle thanks to a 39 amino acid-long, N-terminal mitochondrial targeting sequence (MTS). The MTS, which contains all three PNKD mutations, is then cleaved off the mature proteins before their insertion in the inner mitochondrial membrane. Therefore, mature MR-1S and MR-1L of PNKD patients are identical to those of normal subjects. We found no difference in import efficiency and protein maturation between wild-type and mutant MR-1 variants. These results indicate that PNKD is due to a novel disease mechanism based on a deleterious action of the MTS.
Insights
Paroxysmal non-kinesigenic dyskinesia (PNKD) is linked to mutations in the MR-1 gene. The disease mechanism involves a faulty mitochondrial targeting sequence, not the mature protein itself.
Area of Science:
- Neurogenetics
- Molecular Biology
- Cell Biology
Background:
- Paroxysmal non-kinesigenic dyskinesia (PNKD) is an inherited movement disorder.
- The MR-1 gene, responsible for PNKD, has three splice variants: MR-1L, MR-1M, and MR-1S.
- Previous studies identified two mutations (A7V, A9V) in the N-terminal region of MR-1.
Observation:
- A third mutation (A33P) was identified in a new PNKD patient within the same N-terminal region.
- MR-1M localizes to the Golgi, ER, and plasma membrane, while MR-1L and MR-1S are mitochondrial proteins.
- A 39-amino acid N-terminal mitochondrial targeting sequence (MTS) facilitates import of MR-1L and MR-1S into mitochondria.
Findings:
- The MTS, containing all identified PNKD mutations, is cleaved during protein maturation.
- Mature MR-1S and MR-1L proteins in PNKD patients are identical to those in healthy individuals.
- No differences in import efficiency or maturation were observed between wild-type and mutant MR-1 variants.
Implications:
- PNKD pathogenesis may stem from a detrimental effect of the MTS itself.
- This suggests a novel disease mechanism for PNKD, distinct from previously assumed protein dysfunction.
- Understanding the MTS's role could lead to new therapeutic strategies for PNKD.
More Related Videos

Related Concept Videos
 01:48
ATP Synthase: Mechanism
 01:24
Parkinson Disease l: Introduction
 01:39
Mutations
 01:35
Mutations
Chromosomal Alterations Are Large-Scale Mutations
While point mutations are changes in a single nucleotide in...
 01:12
Alterations in Muscle Tone ll
 01:19
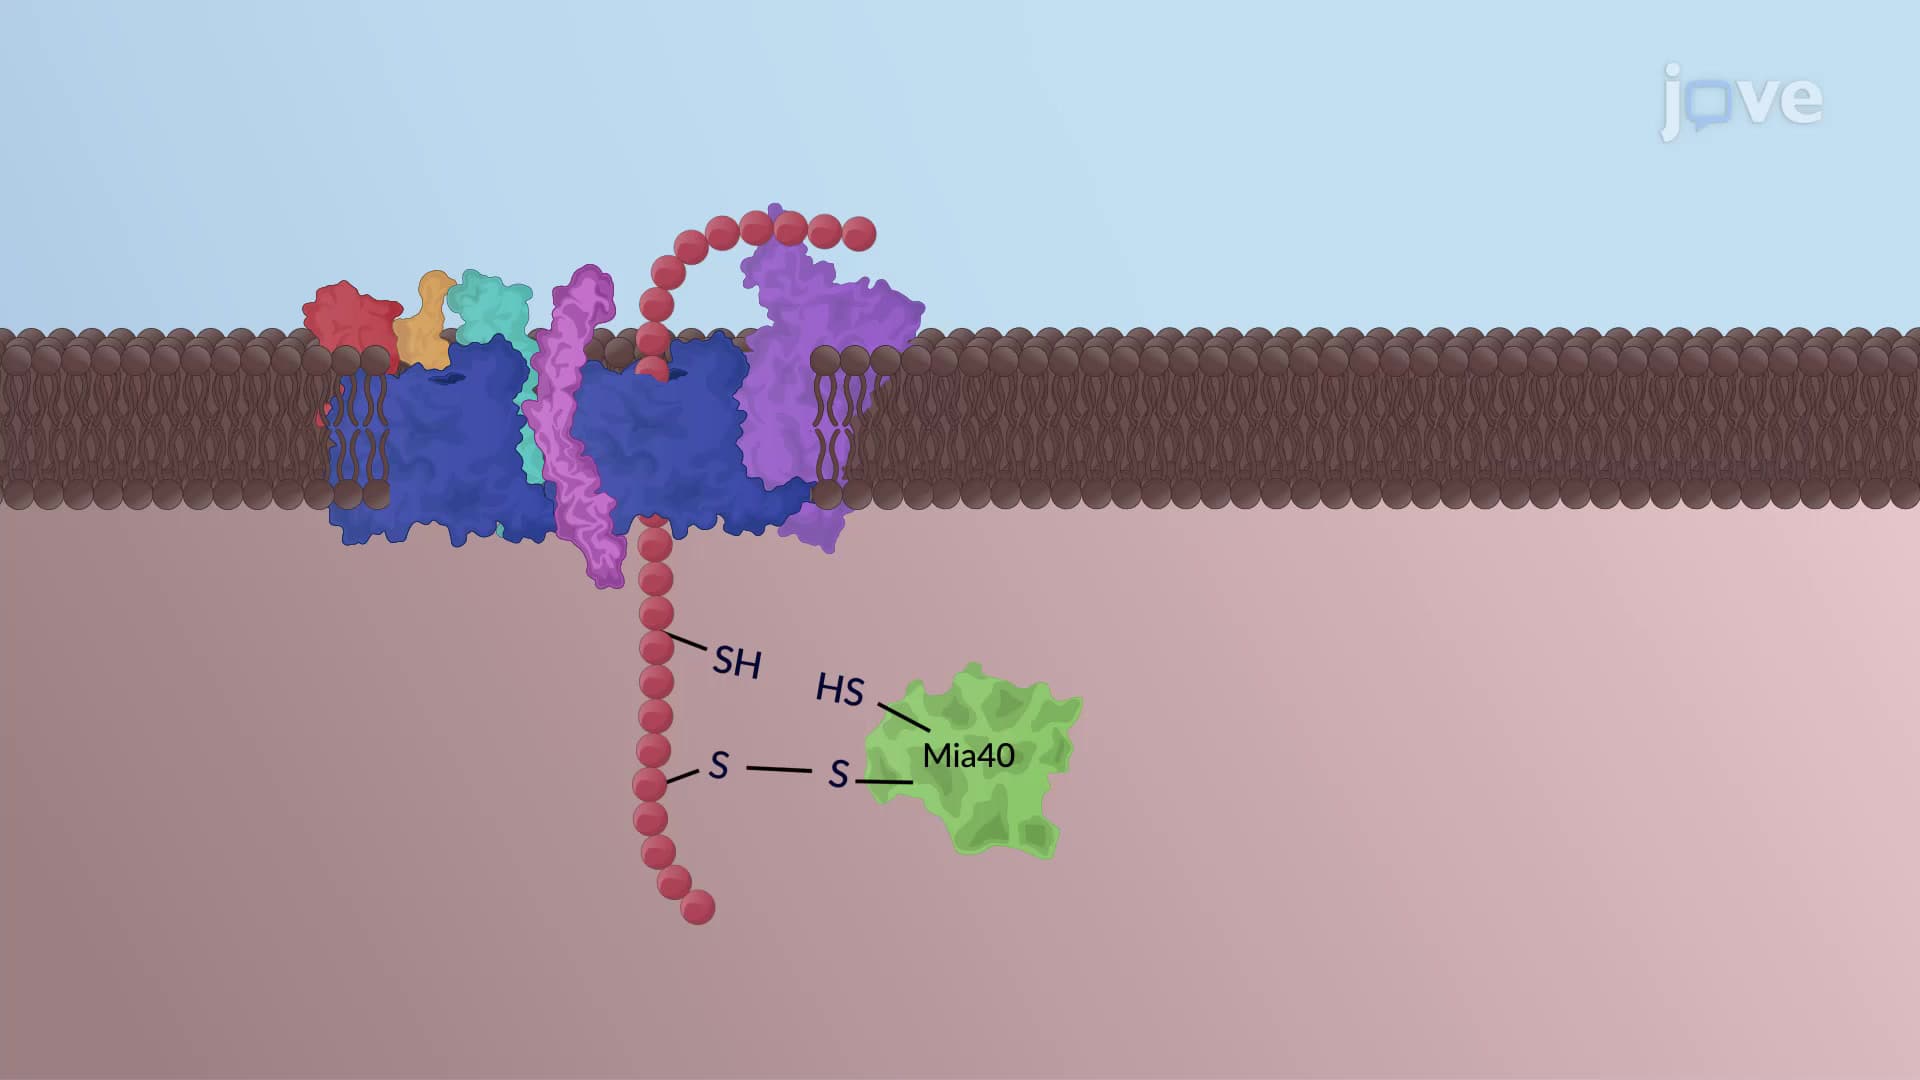
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...