Related Experiment Video
Updated: Jun 26, 2026

Diagonal Method to Measure Synergy Among Any Number of Drugs
Published on: June 21, 2018
Interaction of Oxicam NSAIDs with lipid monolayer: anomalous dependence on drug concentration
Sarathi Kundu1, Hirak Chakraborty, Munna Sarkar
1Surface Physics Division, Saha Institute of Nuclear Physics, 1/AF Bidhannagar, Kolkata 700064, India.

Abstract:
Surface pressure (pi) versus specific molecular area (A) isotherms of Langmuir monolayers of dimyristoylphosphatidylcholine (DMPC) lipid on pure water were studied in pristine form and in presence of three non-steroidal anti-inflammatory drugs, meloxicam (MX), piroxicam (PX) and tenoxicam (TX) in the subphase. Data were taken at three drug/lipid (D/L) ratios of 0.026, 0.05, and 0.1. Integration of drug to the lipid monolayer was measured by the increase in A (Delta A) of DMPC monolayer due to the presence of drugs. All three drugs could be integrated in the monolayer resulting in a positive value of Delta A for D/L ratio of 0.026. Above this D/L value, there is an anomalous, monotonic decrease in Delta A for MX and TX resulting, finally, in negative Delta A values. For PX, however, decrease in Delta A values at D/L of 0.05 is partially compensated at D/L of 0.1. We have tentatively explained these observations by invoking two competing forces in the overall drug-lipid interaction. One of these is an 'in-plane' force that tends to integrate the drug molecule to the plane formed by the lipid monolayer and the other is an 'out-of-plane' force that perturbs the drug and the lipid molecules such that the monolayer plane is no longer well defined.
More Related Videos
 08:18
Self-Nanoemulsification of Healthy Oils to Enhance the Solubility of Lipophilic Drugs
Published on: July 27, 2022
 07:11
Dissipative Microgravimetry to Study the Binding Dynamics of the Phospholipid Binding Protein Annexin A2 to Solid-supported Lipid Bilayers Using a Quartz Resonator
Published on: November 1, 2018
Related Concept Videos
 01:21
Factors Affecting Dissolution: Drug pKa, Lipophilicity and GI pH
A drug's pKa and the pH of the gastrointestinal (GI) tract play crucial roles in drug...
 01:20
Factors Affecting Dissolution: Drug Permeability, Stability and Stereochemistry
 01:14
Time Course of Drug Effect
 01:27
Factors Influencing Drug Absorption: Drug Dissolution
 01:15
Pharmacodynamic Models: Linear Concentration–Effect Model
 01:30
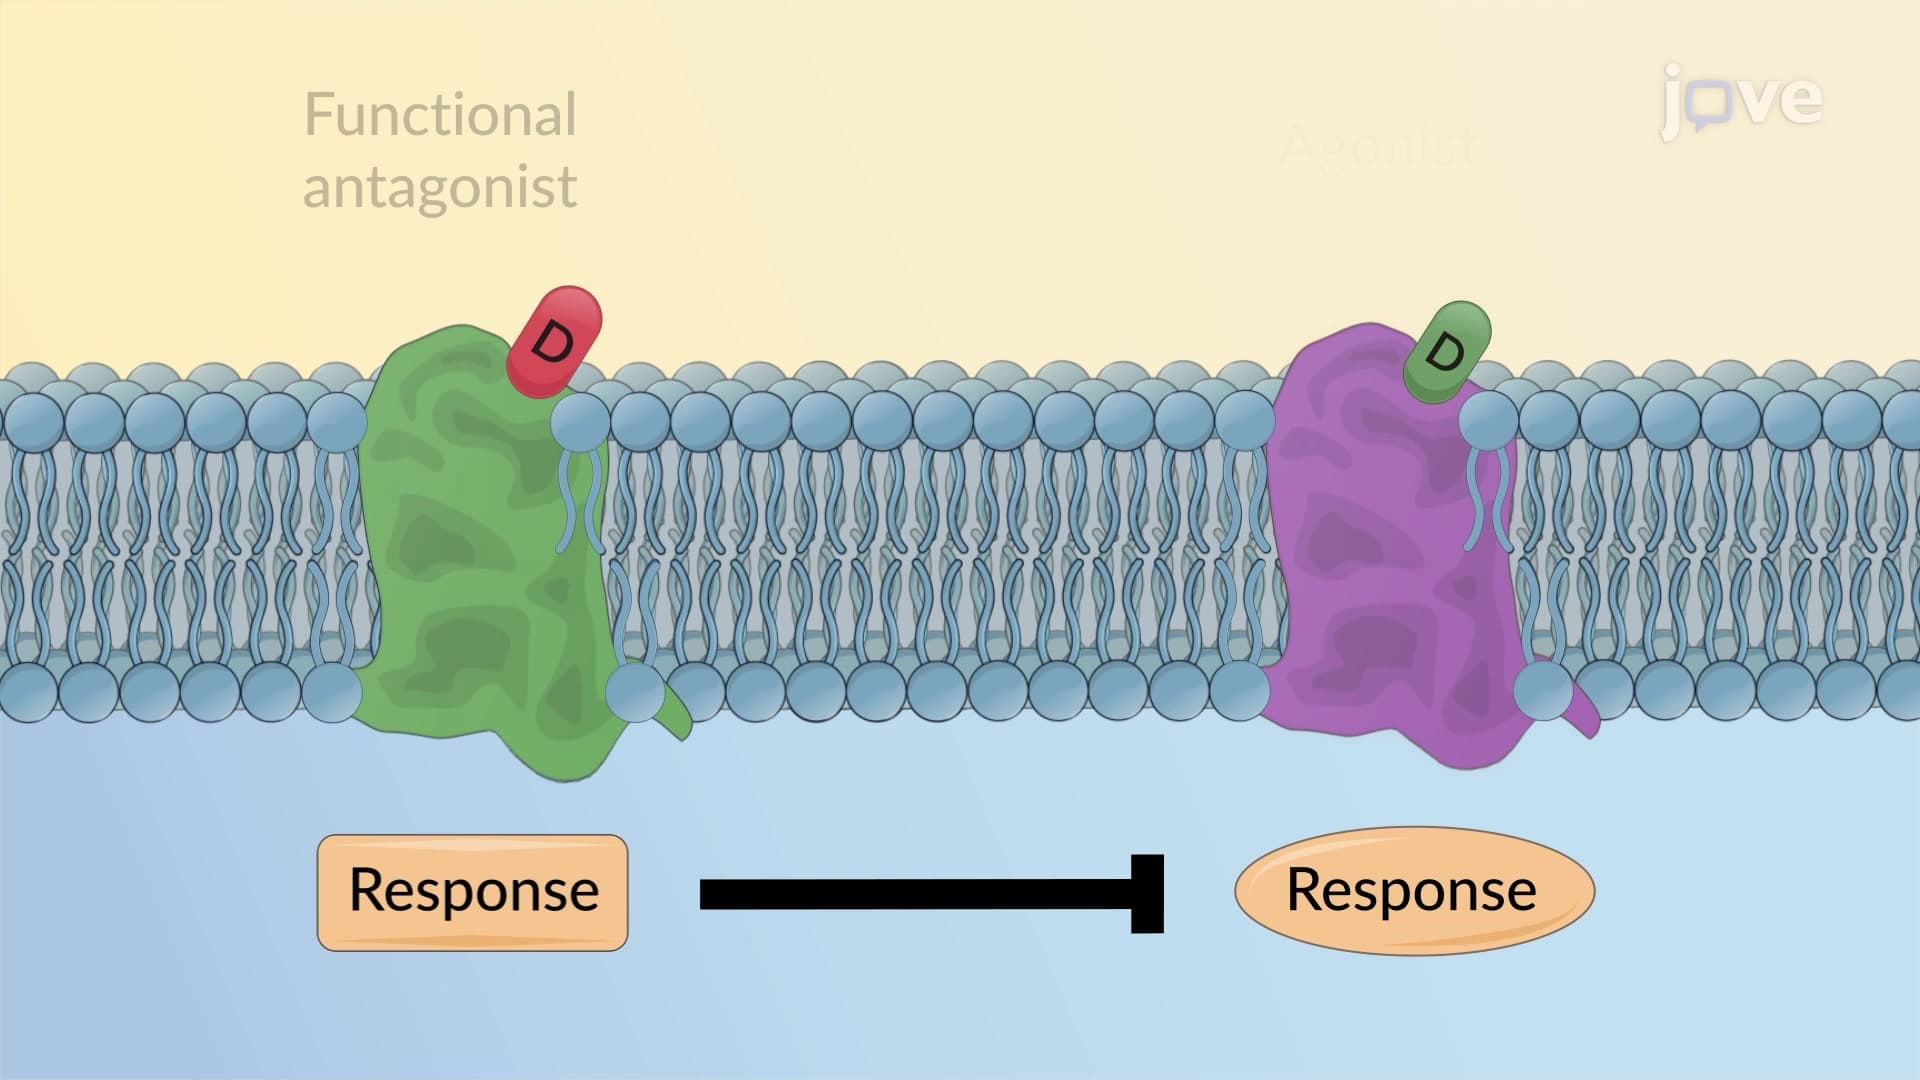
Combined Effects of Drugs: Antagonism
The most common type is receptor antagonism, where one drug acts as an antagonist to block the effects of another drug by...