Related Experiment Video
Updated: Jun 26, 2026

Modeling Mitochondrial Disease Using Brain Organoids: A Focus on Mitochondrial Encephalomyopathy, Lactic Acidosis, and Stroke-like Episodes
Published on: October 10, 2025
Infantile mitochondrial encephalomyopathy with unusual phenotype caused by a novel BCS1L mutation in an isolated
Alberto Blázquez1, Mari Carmen Gil-Borlado, María Morán
1Centro de Investigación, Hospital Universitario 12 de Octubre, Madrid, Spain.

Abstract:
Mutations in BCS1L, a respiratory chain complex III assembly chaperone, constitute a major cause of mitochondrial complex III deficiency and are associated with GRACILE and Björnstad syndromes. Here we describe a 4-year-old infant with hyperlactacidemia, mild liver dysfunction, hypotonia, growth and psychomotor retardation, dysmorphic features and mitochondrial complex III deficiency. Respiratory chain enzyme activities showed an isolated complex III defect in muscle and fibroblasts. Sequencing and polymerase chain reaction-restriction fragment length polymorphism (PCR-RFLP) analysis revealed a novel homozygous BCS1L mutation, c.148A>G, which caused a p.T50A substitution at an evolutionarily conserved BCS1L region. The severity of the complex III enzyme defect correlated with decreased amounts of BCS1L and respiratory chain complex III in the affected tissues. Our findings support a pathogenic role for the novel BCS1L mutation in a patient with a singular clinical phenotype.
More Related Videos
 06:53
Visualization of Mitochondrial Respiratory Function using Cytochrome C Oxidase / Succinate Dehydrogenase (COX/SDH) Double-labeling Histochemistry
Published on: November 23, 2011
 05:45
Isolation of Mitochondria for Mitochondrial Supercomplex Analysis from Small Tissue and Cell Culture Samples
Published on: May 3, 2024
Related Concept Videos
 02:59
Animal Mitochondrial Genetics
 01:43
Electron Transport Chain: Complex III and IV
 01:46
Electron Transport Chain: Complex I and II
ROS generation is regulated and maintained at moderate levels necessary...
 01:20
Inborn Errors of Metabolism
 01:39
Mitochondrial Precursor Proteins
Most of the mitochondrial precursors...
 01:41
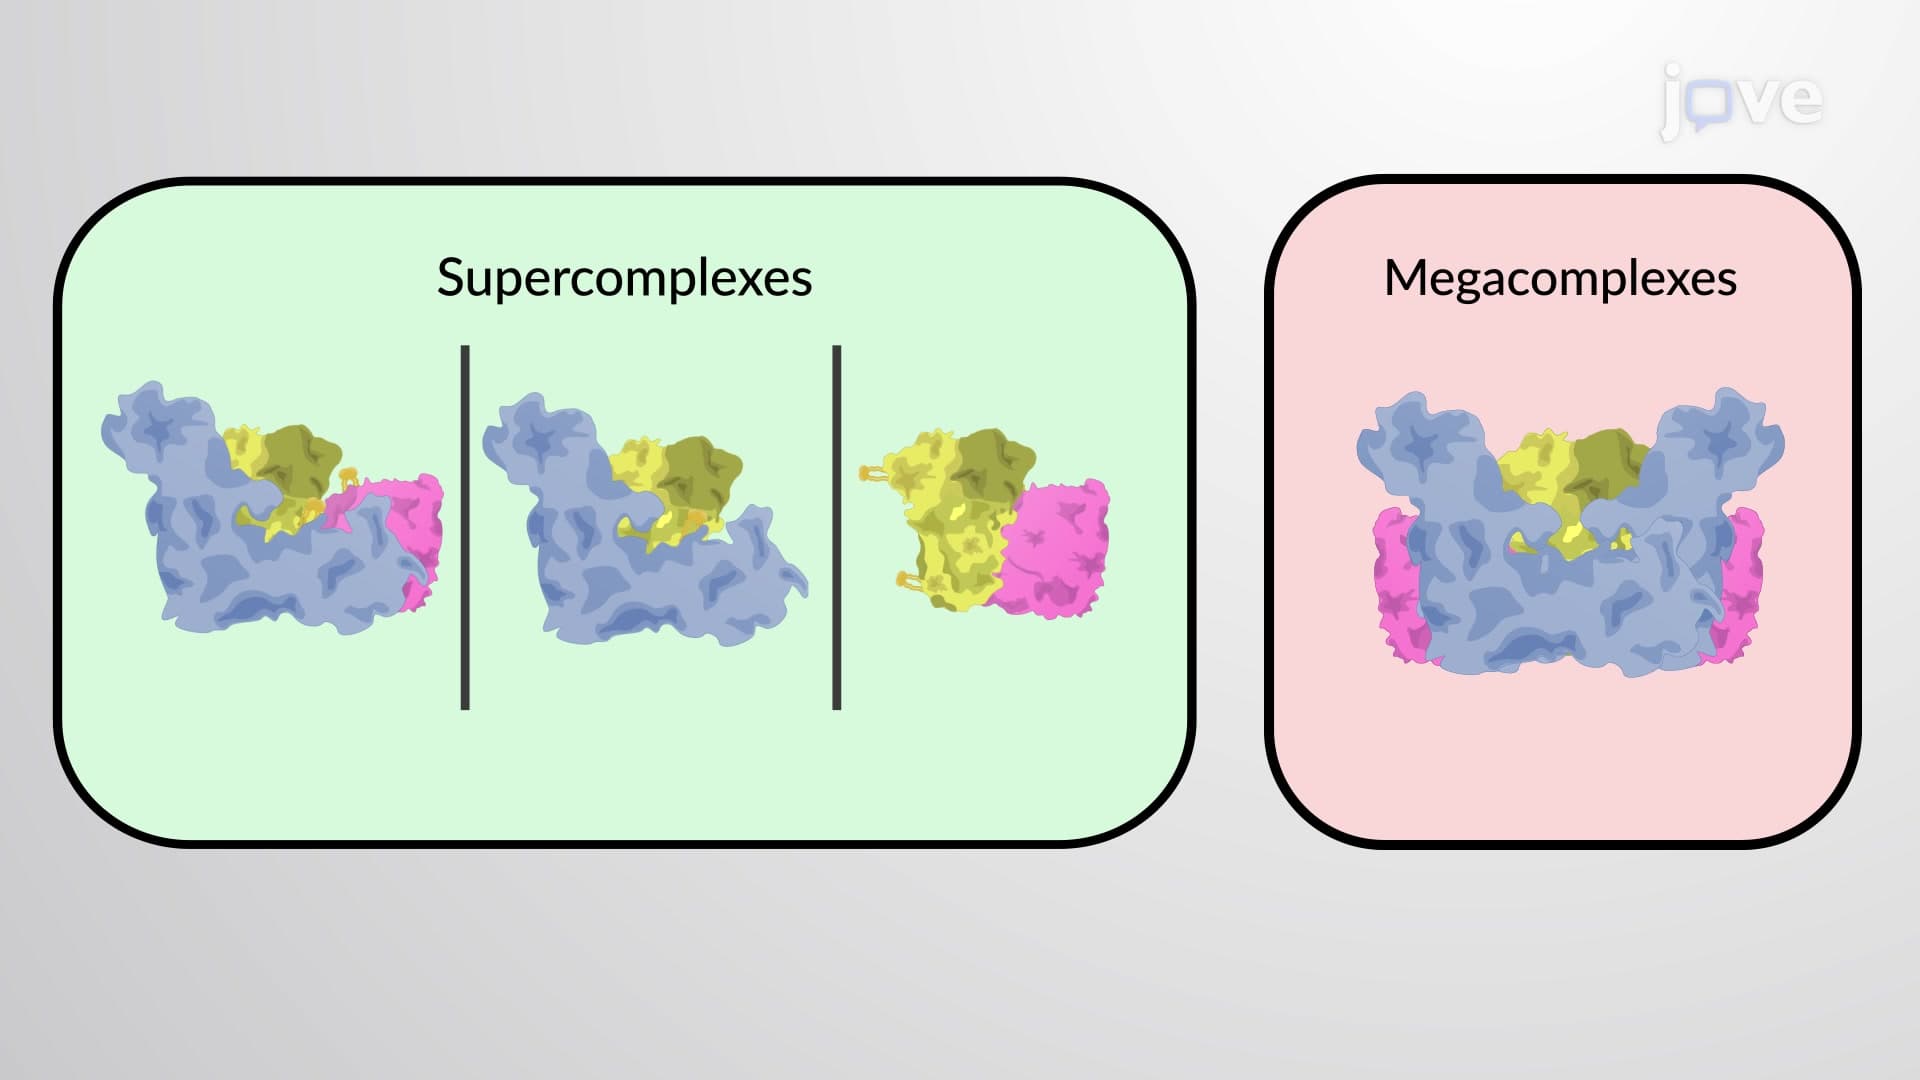
The Supercomplexes in the Crista Membrane