EMMA 2--a MAGE-compliant system for the collaborative analysis and integration of microarray data
Related Experiment Video
Updated: Jun 25, 2026
Informatic Analysis of Sequence Data from Batch Yeast 2-Hybrid Screens
Published on: June 28, 2018
Michael Dondrup1, Stefan P Albaum, Thasso Griebel
1Computational Genomics, Center for Biotechnology, Bielefeld University, 33594 Bielefeld, Germany. mdondrup@cebitec.uni-bielefeld.de

Background:
Understanding transcriptional regulation by genome-wide microarray studies can contribute to unravel complex relationships between genes. Attempts to standardize the annotation of microarray data include the Minimum Information About a Microarray Experiment (MIAME) recommendations, the MAGE-ML format for data interchange, and the use of controlled vocabularies or ontologies. The existing software systems for microarray data analysis implement the mentioned standards only partially and are often hard to use and extend. Integration of genomic annotation data and other sources of external knowledge using open standards is therefore a key requirement for future integrated analysis systems.
Results:
The EMMA 2 software has been designed to resolve shortcomings with respect to full MAGE-ML and ontology support and makes use of modern data integration techniques. We present a software system that features comprehensive data analysis functions for spotted arrays, and for the most common synthesized oligo arrays such as Agilent, Affymetrix and NimbleGen. The system is based on the full MAGE object model. Analysis functionality is based on R and Bioconductor packages and can make use of a compute cluster for distributed services.
Conclusion:
Our model-driven approach for automatically implementing a full MAGE object model provides high flexibility and compatibility. Data integration via SOAP-based web-services is advantageous in a distributed client-server environment as the collaborative analysis of microarray data is gaining more and more relevance in international research consortia. The adequacy of the EMMA 2 software design and implementation has been proven by its application in many distributed functional genomics projects. Its scalability makes the current architecture suited for extensions towards future transcriptomics methods based on high-throughput sequencing approaches which have much higher computational requirements than microarrays.
Related Concept Videos
 02:34
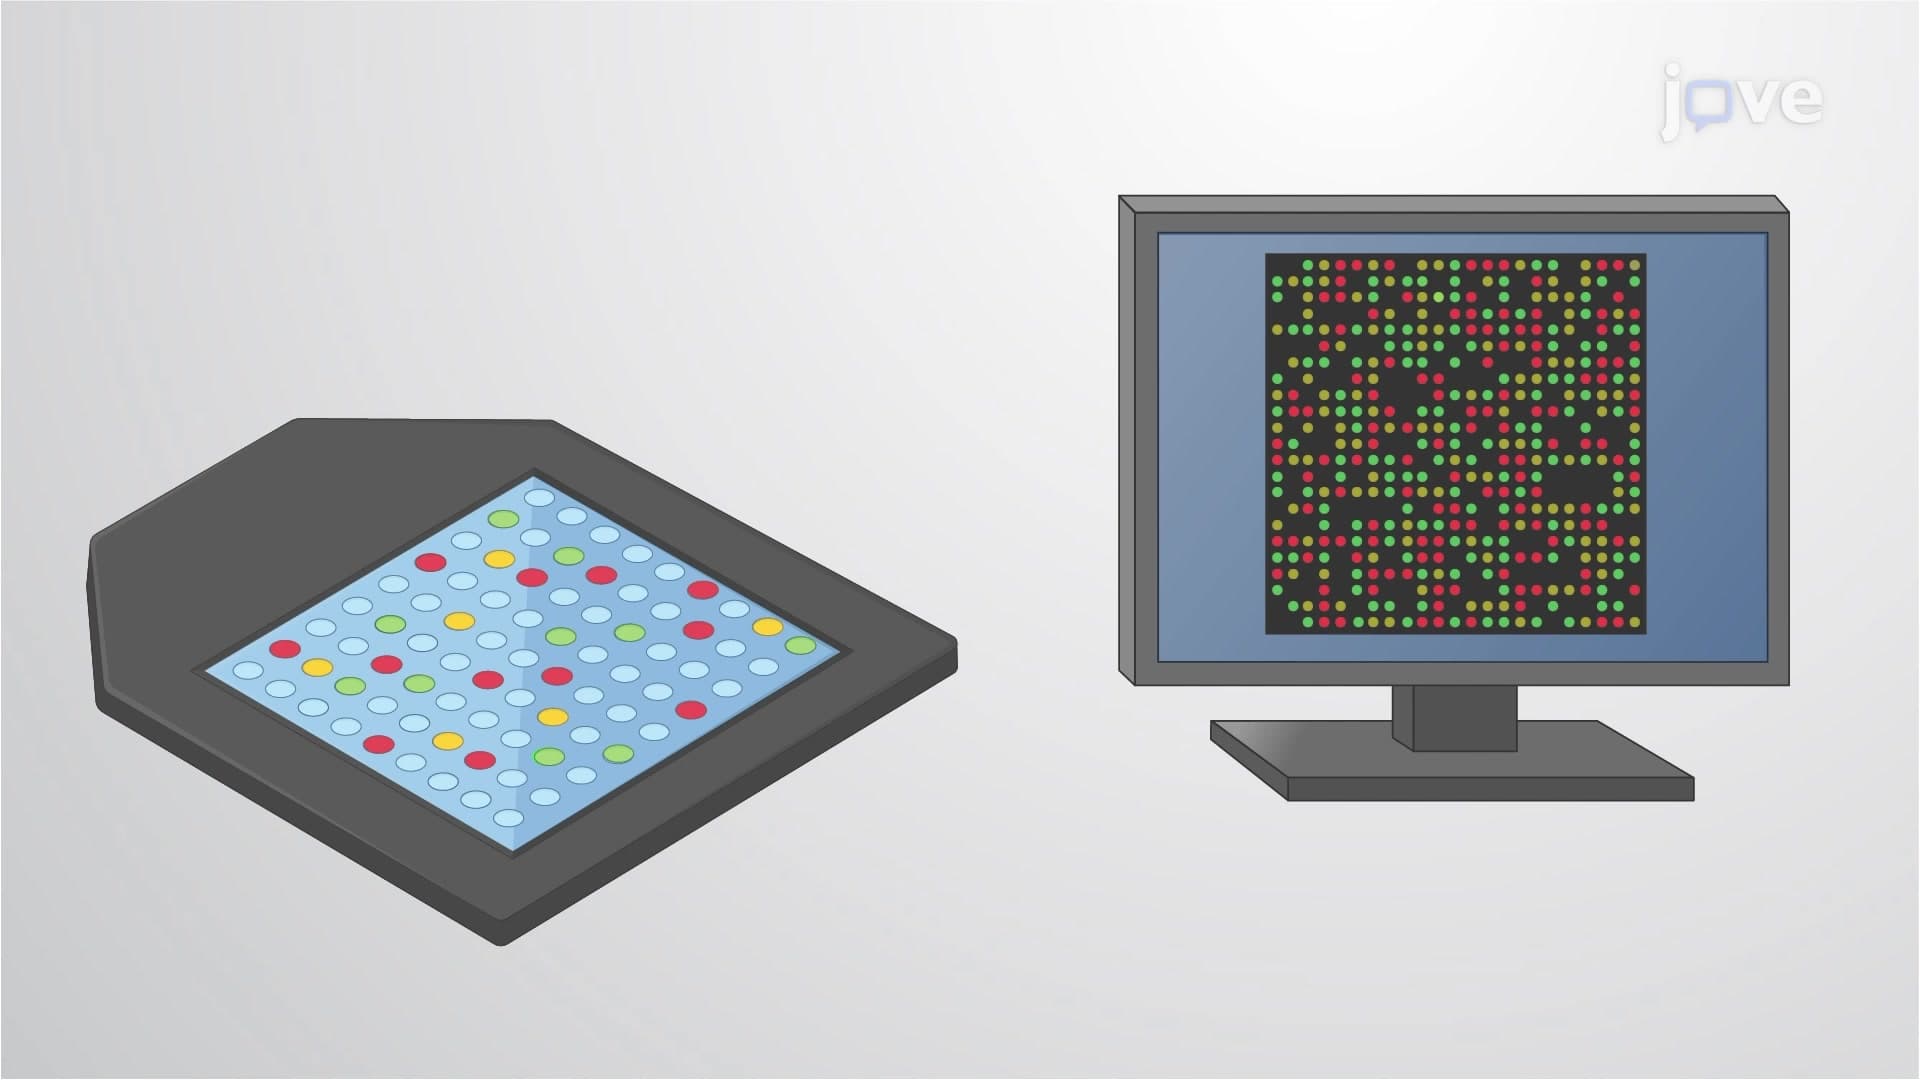
DNA Microarrays
 01:33
Proteomics
Proteomics is the study of proteomes' function. It involves the large-scale systematic study of the proteome to denote the protein complement expressed by a genome. Scientist Mark Wilkins coined the term proteomics...