Predicting protein pK(a) by environment similarity
Related Experiment Video
Updated: Jun 25, 2026

A Protocol for Computer-Based Protein Structure and Function Prediction
Published on: November 3, 2011
Francesca Milletti1, Loriano Storchi, Gabriele Cruciani
1Department of Chemistry, Università degli Studi di Perugia, Italy.

Abstract:
A statistical method to predict protein pK(a) has been developed by using the 3D structure of a protein and a database of 434 experimental protein pK(a) values. Each pK(a) in the database is associated with a fingerprint that describes the chemical environment around an ionizable residue. A computational tool, MoKaBio, has been developed to identify automatically ionizable residues in a protein, generate fingerprints that describe the chemical environment around such residues, and predict pK(a) from the experimental pK(a) values in the database by using a similarity metric. The method, which retrieved the pK(a) of 429 of the 434 ionizable sites in the database correctly, was crossvalidated by leave-one-out and yielded root mean square error (RMSE) = 0.95, a result that is superior to that obtained by using the Null Model (RMSE 1.07) and other well-established protein pK(a) prediction tools. This novel approach is suitable to rationalize protein pK(a) by comparing the region around the ionizable site with similar regions whose ionizable site pK(a) is known. The pK(a) of residues that have a unique environment not represented in the training set cannot be predicted accurately, however, the method offers the advantage of being trainable to increase its predictive power.
More Related Videos
 06:50
Computational Prediction of Amino Acid Preferences of Potentially Multispecific Peptide-Binding Domains Involved in Protein-Protein Interactions
Published on: January 26, 2024
 14:34
A Bilingual Computational Workflow for Identifying Potential PLK1 Inhibitors in American Sign Language and English
Published on: April 3, 2026
Related Concept Videos
 01:13
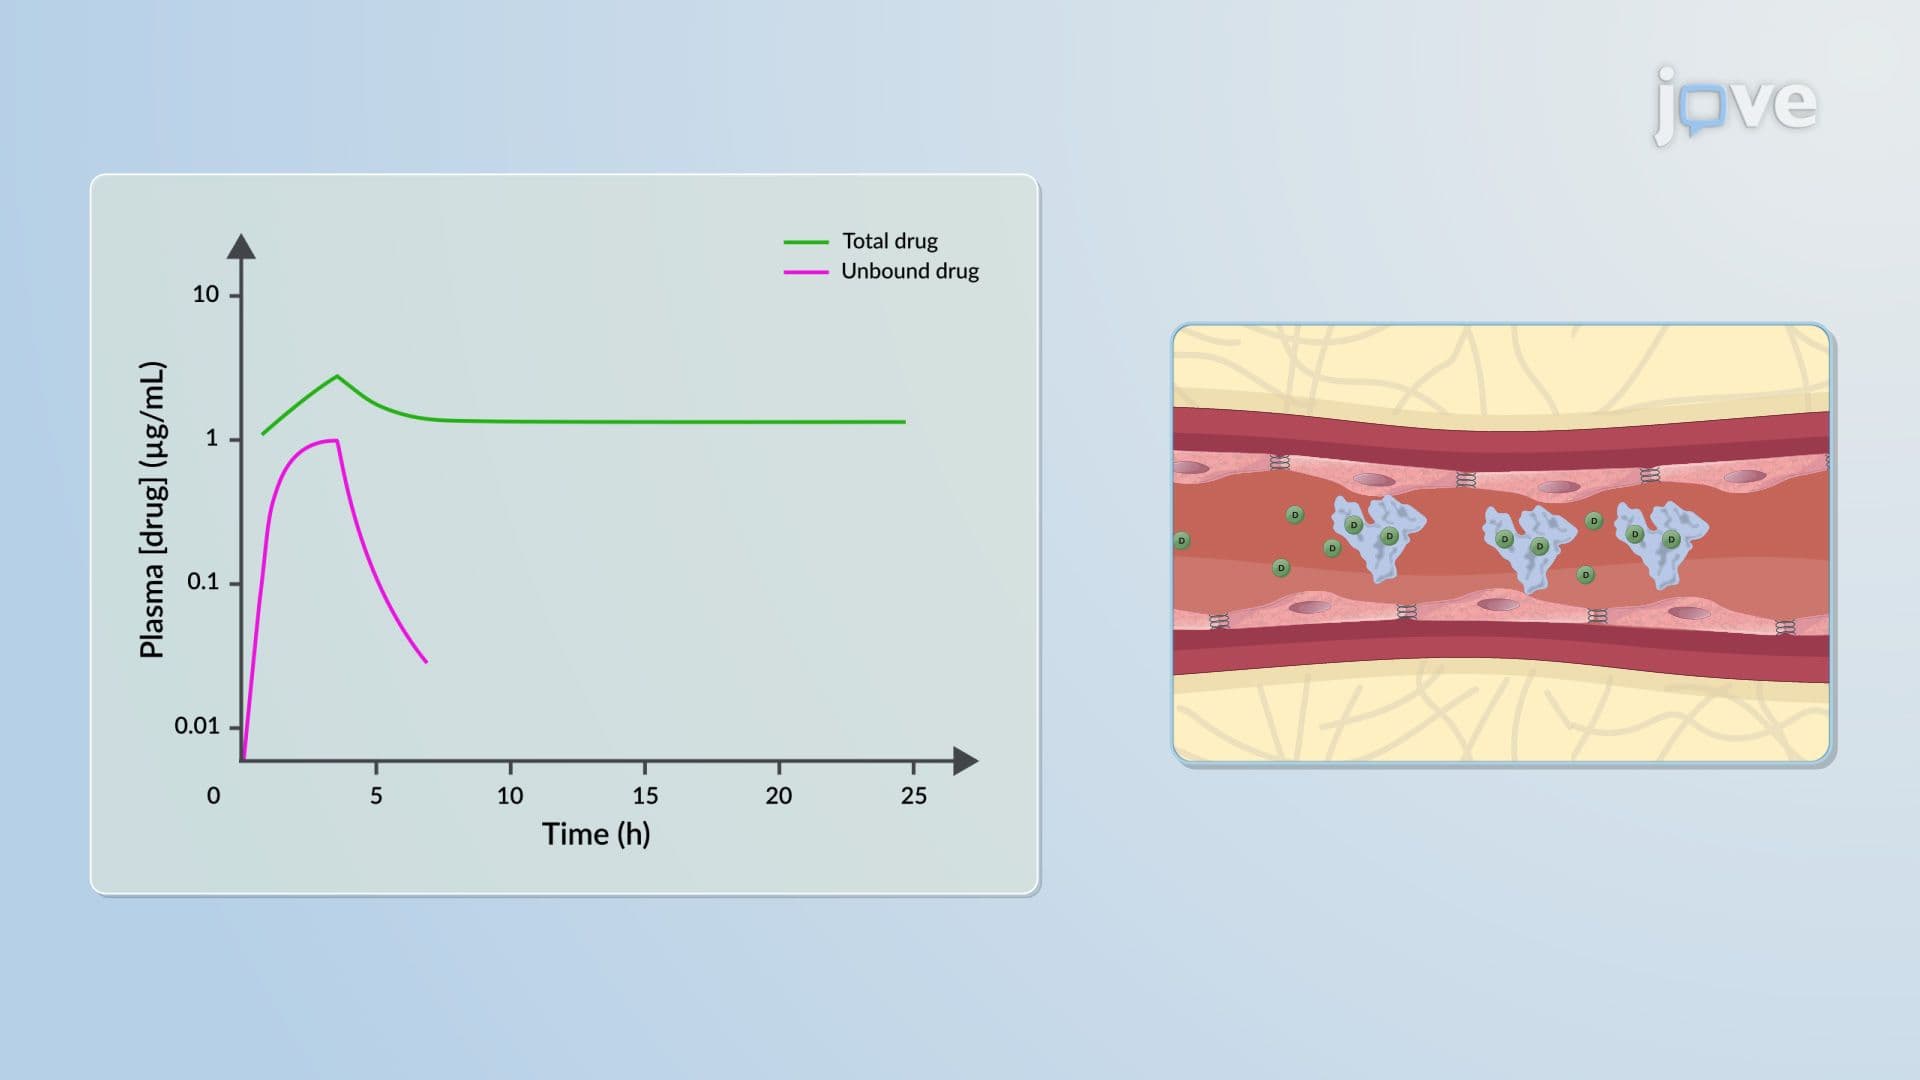
Physiological Pharmacokinetic Models: Assumption with Protein Binding
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 02:26
Protein Networks
These interactions can be represented through maps depicting protein-protein interaction networks, represented as nodes and edges. Nodes are circles that are representative of a protein,...
 02:04
Protein-protein Interfaces
 01:24
Pharmacokinetic–Pharmacodynamic Relationship: Problems
 01:14
Pharmacokinetic–Pharmacodynamic Relationship: Model Components