Related Experiment Video
Updated: Jun 25, 2026


Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
Homology modeling of MCH1 receptor and validation by docking/scoring and protein-aligned CoMFA
Areej Abu-Hammad1, Waleed A Zalloum, Hiba Zalloum
1Department of Pharmaceutical Sciences, Faculty of Pharmacy, University of Jordan, Queen Rania St, Amman, Jordan.

Abstract:
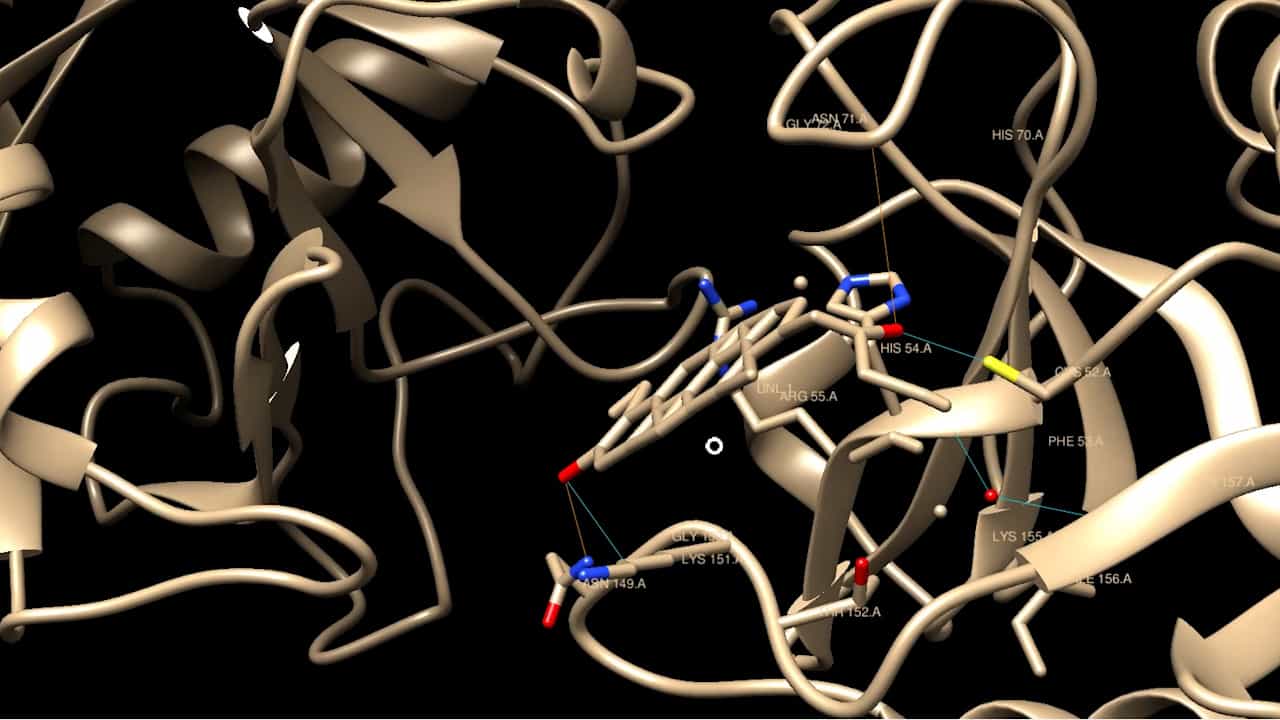
Homology modeling is becoming a valid method for obtaining three-dimensional coordinates for proteins. However, it is hard to judge the qualities of the resulting models warranting robust subsequent validations. In an attempt to evaluate the quality of Melanin-concentrating hormone 1 receptor (MCH1R) homology models, a number of homology structures were scanned for potential binding cavities. Subsequently, a group of 35 benzylpiperidines' MCH1R inhibitors were docked into each of the proposed binding sites via four different scoring functions. The docked structures were utilized to construct corresponding protein-aligned comparative molecular field analysis (CoMFA) models by employing probe-based (H(+), OH, CH(3)) energy grids and genetic partial least squares (G/PLS) statistical analysis. The docking-based alignment succeeded in accessing self-consistent CoMFA models upon employing JAIN scoring function in one of the proposed binding pockets in a particular homology model. Furthermore, a ligand-based pharmacophore model was developed for the same set of inhibitors and was found to agree with the successful docking configuration. Therefore, we proved that the overall procedure of docking, scoring, and CoMFA evaluation can be a useful tool to validate homology models, which can be of value for structure-based design, in-silico screening, and in understanding the structural basis of ligand binding to MCH1R.
Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 01:57
Gene Families
Occasionally these regions can be adapted to take on new roles within the organism, becoming novel genes...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...