Related Experiment Video
Updated: Jun 25, 2026


Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Structural Model Studies for the High-Valent Intermediate Q of Methane Monooxygenase from Broken-Symmetry Density
1Department of Molecular Biology TPC15 The Scripps Research Institute 10550 N. Torrey Pines Rd. La Jolla, California 92037.

Abstract:
Mössbauer isomer shift parameters have been obtained for both density functional theory (DFT) OPBE and OLYP functionals by linear regressions between the measured isomer shifts and calculated electron densities at Fe nuclei for a number of Fe(2+,2.5+) and Fe(2.5+,3+,3.5+,4+) complexes grouped separately. The calculated isomer shifts and quadrupole splittings on the sample Fe complexes from OPBE and OLYP functionals are similar to those of PW91 calculations (J. Comput. Chem. 27 (2006) 1292), however the fit parameters from the linear regressions differ between PW91 and OPBE, OLYP. Four models for the active site structure of the hydroxylase component of soluble methane monooxygenase (MMOH) have been studied, using three DFT functionals OPBE, OLYP, and PW91, incorporated with broken-symmetry methodology and the conductor-like screening (COSMO) solvation model. The calculated properties, including optimized geometries, electronic energies, pK(a)'s, Fe net spin populations, and Mössbauer isomer shifts and quadrupole splittings, have been reported and compared with available experimental values. The high-spin antiferromagnetically (AF) coupled Fe(4+) sites are correctly predicted by OPBE and OLYP methods for all active site models. PW91 potential overestimates the Fe-ligand covalencies for some of the models because of spin crossover. Our calculations and data analysis support the structure (our current model II shown in Figure 8) proposed by Friesner and Lippard's group (J. Am. Chem. Soc. 123 (2001) 3836-3837), which contains an Fe(4+)(μ-O)(2)Fe(4+) center, one axial water which also H-bonds to both side chains of Glu243 and Glu114, and one bidentate carboxylate group from the side chain of Glu144, which is likely to represent the active site of MMOH-Q. A new model structure (model IV shown in Figure 9), which has a terminal hydroxo and a protonated His147 which is dissociated from a nearby Fe, is more asymmetric in its Fe(μ-O)(2)Fe diamond core, and is another very good candidate for intermediate Q.
More Related Videos
 05:51
Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 02:40
MO Theory and Covalent Bonding
 02:00
Molecular Models
 02:16
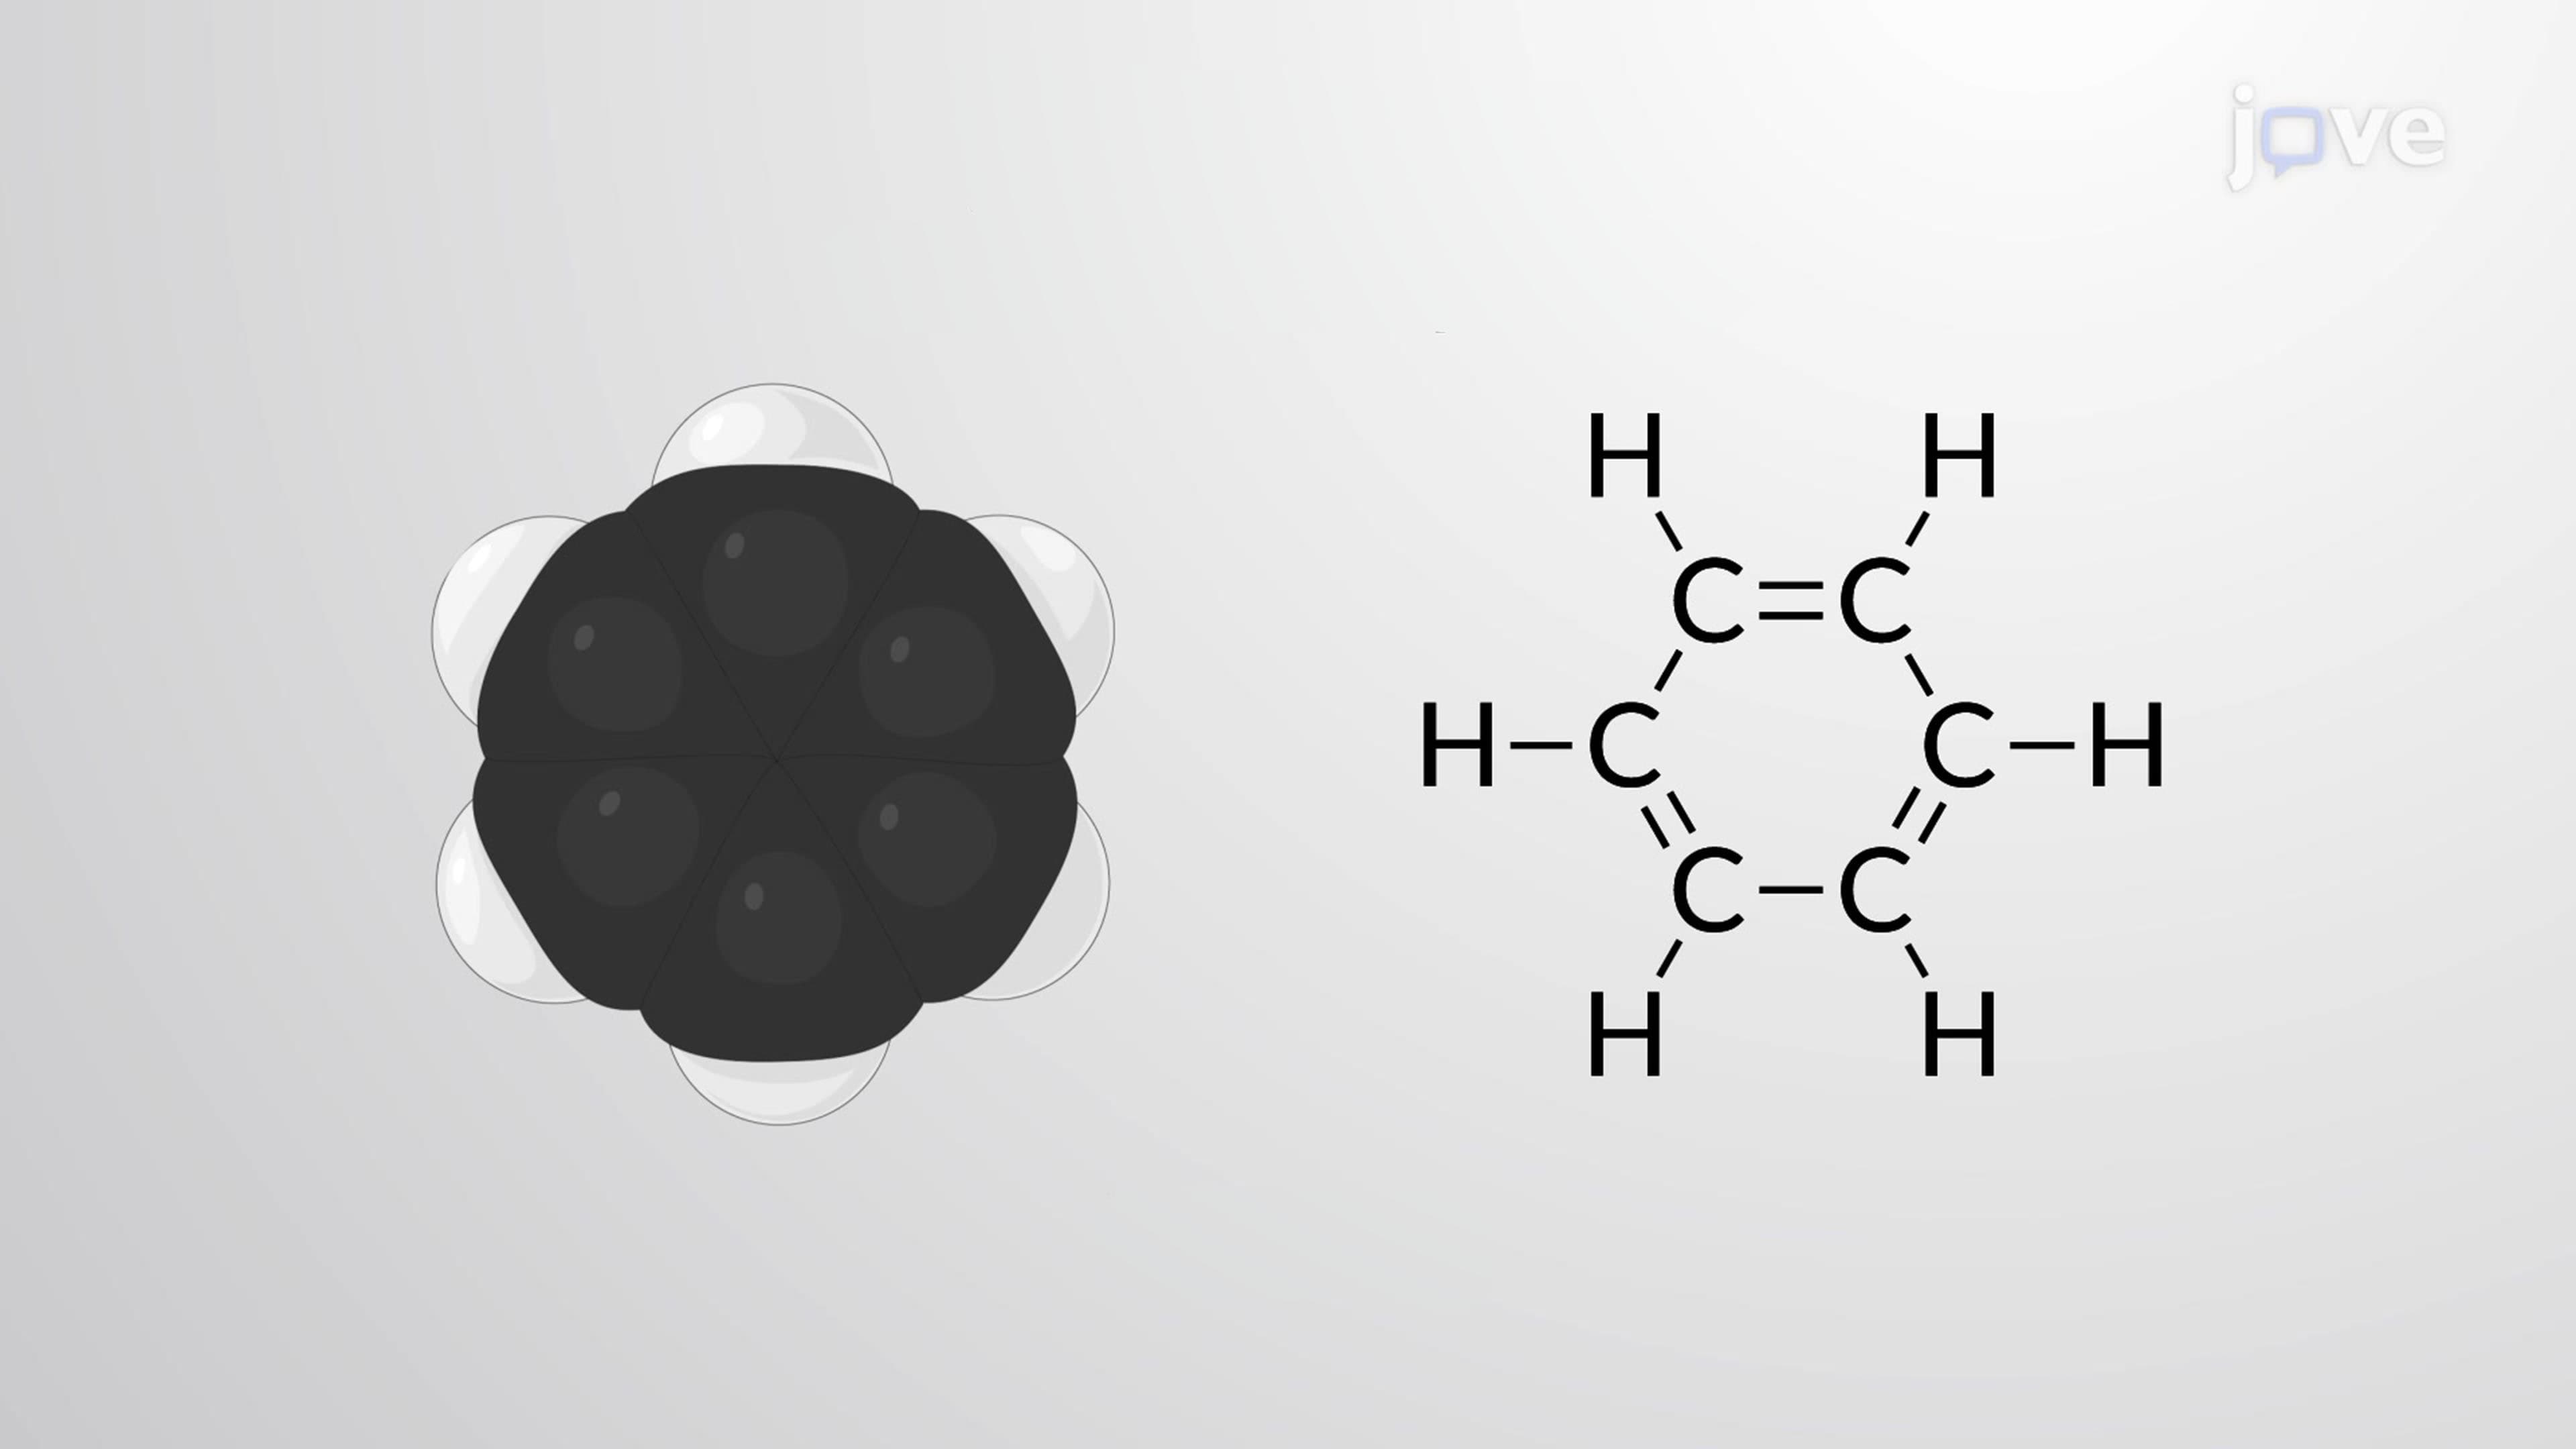
Resonance and Hybrid Structures
Resonance Structures and Resonance Hybrids
The Lewis structure of a nitrite anion (NO2−) may actually be drawn in two different ways, distinguished by the locations of the N–O and N=O bonds.
 03:51
Molecular Orbital Theory II
 03:24
Hybridization of Atomic Orbitals I
 02:52
Resonance