Related Experiment Video
Updated: Jun 24, 2026

Realistic Membrane Modeling Using Complex Lipid Mixtures in Simulation Studies
Published on: September 1, 2023
Mesoscale simulation of polymer reaction equilibrium: Combining dissipative particle dynamics with reaction ensemble
Martin Lísal1, John K Brennan, William R Smith
1E. Hala Laboratory of Thermodynamics, Institute of Chemical Process Fundamentals of the ASCR, v. v. i., 165 02 Prague 6-Suchdol, Czech Republic. lisal@icpf.cas.cz

Abstract:
We present an alternative formulation of the reaction ensemble dissipative particle dynamics (RxDPD) method [M. Lisal, J. K. Brennan, and W. R. Smith, J. Chem. Phys. 125, 16490 (2006)], a mesoscale simulation technique for studying polymer systems in reaction equilibrium. The RxDPD method combines elements of dissipative particle dynamics (DPD) and reaction ensemble Monte Carlo (RxMC), and is primarily targeted for the prediction of the system composition, thermodynamic properties, and phase behavior of reaction equilibrium polymer systems. The alternative formulation of the RxDPD method is demonstrated by considering a supramolecular diblock copolymer (SDC) melt in which two homopolymers, A(n) and B(m), can reversibly bond at terminal binding sites to form a diblock copolymer, A(n)B(m). We consider the effect of the terminal binding sites and the chemical incompatibility between A- and B-segments on the phase behavior. Both effects are found to strongly influence the resulting phase behavior. Due to the reversible nature of the binding, the SDC melt can be treated as the reaction equilibrium system A(n)+B(m)right harpoon over left harpoonA(n)B(m). To simulate the A(n)+B(m)right harpoon over left harpoonA(n)B(m) melt, the system contains, in addition to full A(n), B(m), and A(n)B(m) polymers, two fractional polymers: one fractional polymer either fA(n) or fB(m), and one fractional polymer fA(n)B(m), which have fractional particles at the ends of the polymer chains. These fractional particles are coupled to the system via a coupling parameter. The time evolution of the system is governed by the DPD equations of motion, accompanied by random changes in the coupling parameter. Random changes in the coupling parameter mimic forward and reverse reaction steps as in the RxMC approach, and they are accepted with a probability derived from the expanded ensemble grand canonical partition function. Unlike the original RxDPD method that considers coupling of entire fractional polymers to the system, the expanded ensemble framework allows a stepwise coupling, thus greatly increasing the efficiency of the RxDPD approach. The RxDPD technique rigorously satisfies thermodynamic equilibrium, but not the hydrodynamic behavior. However, the approximate treatment of the hydrodynamics can be minimized by simulating a large number of particles.
More Related Videos

Related Concept Videos
 01:17
Ziegler–Natta Chain-Growth Polymerization: Overview
 01:26
Reaction Mechanisms: The Steady-State Approximation
 00:53
Olefin Metathesis Polymerization: Acyclic Diene Metathesis (ADMET)
Similar to cross-metathesis, ADMET also involves the formation of metallacyclobutane intermediate by [2+2] cycloaddition of one of the double bonds of a terminal diene with...
 01:08
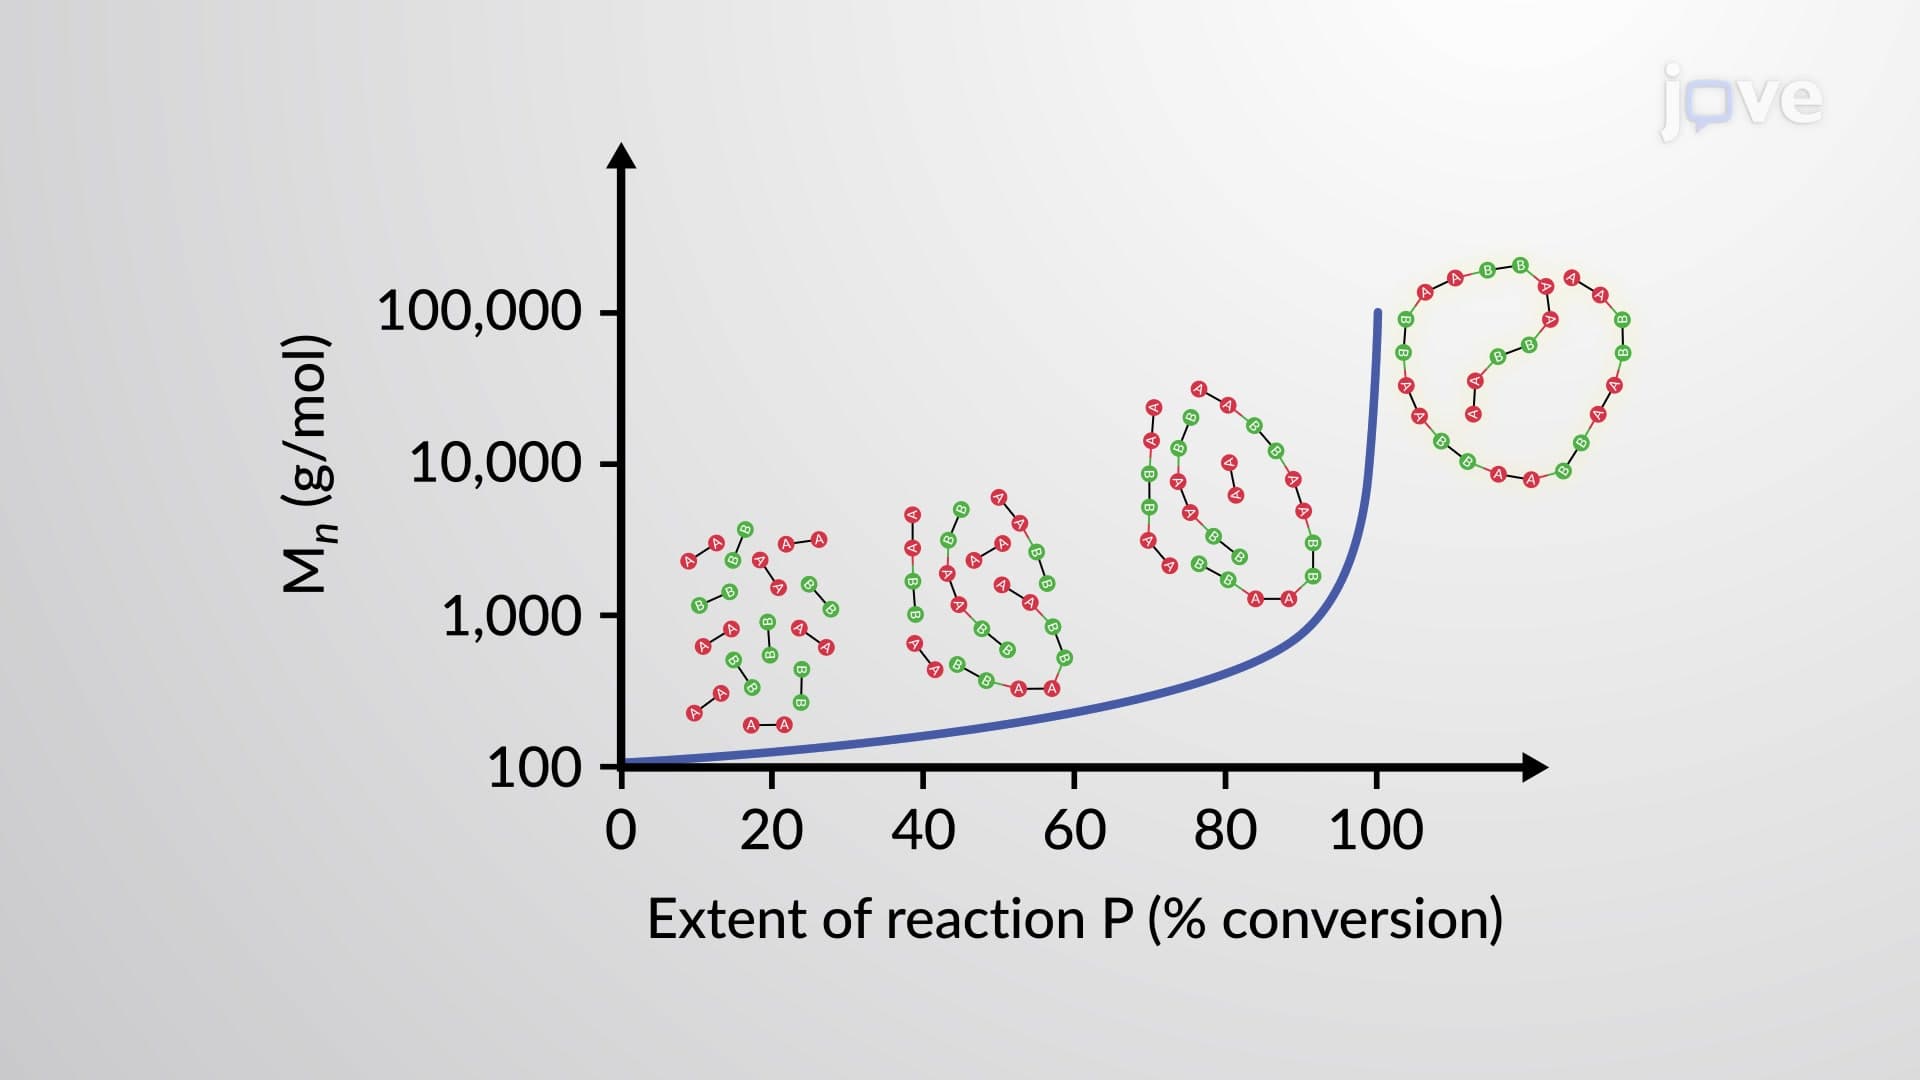
Molecular Weight of Step-Growth Polymers
As the step-growth polymerization involves step-wise condensation of monomers, the molecular weight also builds up eventually. Consequently, high molecular weight polymers are obtained at the late stages of the polymerization, where 99% of monomers have been consumed.
The extent of the...
 01:03
Step-Growth Polymerization: Overview
Many natural and synthetic polymers are produced by...
 01:29
Reaction Mechanisms: Rate-limiting Step Approximation