Related Experiment Video
Updated: Jun 24, 2026
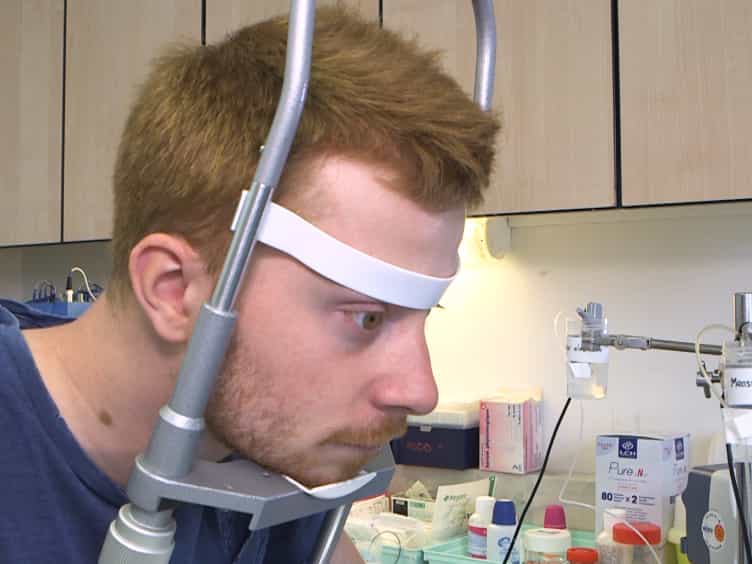
Standardized Measurement of Nasal Membrane Transepithelial Potential Difference (NPD)
Published on: September 13, 2018
Phenotypic characterisation of patients with intermediate sweat chloride values: towards validation of the European
C Goubau1, M Wilschanski, V Skalická
1Department of Paediatrics, Paediatric Pulmonology, University Hospital of Leuven, Herestraat 49, 3000 Leuven, Belgium.

Background:
In patients with symptoms suggestive of cystic fibrosis (CF) and intermediate sweat chloride values (30-60 mmol/l), extensive CFTR gene mutation analysis and nasal potential difference (NPD) measurement are used as additional diagnostic tests and a positive result in either test provides evidence of CFTR dysfunction. To define the phenotype of such patients and confirm the validity of grouping them, patients with intermediate sweat chloride values in whom either additional CF diagnostic test was abnormal were compared with subjects in whom this was not the case and patients with classic CF.
Methods:
The phenotypic features of four groups were compared: 59 patients with CFTR dysfunction, 46 with an intermediate sweat chloride concentration but no evidence of CFTR dysfunction (CF unlikely), 103 patients with CF and pancreatic sufficiency (CF-PS) and 62 with CF and pancreatic insufficiency (CF-PI).
Results:
The CFTR dysfunction group had more lower respiratory tract infections (p = 0.01), more isolation of CF pathogens (p<0.001) and clubbing (p = 0.001) than the CF unlikely group, but less frequent respiratory tract infections with CF pathogens than the CF-PS group (p = 0.05). Patients in the CF-PS group had a milder phenotype than those with PI. Many features showed stepwise changes through the patient groups.
Conclusion:
Patients with intermediate sweat chloride values and two CFTR mutations or an abnormal NPD measurement have a CF-like phenotype compatible with CFTR dysfunction and, as a group, differ phenotypically from patients with intermediate sweat chloride values in whom further CF diagnostic tests are normal as well as from CF-PS and CF-PI patients.
Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic sinusitis...
 01:27
Chronic Obstructive Pulmonary Disease-IV: Assessement and Diagnostic Studies
Medical History