
Vibrational analysis as a powerful tool in structure elucidation of polyarsenicals: a DFT-based investigation of
Graziano Guella1, Ines Mancini, Gino Mariotto
1Dipartimento di Fisica, Università di Trento, Via Sommarive 14, 38050 Povo, Trento, Italy. graziano.guella@unitn.it

Abstract:
A detailed experimental and theoretical vibrational analysis of arsenicin A, the first polyarsenical isolated from Nature, is reported. By exploiting the dissimilar and complementary selection rules of infrared (IR) absorption and Raman scattering transitions, and by carrying out density functional theory (DFT) calculations on several candidate structures, we are able to determine the structure of arsenicin A. We demonstrate that vibrational spectroscopy can be a very useful tool in structure elucidation in cases where mass spectrometry (MS) measurements lead to unreliable information, and nuclear magnetic resonance (NMR) investigations are hindered by the presence of silent nuclei, and/or elements of symmetry, in the molecule. Our approach allows to establish a reliable methodology, based on molecular vibrational parameters, for the structural elucidation of any unknown organic compounds of medium-small size on which, nowadays, accurate theoretical calculations can be carried out in reasonable times, without resorting to large-dimension computing resources.
Related Concept Videos
 01:18
Aromatic Hydrocarbon Anions: Structural Overview
Due to the absence of continuous overlap of p...
 01:08
IR Spectrum Peak Splitting: Symmetric vs Asymmetric Vibrations
 01:14
NMR Spectroscopy of Aromatic Compounds
 01:24
IR Spectroscopy: Molecular Vibration Overview
Stretching vibrations are vibrational motions that occur along the bond line, changing the bond length or distance between two bonded atoms. They are further distinguished as symmetric or asymmetric. In symmetric stretching, the...
 01:29
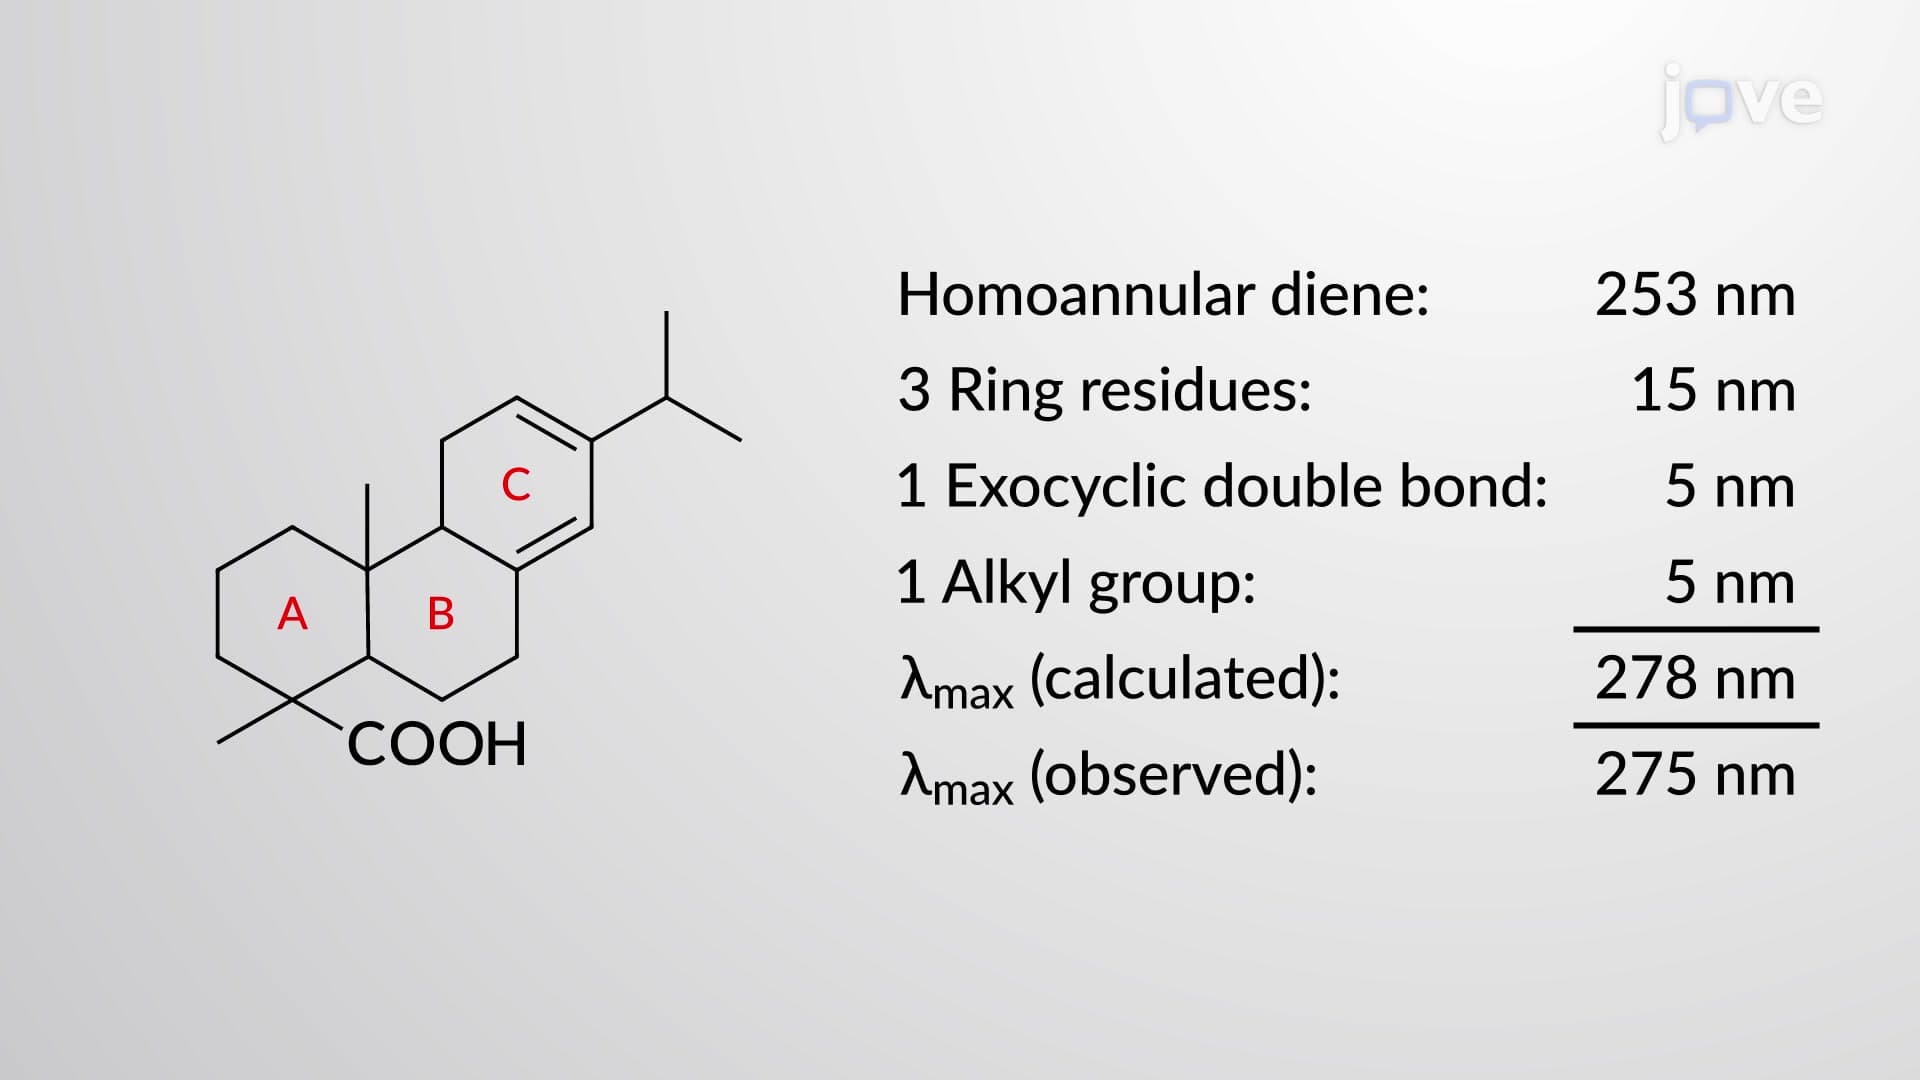
UV–Vis Spectroscopy: Woodward–Fieser Rules
 01:14
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds