Modified regional self-interaction corrected time-dependent density functional theory for core excited-state calculations
Related Experiment Video
Updated: Jun 23, 2026
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Ayako Nakata1, Takao Tsuneda, Kimihiko Hirao
1Department of Applied Chemistry, School of Engineering, The University of Tokyo, Tokyo 113-8656, Japan.

Abstract:
A modified regional self-interaction correction (mRSIC) method is proposed for obtaining accurate core-excitation energies in time-dependent density functional theory (TDDFT) calculations. The mRSIC method is an improvement of the RSIC method (Tsuneda et al. J Comput Chem 2003, 24, 1592). It takes into account the energy contributions from 2s and higher atomic orbitals that the RSIC method neglects. Furthermore, mRSIC improves the poor description for the nuclear-electron cusp of Gaussian basis functions. The mRSIC method was combined with a long-range correction (LC) scheme, which has been proved to give accurate valence-, Rydberg-, and charge transfer (CT)-excitation energies. In so doing, it dramatically improved the accuracy of the calculated core-excitation energies and did not affect the already accurate values of valence-, Rydberg-, and CT-excitation energies produced by the LC functionals. These results mean that the combined scheme is accurate for all excitation energy forms.
More Related Videos
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 03:51
Molecular Orbital Theory II
 02:35
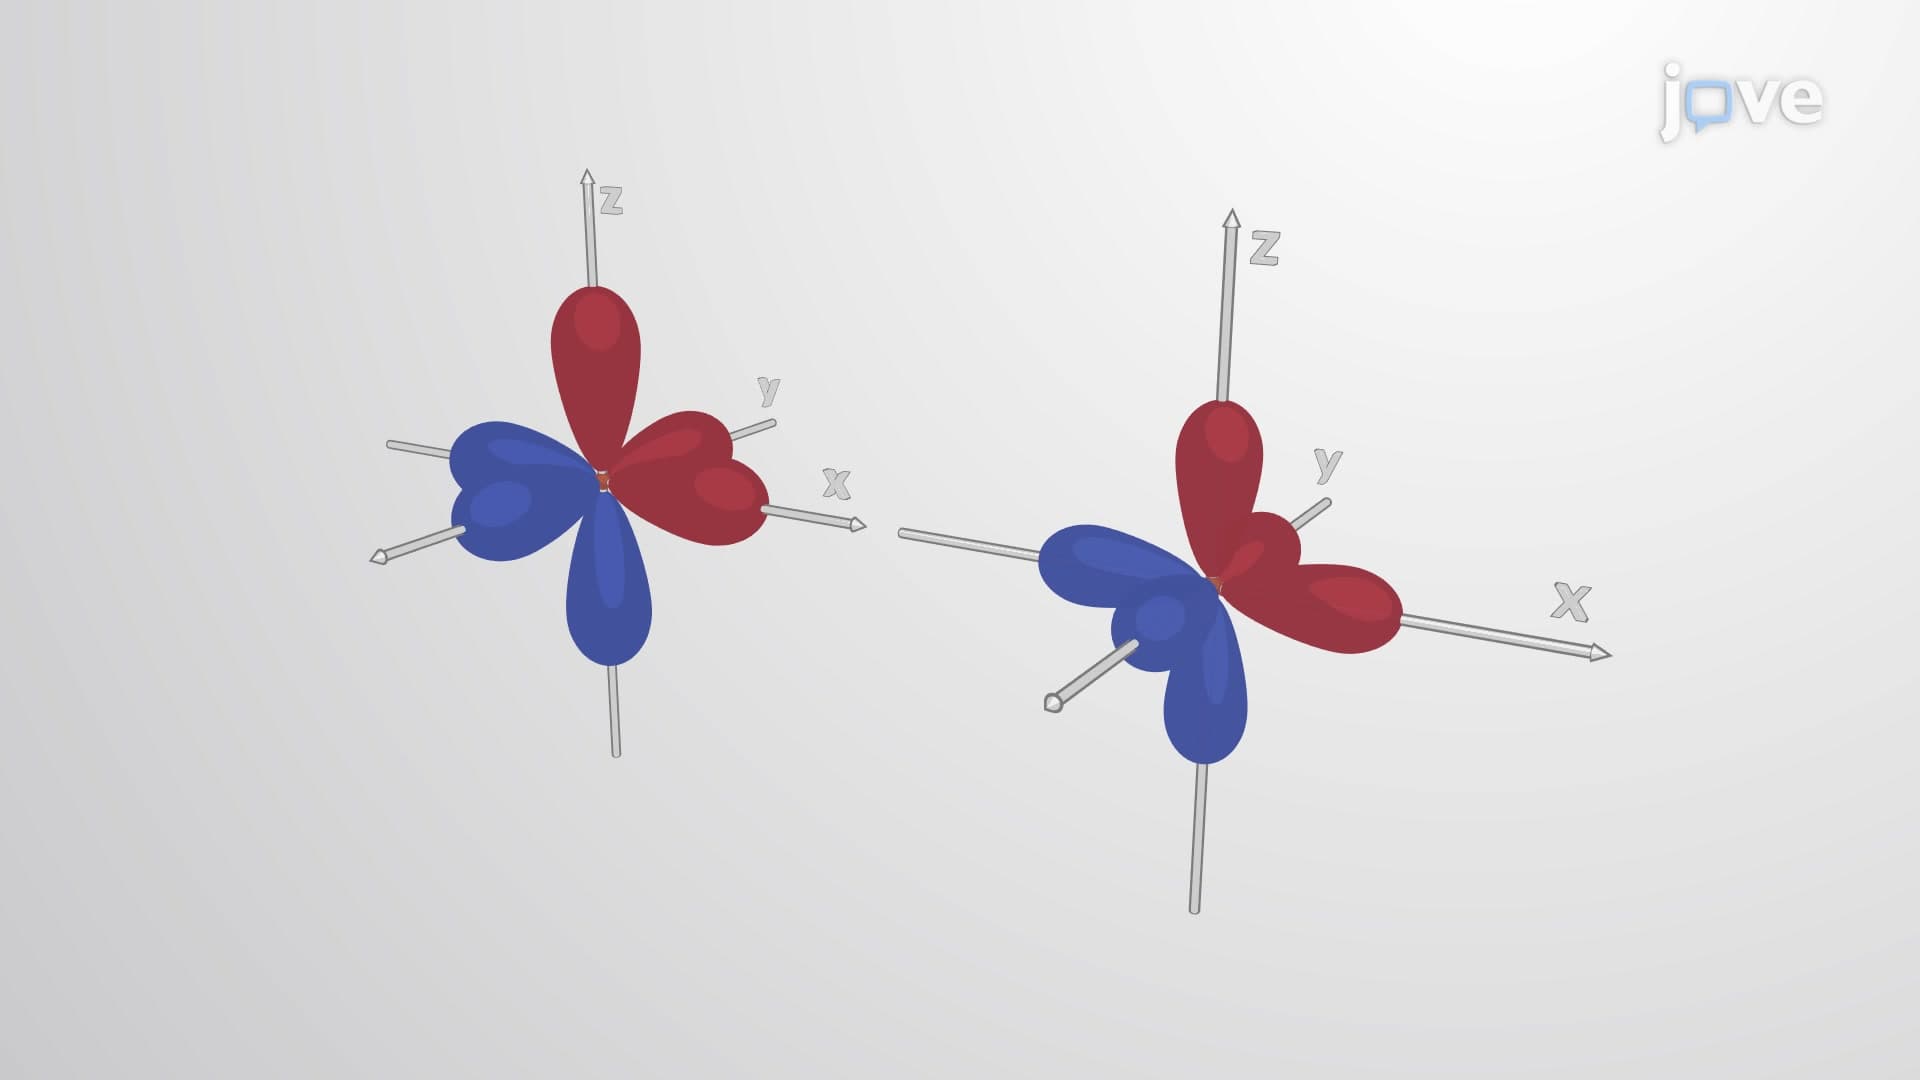
Molecular Orbital Theory I
 02:38
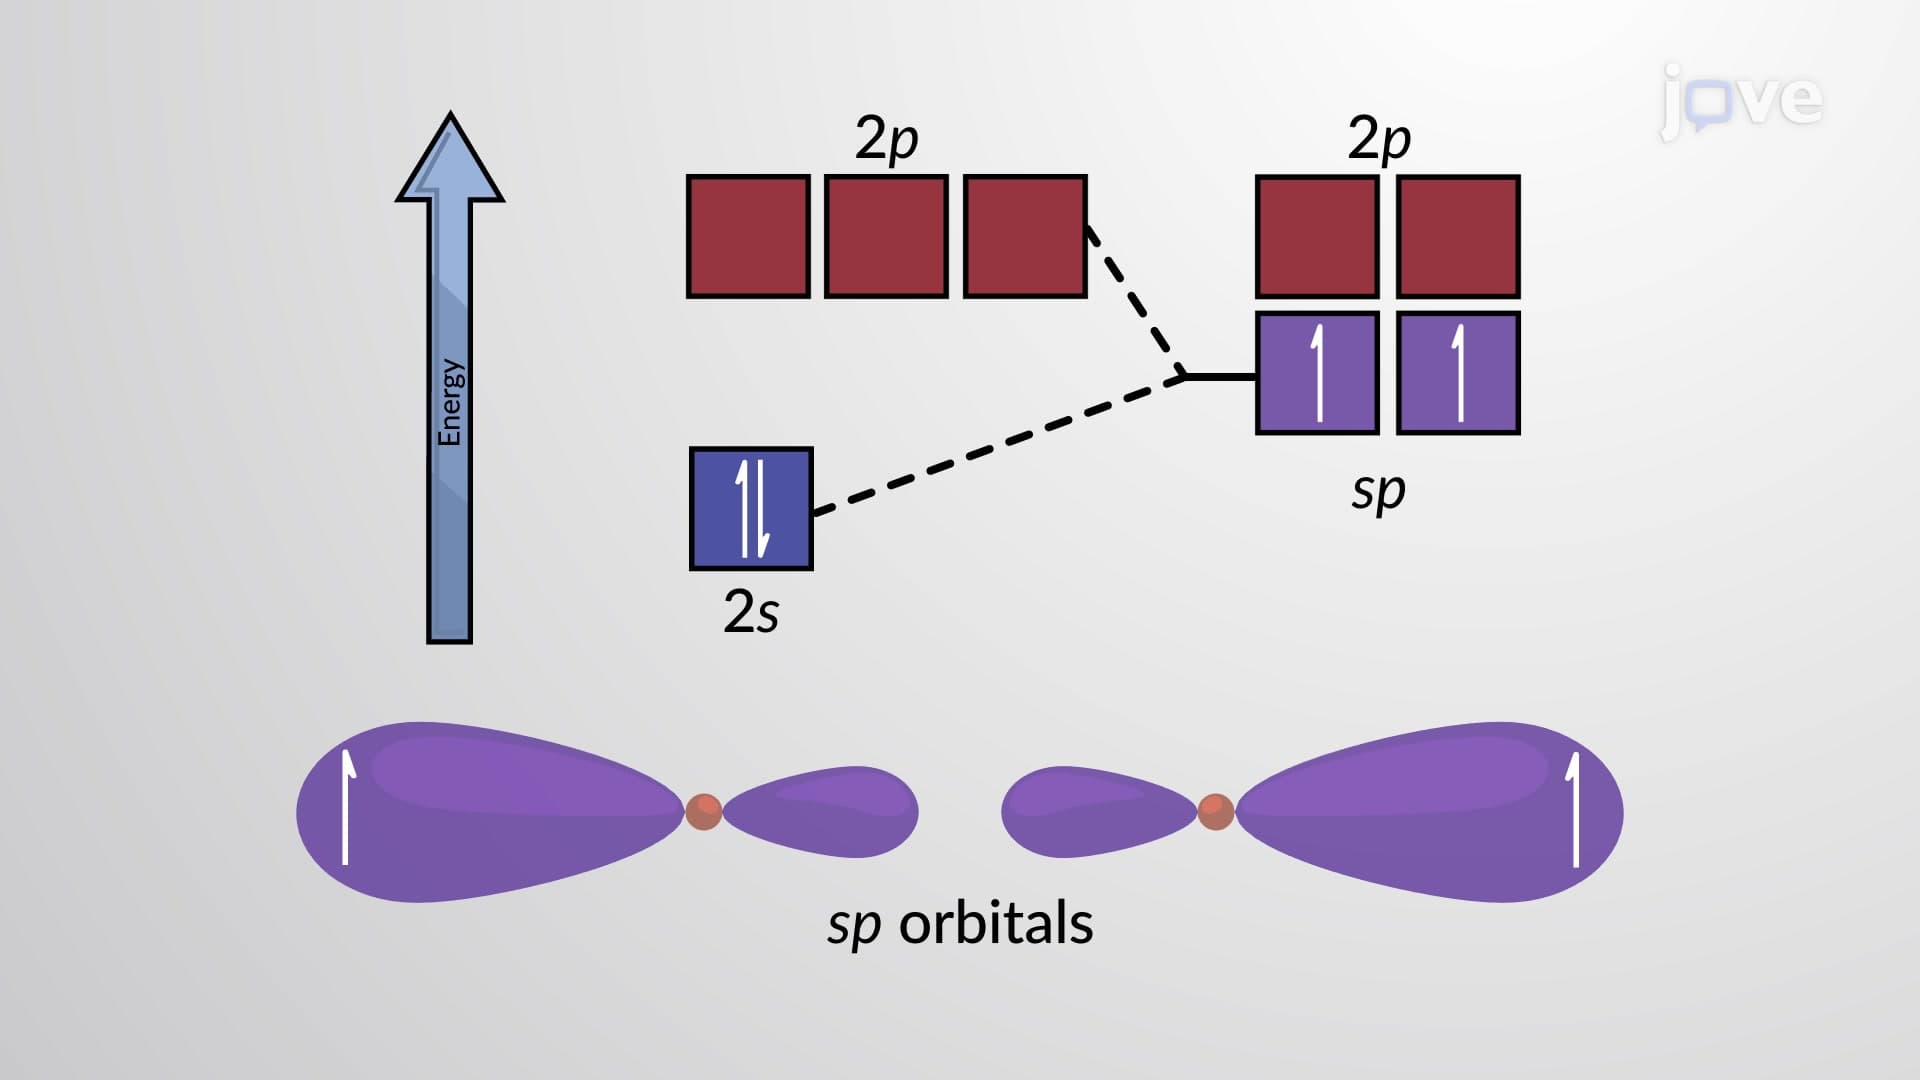
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 02:28
Electronic Structure of Atoms
An atom comprises protons and neutrons, which are contained inside the dense, central core called the nucleus, with electrons present around the nucleus. Taking into account the wave–particle duality of electrons and the uncertainty in position around the nucleus, quantum mechanics provides a more accurate model for the atomic structure. It describes atomic orbitals as the regions around the nucleus where electrons of discrete energy exist, characterized by four quantum numbers: n, l, ml, and...
 01:27
π Electron Effects on Chemical Shift: Overview
 01:12
Fermi Level Dynamics
Electron affinity in semiconductors refers to the energy gap between the minimum of its conduction band and the vacuum level and it is a critical parameter in determining how easily a semiconductor can accept additional electrons.
The work...