Related Experiment Video
Updated: Jun 23, 2026

Determining the Serum Stability of Human Adenosine Deaminase 1 Enzyme
Published on: September 27, 2024
Structure-activity relationships of truncated adenosine derivatives as highly potent and selective human A3 adenosine
Shantanu Pal1, Won Jun Choi, Seung Ah Choe
1Department of Bioinspired Science and Division of Life and Pharmaceutical Sciences, Laboratory of Medicinal Chemistry, College of Pharmacy, Ewha Womans University, Seoul 120-750, Republic of Korea.

Abstract:
On the basis of potent and selective binding affinity of truncated 4'-thioadenosine derivatives at the human A(3) adenosine receptor (AR), their bioisosteric 4'-oxo derivatives were designed and synthesized from commercially available 2,3-O-isopropylidene-D-erythrono lactone. The derivatives tested in AR binding assays were substituted at the C2 and N(6) positions. All synthesized nucleosides exhibited potent and selective binding affinity at the human A(3) AR. They were less potent than the corresponding 4'-thio analogues, but showed still selective to other subtypes. The 2-Cl series generally were better than the 2-H series in view of binding affinity and selectivity. Among compounds tested, compound 5d (X=Cl, R=3-bromobenzyl) showed the highest binding affinity (K(i)=13.0+/-6.9 nM) at the hA(3) AR with high selectivity (at least 88-fold) in comparison to other AR subtypes. Like the corresponding truncated 4'-thio series, compound 5d antagonized the action of an agonist to inhibit forskolin-stimulated adenylate cyclase in hA(3) AR-expressing CHO cells. Although the 4'-oxo series were less potent than the 4'-thio series, this class of human A(3) AR antagonists is also regarded as another good template for the design of A(3) AR antagonists and for further drug development.
More Related Videos
 10:29
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
 07:16
Methods for the Discovery of Novel Compounds Modulating a Gamma-Aminobutyric Acid Receptor Type A Neurotransmission
Published on: August 16, 2018
Related Concept Videos
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of the aromatic...
 01:17
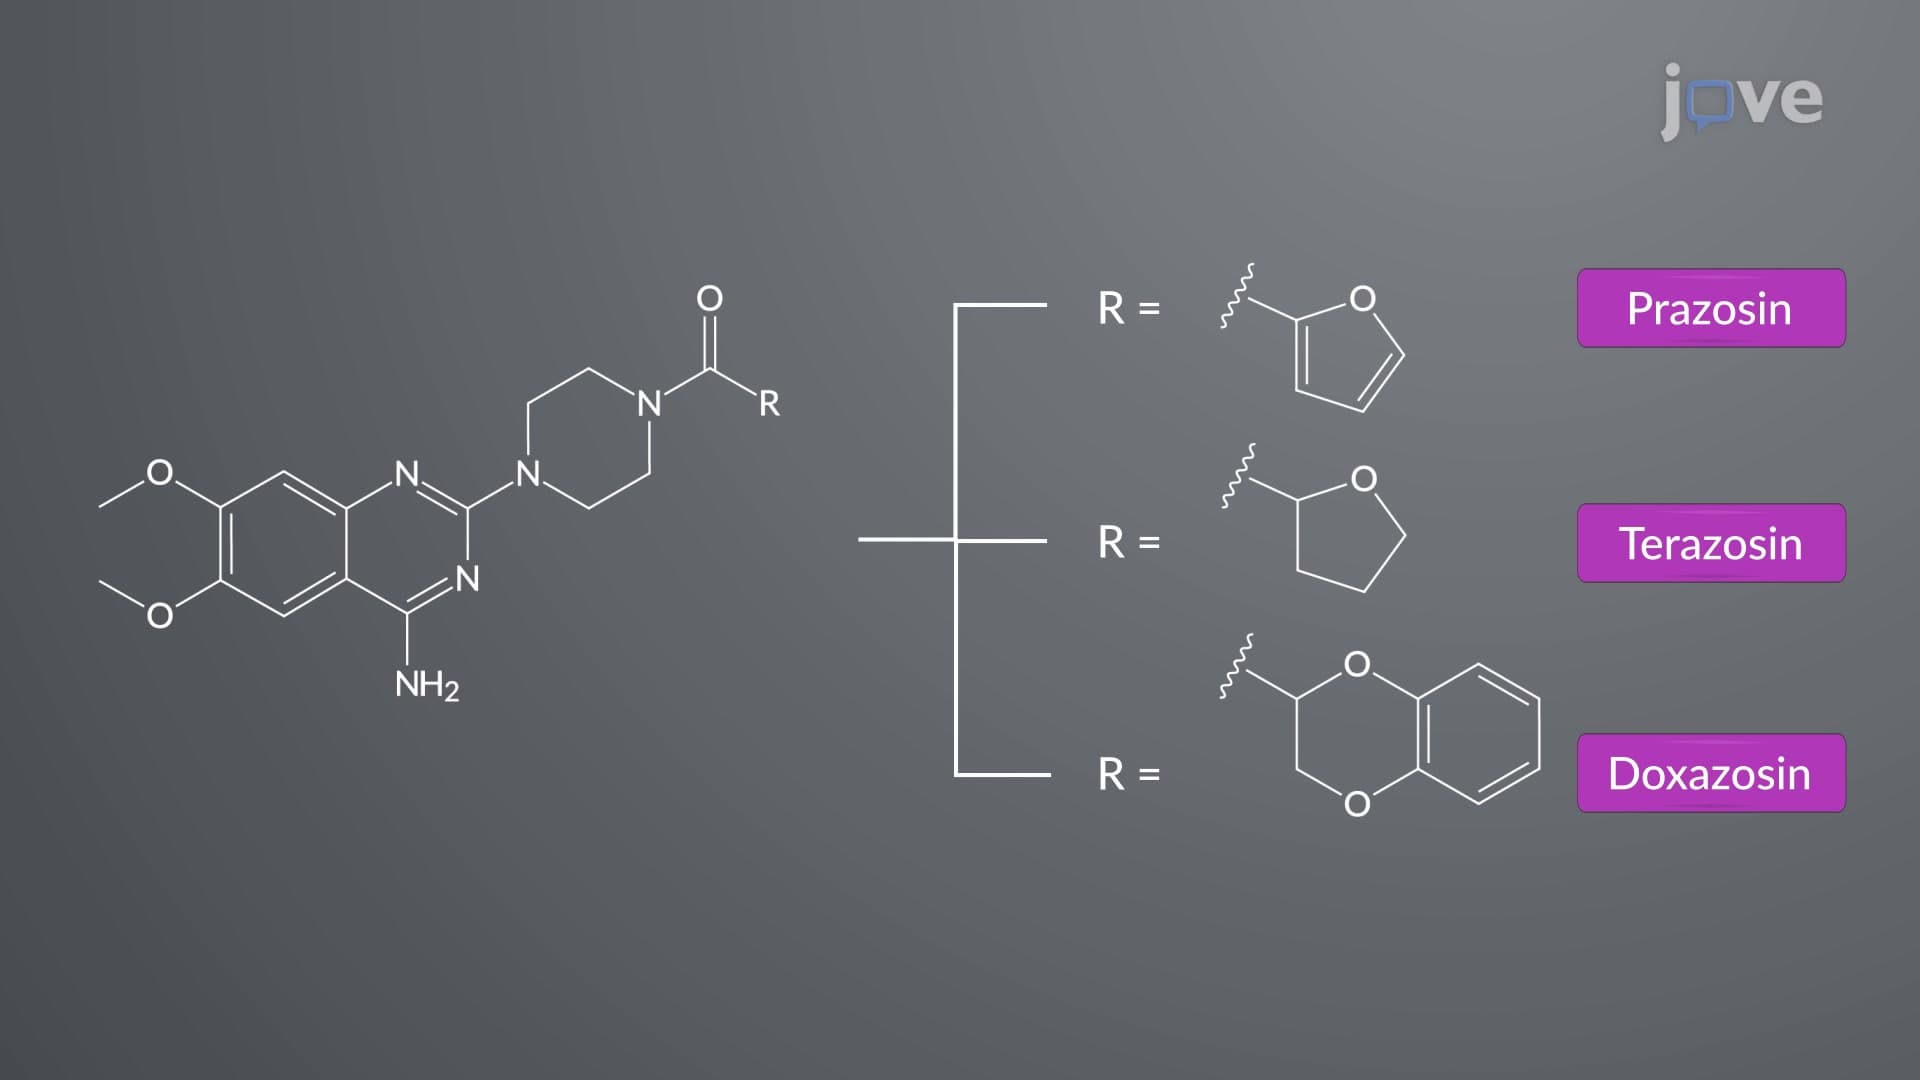
Adrenergic Antagonists: Chemistry and Classification of ɑ-Receptor Blockers
Nonselective α-blockers: Nonselective α-blockers contain haloalkylamine or imidazoline moieties. Phenoxybenzamine, with a haloalkylamine...
 01:22
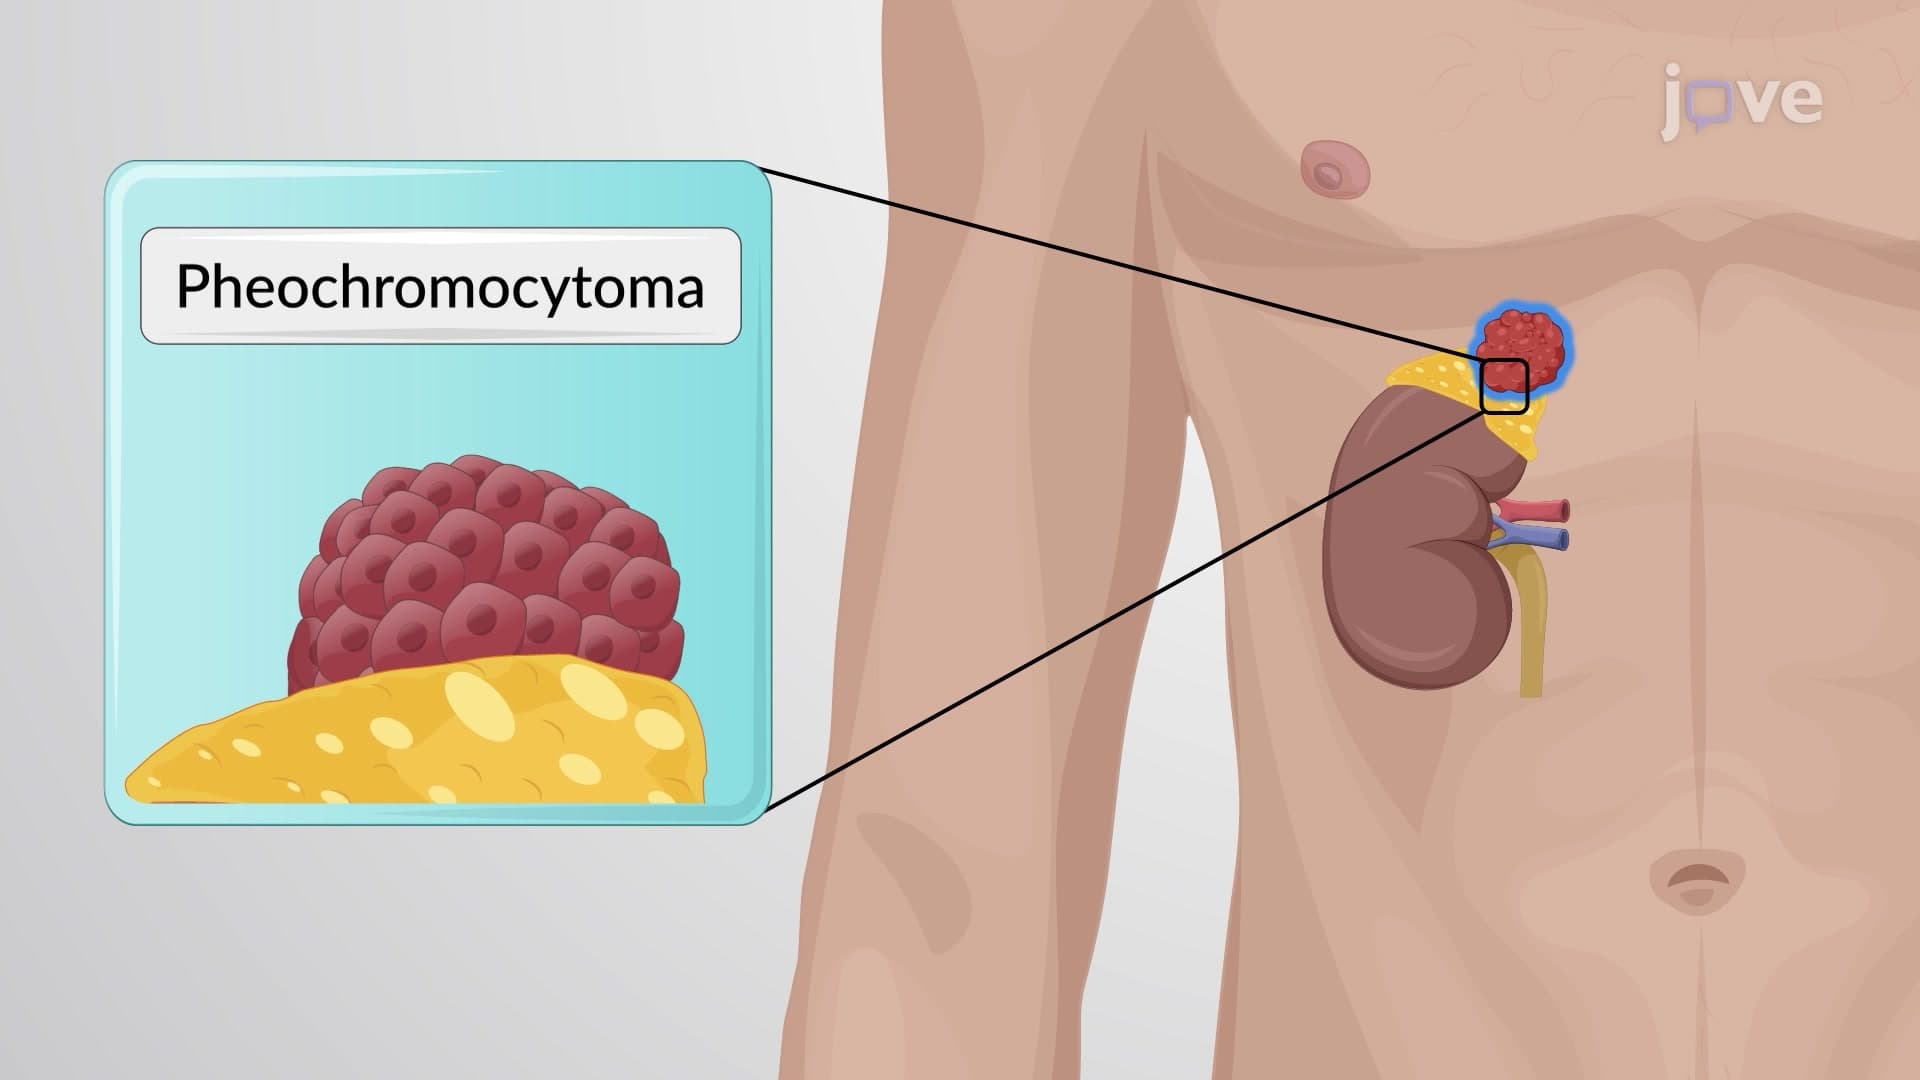
Adrenergic Antagonists: Pharmacological Actions of ɑ-Receptor Blockers
α1-blockers: These drugs inhibit α1-adrenoceptors on smooth muscle cells, resulting in vasodilation. This vasodilation lowers blood pressure, making α1-blockers valuable in treating hypertension. Additionally, α1-blockers effectively address urinary obstruction...
 01:31
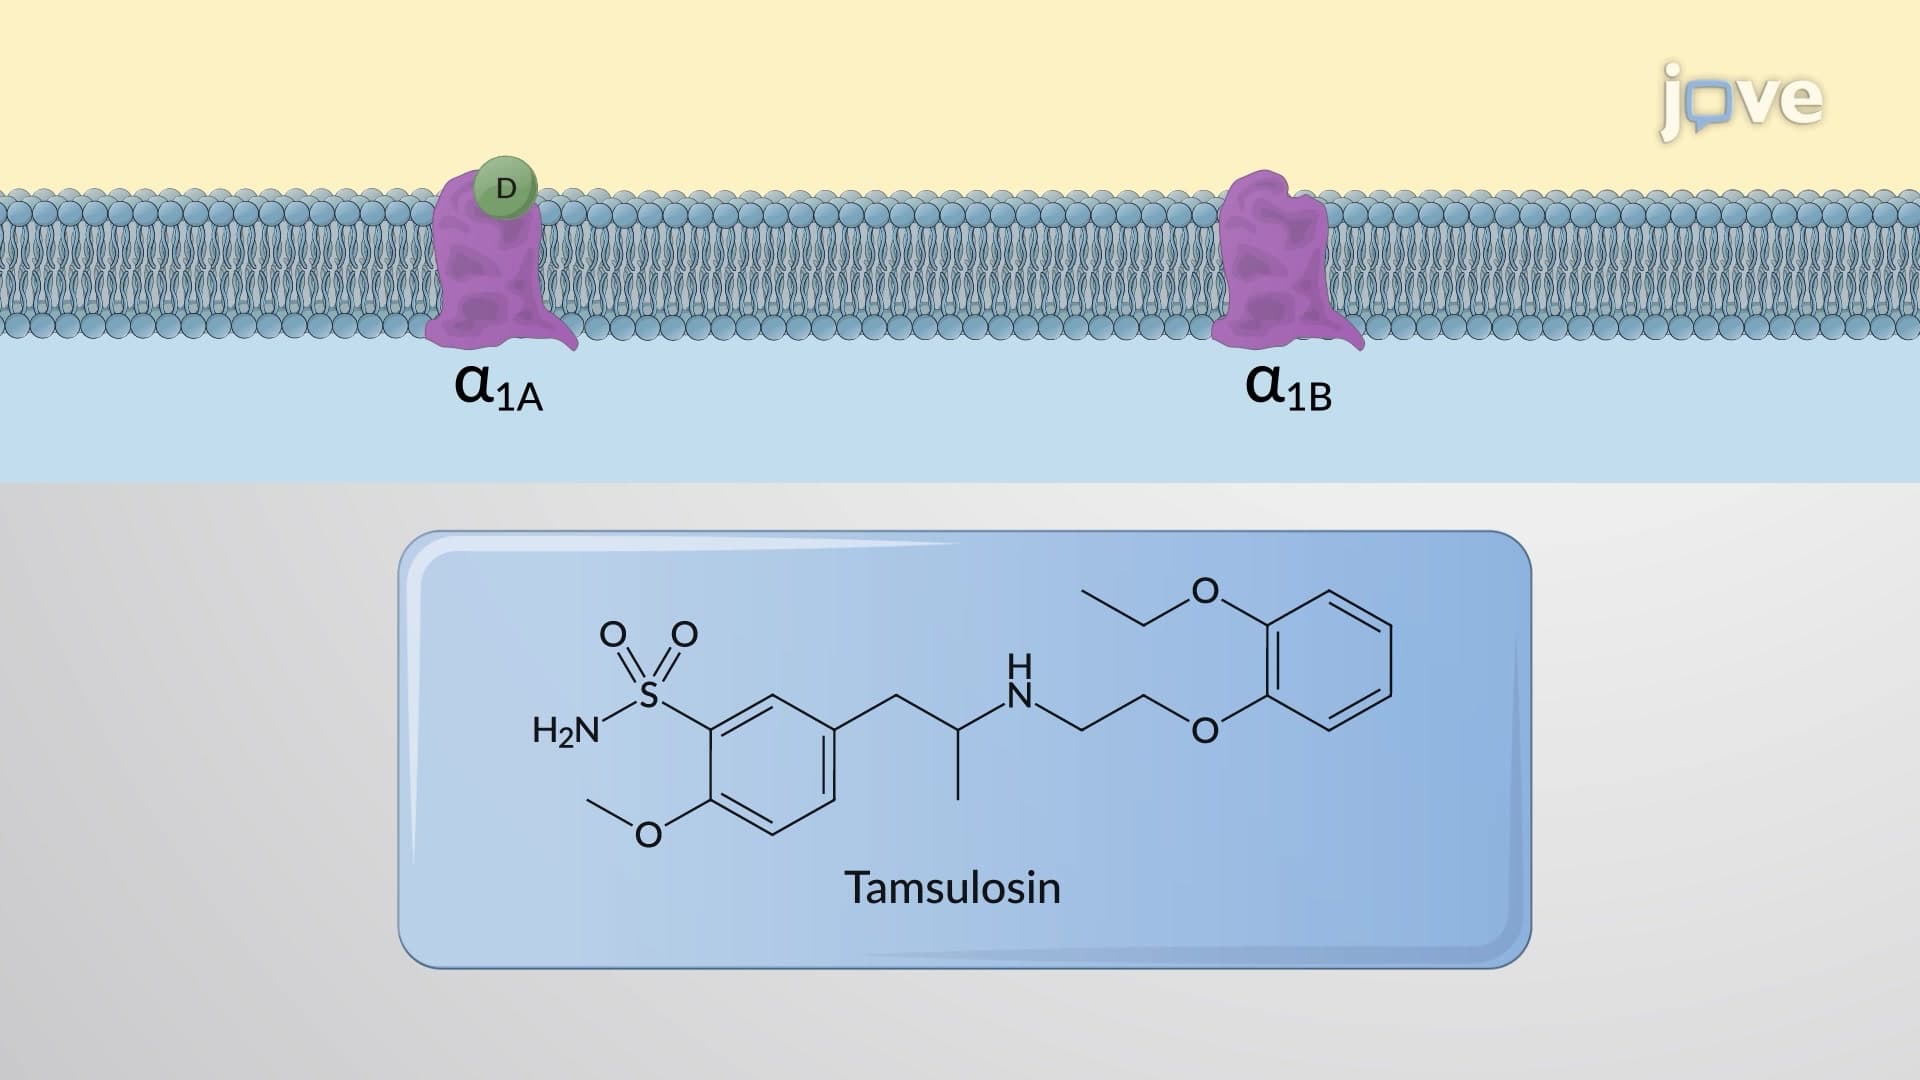
Adrenergic Receptors: ɑ Subtype
Adrenaline ≥ Noradrenaline >> Isoprenaline
α-adrenoceptors are further divided into α1 and α2-adrenoceptors.
α1-Adrenoceptors: These receptors are located postsynaptically on the effector organs and cause constriction of smooth muscle mediated by activation of phospholipase C—inositol-1,4,5-trisphosphate...
 01:29
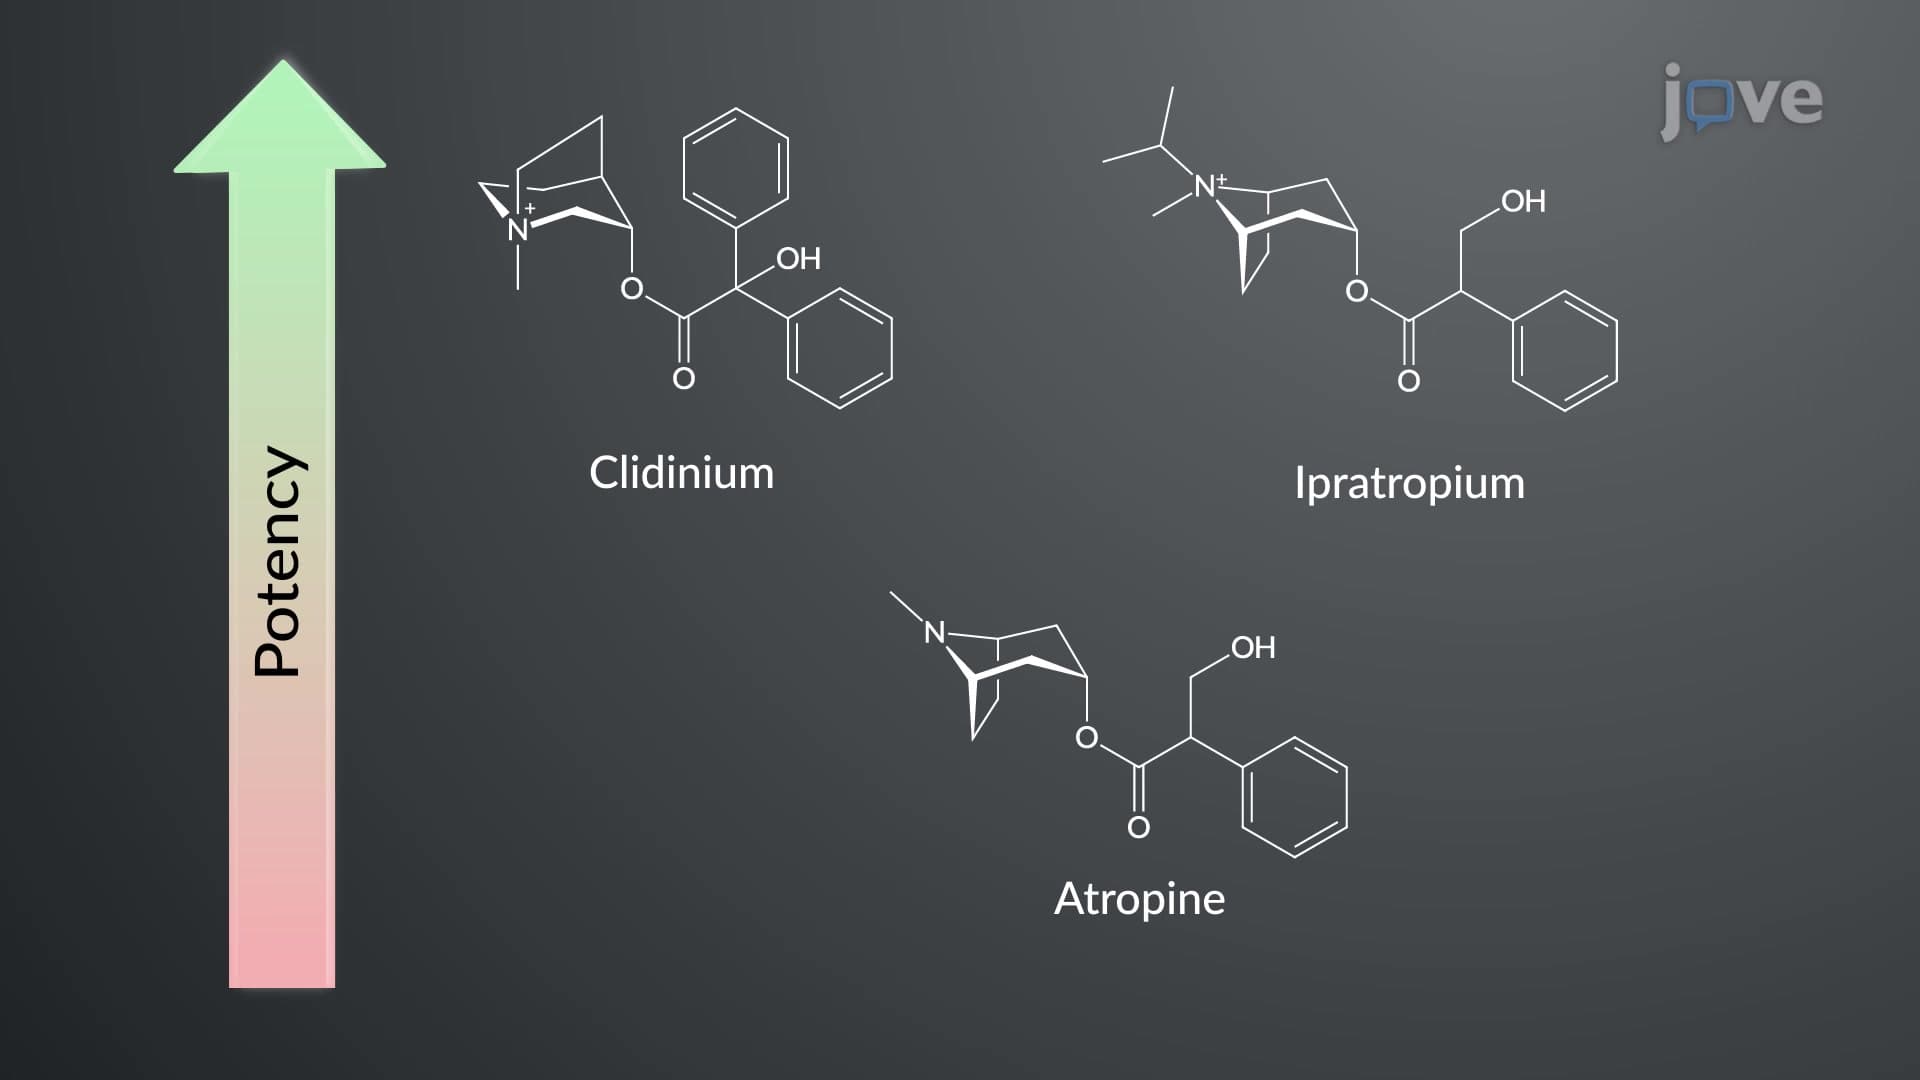
Cholinergic Antagonists: Chemistry and Structure-Activity Relationship
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence its...