Related Experiment Video
Updated: Jun 23, 2026


Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Deleterious effects of long-range self-repulsion on the density functional description of O2 sticking on aluminum
Ester Livshits1, Roi Baer, Ronnie Kosloff
1Institute of Chemistry and the Fritz Haber Center for Molecular Dynamics, the Hebrew University of Jerusalem, Jerusalem 91904 Israel.

Abstract:
Density functional theory (DFT) with semilocal functionals such as the local-density and generalized gradients approximations predicts that the dissociative adsorption of oxygen on Al (111) goes through without a barrier in stark contradiction to experimental findings. This problem motivated our study of the reaction of oxygen colliding with a small aluminum cluster Al(5). We found semilocal functionals predict a minute barrier to sticking, associated with smeared long-range charge transfer from the metal to the oxygen. Hybrid B3LYP predicts a larger barrier while the range-separated the Baer-Neuhauser-Livshits (BNL, Phys. Chem. Chem. Phys. 2007, 9, 2932.) functional finds a more prominent barrier. BNL predicts short-ranged and more abrupt charge transfer from the surface to the oxygen. We conclude that spurious self-repulsion inherent in semilocal functionals causes early electron-transfer, long-range attraction toward the surface and low reaction barriers for these systems. The results indicate that the missing DFT barrier for O(2) sticking on Al (111) may be due to spurious self-repulsion.
Related Concept Videos
 01:27
π Electron Effects on Chemical Shift: Overview
 03:51
Molecular Orbital Theory II
 02:40
MO Theory and Covalent Bonding
 01:27
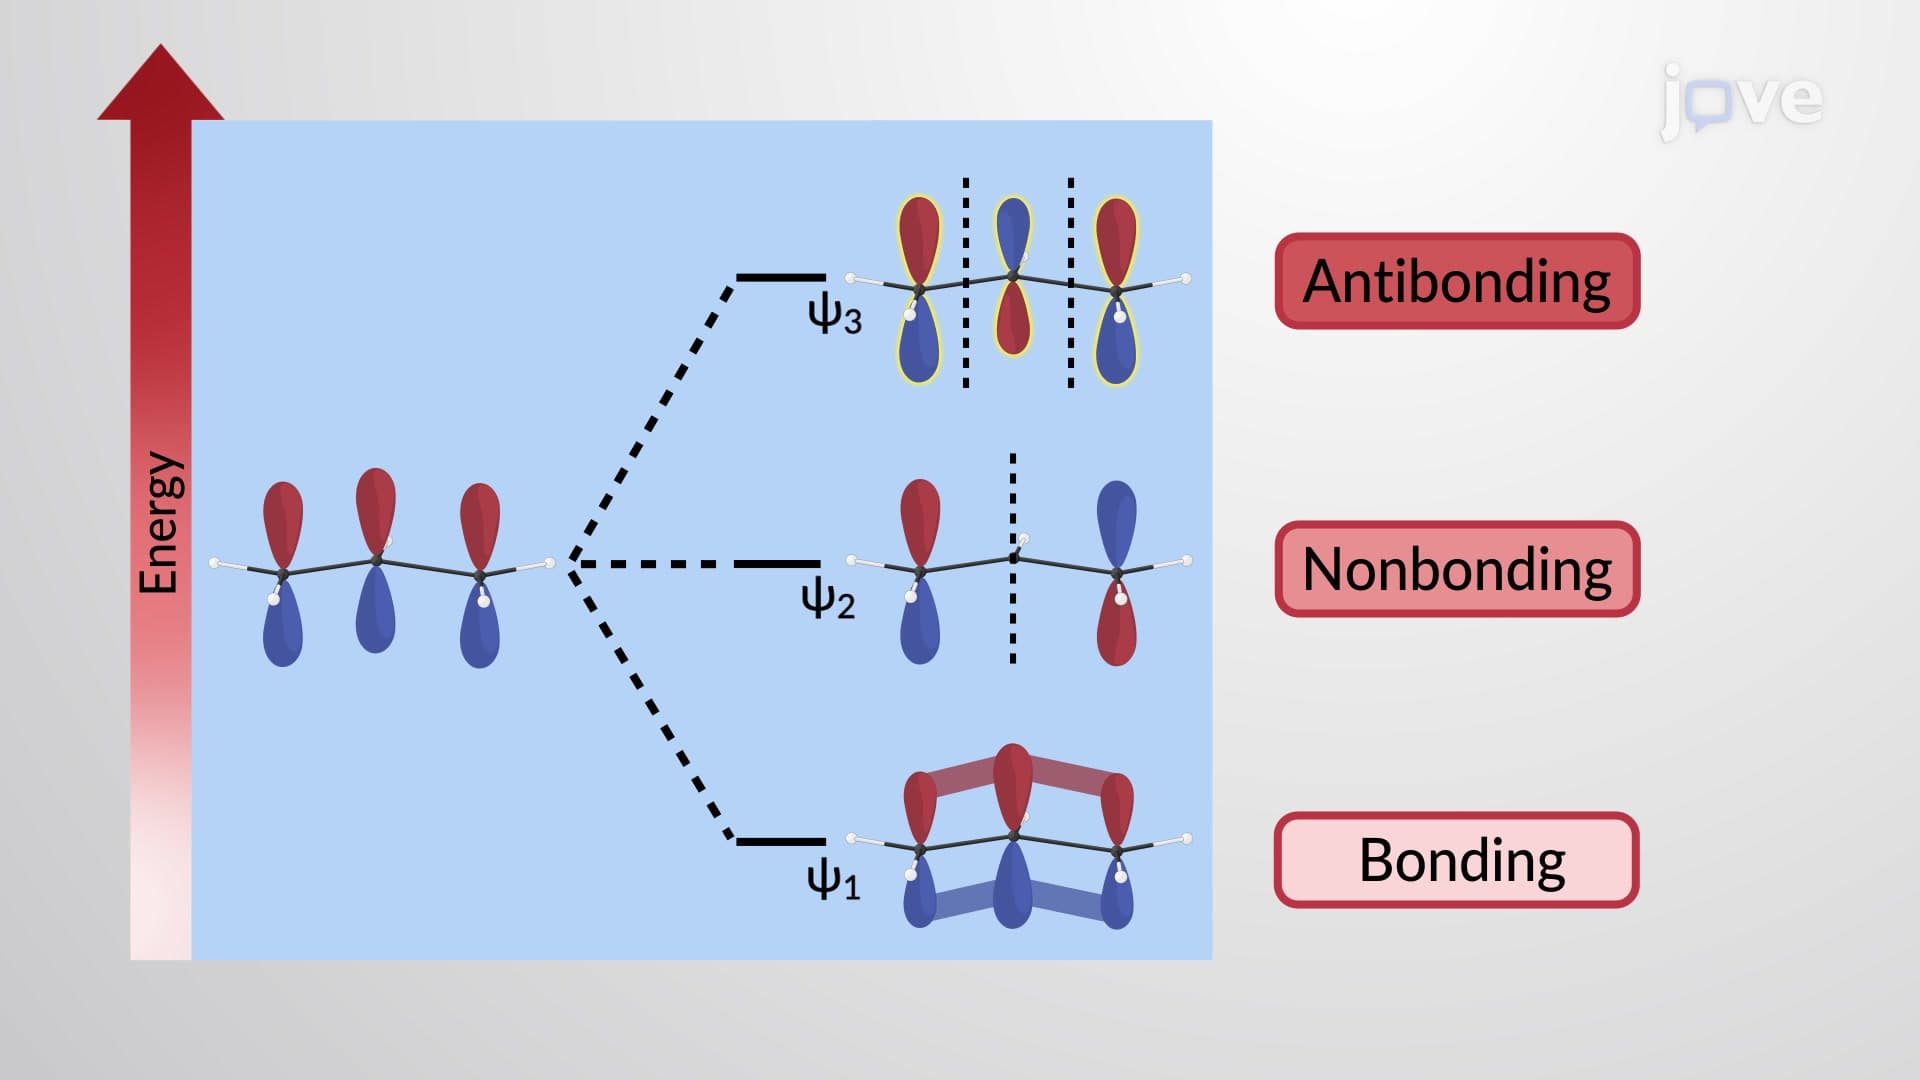
π Molecular Orbitals of the Allyl Radical
The allyl systems have identical molecular orbitals but differ in the number of π electrons.
 01:24
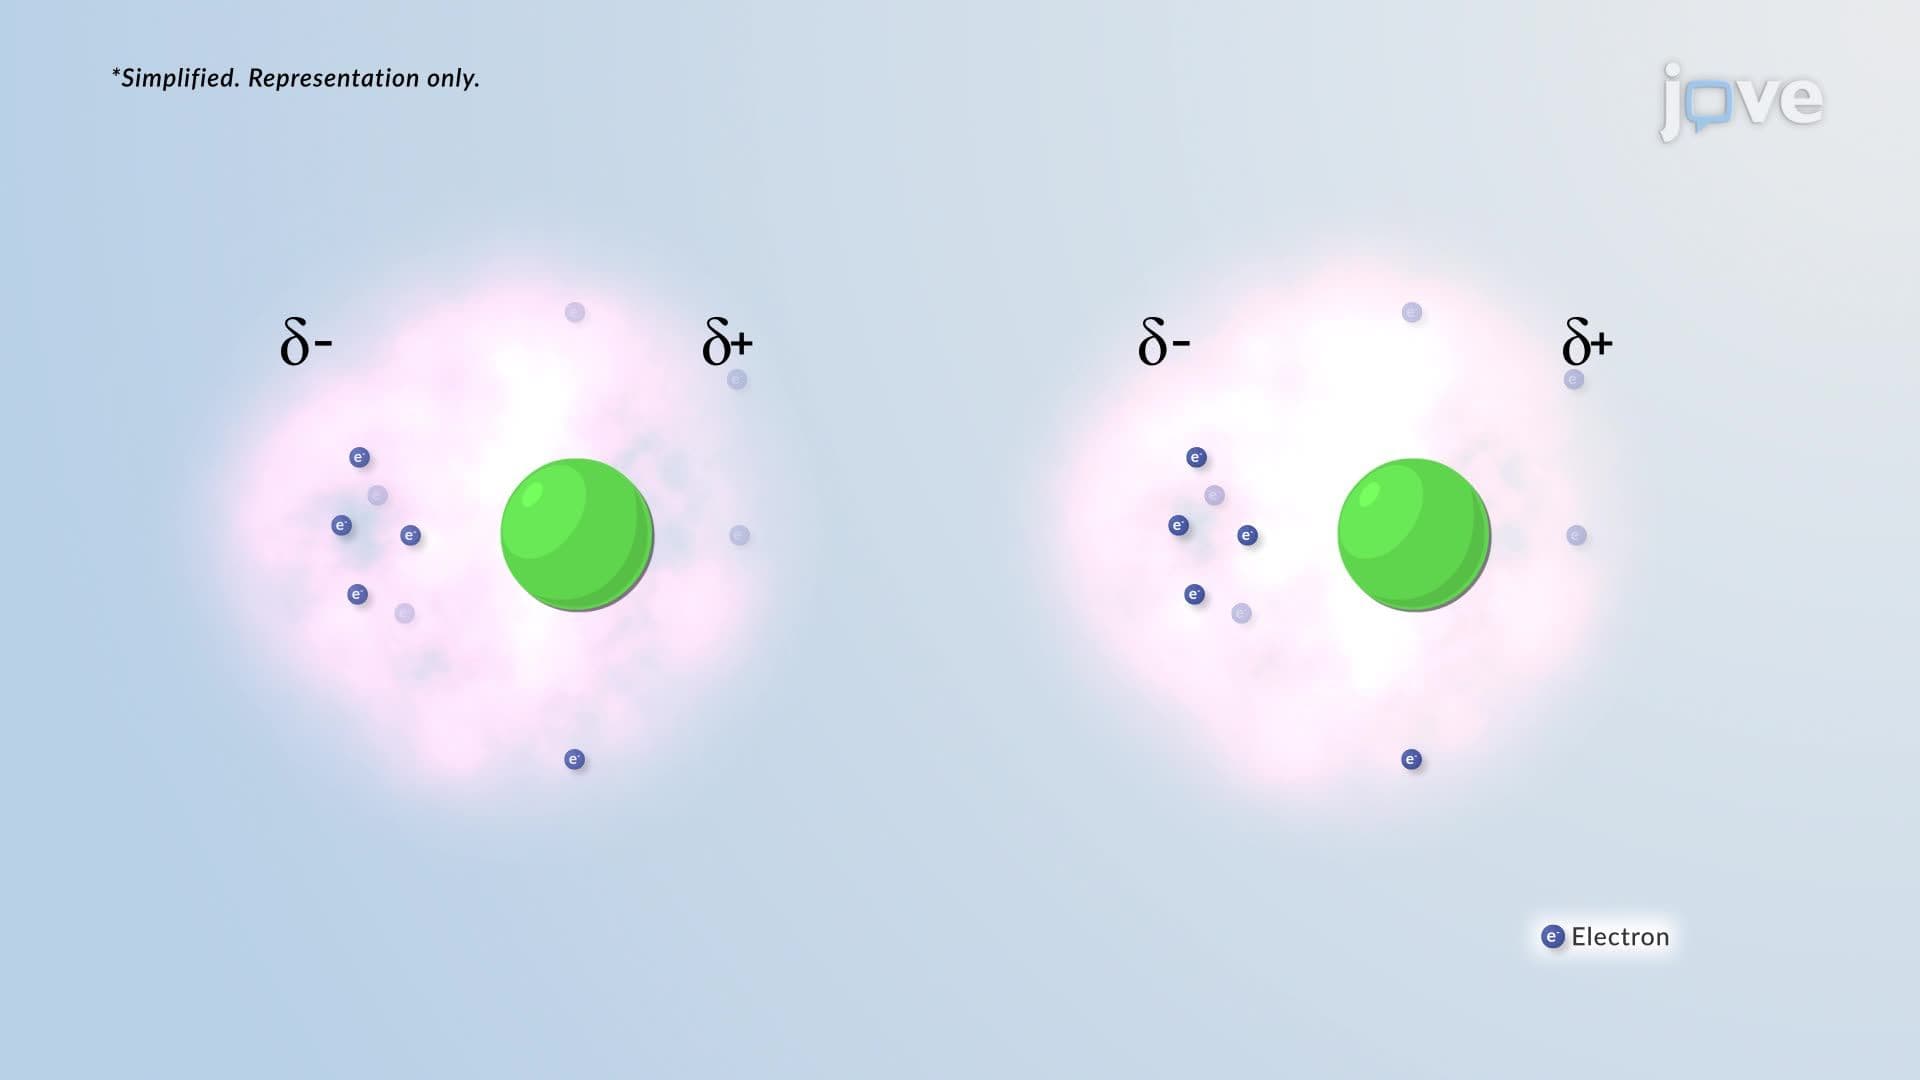
Van der Waals Interactions
 01:18
π Molecular Orbitals of the Allyl Cation and Anion