Related Experiment Video
Updated: Jun 23, 2026

An All-Human Hepatic Culture System for Drug Development Applications
Published on: October 20, 2023
Structure-based prediction of the nonspecific binding of drugs to hepatic microsomes
Haiyan Li1, Jin Sun, Xiaofan Sui
1Department of Biopharmaceutics, School of Pharmacy, Shenyang Pharmaceutical University, No. 103 of Wenhua Road, P.O. Box. 59, Shenyang, 110016, China.

Abstract:
For the accurate prediction of in vivo hepatic clearance or drug-drug interaction potential through in vitro microsomal metabolic data, it is essential to evaluate the fraction unbound in hepatic microsomal incubation media. Here, a structure-based in silico predictive model of the nonspecific binding (fu(mic), fraction unbound in hepatic microsomes) for 86 drugs was successfully developed based on seven selected molecular descriptors. The R(2) of the predicted and observed log((1 - fu(mic))/fu(mic)) for the training set (n = 64) and test set (n = 22) were 0.82 and 0.85, respectively. The average fold error (AFE, calculated by fu(mic) rather than log((1 - fu(mic))/fu(mic))) of the in silico model was 1.33 (n = 86). The predictive capability of fu(mic) for neutral drugs compared well to that for basic compounds (R(2) = 0.82, AFE = 1.18 and fold error values were all below 2, except for felodipine and progesterone) in our model. This model appears to perform better for neutral compounds when compared to models previously published in the literature. Therefore, this in silico model may be used as an additional tool to estimate fu(mic) and for predicting in vivo hepatic clearance and inhibition potential from in vitro hepatic microsomal studies.

Related Concept Videos
 01:07
Physiological Pharmacokinetic Models: Incorporating Hepatic Transporter-Mediated Clearance
A recent model describes pravastatin's hepatobiliary excretion, mediated...
 01:29
Drug Distribution: Plasma Protein Binding
 01:09
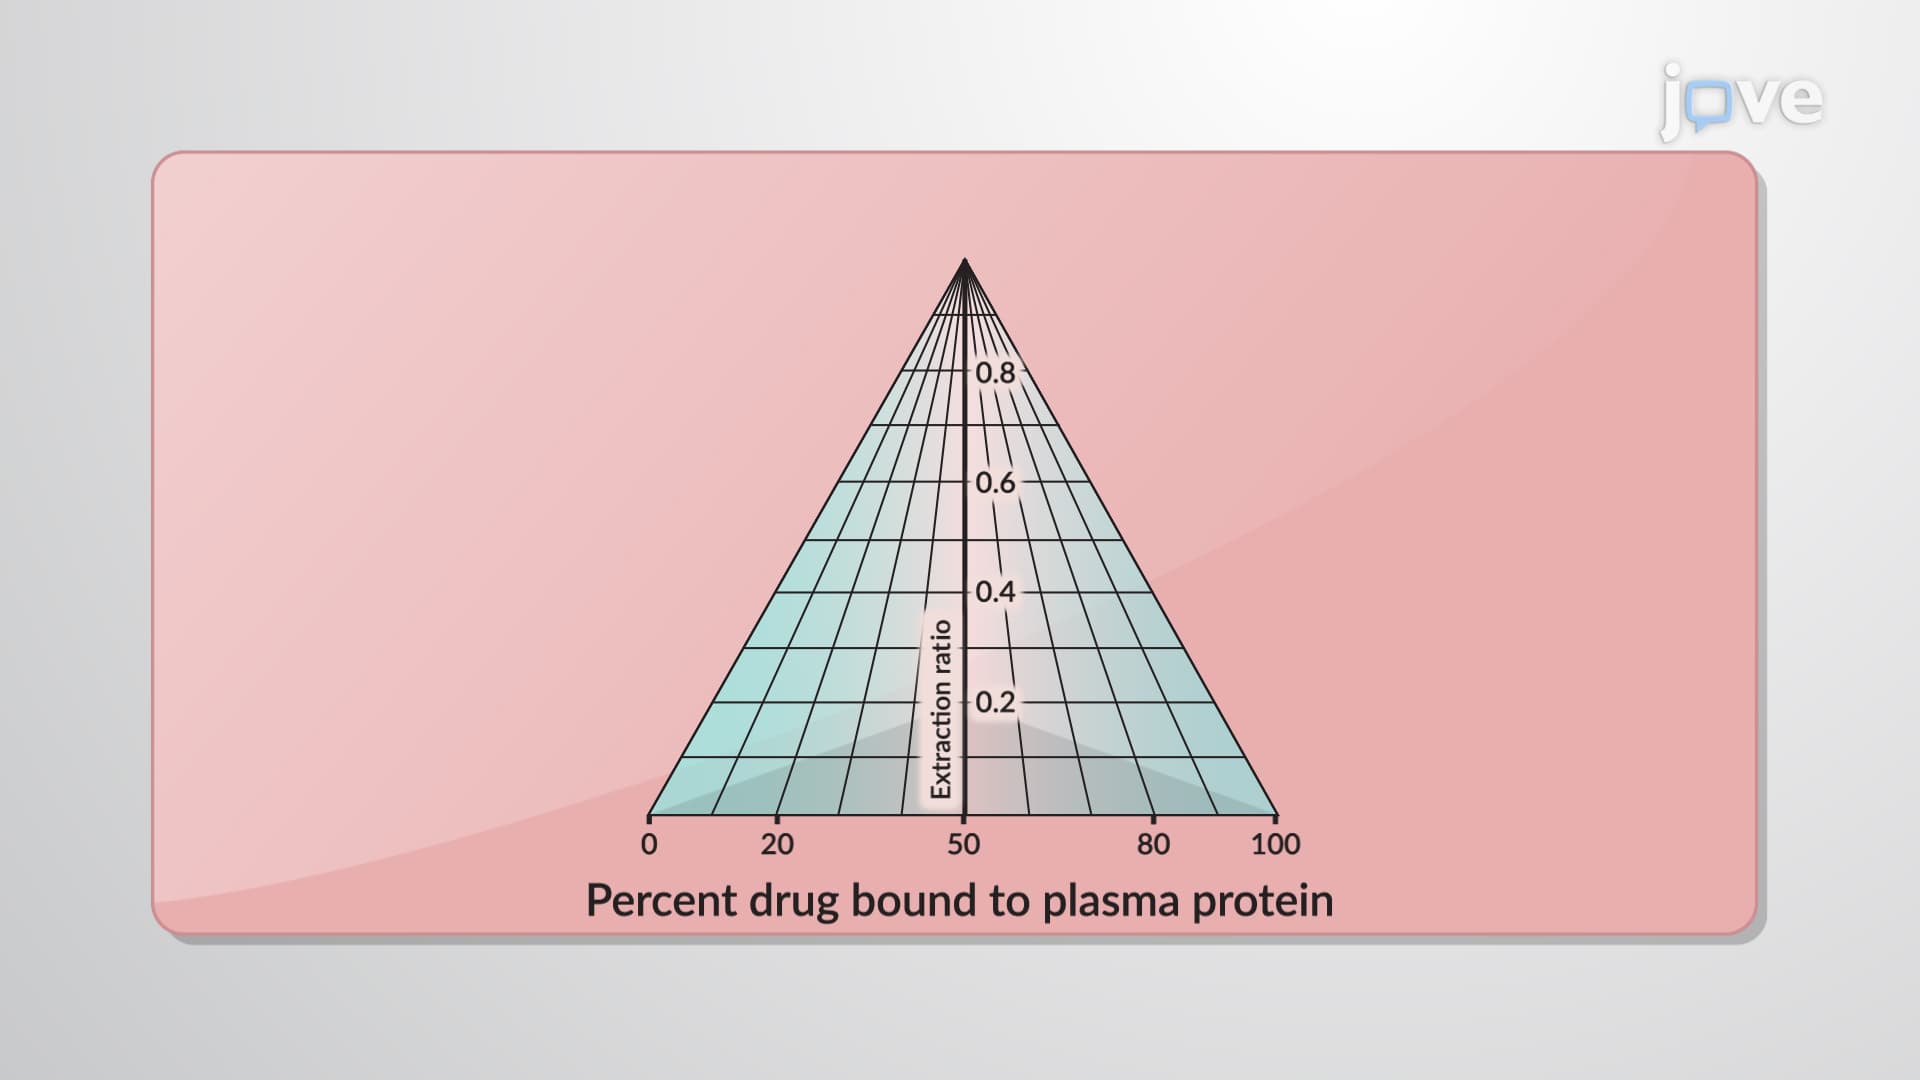
Hepatic Drug Clearance: Effect of Protein Binding
For low-extraction-ratio drugs that are less than 80% protein-bound, minor changes in protein binding...
 01:30
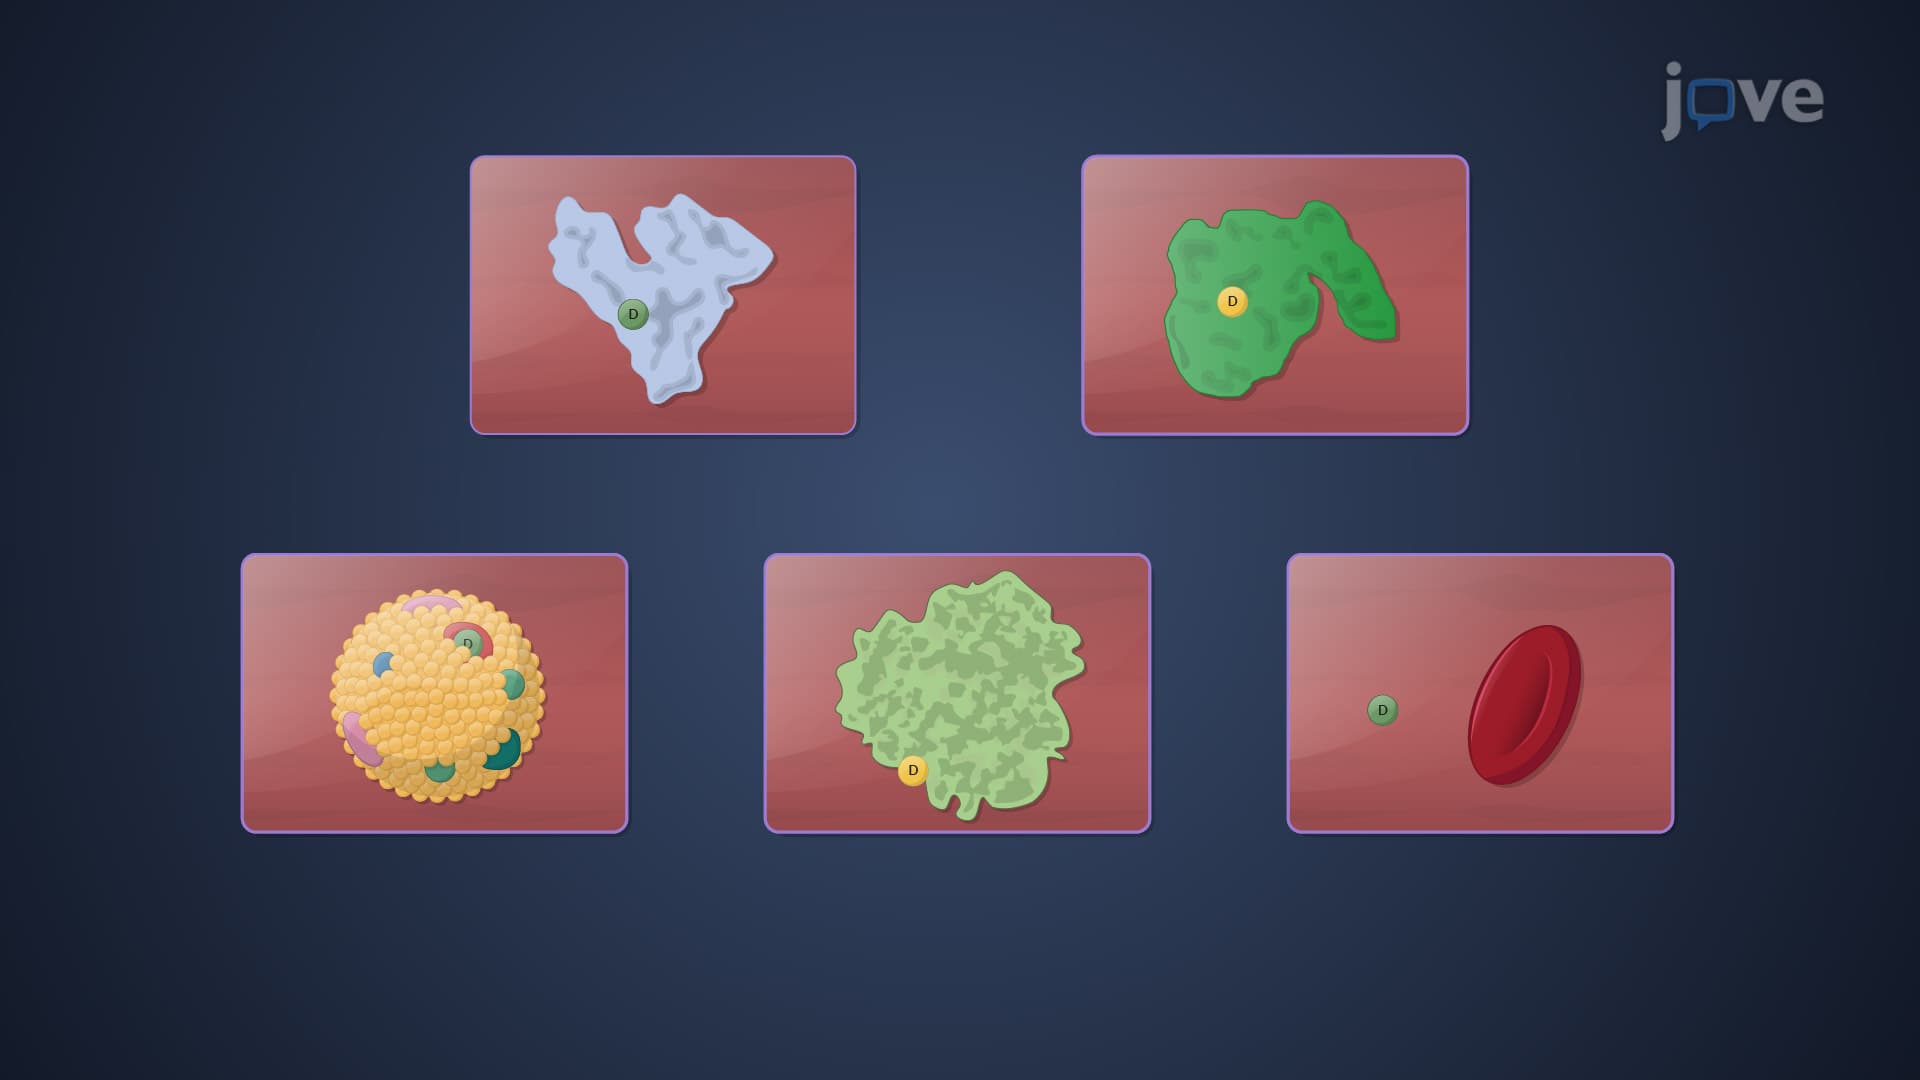
Drug Binding to Blood Components
HSA is the most abundant plasma protein and is vital in drug binding. It contains distinct drug-binding sites, with different drugs exhibiting affinity for specific sites. There are three main drug-binding domains for HSA: sites I, II, and III. These domains are further...
 01:14
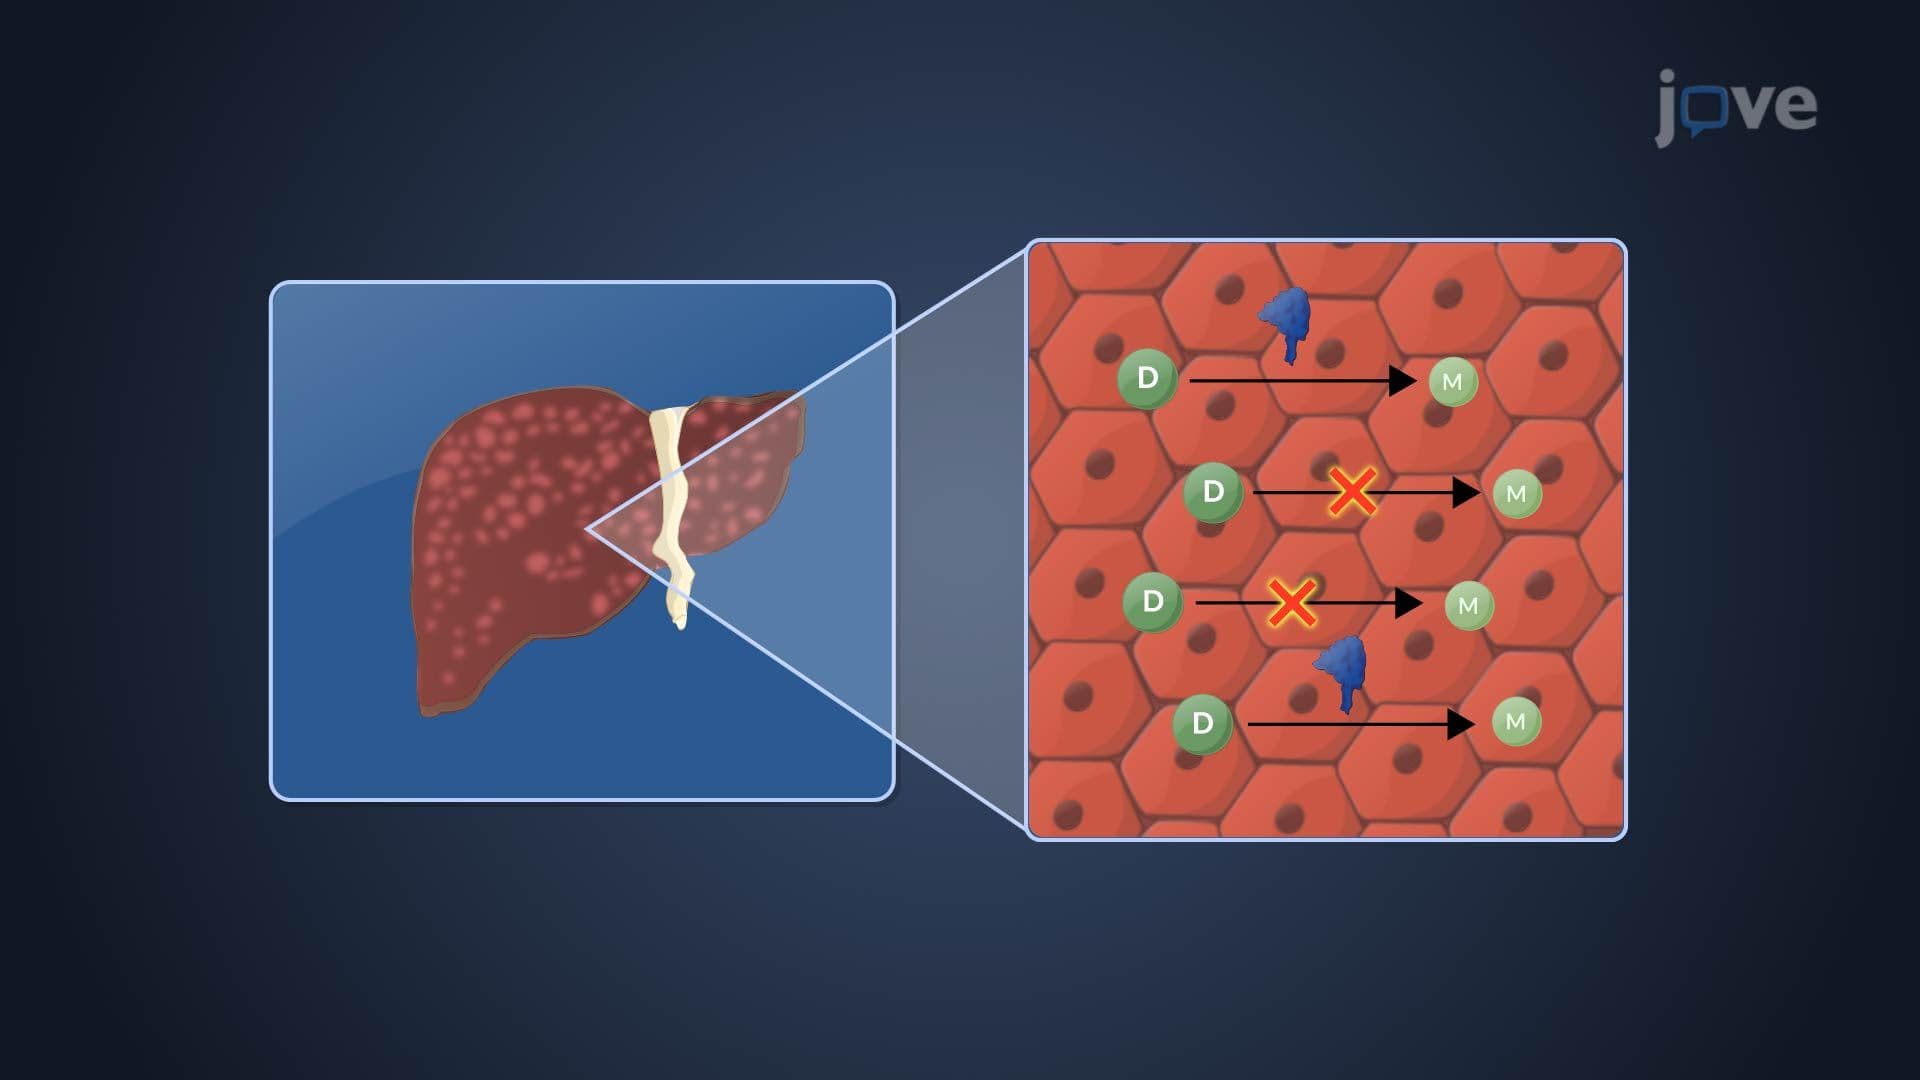
Effect of Hepatic Disease on Pharmacokinetics: Active Drug, Metabolite and Fraction of Metabolized Drug
 01:13
Physiological Pharmacokinetic Models: Assumption with Protein Binding