Related Experiment Video
Updated: Jun 23, 2026
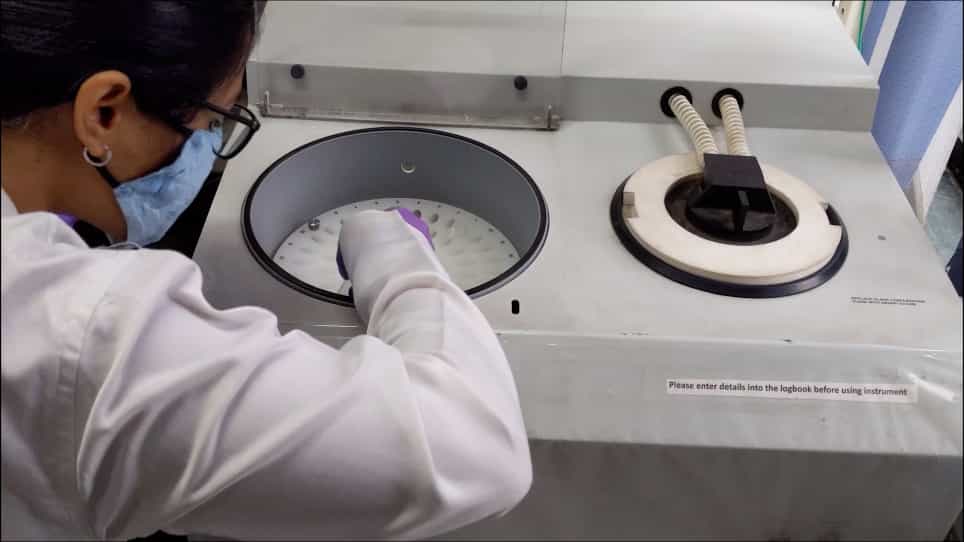

Comprehensive Workflow of Mass Spectrometry-based Shotgun Proteomics of Tissue Samples
Published on: November 13, 2021
A HUPO test sample study reveals common problems in mass spectrometry-based proteomics
Alexander W Bell1, Eric W Deutsch, Catherine E Au
1Department of Anatomy and Cell Biology, McGill University, Montreal, Canada.

We performed a test sample study to try to identify errors leading to irreproducibility, including incompleteness of peptide sampling, in liquid chromatography-mass spectrometry-based proteomics. We distributed an equimolar test sample, comprising 20 highly purified recombinant human proteins, to 27 laboratories. Each protein contained one or more unique tryptic peptides of 1,250 Da to test for ion selection and sampling in the mass spectrometer. Of the 27 labs, members of only 7 labs initially reported all 20 proteins correctly, and members of only 1 lab reported all tryptic peptides of 1,250 Da. Centralized analysis of the raw data, however, revealed that all 20 proteins and most of the 1,250 Da peptides had been detected in all 27 labs. Our centralized analysis determined missed identifications (false negatives), environmental contamination, database matching and curation of protein identifications as sources of problems. Improved search engines and databases are needed for mass spectrometry-based proteomics.
We performed a test sample study to try to identify errors leading to irreproducibility, including incompleteness of peptide sampling, in liquid chromatography-mass spectrometry-based proteomics. We distributed an equimolar test sample, comprising 20 highly purified recombinant human proteins, to 27 laboratories. Each protein contained one or more unique tryptic peptides of 1,250 Da to test for ion selection and sampling in the mass spectrometer. Of the 27 labs, members of only 7 labs initially reported all 20 proteins correctly, and members of only 1 lab reported all tryptic peptides of 1,250 Da. Centralized analysis of the raw data, however, revealed that all 20 proteins and most of the 1,250 Da peptides had been detected in all 27 labs. Our centralized analysis determined missed identifications (false negatives), environmental contamination, database matching and curation of protein identifications as sources of problems. Improved search engines and databases are needed for mass spectrometry-based proteomics.
Related Concept Videos
 01:33
Peptide Identification Using Tandem Mass Spectrometry
This technique helps gather information regarding the protein from which the peptide was obtained and to study the peptides’ amino acid sequence. Identifying peptides from a complex mixture is an important component of the growing field of...
 01:21
Mass Spectrometry: Complex Analysis
GC–MS is a powerful hyphenated method commonly used in forensics and environmental...
 01:15
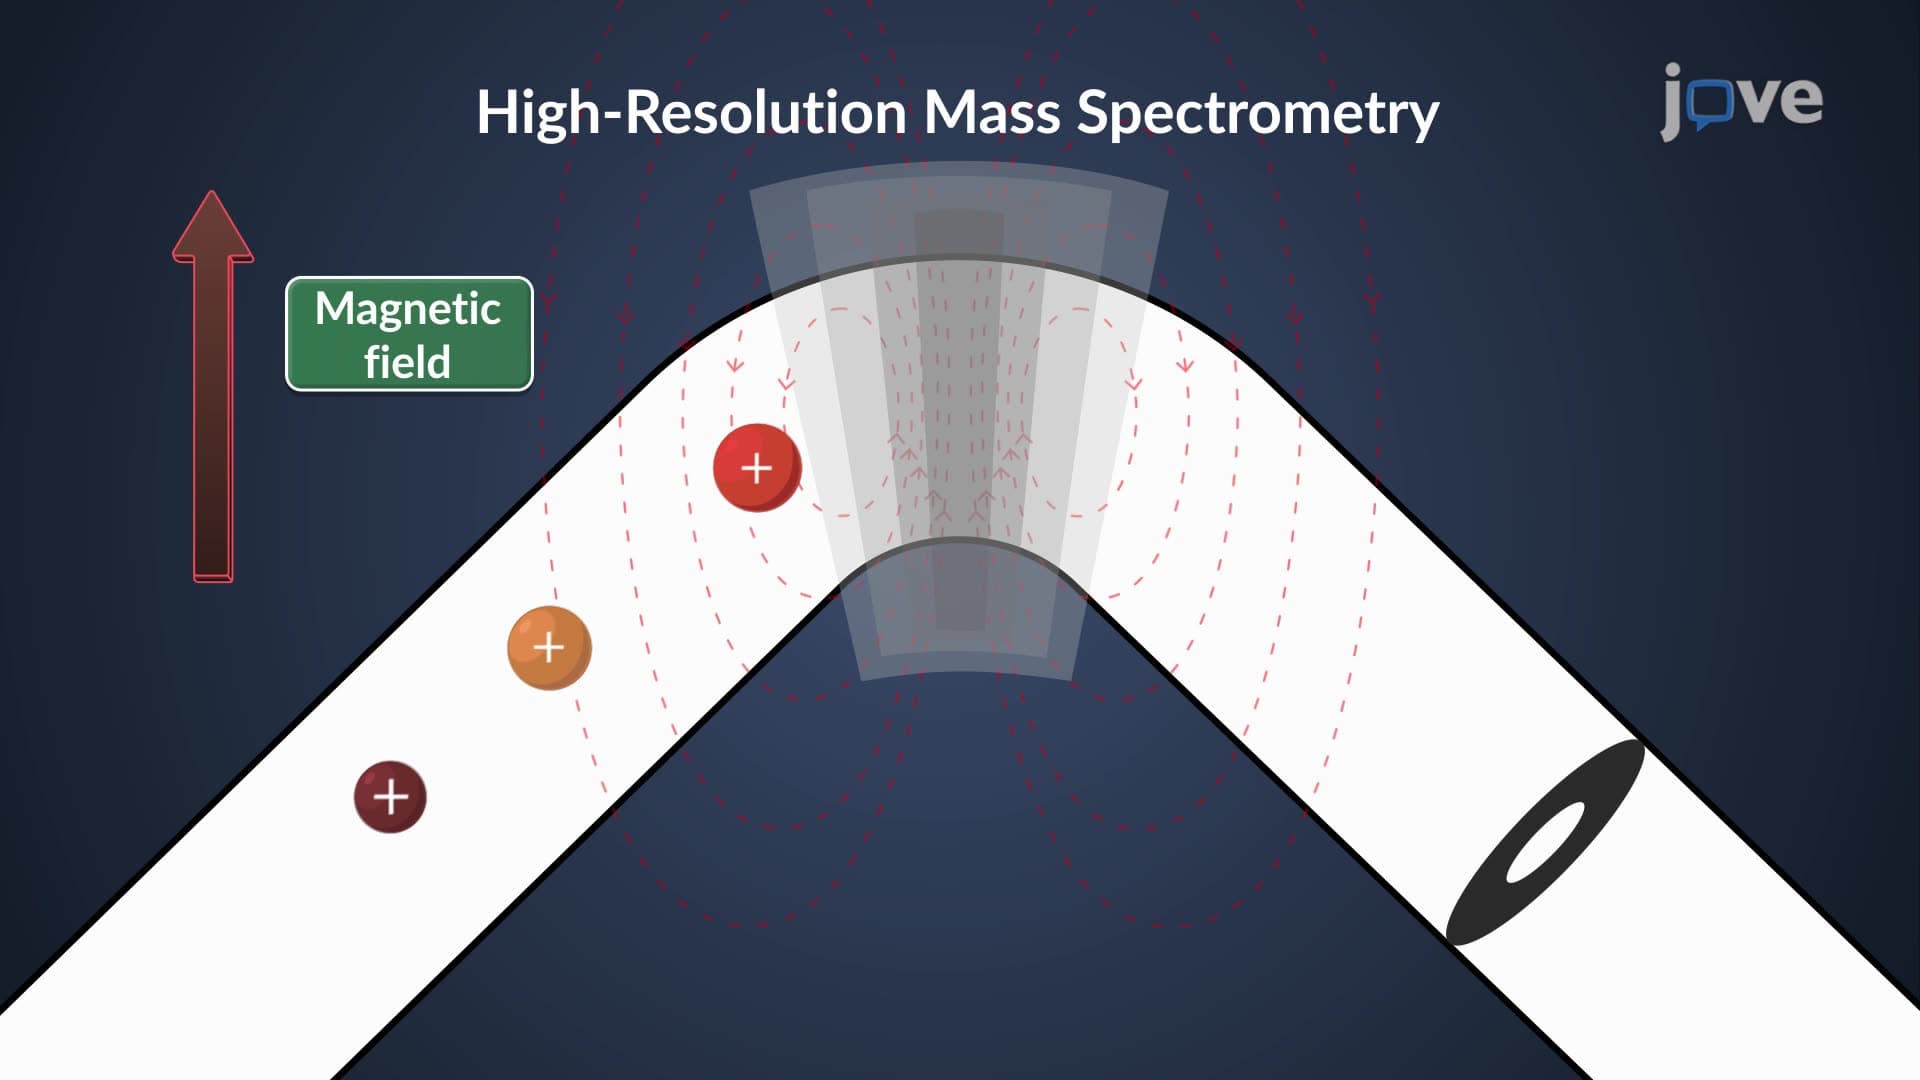
High-Resolution Mass Spectrometry (HRMS)
 01:33
Proteomics
Proteomics is the study of proteomes' function. It involves the large-scale systematic study of the proteome to denote the protein complement expressed by a genome. Scientist Mark Wilkins coined the term proteomics...