Related Experiment Video
Updated: Jun 22, 2026
Atomic Scale Structural Studies of Macromolecular Assemblies by Solid-state Nuclear Magnetic Resonance Spectroscopy
Published on: September 17, 2017
Assessing the accuracy of protein structures by quantum mechanical computations of 13C(alpha) chemical shifts
Jorge A Vila1, Harold A Scheraga
1Baker Laboratory of Chemistry and Chemical Biology, Cornell University, Ithaca, New York 14853-1301, USA.

Abstract:
Two major techniques have been used to determine the three-dimensional structures of proteins: X-ray diffraction and NMR spectroscopy. In particular, the validation of NMR-derived protein structures is one of the most challenging problems in NMR spectroscopy. Therefore, researchers have proposed a plethora of methods to determine the accuracy and reliability of protein structures. Despite these proposals, there is a growing need for more sophisticated, physics-based structure validation methods. This approach will enable us to (a) characterize the "quality" of the NMR-derived ensemble as a whole by a single parameter, (b) unambiguously identify flaws in the sequence at a residue level, and (c) provide precise information, such as sets of backbone and side-chain torsional angles, that we can use to detect local flaws. Rather than reviewing all of the existing validation methods, this Account describes the contributions of our research group toward a solution of the long-standing problem of both global and local structure validation of NMR-derived protein structures. We emphasize a recently introduced physics-based methodology that makes use of observed and computed (13)C(alpha) chemical shifts (at the density functional theory (DFT) level of theory) for an accurate validation of protein structures in solution and in crystals. By assessing the ability of computed (13)C(alpha) chemical shifts to reproduce observed (13)C(alpha) chemical shifts of a single structure or ensemble of structures in solution and in crystals, we accomplish a global validation by using the conformationally averaged root-mean-square deviation, ca-rmsd, as a scoring function. In addition, the method enables us to provide local validation by identifying a set of individual amino acid conformations for which the computed and observed (13)C(alpha) chemical shifts do not agree within a certain error range and may represent a nonreliable fold of the protein model. Although it is computationally intensive, our validation method has several advantages, which we illustrate through a series of applications. This method makes use of the (13)C(alpha) chemical shifts, not shielding, that are ubiquitous to proteins and can be computed precisely from the phi, psi, and chi torsional angles. There is no need for a priori knowledge of the oligomeric state of the protein, and no knowledge-based information or additional NMR data are required. The primary limitation at this point is the computational cost of such calculations. However, we anticipate that enhancements in the speed of calculating these chemical shifts coupled with the ever-increasing computational power should soon make this a standard method accessible to the general NMR community.

Related Concept Videos
 01:02
¹H NMR: Interpreting Distorted and Overlapping Signals
As Δν decreases and the signals move closer, the doublets appear increasingly distorted. The intensities of the inner lines increase at the cost of those of the outer lines as the signals are slanted or...
 01:10
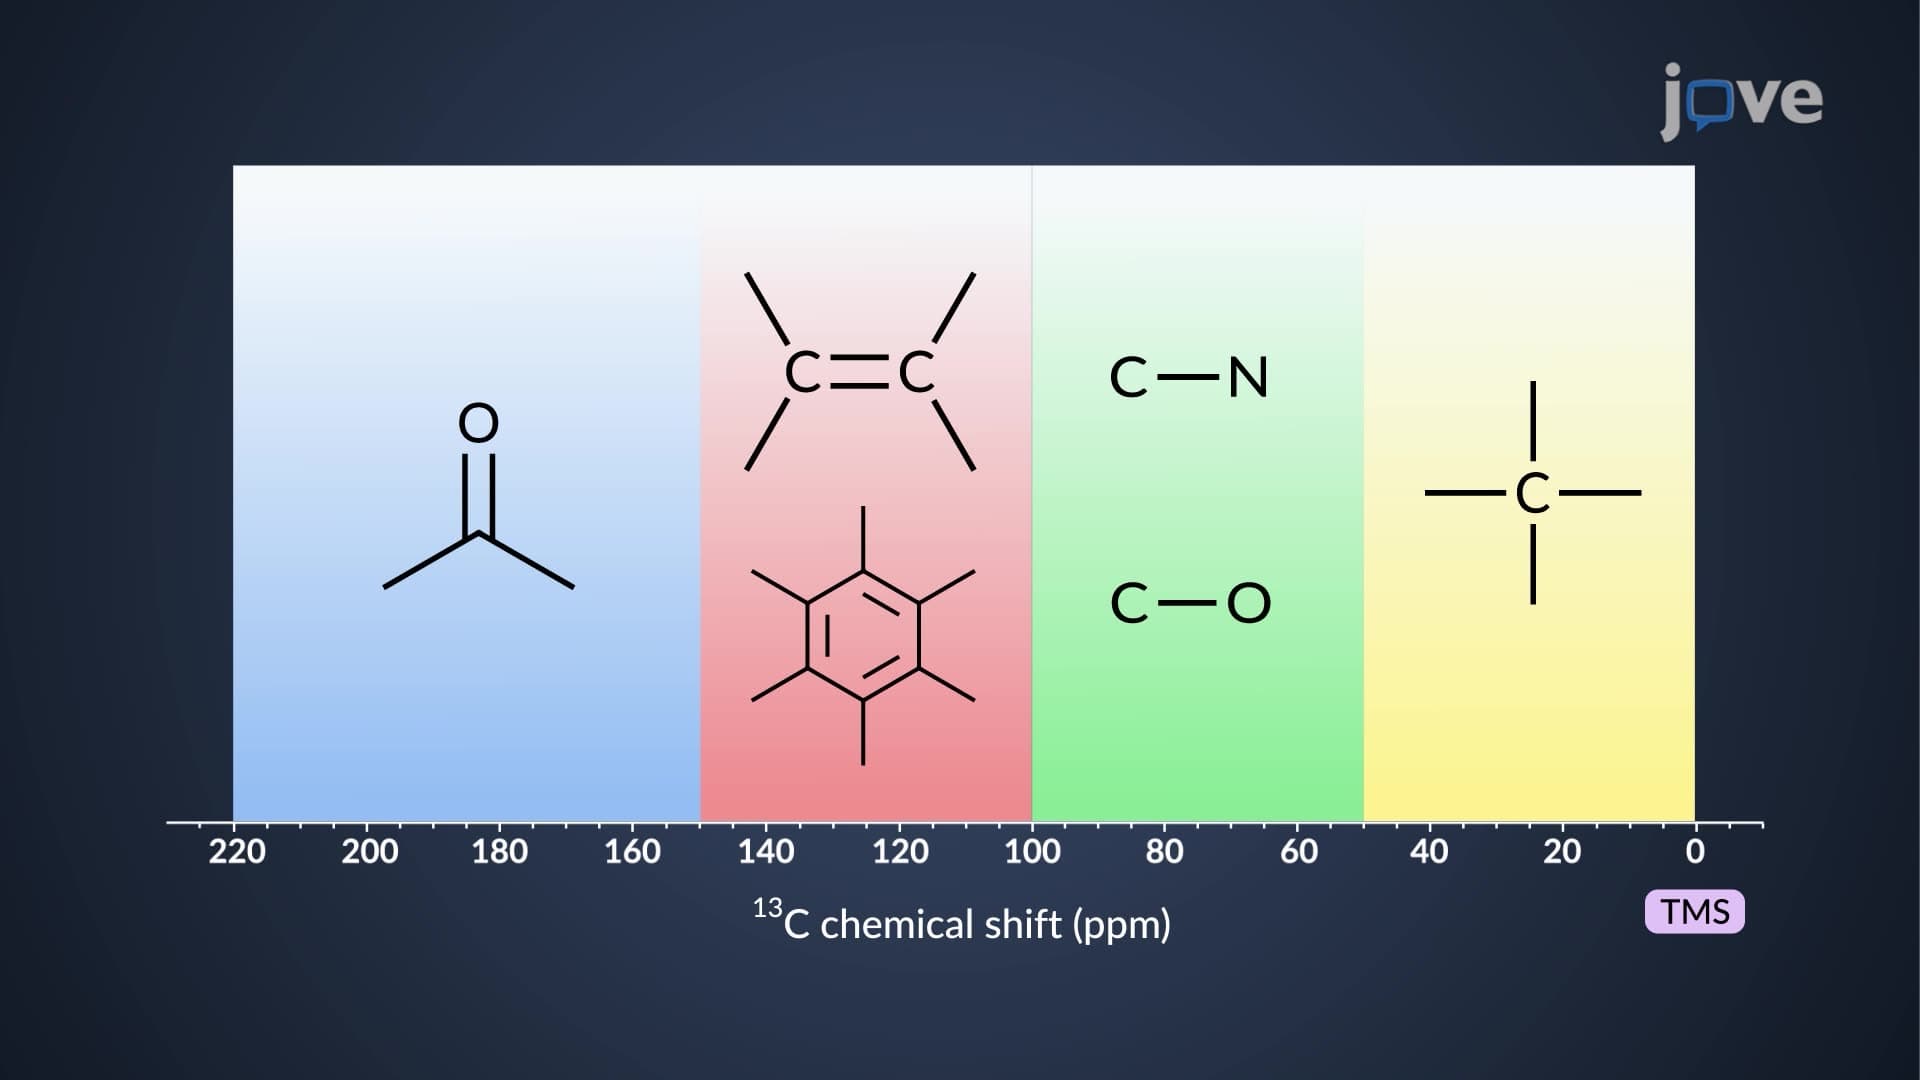
Carbon-13 (¹³C) NMR: Overview
 01:19
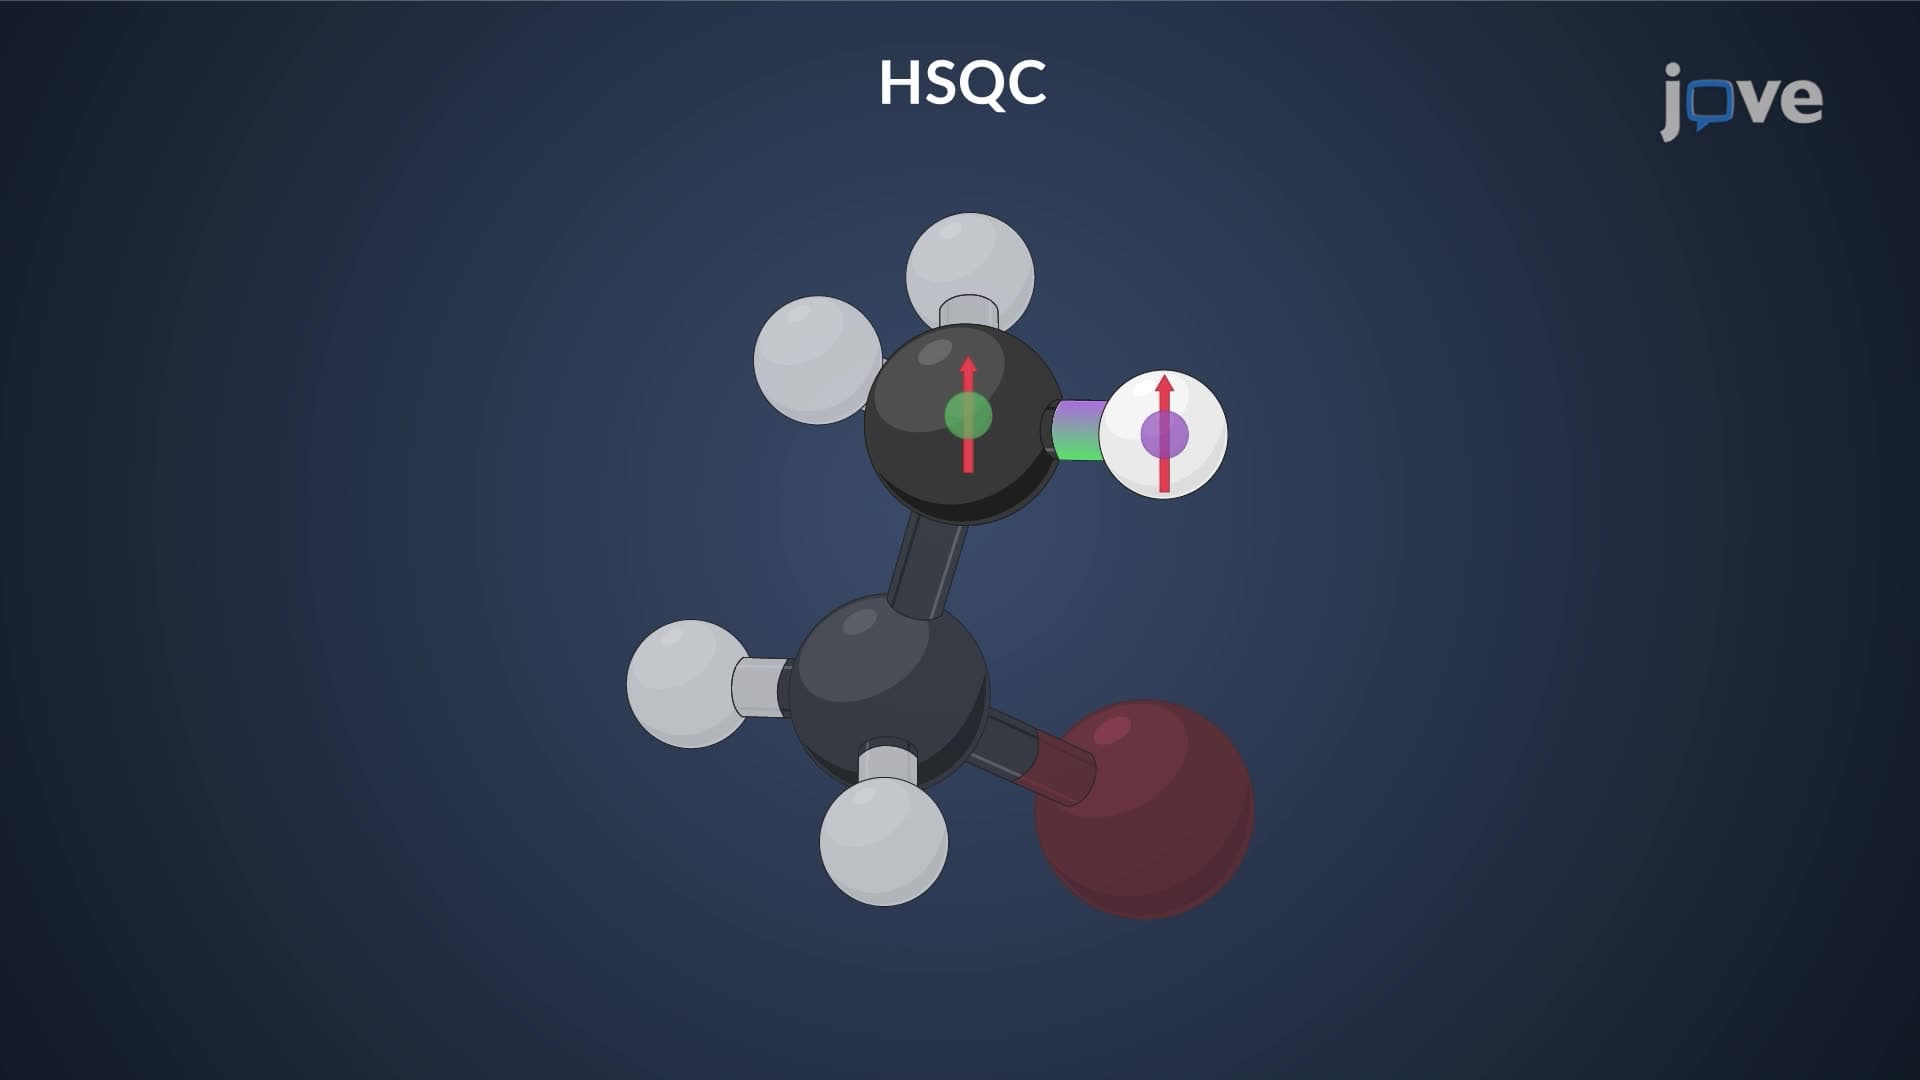
2D NMR: Heteronuclear Single-Quantum Correlation Spectroscopy (HSQC)
 01:24
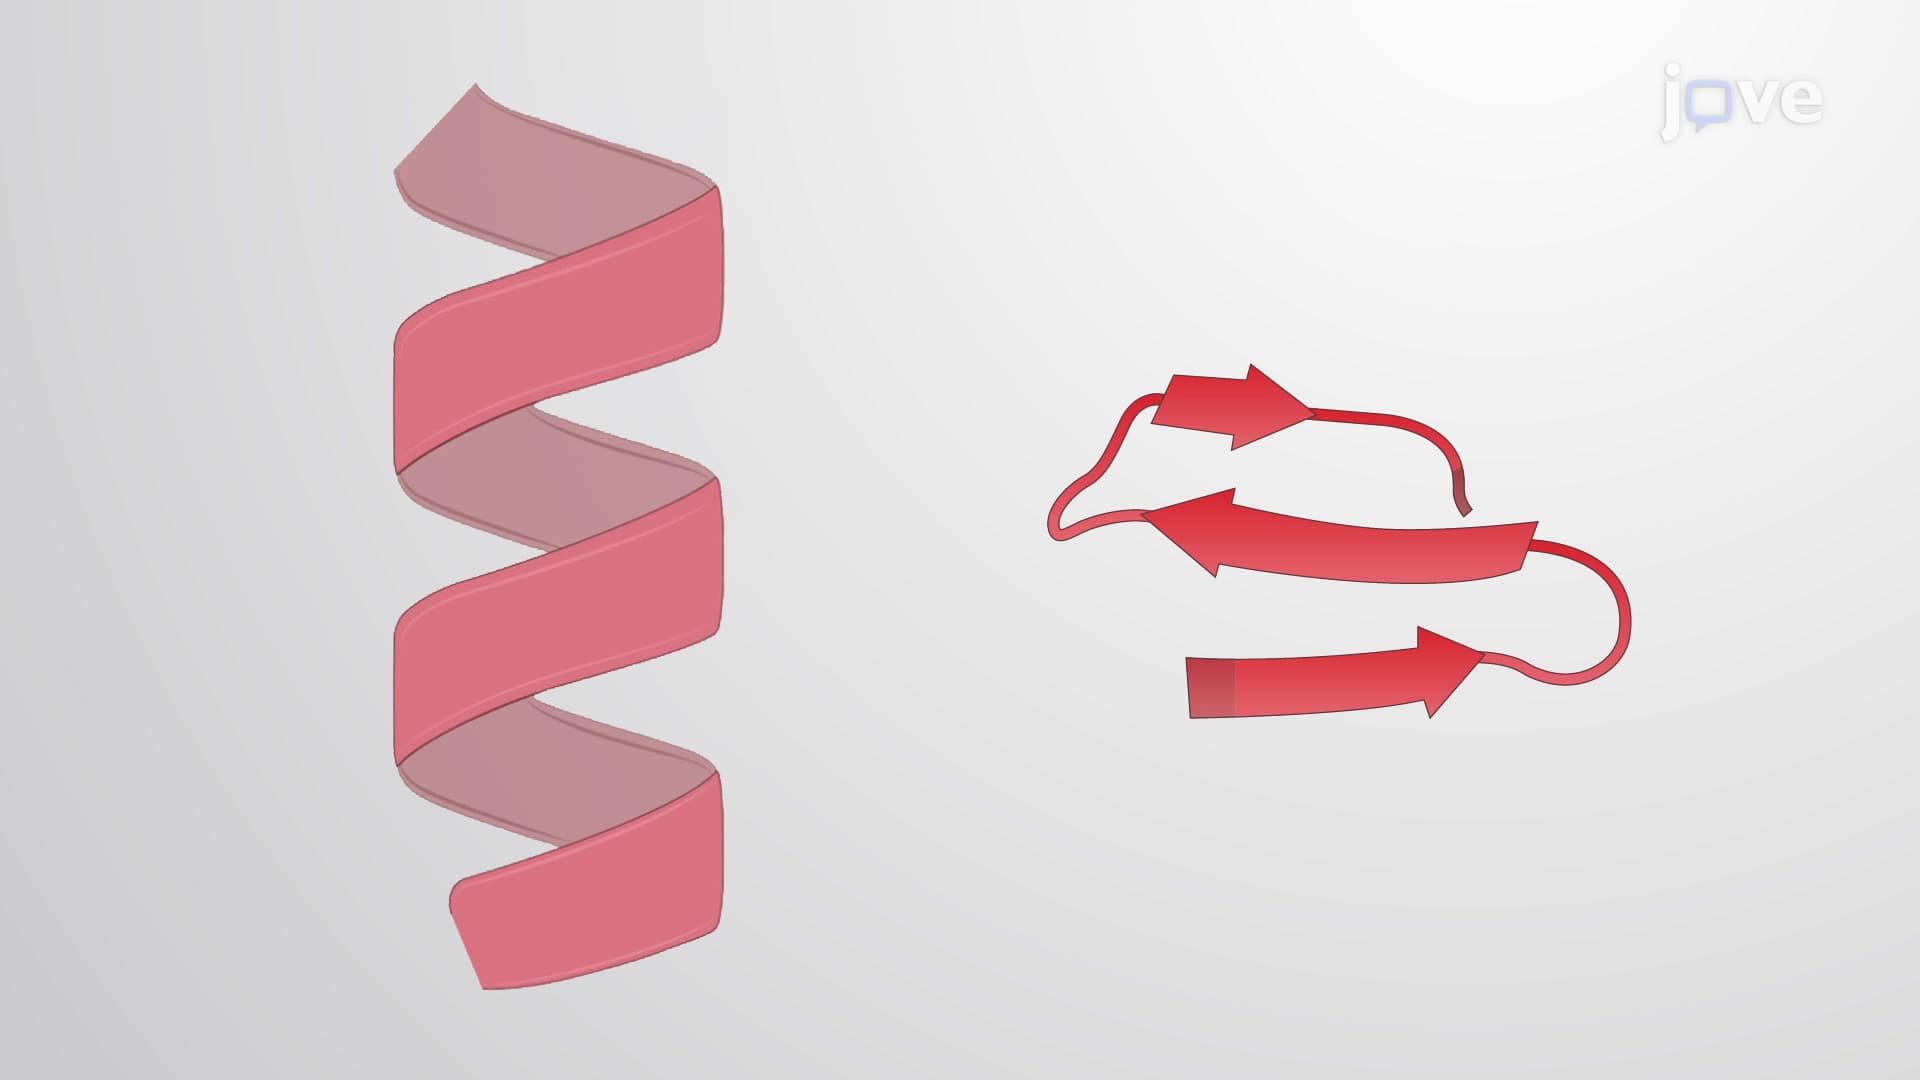
Protein Organization
The primary structure of a protein is its amino acid sequence.
 01:20
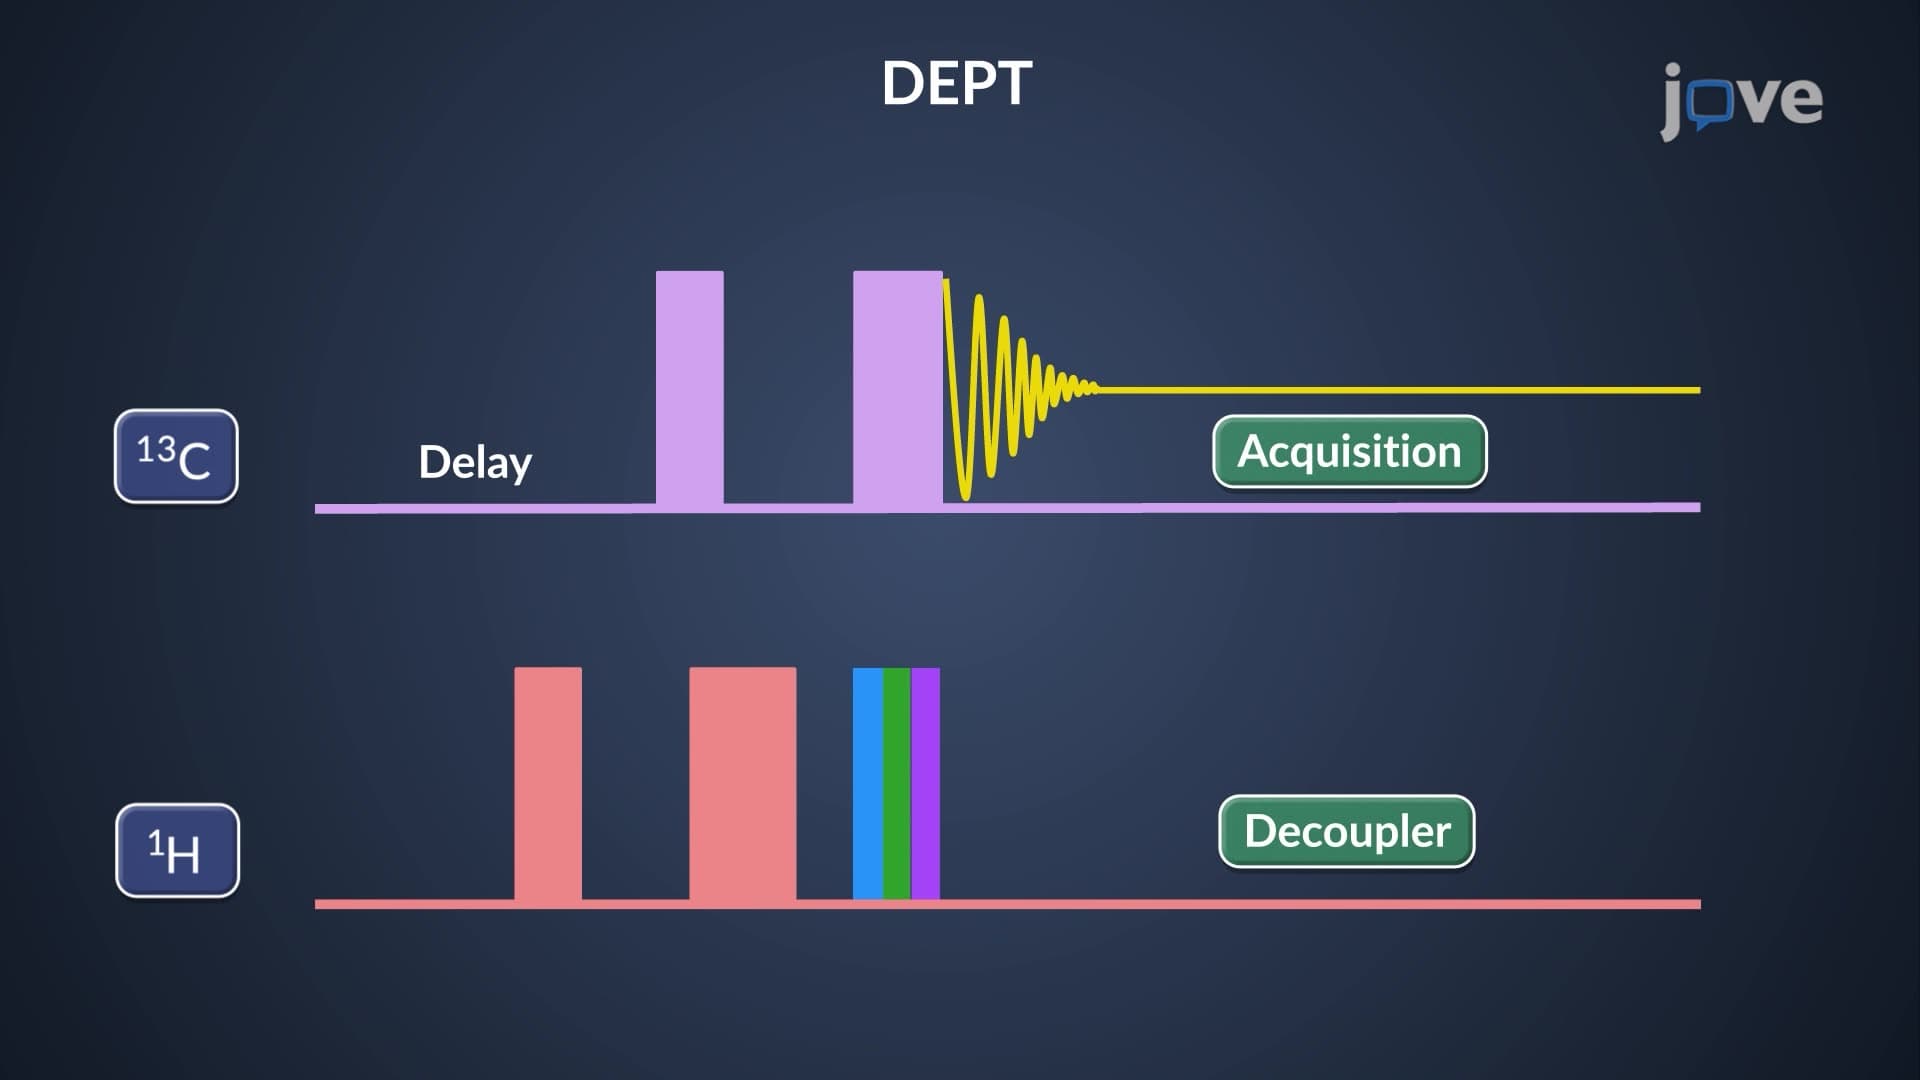
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
 01:19
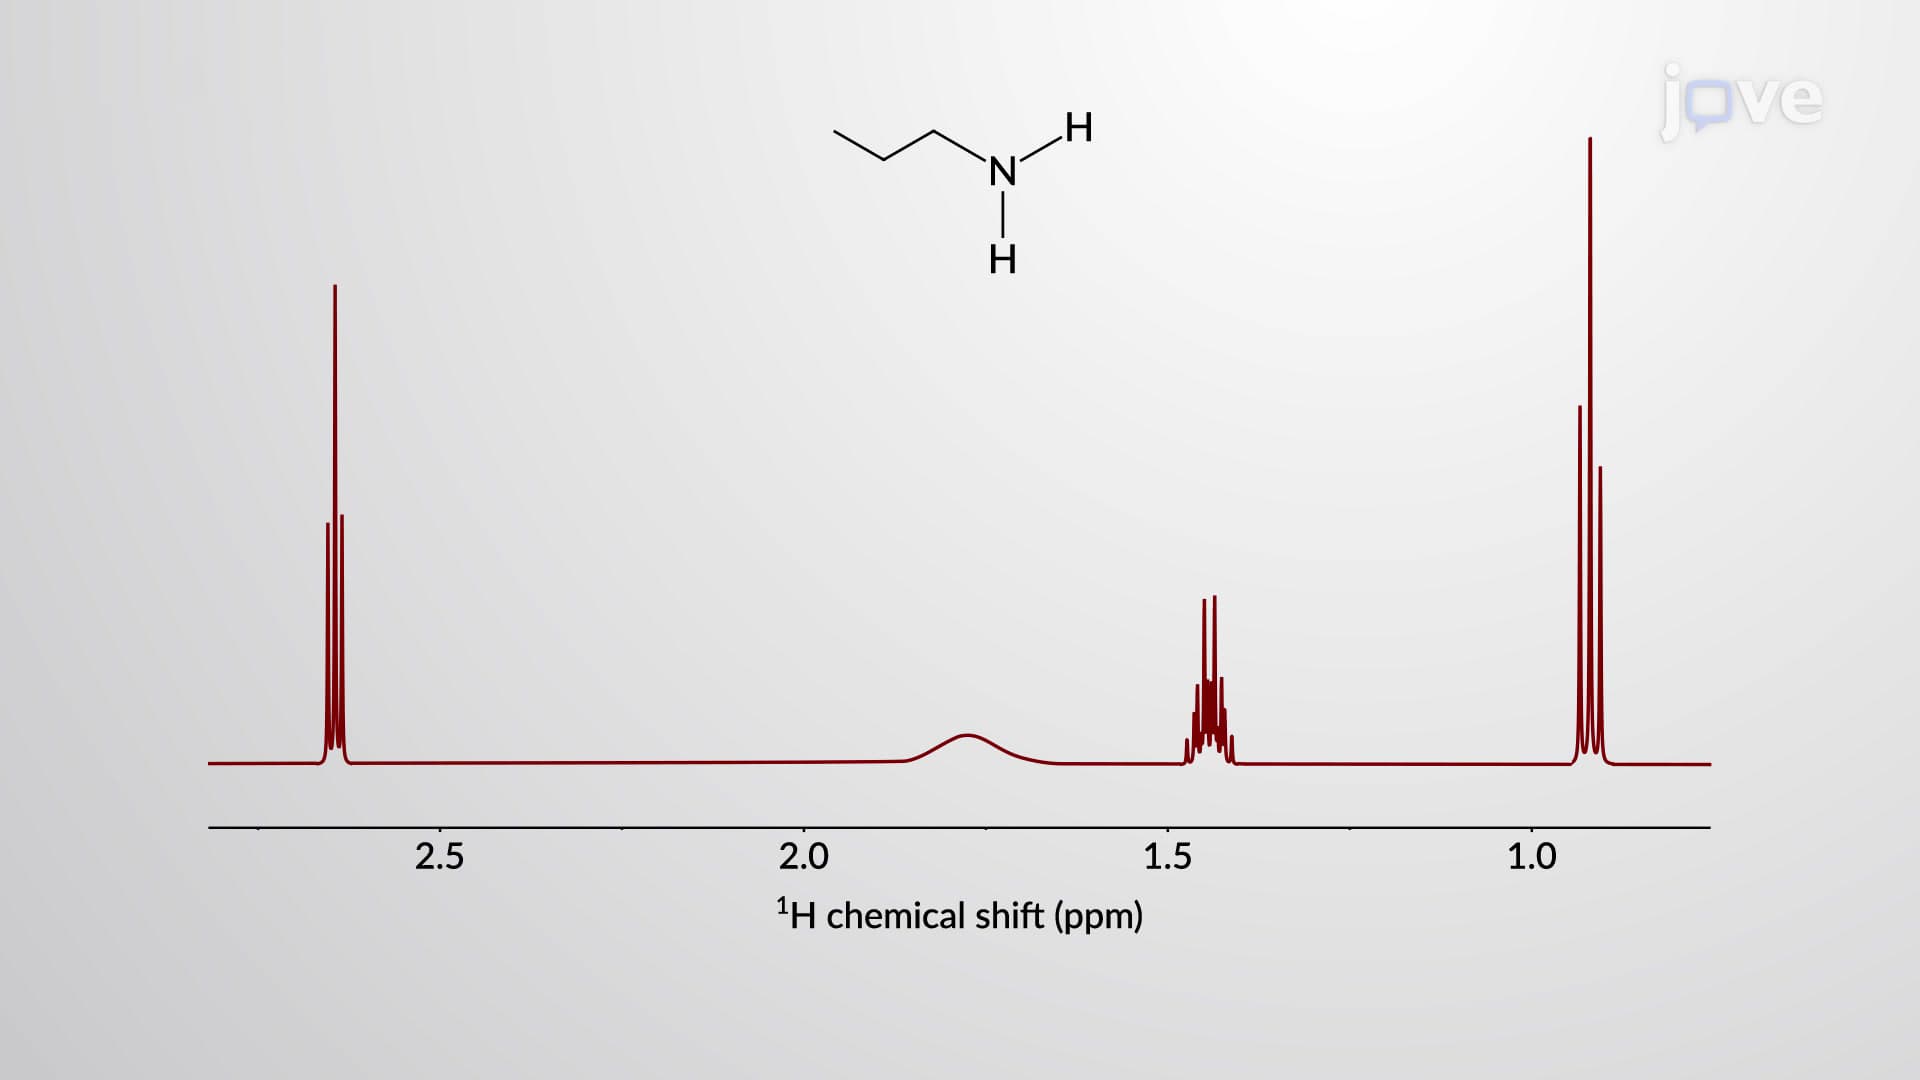
NMR Spectroscopy Of Amines