Related Experiment Video
Updated: Jun 21, 2026

Isolation of Primary Mouse Hepatocytes for Nascent Protein Synthesis Analysis by Non-radioactive L-azidohomoalanine Labeling Method
Published on: October 23, 2018
D-amphetamine toxicity in freshly isolated rat hepatocytes: a possible role of CYP3A
Vessela Vitcheva1, Magdalena Kondeva-Burdina, Mitka Mitcheva
1Laboratory of Drug Metabolism and Drug Toxicity, Department of Pharmacology, Pharmacotherapy, and Toxicology, Faculty of Pharmacy, Medical University Sofia, Sofia, Bulgaria. vesselavitcheva@yahoo.com

Abstract:
The aim of this study was to trace D-amphetamine toxicity in isolated rat hepatocytes and to elucidate a possible involvement of CYP3A in the mechanisms of its toxicity. To this end, male Wistar rats were treated with nifedipine (5 mg kg(-1) i.p., 5 days), a substrate and inducer of CYP3A. Hepatocytes isolated from nifedipine-treated and control rats were incubated with D-amphetamine at a concentration of 100 micromol L(-1), which was determined to be an average toxic concentration (TC50) for the compound. To evaluate the possible toxic effects of D-amphetamine on freshly isolated rat hepatocytes, we assessed the following parameters: cell viability, lactate dehydrogenase (LDH) activity, and glutathione (GSH).The results showed that nifedipine potentiated amphetamine cytotoxicity in vitro, as follows: cell viability dropped by 65% (p>0.001), GSH by 80% (p>0.001), and LDH activity increased by 190% (p>0.001). To clarify the role of nifedipine in amphetamine cytotoxicity, we used amiodarone, a substrate and an inhibitor of CYP3A. Pre-incubation of nifedipine-treated hepatocytes with amiodarone (14 micromol L(-1)) significantly lowered amphetamine cytotoxicity. Our results confirmed the toxicity of D-amphetamine in isolated rat hepatocytes and the involvement of CYP3A in its metabolism and hepatotoxicity.
Insights
This study investigated D-amphetamine toxicity in rat liver cells, finding that CYP3A enzymes play a key role in its harmful effects. Nifedipine treatment increased D-amphetamine
Area of Science:
- Hepatology
- Toxicology
- Pharmacology
Background:
- D-amphetamine is a stimulant with known toxic effects.
- Cytochrome P450 3A (CYP3A) enzymes are crucial for drug metabolism and can influence toxicity.
- Understanding the specific mechanisms of D-amphetamine hepatotoxicity is important for clinical safety.
Purpose of the Study:
- To investigate the toxicity of D-amphetamine in isolated rat hepatocytes.
- To determine the potential involvement of CYP3A in D-amphetamine-induced liver injury.
- To assess how CYP3A induction affects D-amphetamine cytotoxicity.
Main Methods:
- Isolated rat hepatocytes were used to study D-amphetamine toxicity.
- Hepatocytes from rats pre-treated with nifedipine (a CYP3A inducer) were exposed to D-amphetamine.
- Cell viability, lactate dehydrogenase (LDH) release, and glutathione (GSH) levels were measured.
- Amiodarone (a CYP3A inhibitor) was used to confirm the role of CYP3A.
Main Results:
- D-amphetamine significantly reduced cell viability and glutathione levels while increasing LDH release in isolated hepatocytes.
- Pre-treatment with nifedipine potentiated D-amphetamine's toxic effects, indicating increased hepatotoxicity.
- Inhibition of CYP3A with amiodarone significantly reduced the potentiation of D-amphetamine cytotoxicity caused by nifedipine.
Conclusions:
- D-amphetamine exhibits significant toxicity in isolated rat hepatocytes.
- The cytochrome P450 3A enzyme system is involved in the metabolism and subsequent hepatotoxicity of D-amphetamine.
- CYP3A induction can exacerbate D-amphetamine-induced liver injury.
Related Concept Videos
 01:22
Effect of Hepatic Disease on Pharmacokinetics: Pathophysiologic Assessment and Liver Function Test
 01:28
Pharmacogenetics of Phase I Enzymes: Cytochrome P450 Isozymes
 01:07
Physiological Pharmacokinetic Models: Incorporating Hepatic Transporter-Mediated Clearance
A recent model describes pravastatin's hepatobiliary excretion, mediated...
 01:14
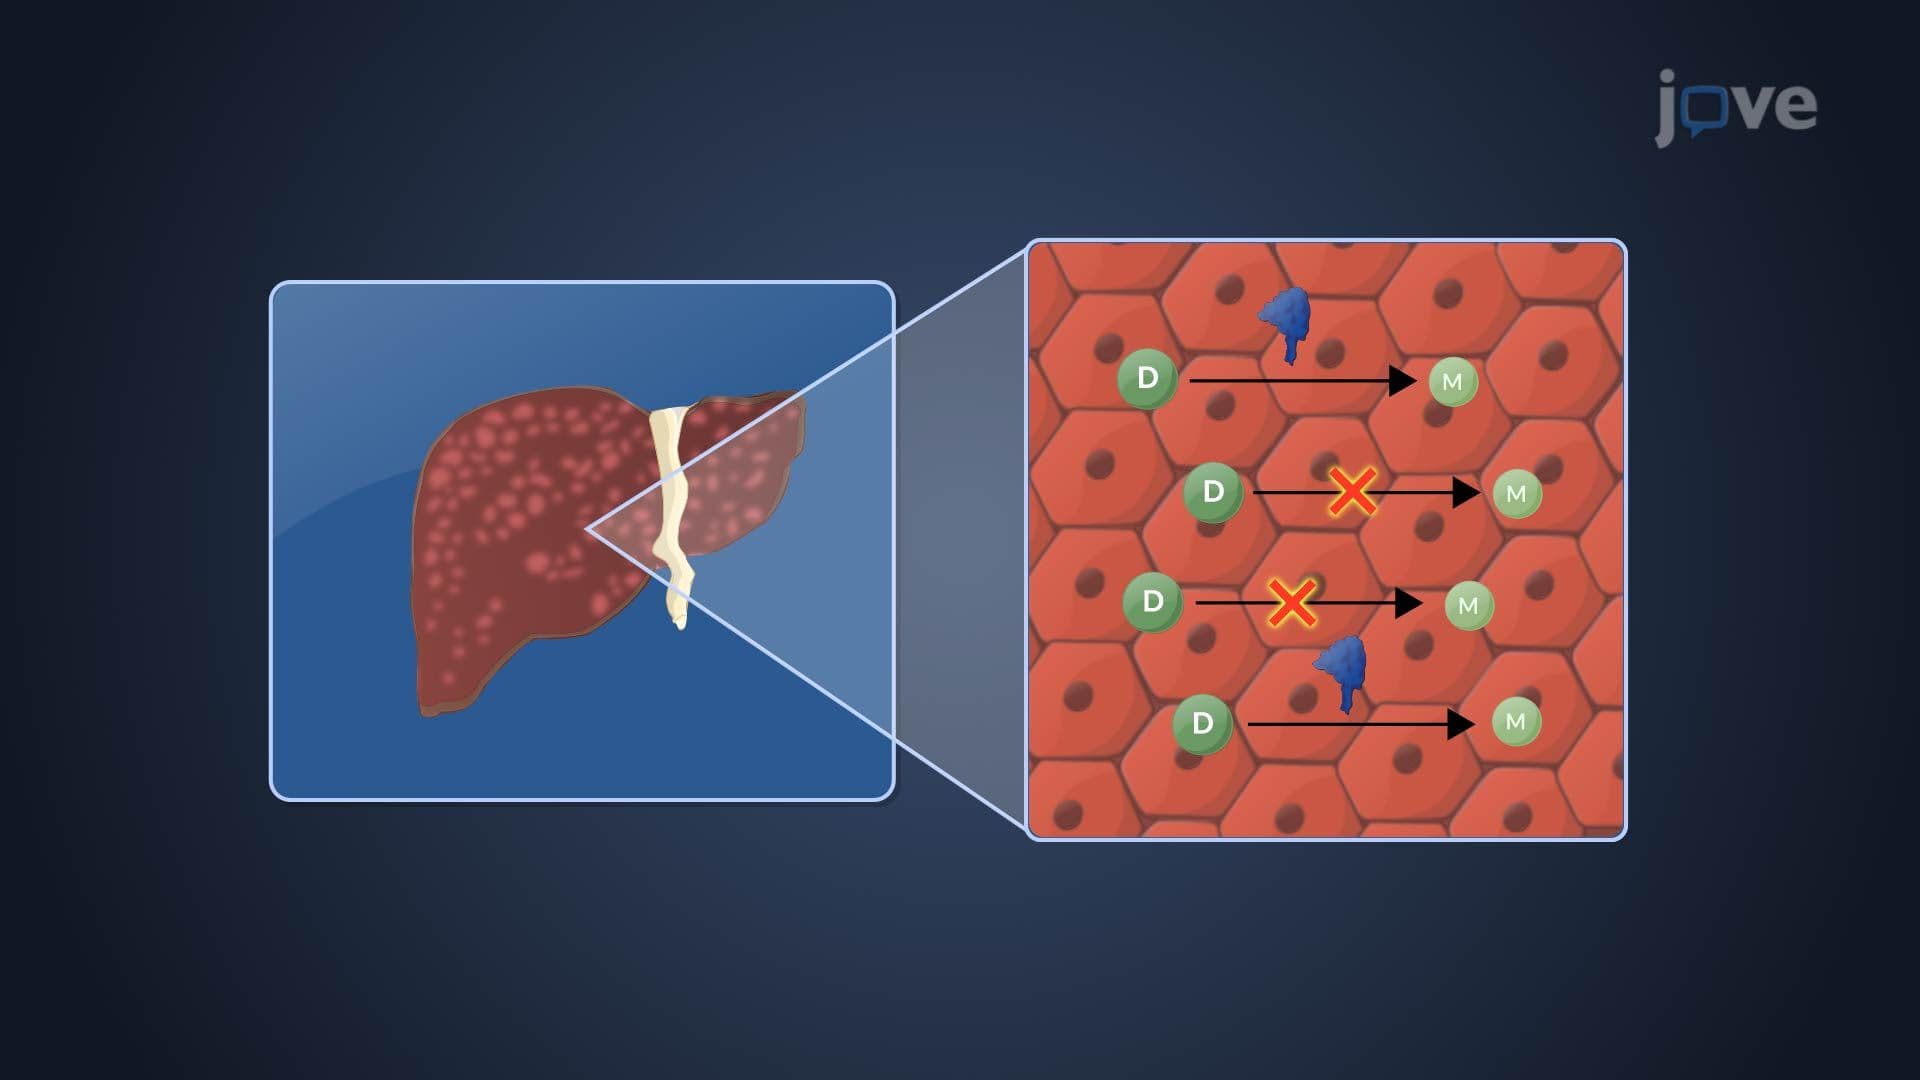
Effect of Hepatic Disease on Pharmacokinetics: Active Drug, Metabolite and Fraction of Metabolized Drug
 01:24
Drug Toxicity: Risk factors
 01:16
Drug toxicity: Idiosyncratic Reactions