Related Experiment Video
Updated: Jun 21, 2026


Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
Degenerate intermolecular and intramolecular proton-transfer reactions: electronic structure of the transition states

Abstract:
Density functional theory (DFT) calculations are performed on a series of double and single proton-transfer reactions to study the variation in polarizations in complexes during the dynamics of proton transfer from one isoenergetic, hydrogen-bonded ground-state structure to the other. The isotropic average polarizability (alpha(av)) shows an interesting single-humped profile with a maxima coinciding with the transition state of the reaction. Similar profiles are also computed at Nd:YaG frequencies. The origin of the maximal polarizability at the transition state is traced to maximal charge separation and large D (donor)-A (acceptor) distances. Maximal polarizability for the transition state suggests an interesting, novel, and less memory extensive computational tool to locate the transition state for hydrogen-transfer reactions in hydrogen-bonded complexes.
More Related Videos

Related Concept Videos
 02:13
Energy Diagrams, Transition States, and Intermediates
 01:25
Deactivation Processes: Jablonski Diagram
 01:16
UV–Vis Spectroscopy: Molecular Electronic Transitions
 01:25
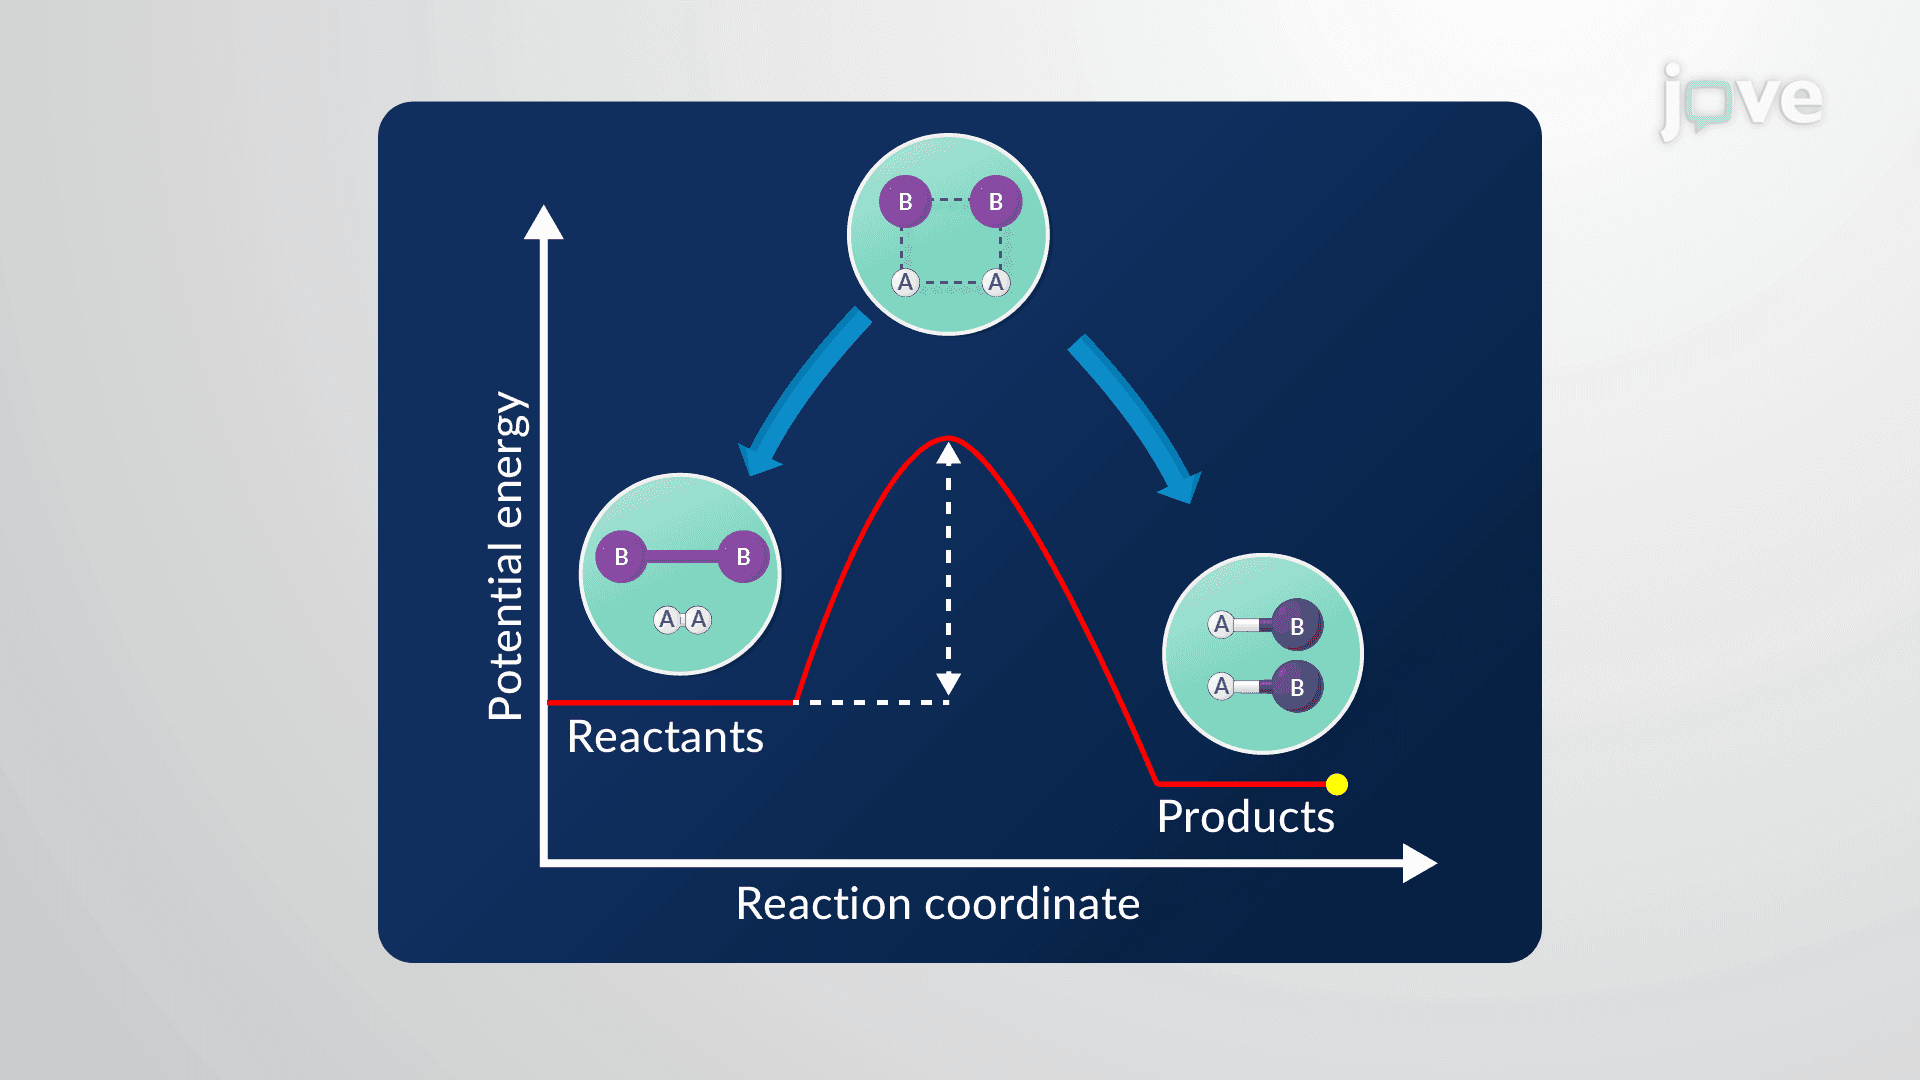
Transition State Theory
 02:16
Resonance and Hybrid Structures
Resonance Structures and Resonance Hybrids
The Lewis structure of a nitrite anion (NO2−) may actually be drawn in two different ways, distinguished by the locations of the N–O and N=O bonds.
 02:26
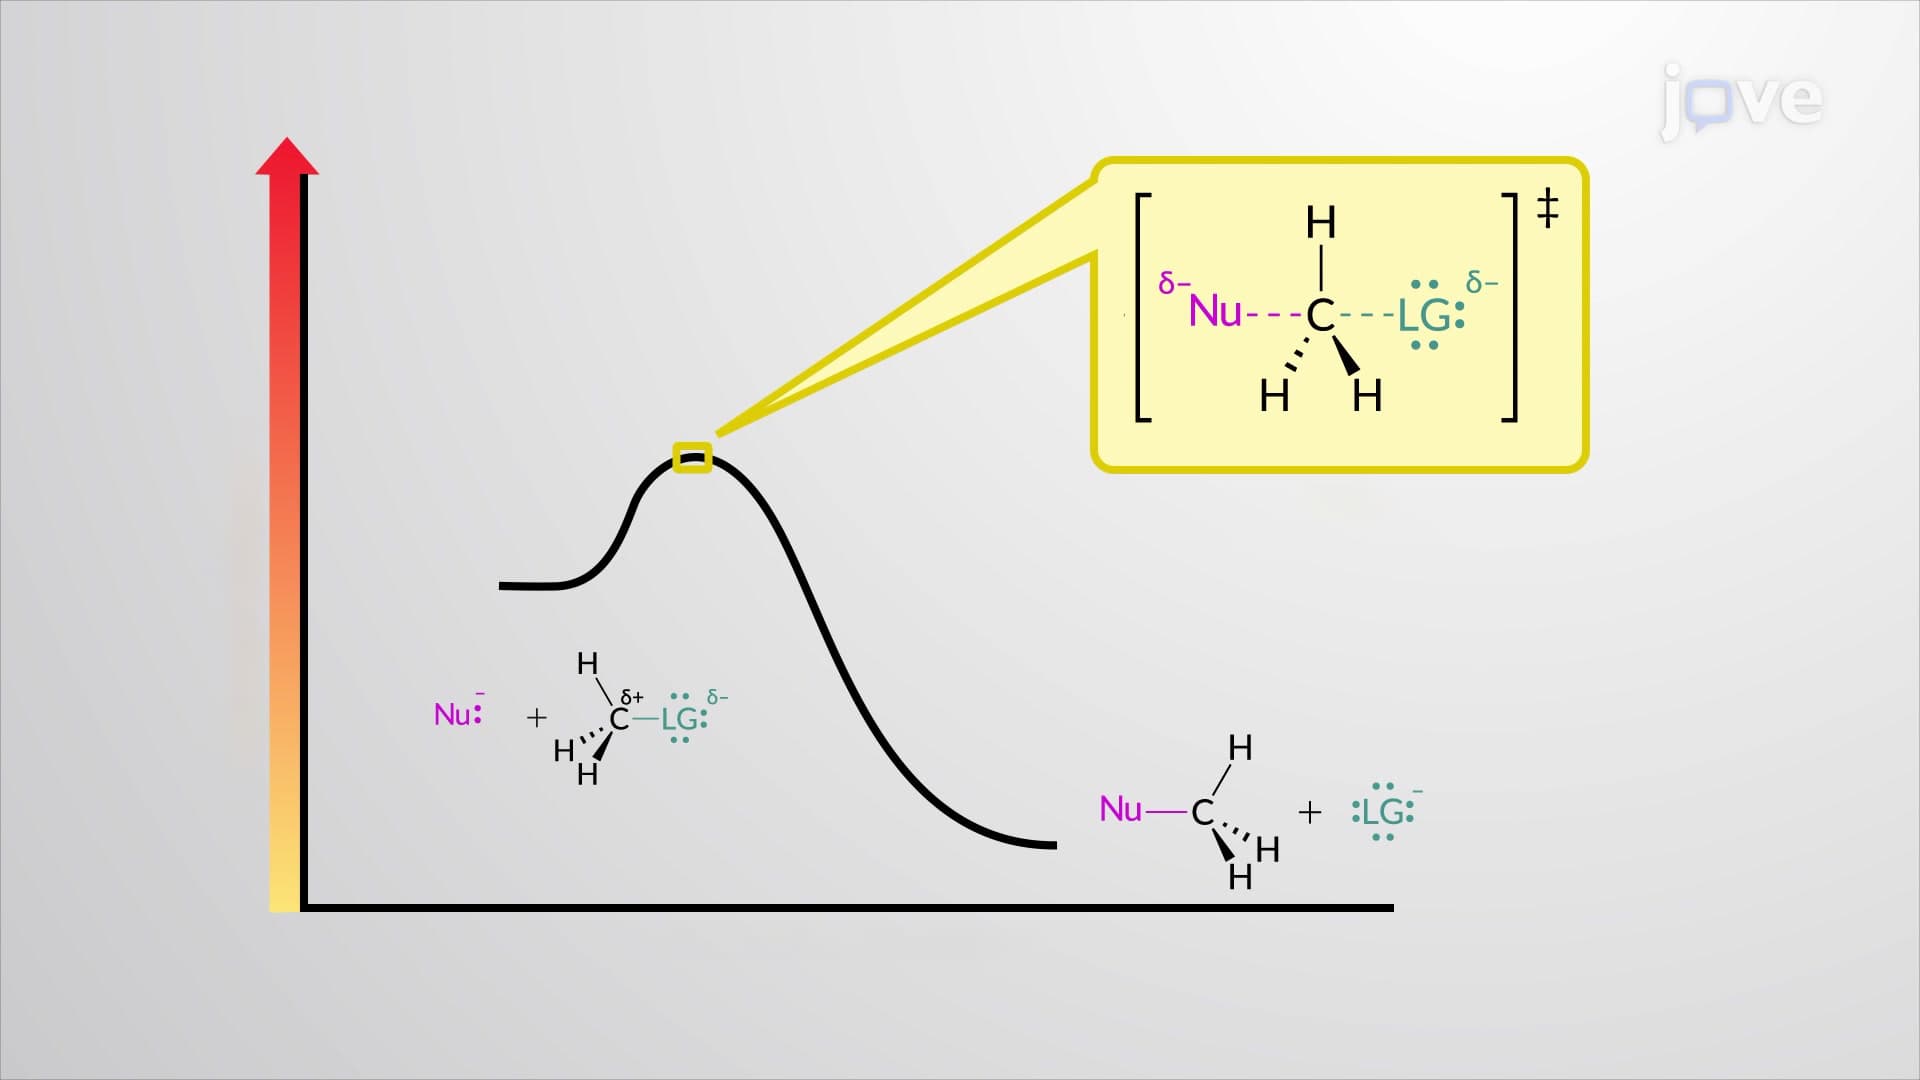
SN2 Reaction: Transition State
When the nucleophile approaches the electrophilic carbon with its lone pairs, the halide acts as a leaving group and moves away with the electron-pair bonded to the carbon. Dotted partial bonds represent the bonds being formed or broken...