Related Experiment Video
Updated: Jun 21, 2026

An In Vitro Approach to Study Mitochondrial Dysfunction: A Cybrid Model
Published on: March 9, 2022
Clinical and molecular aspects of diseases of mitochondrial DNA instability
1Department of Anesthesiology, Chang Gung Memorial Hospital, Taipei, Chang Gung University College of Medicine, Taoyuan, Taiwan. ccm.mao@googlemail.com

Abstract:
Mitochondria within human cells contain vast numbers of mitochondrial DNA (mtDNA), which are small, circular, and double-stranded. The proper functions of mtDNA depend totally on specific proteins that are encoded by the nucleus and then imported into mitochondria. Thus instability of mtDNA can stem from the mtDNA itself, or secondarily from abnormalities in nuclear DNA. In this review, we will first introduce mtDNA, including its characteristics, replication, transcription, translation, and the proteins involved in its metabolism, in particular DNA polymerase gamma (POLG), DNA helicase Twinkle (Twinkle), and mitochondrial transcription factor A (TFAM). Secondly, we will stress the importance of mitochondrial nucleoid structures in the protection and facilitation of mtDNA metabolism, and report on the few known protein components of nucleoid, especially Twinkle, TFAM, and the recently discovered ATAD3. Based on this information, mtDNA instabilities will be categorized in accordance with their molecular etiologies, those that are caused by primary defects of mtDNA, and those by secondary effects from abnormalities in nuclear DNA. The former includes large defects or point mutations of mtDNA. The latter involves the nuclear genes of POLG1, Twinkle, ANT1, TK2, dGK, and TP. With the comprehensive categorization in this review, links are provided between the molecular and clinical aspects of mitochondrial DNA diseases. This report should help medical staff understand the complexity of these diseases and encourage them in further investigations. (
Insights
Mitochondrial DNA (mtDNA) instability arises from defects within the mtDNA itself or nuclear DNA abnormalities affecting mtDNA metabolism. This review categorizes these instabilities, linking molecular causes to clinical aspects of mitochondrial diseases.
Area of Science:
- Cellular Biology
- Genetics
- Biochemistry
Background:
- Mitochondria contain circular mitochondrial DNA (mtDNA) crucial for cellular function.
- mtDNA relies on nuclear-encoded proteins for replication, transcription, and translation.
- Instability in mtDNA can originate from intrinsic mtDNA defects or secondary nuclear DNA abnormalities.
Purpose of the Study:
- To review the characteristics of mtDNA and its associated proteins.
- To emphasize the role of mitochondrial nucleoid structures in mtDNA metabolism.
- To categorize mtDNA instabilities based on molecular etiologies and link them to clinical manifestations.
Main Methods:
- Literature review of mtDNA characteristics, metabolism, and nucleoid components.
- Categorization of mtDNA instabilities based on primary mtDNA defects or secondary nuclear gene abnormalities.
- Correlation of molecular defects with clinical aspects of mitochondrial DNA diseases.
Main Results:
- Detailed introduction to mtDNA, including its replication, transcription, translation, and key proteins like POLG, Twinkle, and TFAM.
- Emphasis on mitochondrial nucleoid structure and its protein components (Twinkle, TFAM, ATAD3).
- Classification of mtDNA instabilities into primary mtDNA defects (large deletions, point mutations) and secondary nuclear defects (e.g., POLG1, Twinkle, ANT1, TK2, dGK, TP gene mutations).
Conclusions:
- mtDNA instabilities have diverse molecular origins, stemming from either mtDNA itself or nuclear DNA.
- Understanding these molecular etiologies is crucial for diagnosing and managing mitochondrial DNA diseases.
- This review provides a framework connecting molecular defects to clinical presentations, aiding medical professionals.

Related Concept Videos
 01:45
Mitochondrial Membranes
 02:59
Animal Mitochondrial Genetics
 01:37
Mitochondria
 01:19
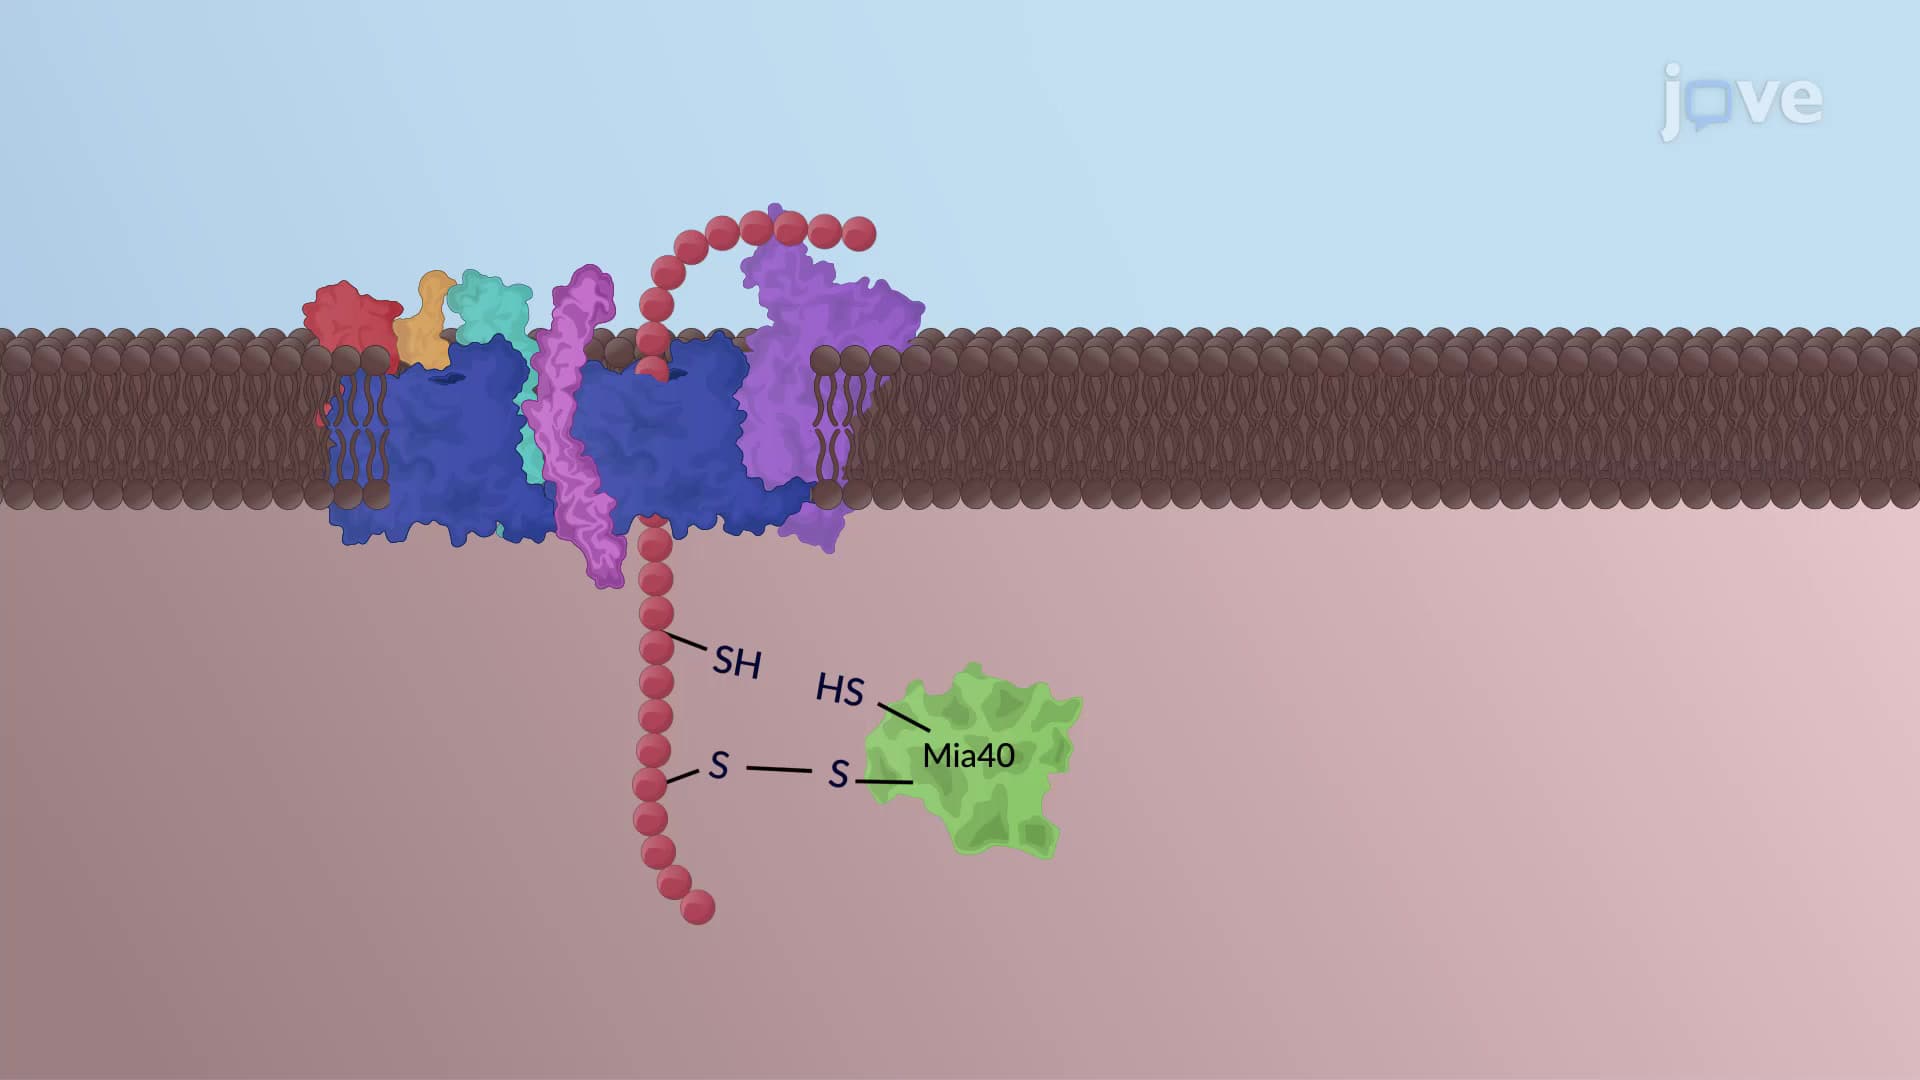
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...
 01:46
Electron Transport Chain: Complex I and II
ROS generation is regulated and maintained at moderate levels necessary...
 01:39
Mitochondrial Precursor Proteins
Most of the mitochondrial precursors...