A fast and efficient gene-network reconstruction method from multiple over-expression experiments
Related Experiment Video
Updated: Jun 20, 2026
JUMPn: A Streamlined Application for Protein Co-Expression Clustering and Network Analysis in Proteomics
Published on: October 19, 2021
Dejan Stokić1, Rudolf Hanel, Stefan Thurner
1Complex Systems Research Group, Medical University of Vienna, A-1090, Austria. stokic@swissquant.ch

Background:
Reverse engineering of gene regulatory networks presents one of the big challenges in systems biology. Gene regulatory networks are usually inferred from a set of single-gene over-expressions and/or knockout experiments. Functional relationships between genes are retrieved either from the steady state gene expressions or from respective time series.
Results:
We present a novel algorithm for gene network reconstruction on the basis of steady-state gene-chip data from over-expression experiments. The algorithm is based on a straight forward solution of a linear gene-dynamics equation, where experimental data is fed in as a first predictor for the solution. We compare the algorithm's performance with the NIR algorithm, both on the well known E. coli experimental data and on in-silico experiments.
Conclusion:
We show superiority of the proposed algorithm in the number of correctly reconstructed links and discuss computational time and robustness. The proposed algorithm is not limited by combinatorial explosion problems and can be used in principle for large networks.
Related Concept Videos
 02:05
Gene Evolution - Fast or Slow?
In contrast, regions which code...
02:05Gene Evolution - Fast or Slow?
In contrast, regions which code...
 02:34
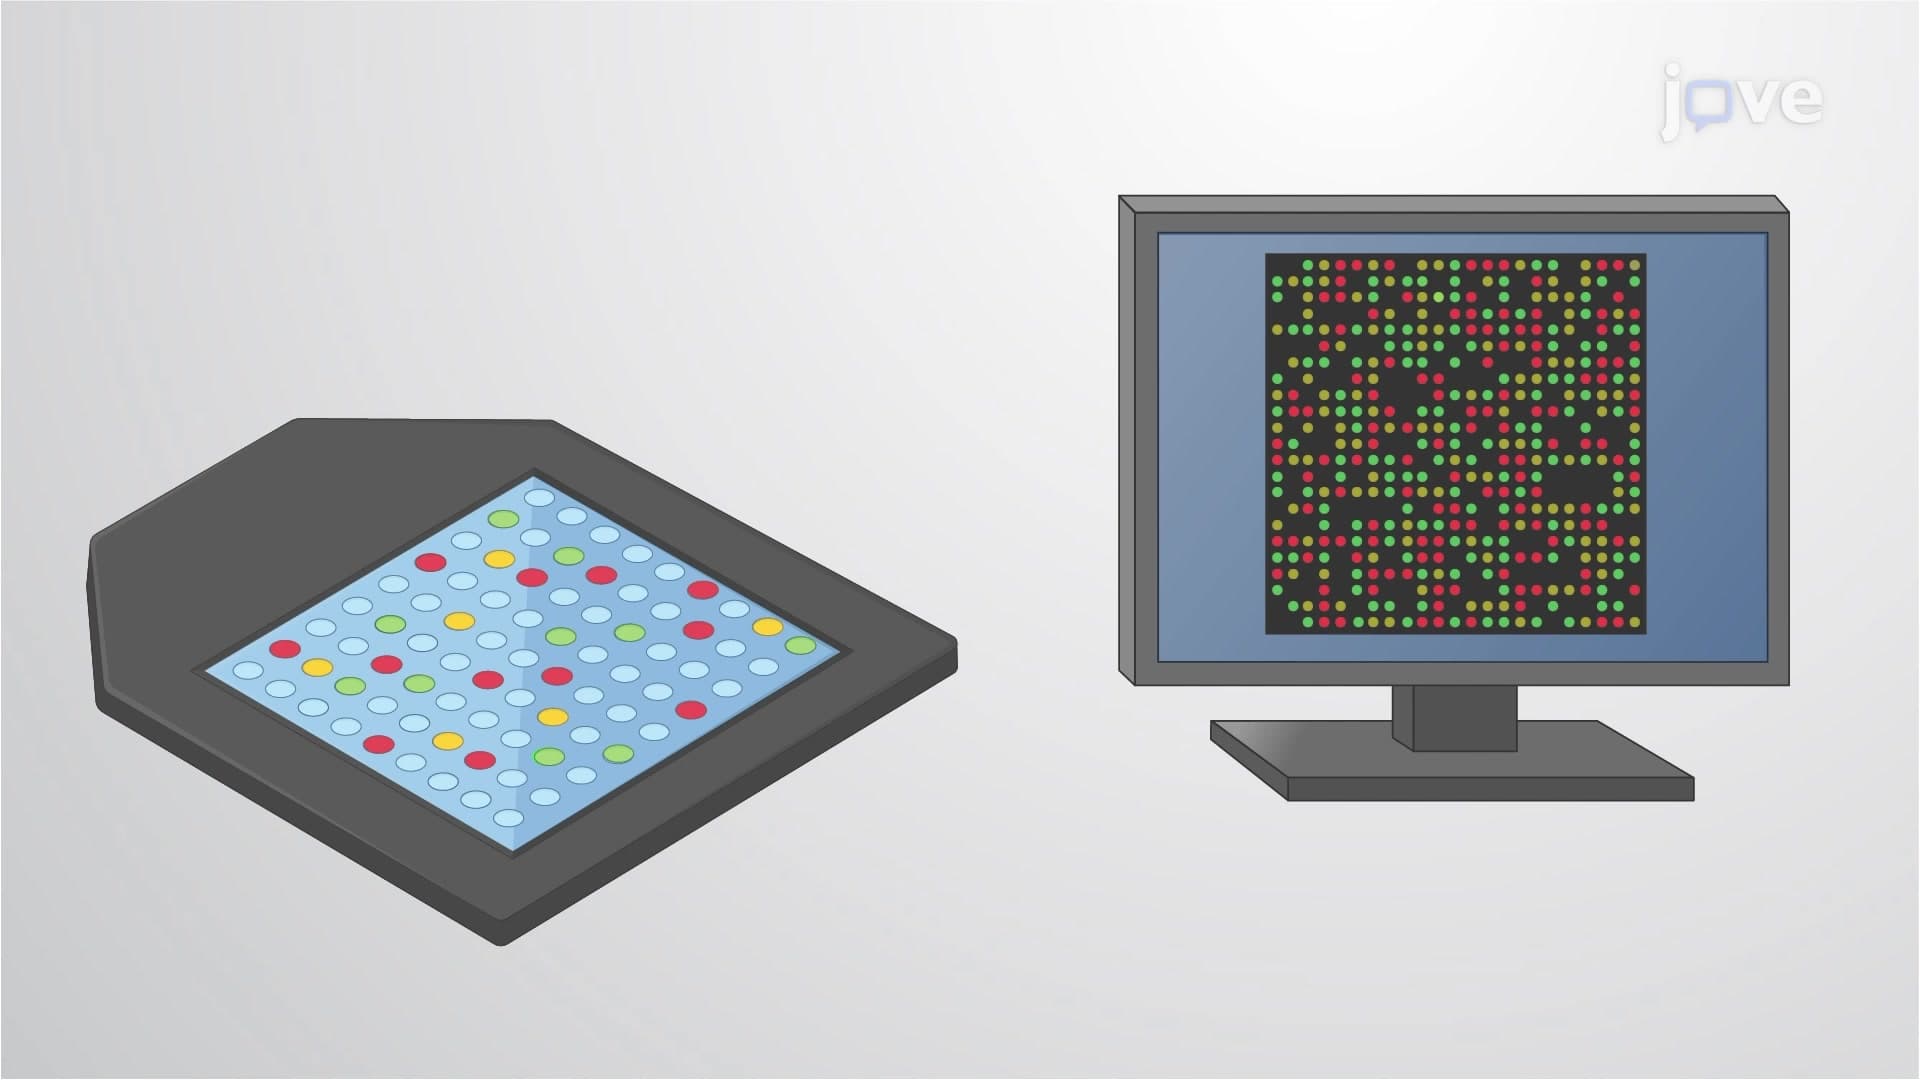
DNA Microarrays