Related Experiment Video
Updated: Jun 20, 2026

X-Ray Crystallography to Study the Oligomeric State Transition of the Thermotoga maritima M42 Aminopeptidase TmPep1050
Published on: May 13, 2020
Structural and mechanistic insights into the oxy form of tyrosinase from molecular dynamics simulations
Robert J Deeth1, Christian Diedrich
1Inorganic Computational Chemistry Group, Department of Chemistry, University of Warwick, Coventry, UK. r.j.deeth@warwick.ac.uk

Abstract:
The first, long time scale (16-ns) ligand field molecular dynamics (LFMD) simulations of the oxy form of tyrosinase are reported. The calculations use our existing type 3 copper force field for the peroxido-bridged [Cu(2)O(2)](2+) unit which is here translated from MMFF into the AMBER format together with a new charge scheme. The protein secondary and tertiary structures are not significantly altered by removing the 'caddie' protein, ORF378, which must be bound to tyrosinase before crystals will grow. A comprehensive principal component analysis of the Cartesian coordinates from the final 8 ns shows that the protein backbone is relatively rigid. However, the significant butterfly fold of the [Cu(2)O(2)](2+) moiety observed in the X-ray structure, presumably due to the caddie protein tyrosine at the active site, is absent in the simulations. LFMD gives a clear and persistent distinction between equatorial and axial Cu-N distances, with the latter about 0.2 A longer and remaining syn to each other. However, the two coordination spheres display important differences. LFMD simulations of the symmetric model complex [mu-eta(2):mu(2)-O(2){Cu(Meim)(3)}(2)](2+) (Meim is 5-methyl-1H-imidazole) provide a mechanism for syn-anti interchange of axial ligands which suggests, in combination with the old experimental X-ray data, the new LFMD simulations and traditional coordination chemistry arguments, that His(54) on Cu(A) is 'insipiently axial' and that a combination of a butterfly distortion of the [Cu(2)O(2)](2+) group and a rotation of the Cu(A)(His)(3) moiety converts the vacant, initially axial, binding site on Cu(A) into a much more favourable equatorial site.
More Related Videos
 05:57
Synthesizing Amino Acids Modified with Reactive Carbonyls in Silico to Assess Structural Effects Using Molecular Dynamics Simulations
Published on: April 26, 2024
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 01:48
ATP Synthase: Mechanism
 01:13
Introduction to Mechanisms of Enzyme Catalysis
01:13Introduction to Mechanisms of Enzyme Catalysis
 01:18
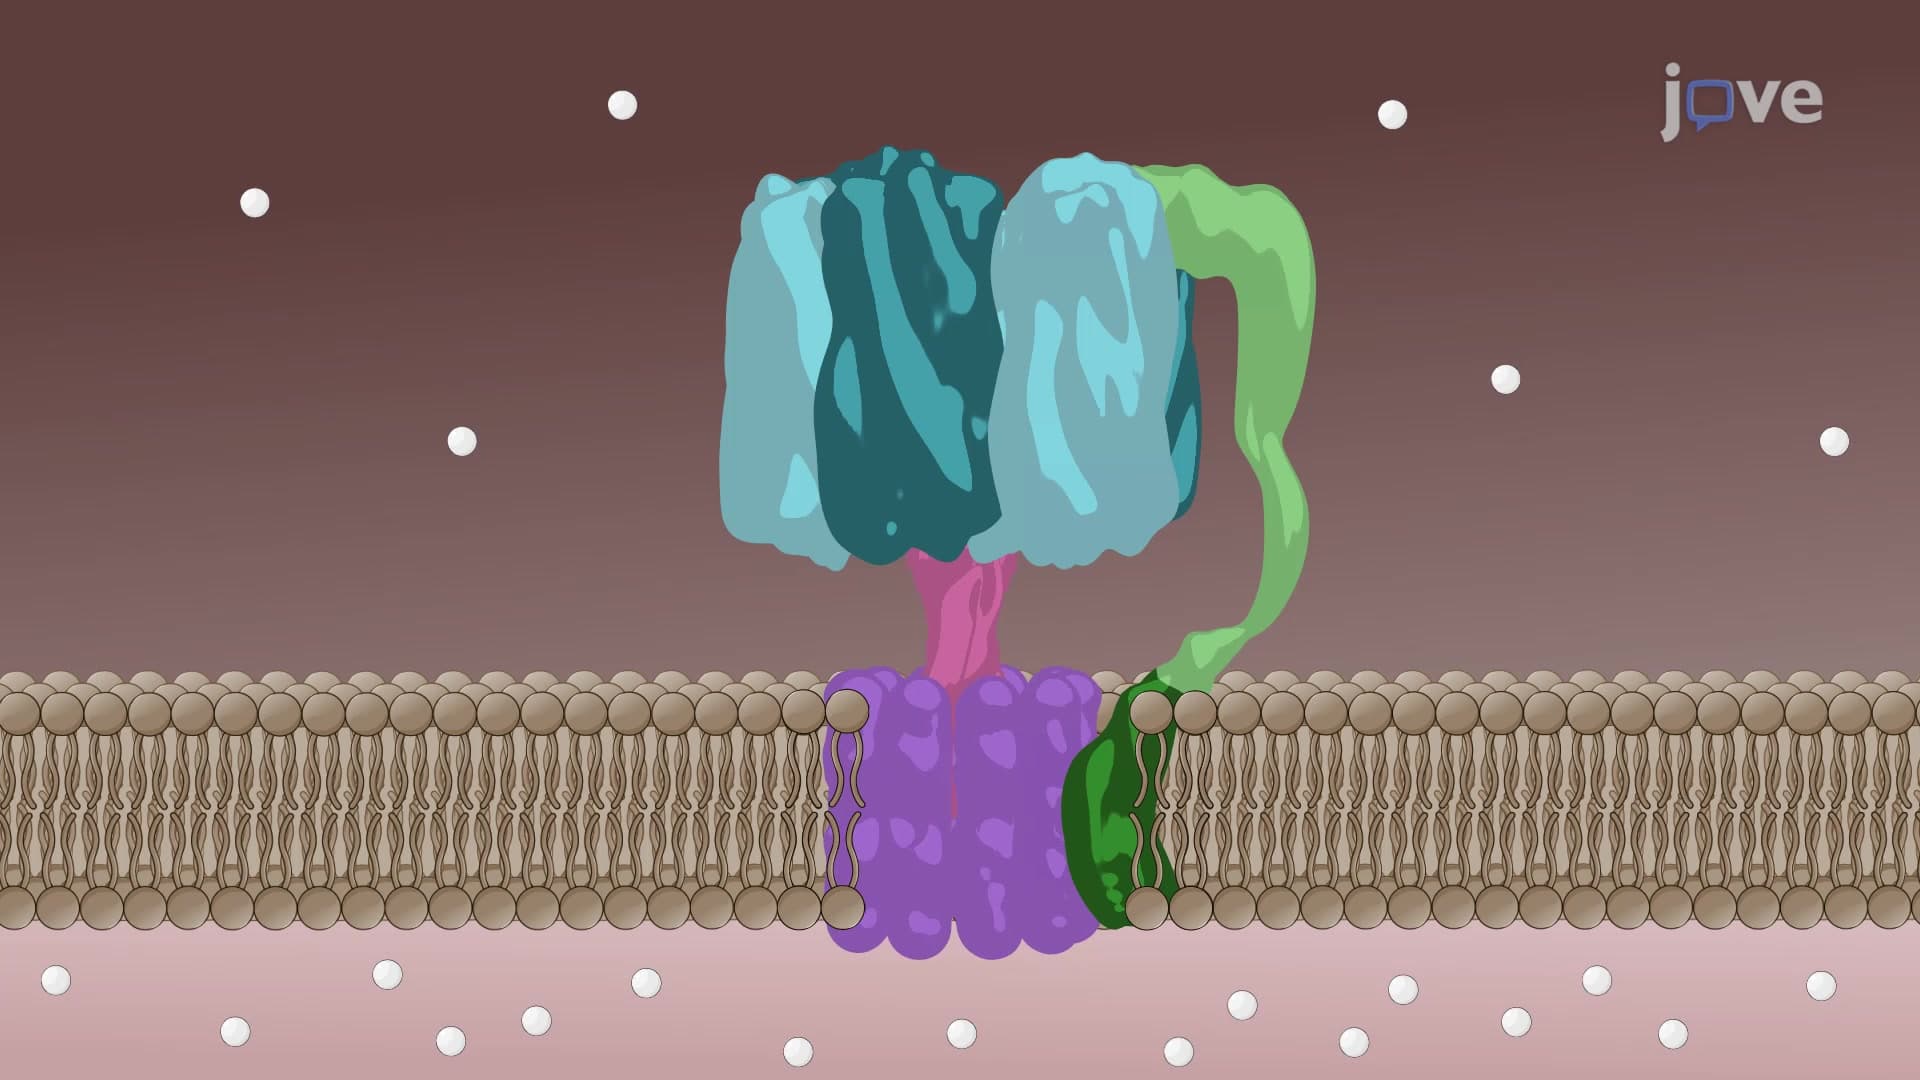
ATP Synthase: Structure
 02:15
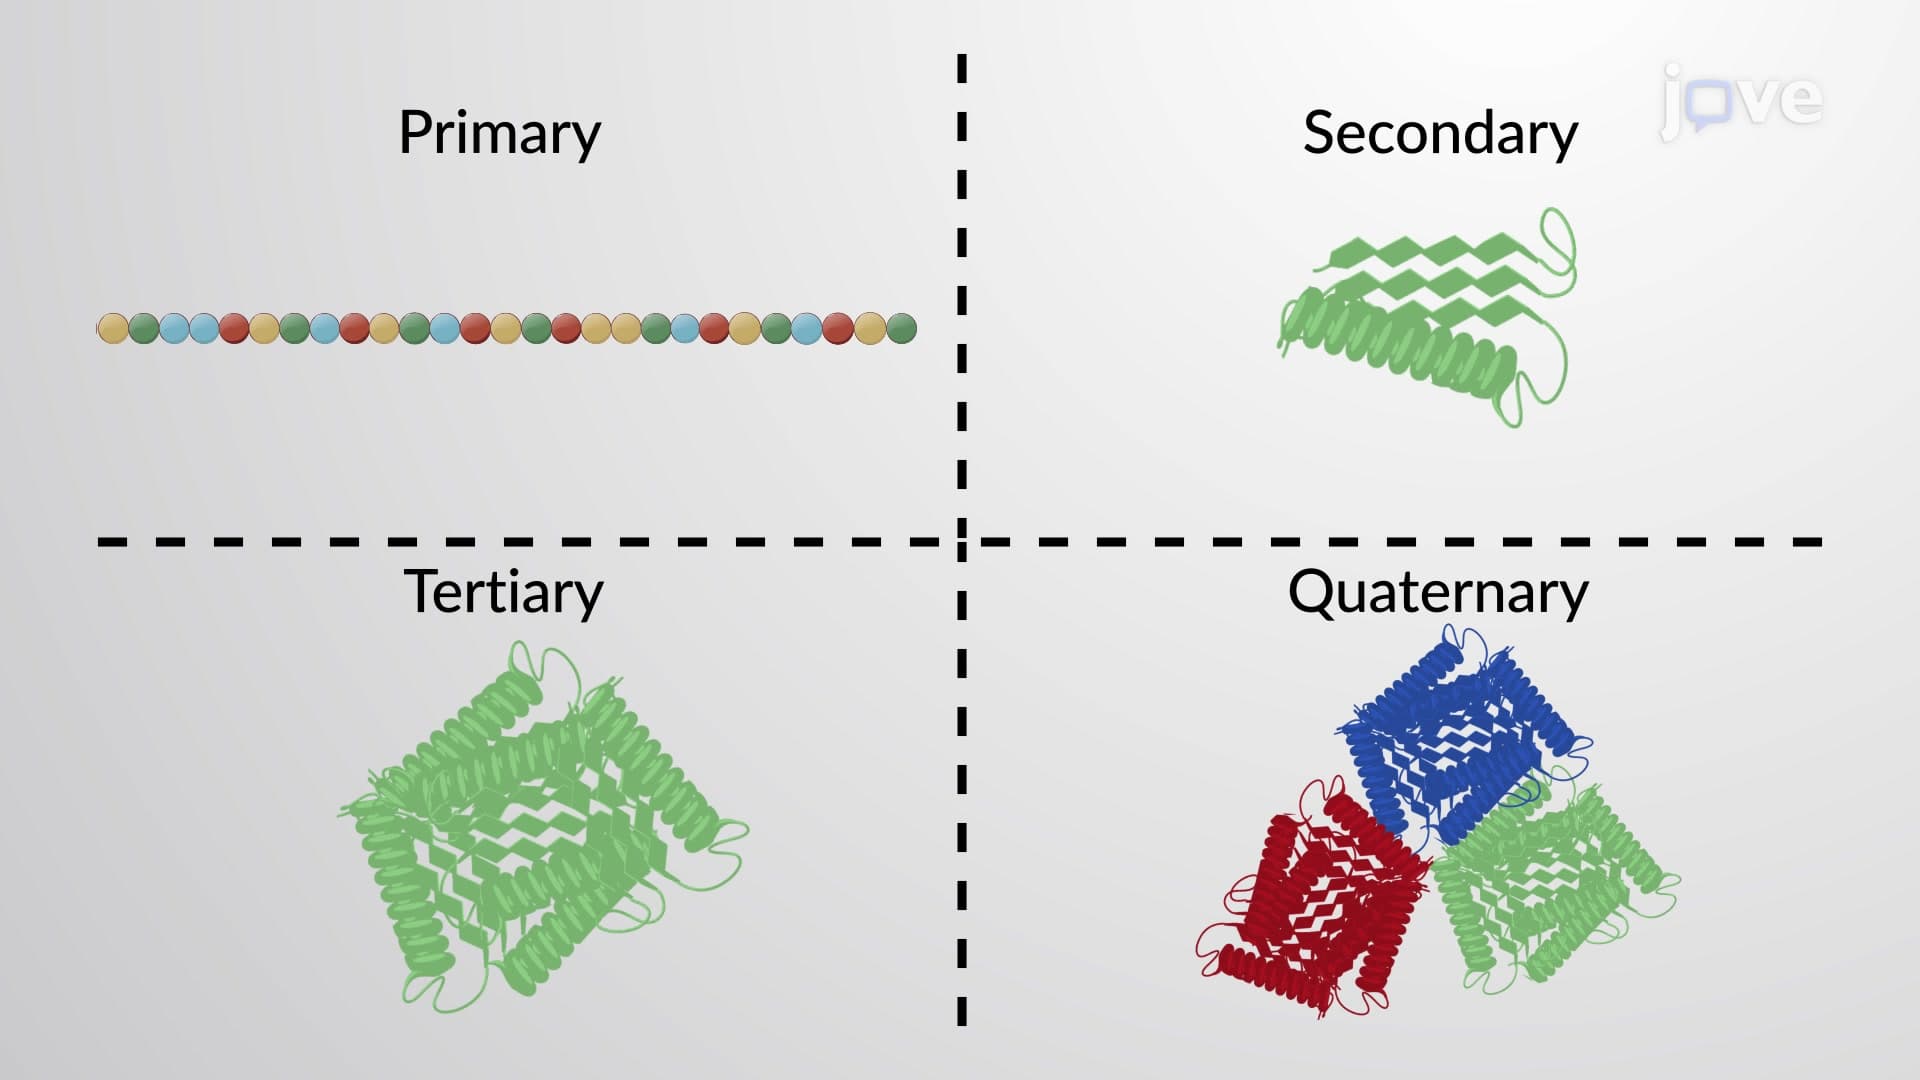
Protein and Protein Structure
A protein's shape is critical to its function. For example, an enzyme can...
 02:21
Globular and Fibrous Proteins
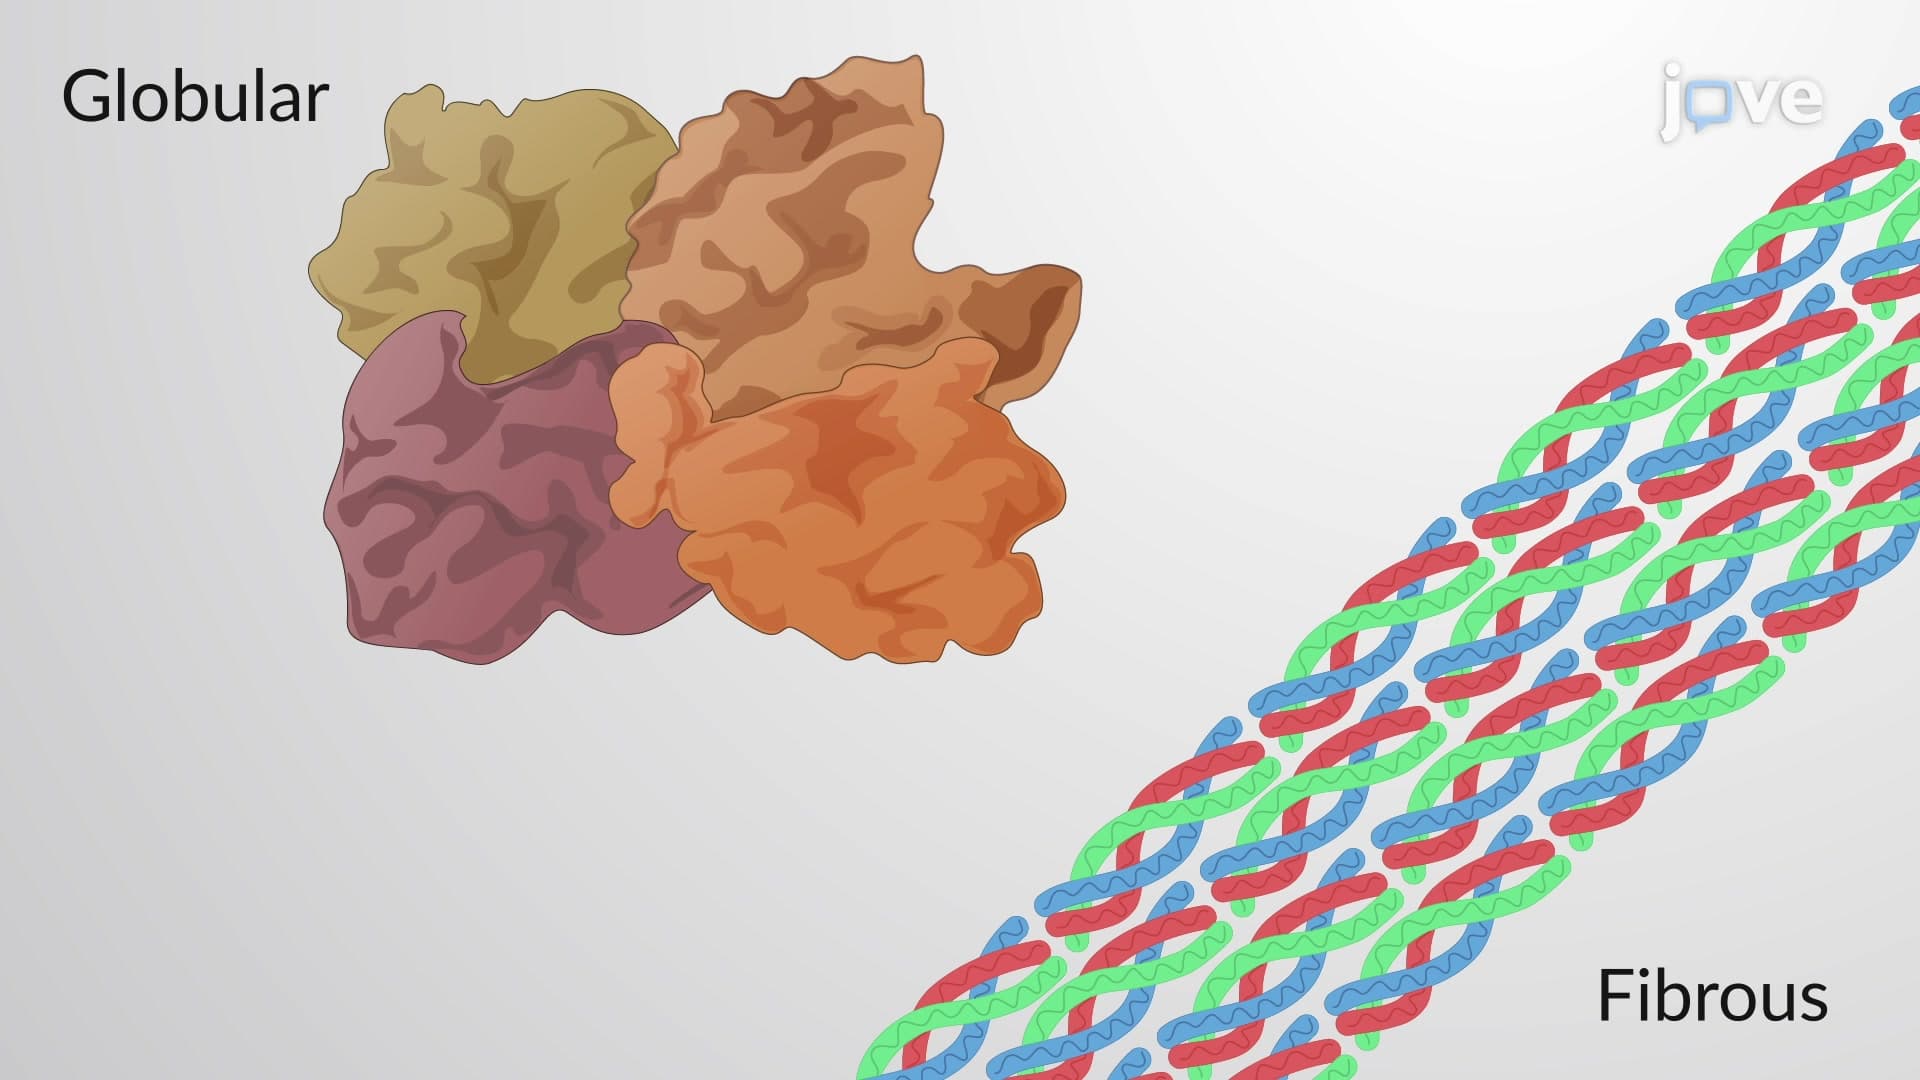
Globular proteins are also known as spheroproteins and typically are approximately round in shape. They contain a mix of amino acid types and contain differing sequences in their primary structures. Globular proteins have many different functions, such as enzymes, cellular messengers, and molecular transporters. These roles often require the proteins to be...