Related Experiment Video
Updated: Jun 20, 2026

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
On the variational computation of a large number of vibrational energy levels and wave functions for medium-sized
Edit Mátyus1, Ján Simunek, Attila G Császár
1Laboratory of Molecular Spectroscopy, Institute of Chemistry, Eötvös University, P.O. Box 32, H-1518 Budapest 112, Hungary.

Abstract:
In a recent publication [J. Chem. Phys. 127, 084102 (2007)], the nearly variational DEWE approach (DEWE denotes Discrete variable representation of the Watson Hamiltonian using the Eckart frame and an Exact inclusion of a potential energy surface expressed in arbitrarily chosen coordinates) was developed to compute a large number of (ro)vibrational eigenpairs for medium-sized semirigid molecules having a single well-defined minimum. In this publication, memory, CPU, and hard disk usage requirements of DEWE, and thus of any DEWE-type approach, are carefully considered, analyzed, and optimized. Particular attention is paid to the sparse matrix-vector multiplication, the most expensive part of the computation, and to rate-determining steps in the iterative Lanczos eigensolver, including spectral transformation, reorthogonalization, and restart of the iteration. Algorithmic improvements are discussed in considerable detail. Numerical results are presented for the vibrational band origins of the (12)CH(4) and (12)CH(2)D(2) isotopologues of the methane molecule. The largest matrix handled on a personal computer during these computations is of the size of (4x10(8))x(4x10(8)). The best strategy for determining vibrational eigenpairs depends largely on the actual details of the required computation. Nevertheless, for a usual scenario requiring a large number of the lowest eigenpairs of the Hamiltonian matrix the combination of the thick-restart Lanczos method, shift-fold filtering, and periodic reorthogonalization appears to result in the computationally most feasible approach.
More Related Videos
 06:37
Analyzing Melts and Fluids from Ab Initio Molecular Dynamics Simulations with the UMD Package
Published on: September 17, 2021
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to the...
 01:26
The Van der Waals Equation
 04:01
Real Gases: Effects of Intermolecular Forces and Molecular Volume Deriving Van der Waals Equation
 01:10
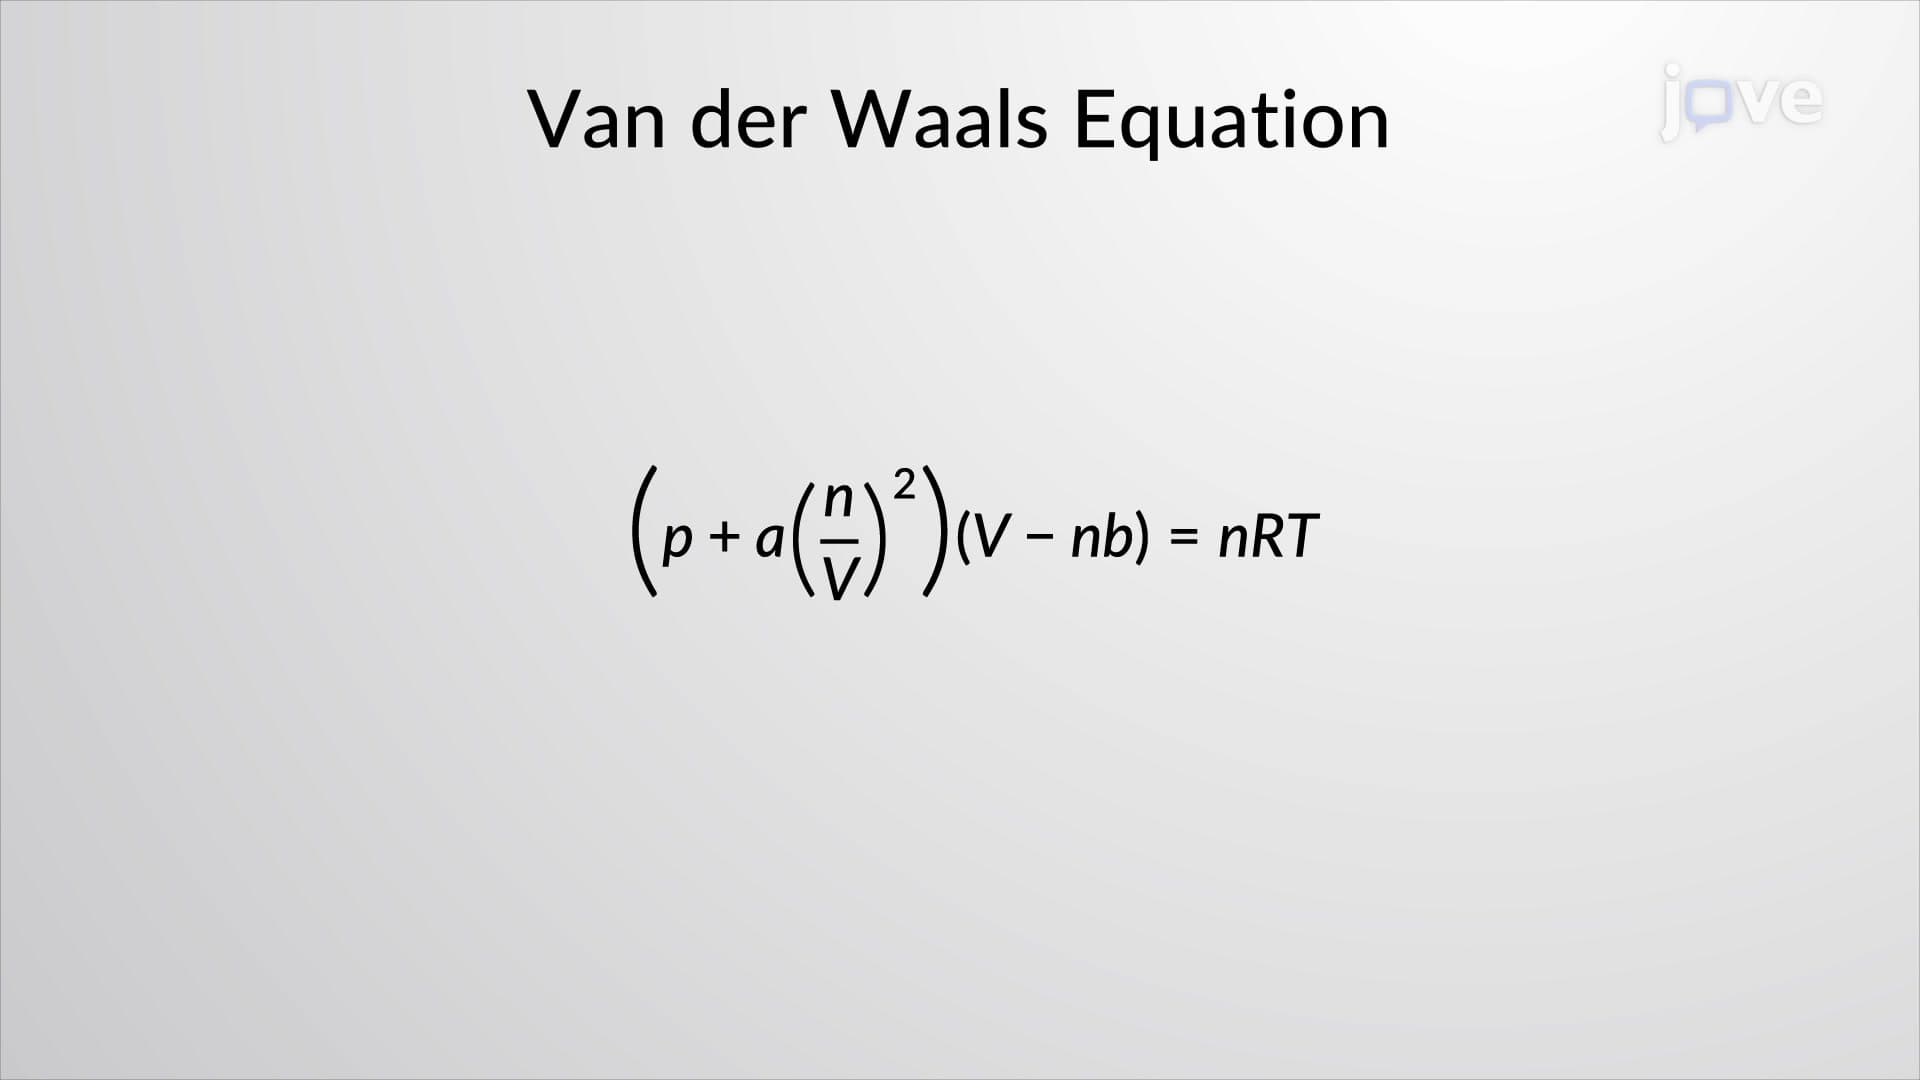
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 01:16
UV–Vis Spectroscopy: Molecular Electronic Transitions
 01:14
Molecular Spectroscopy: Absorption and Emission