Related Experiment Video
Updated: Jun 20, 2026

Isolating Free Carbenes, their Mixed Dimers and Organic Radicals
Published on: April 19, 2019
The contributions of through-bond interactions to the singlet-triplet energy difference in 1,3-dehydrobenzene
Haiyan Wei1, David A Hrovat, Yirong Mo
1Department of Chemistry and Center for Advanced Scientific Computing and Modeling, University of North Texas, 1155 Union Circle, #305070, Denton, Texas 76203-5070, USA.

Abstract:
Separation of the through-space (TS) from the through-bond (TB) interactions between the two atomic orbitals at C(1) and C(3) of 1,3-dehydrobenzene (1) has been achieved by carrying out ab initio, valence-bond, self-consistent-field (VBSCF) calculations. The results indicate that, at the CCSD(T)/cc-pVTZ optimized geometry of the singlet state of 1, the stabilization provided by TB interactions contributes 10% more than the stabilization provided by the TS interactions to the adiabatic singlet-triplet energy difference. The highest occupied MO of 1 contains a contribution from a hybrid AO at C(2), which has the same phase as the smaller lobes of the AOs at C(1) and C(3). Consequently, TB interactions in 1 increase with decreasing values of the C(1)-C(3) distance. The origin of this hybrid AO at C(2) and the contributions of hyperconjugation to TB and TS interactions in 1 are described and discussed.
Related Concept Videos
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 01:37
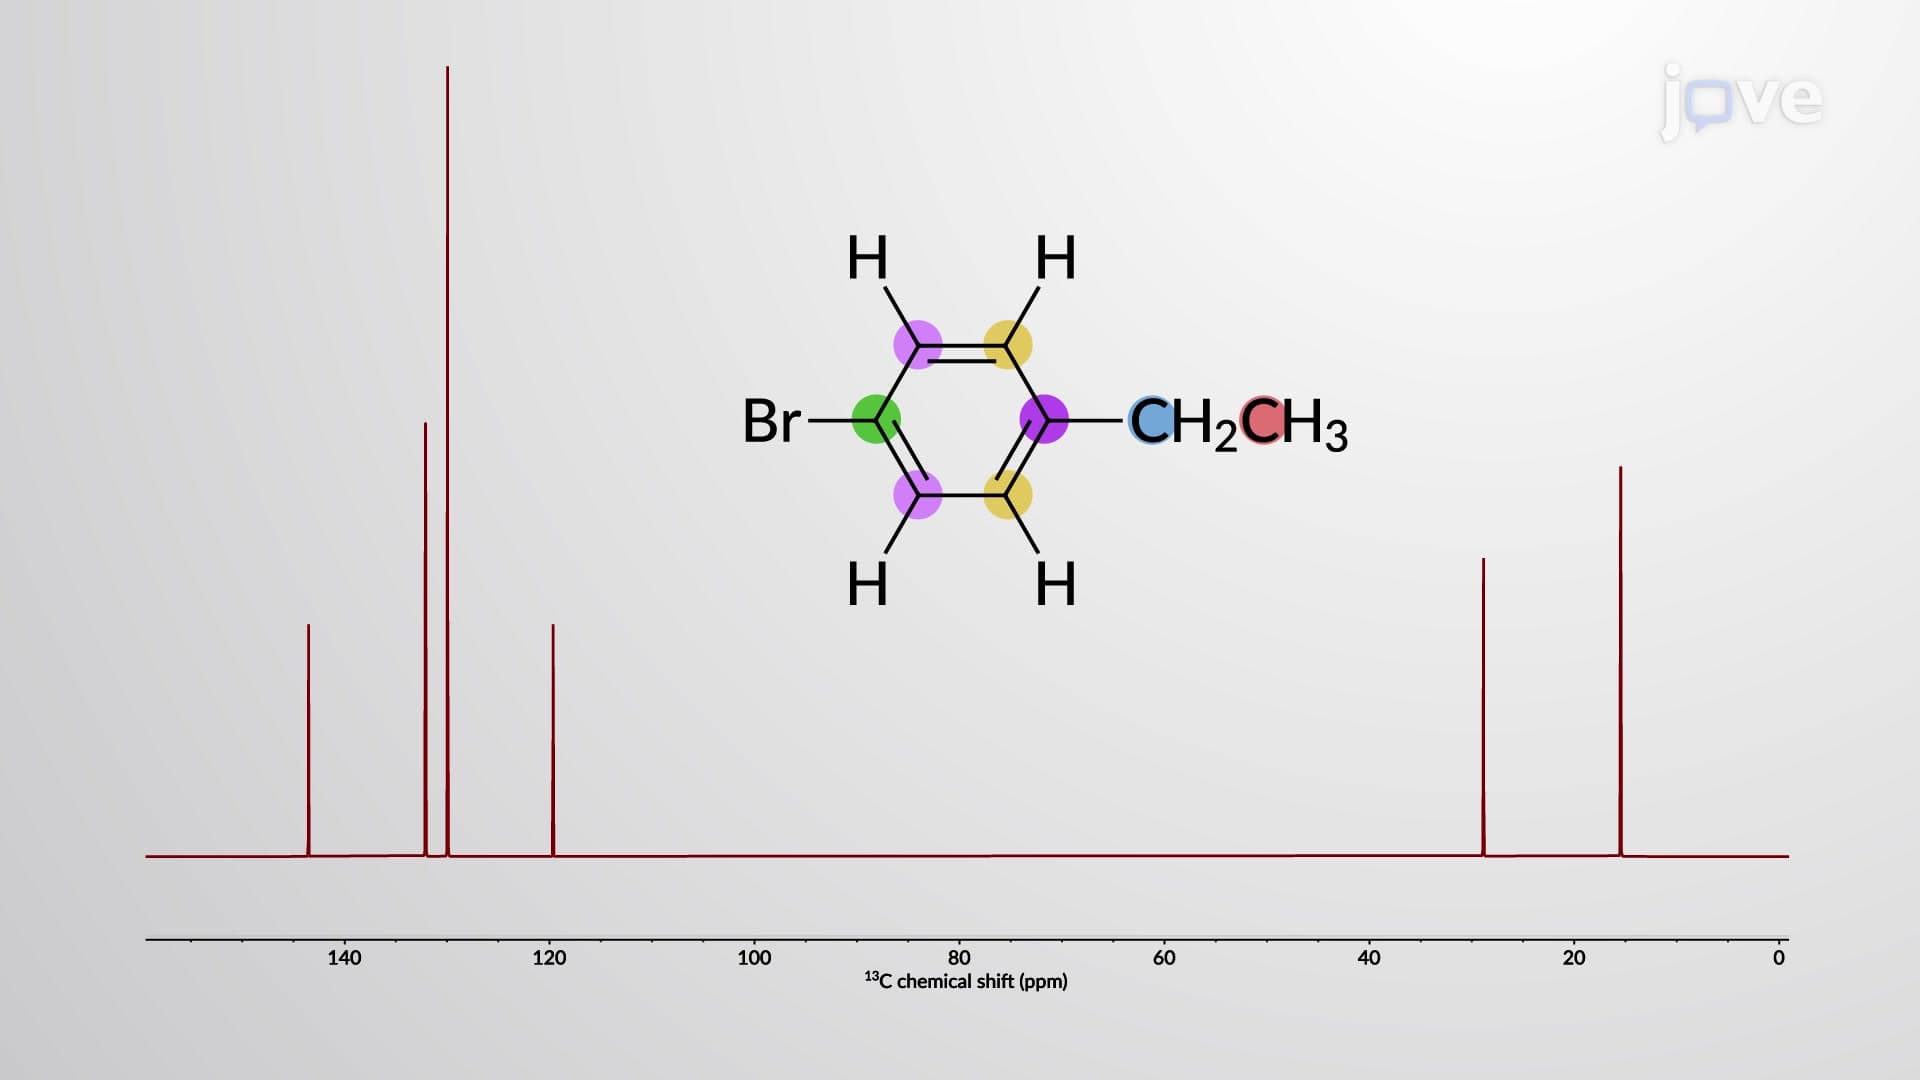
NMR Spectroscopy of Benzene Derivatives
 01:18
Structure of Benzene: Molecular Orbital Model
 03:51
Molecular Orbital Theory II
 01:17
Spin–Spin Coupling: One-Bond Coupling
 03:35
Hybridization of Atomic Orbitals II